
加速された金属酸化物の熱化学予測のための機械学習
Nickolas A Joyner1, Sarah Sprouse1, Haizley Herndon1
1Department of Chemistry and Biochemistry, The University of Alabama, Shelby Hall, Tuscaloosa, Alabama 35487-0336, United States.
Journal of chemical theory and computation
|February 20, 2026

まとめ
機械学習モデルは,金属酸化物のエネルギーを正確に予測し,大量結合エネルギーの計算を迅速に可能にします. このフレームワークは,標準化されたクラスタリングエネルギー (NCE) を使用して,計算コストを削減してDFTレベルの精度を達成します.
科学分野:
- コンピューティング・マテリアル・サイエンス・サイエンス
- 化学における機械学習
- 固体物理 固体物理学
背景:
- 凝固エネルギーなどの物質の性質を正確に予測することは,新しい材料を発見するために非常に重要です.
- 密度関数理論 (DFT) などの伝統的な電子構造計算は,計算的に高価です.
- 機械学習 (ML) は,これらの予測を加速するための潜在的な経路を提供します.
研究 の 目的:
- 金属 (II) オキシドの正常化されたクラスタリングエネルギー (NCE) を予測するための機械学習の枠組みを開発する.
- NCEの予測を抽象化して,凝固した質量エネルギーを決定する.
- DFT計算と比較した異なるMLモデルの正確性と効率性を評価する.
主な方法:
- DFTで最適化されたM(II) O構造 (アルカリ土と3D移行金属) のデータセットをまとめました.
- 構造は,スムーズ・オーバーラップ・オブ・アトム・ポジション (SOAP) ディスクリプターを使用してコード化されました.
- カーネルリッジ回帰 (KRR),人工ニューラルネットワーク (ANN),ツリーベースのモデルが訓練され,評価されました.
主要な成果:
- KRRは,NCE (MAE = 0.619 kcal/mol,R2 = 0.994) の予測誤差が最も低い値を達成し,次に ANN.
- いくつかの酸化物に対する予測された大量結合エネルギーは,DFT値の~1.5 kcal/mol以内でした.
- 多様な構造を組み込むことで,モデルの移転性が向上し,正確な熱化学予測のためのMLの潜在能力を実証しました.
結論:
- 控えめなクラスターデータで訓練された機械学習モデルは,NCEと大量熱化学を正確に予測できます.
- このMLアプローチは,従来のDFT計算と比較して,計算コストを大幅に削減します.
- この研究は,ML主導の材料特性予測のための複雑な電子構造のエンコーディングの有効性を強調しています.
さらに関連する動画
関連する概念動画
 02:24
Predicting Reaction Outcomes
11.1K
Kinetics describes the rate and path by which a reaction occurs. In contrast, thermodynamics deals with state functions and describes the properties, behavior, and components of a system. It is not concerned with the path taken by the process and cannot address the rate at which a reaction occurs. Although it does provide information about what can happen during a reaction process, it does not describe the detailed steps of what appears on an atomic or a molecular level. On the other hand,...
11.1K
 02:55
Thermochemical Equations
36.2K
For a chemical reaction (the system) carried out at constant pressure – with the only work done caused by expansion or contraction – the enthalpy of reaction (also called the heat of reaction, ΔHrxn) is equal to the heat exchanged with the surroundings (qp).
36.2K
 01:16
Thermal Sigmatropic Reactions: Overview
2.6K
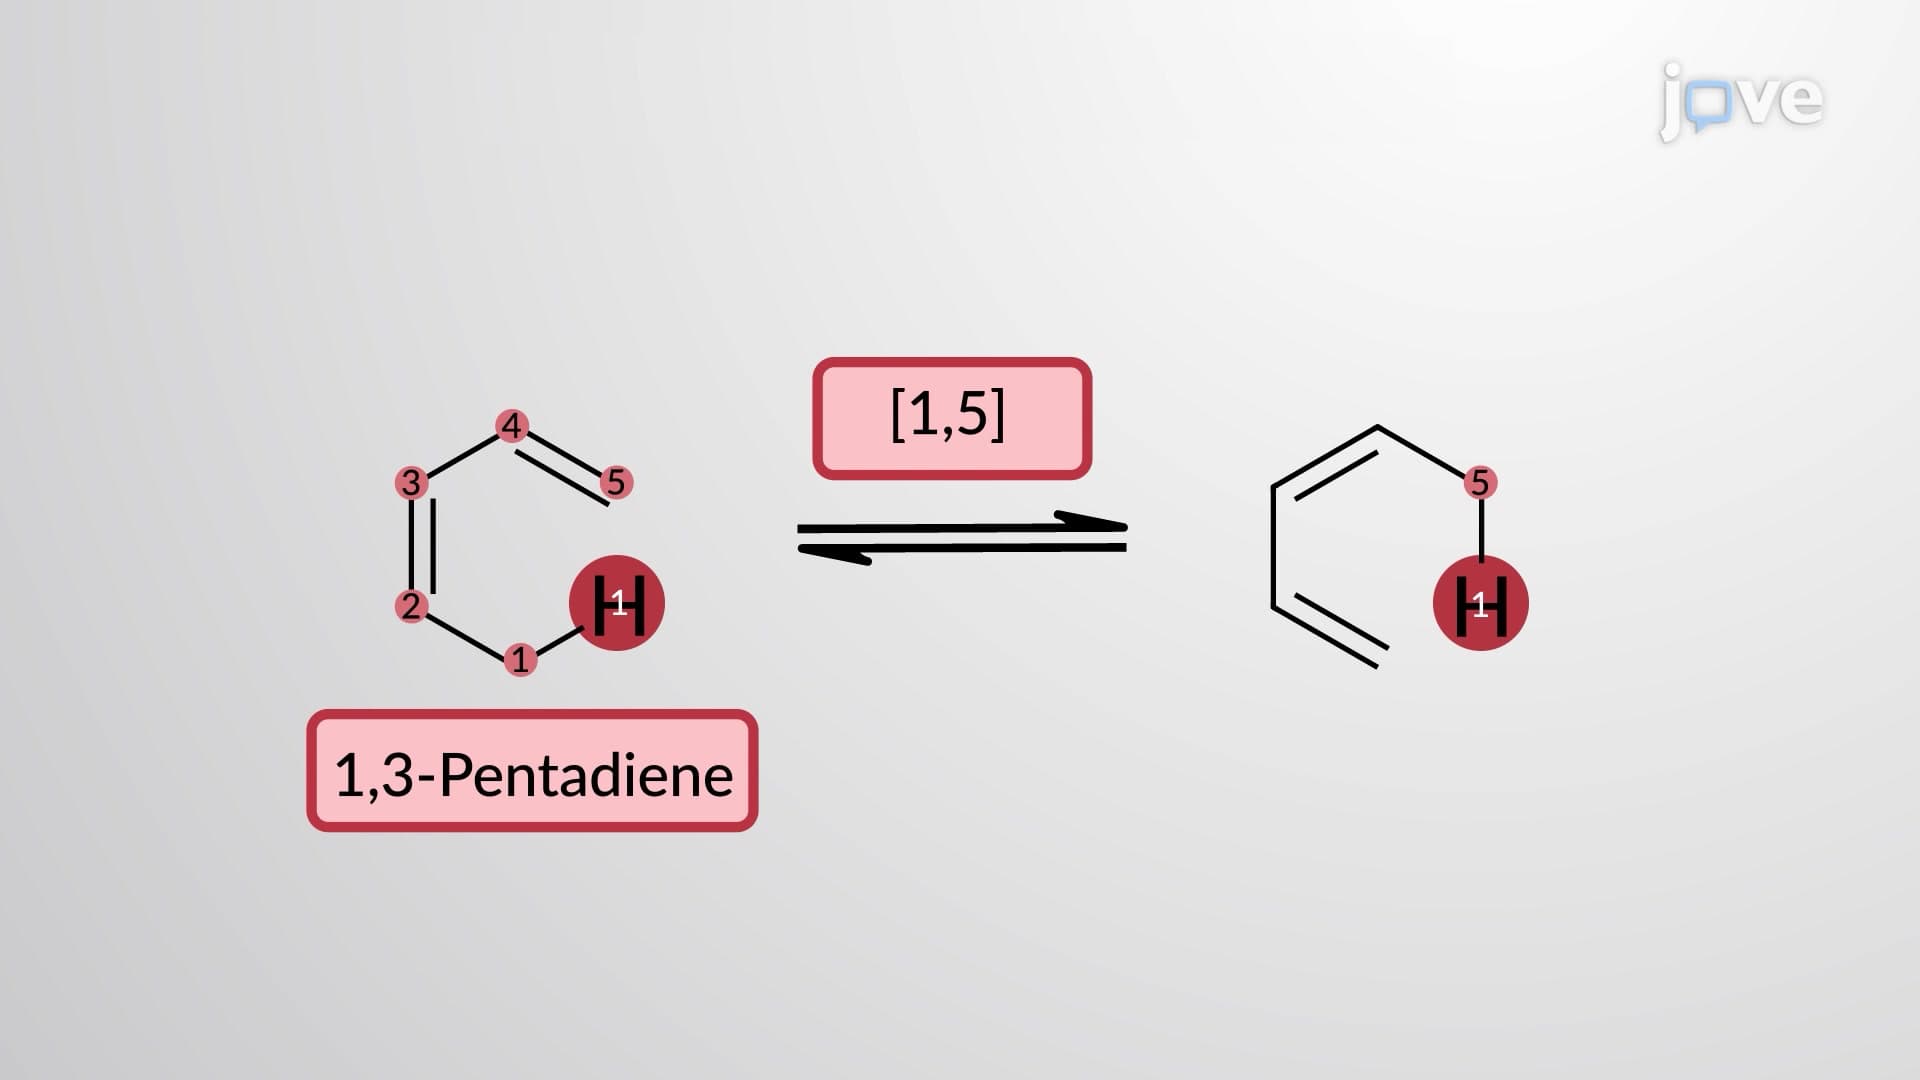
Sigmatropic rearrangements are a class of pericyclic reactions in which a σ bond migrates from one part of a π system to another. These are intramolecular rearrangements where the total number of σ and π bonds remain unchanged.
Sigmatropic shifts are classified based on an order term [i, j ], where i and j indicate the number of atoms across which each end of the σ bond migrates. Below are examples of a [3,3] sigmatropic shift in 1,5-hexadiene, referred...
Sigmatropic shifts are classified based on an order term [i, j ], where i and j indicate the number of atoms across which each end of the σ bond migrates. Below are examples of a [3,3] sigmatropic shift in 1,5-hexadiene, referred...
2.6K