非常长链乙基-CoA脱酶缺乏的分子异质性导致儿科心肌病和突然死亡
A Mathur1, H F Sims, D Gopalakrishnan
1Departments of Pediatrics, Medicine and Molecular Biology and Pharmacology, Washington University School of Medicine, St Louis Children's Hospital, St Louis, MO, USA.
Circulation
|March 17, 1999

概括
非常长链乙基-CoA脱酶 (VLCAD) 的遗传缺陷导致婴儿心肌病. 虽然往往是致命的,VLCAD缺乏和心脏问题是可逆的治疗.
科学领域:
- 遗传学 是一个遗传学.
- 代谢障碍 代谢障碍 代谢障碍
- 心脏病学 心脏病学
背景情况:
- 初级心肌病 (CM) 越来越多地与遗传缺陷有关.
- 非常长链乙基-CoA脱酶 (VLCAD) 通过脂肪酸β-氧化对心脏能量产生至关重要.
研究的目的:
- 研究VLCAD基因突变在患有心肌病和脂肪酸氧化障碍特征的患者中的作用.
主要方法:
- 使用单链形态差异分析对基因组DNA进行选.
- 进行了VLCAD基因的DNA测序和突变分析.
- 使用RNA斑点分析和VLCAD酶检测.
主要成果:
- 在18名患者中确定了21种不同的VLCAD基因突变;80%与CM有关.
- 严重的婴儿CM (67%) 和肝功能障碍 (33%) 是常见的表现.
- 框架转移或拼接部位突变导致VLCAD mRNA和酶活性降低.
结论:
- 婴儿心肌病是VLCAD缺乏的主要临床表现.
- VLCAD基因突变是多种多样的.
- 与VLCAD缺乏相关的代谢障碍和心肌病症都可以逆转,尽管最初的死亡率很高.

相关概念视频
 01:26
Cross-bridge Cycle
As muscle contracts, the overlap between the thin and thick filaments increases, decreasing the length of the sarcomere—the contractile unit of the muscle—using energy in the form of ATP. At the molecular level, this is a cyclic, multistep process that involves binding and hydrolysis of ATP, and movement of actin by myosin.
 01:46
Electron Transport Chain: Complex I and II
The mitochondrial electron transport chain (ETC) is the main energy generation system in the eukaryotic cells. However, mitochondria also produce cytotoxic reactive oxygen species (ROS) due to the large electron flow during oxidative phosphorylation. While Complex I is one of the primary sources of superoxide radicals, ROS production by Complex II is uncommon and may only be observed in cancer cells with mutated complexes.
ROS generation is regulated and maintained at moderate levels necessary...
ROS generation is regulated and maintained at moderate levels necessary...
 01:48
ATP Synthase: Mechanism
In animals, the mitochondrial F1F0 ATP synthase is the key protein that synthesizes ATP molecules through a complex catalytic mechanism. While the nuclear genome encodes the majority of ATP synthase subunits, the mitochondrial genome encodes some of the enzyme's most critical components. The formation of this multi-subunit enzyme is a complex multi-step process regulated at the level of transcription, translation, and assembly. Defects in one or more of these steps can result in decreased ATP...
 01:18
ATP Synthase: Structure
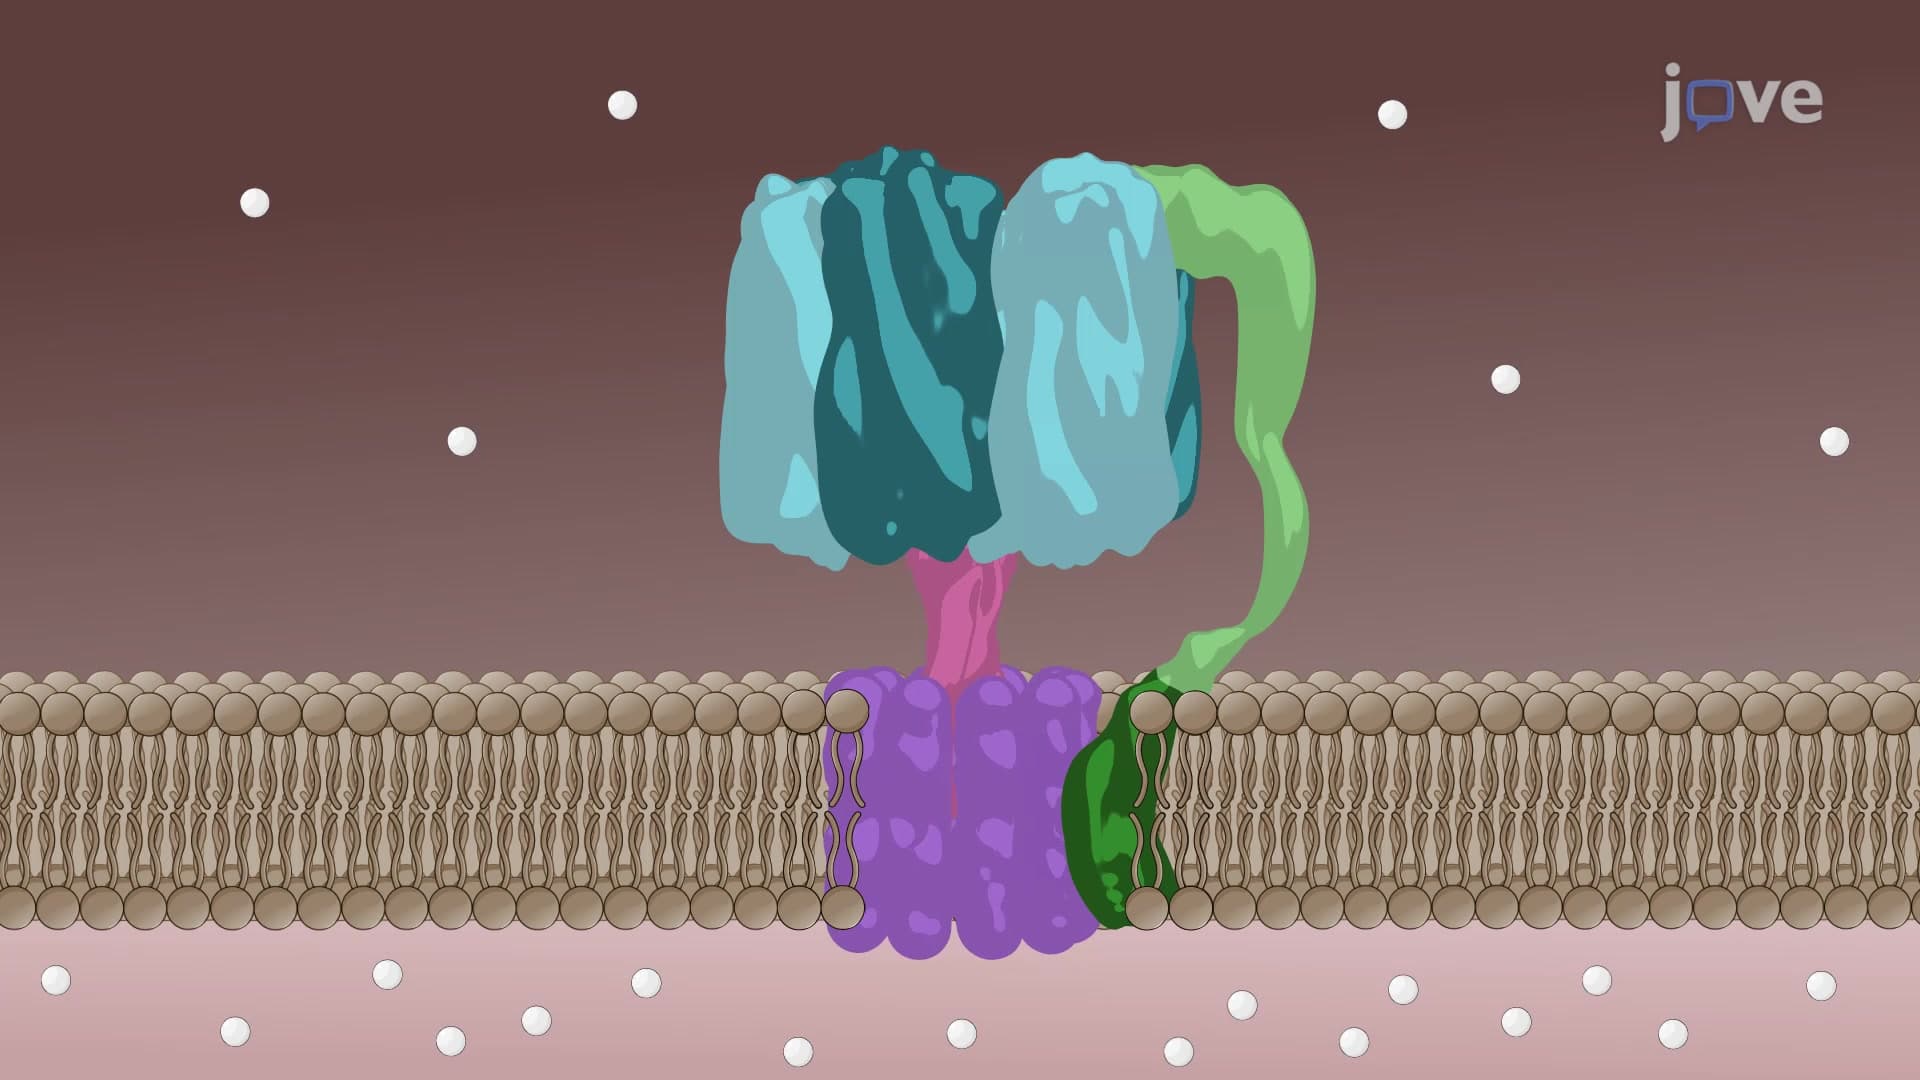
ATP synthase or ATPase is among the most conserved proteins found in bacteria, mammals, and plants. This enzyme can catalyze a forward reaction in response to the electrochemical gradient, producing ATP from ADP and inorganic phosphate. ATP synthase can also work in a reverse direction by hydrolyzing ATP and generating an electrochemical gradient. Different forms of ATP synthases have evolved special features to meet the specific demands of the cell. Based on their specific feature, ATP...
 01:20
Inborn Errors of Metabolism
Phenylketonuria (PKU) is a protein metabolism disorder characterized by high blood levels of the amino acid phenylalanine. This results from a mutation in the gene responsible for phenylalanine hydroxylase, an enzyme that converts phenylalanine into tyrosine. When this enzyme is deficient, phenylalanine builds up in the blood, leading to symptoms such as vomiting, rashes, seizures, growth deficiency, and severe mental retardation. An early diagnosis and a diet restricting phenylalanine intake...
 01:25
Cardiomyopathy I: Introduction and Classification
Cardiomyopathy, or CMP, is a group of diseases affecting the myocardial structure, impairing its ability to pump blood effectively. This condition can lead to arrhythmias, heart failure, or sudden cardiac death.Cardiomyopathies are classified into primary and secondary categories:Primary Cardiomyopathy refers to conditions involving only the heart muscle that are often idiopathic (of unknown cause) or genetic. They primarily affect the myocardium without the involvement of other systemic...