相关实验视频
Updated: Jul 15, 2026
10:05

In Vitro Analysis of PDZ-dependent CFTR Macromolecular Signaling Complexes
Published on: August 13, 2012
与囊性纤维化相关的突变中异常的CFTR依赖的HCO3-运输
J Y Choi1, D Muallem, K Kiselyov
1Department of Physiology, The University of Texas Southwestern Medical Center, Dallas 75390, USA.
Nature
|March 10, 2001

概括
囊性纤维化 (CF) 由囊性纤维化跨膜导电调节器 (CFTR) 的突变引起. 损害CFTR二碳酸盐 (HCO3-) 运输,而不仅仅是化物 (Cl-) 通道功能,对于CF病变发生至关重要.
科学领域:
- 生物化学 生物化学
- 细胞生物学 细胞生物学
- 遗传学 是一个遗传学.
背景情况:
- 囊性纤维化 (CF) 是一种遗传性疾病,由囊性纤维化跨膜导电性调节器 (CFTR) 基因的突变引起.
- CFTR的主要已知的功能是作为循环AMP调节的化物 (Cl-) 通道,但一些CF引起的突变不会损害该通道的活动.
- 此外,CFTR还调节其他载体,特别是二碳酸盐 (HCO3-) 运输,这对于维持液体pH和分泌表皮质粘度至关重要.
研究的目的:
- 研究CFTR依赖的二碳酸盐 (HCO3-) 运输在囊性纤维化 (CF) 中的作用.
- 为了检查不同的CFTR突变,特别是那些与胰腺功能不充分或充分相关的突变,如何影响HCO3-运输.
- 阐明HCO3-运输在分泌表皮质功能中的重要性及其对CF病理生理学的贡献.
主要方法:
- 对具有保留或正常Cl-通道活性的CFTR突变的分析.
- 在各种CFTR突变体中评估Cl(-) 合的HCO3-运输能力.
- HCO3-运输功能与临床CF表型的相关性,特别是胰腺功能不足和充分性.
主要成果:
- 与胰腺功能不充分相关的CFTR突变表明完全缺乏HCO3-运输.
- 与胰腺充分性相关的CFTR突变显著减少了HCO3-运输.
- 这些发现强调,受损的HCO3-运输是CF的关键特征,无论Cl-通道功能如何保持.
结论:
- 通过CFTR介导的二碳酸盐 (HCO3-) 运输对于分泌表皮的正常功能至关重要.
- CFTR突变体对HCO3-运输的缺陷对囊性纤维化 (CF) 的疾病机制有显著的贡献.
- 向或理解HCO3-运输可能为CF患者提供新的治疗途径,特别是那些具有特定突变类型的患者.
相关概念视频
 01:15
Mechanisms of Membrane-bending
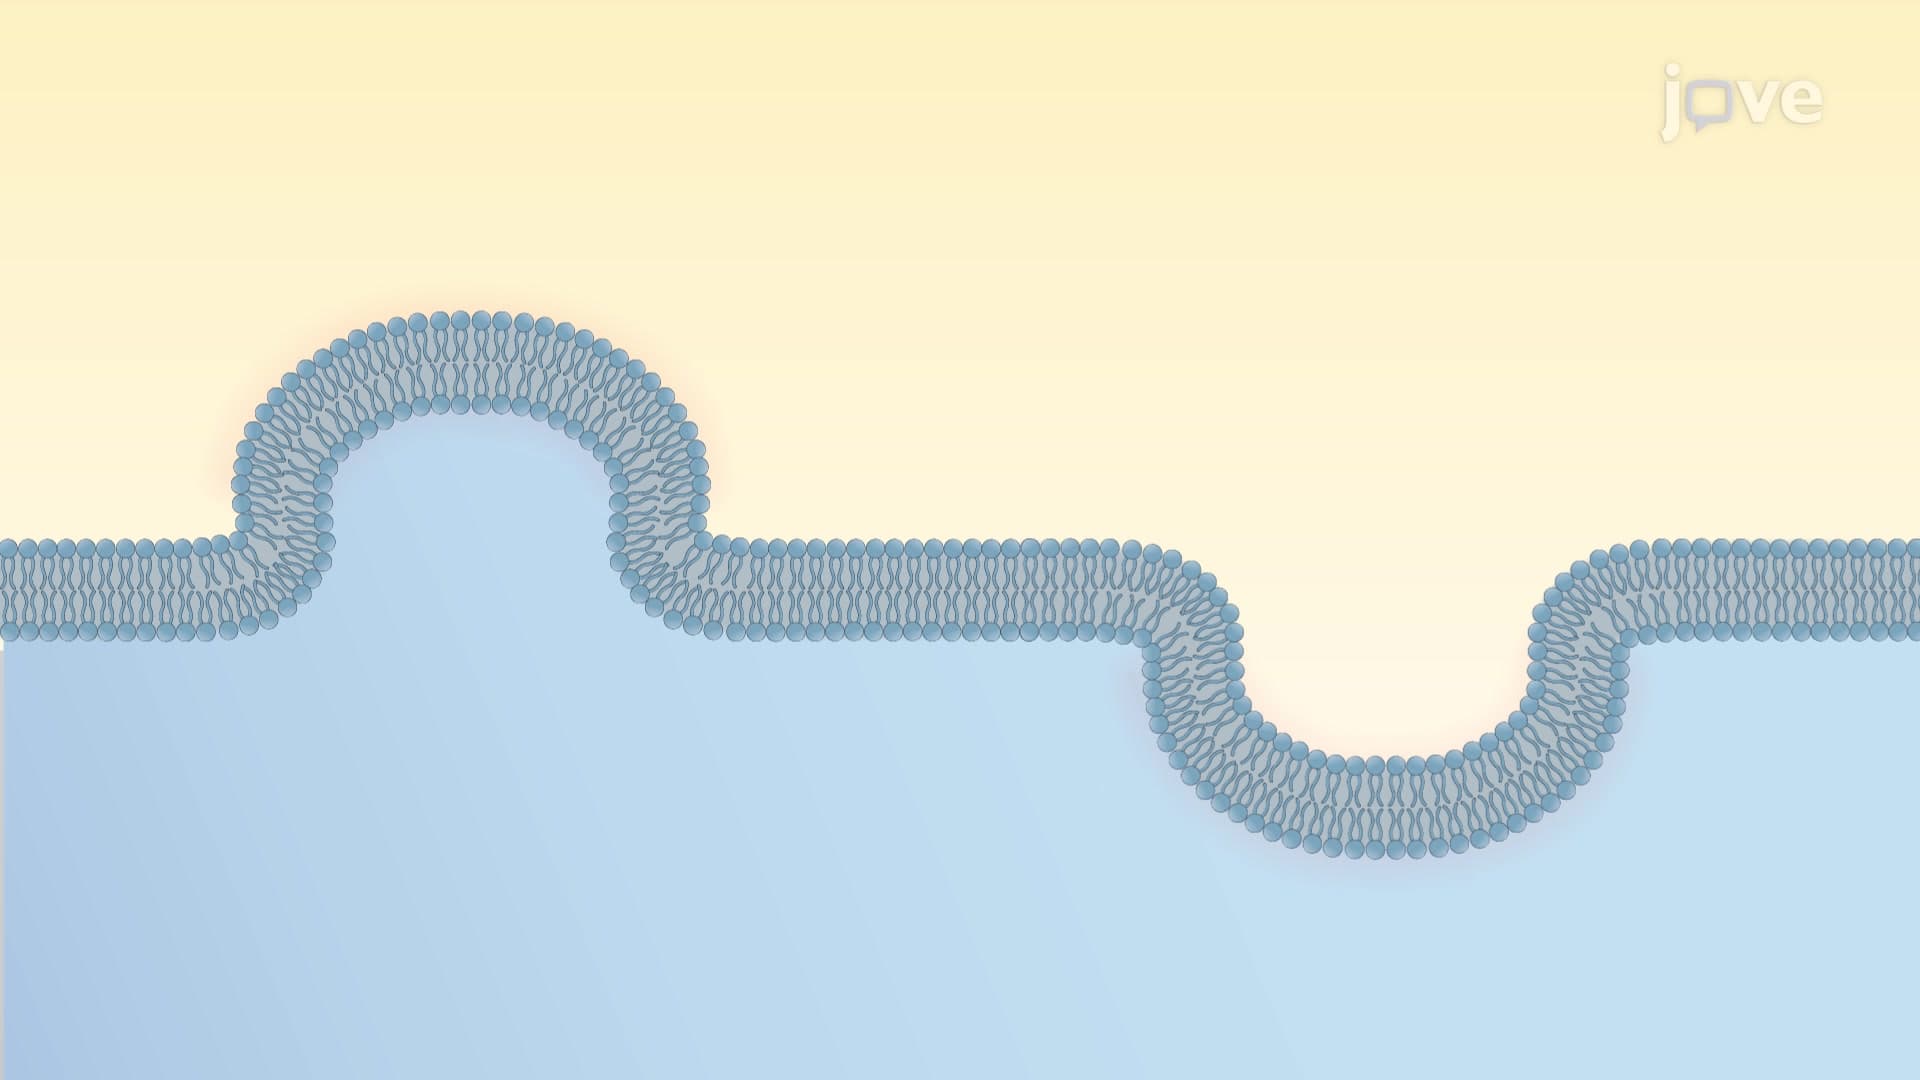
The living membranes are flexible due to their fluid mosaic nature; however, their bending into different shapes is an active process regulated by specific lipids and proteins. The membrane bending can be transient as seen in vesicles or stable for a long time as in microvilli. Cells regulate the size, location, and duration of the membrane curvature.
Membrane bending can happen due to intrinsic changes in lipid composition or extrinsic association with different proteins. The proteins involved...
Membrane bending can happen due to intrinsic changes in lipid composition or extrinsic association with different proteins. The proteins involved...
 01:23
Transcellular Transport of Solutes
Transcellular transport of solutes is the movement of substances like monosaccharides and amino acids through polarized cells. This transport mechanism is primarily seen in epithelial and endothelial cells aided by membrane transport proteins such as channels and transporters. The tight junctions between these cells confine the membrane proteins to the two sides of the cell. The epithelial cells have distinct apical and basolateral domains. In contrast, the endothelial cells show the luminal...
 01:45
Vesicular Tubular Clusters
After budding out from the ER membrane, some COPII vesicles lose their coat and fuse with one another to form larger vesicles and interconnected tubules called vesicular tubular clusters or VTCs. These clusters constitute a compartment at the ER-Golgi interface known as ERGIC (Endoplasmic Reticulum Golgi Intermediate Compartment). The ERGIC is a mobile membrane-bound cargo transport system that sorts proteins secreted from ER and delivers them to the Golgi.
With the help of motor proteins such...
With the help of motor proteins such...
 01:19
Membrane Asymmetry Regulating Transporters
Enzymes like flippase, floppase, and scramblase transfer phospholipids from one layer to another in the membrane, thereby affecting membrane asymmetry.
Flippase
Eukaryotic flippases are type-IV P-type ATPases or P4-ATPases belonging to P-type ATPase family proteins that are membrane-bound pumps involved in the ATP-mediated transport of ions and molecules across the membrane. Flippases flip specific phospholipids from the outer to the inner leaflet of a membrane. All P4-ATPases have one...
Flippase
Eukaryotic flippases are type-IV P-type ATPases or P4-ATPases belonging to P-type ATPase family proteins that are membrane-bound pumps involved in the ATP-mediated transport of ions and molecules across the membrane. Flippases flip specific phospholipids from the outer to the inner leaflet of a membrane. All P4-ATPases have one...
 01:23
Cystic Fibrosis: Pathogenesis
Cystic fibrosis (CF), an autosomal recessive disorder, significantly affects the function of exocrine glands. This genetically inherited disease is characterized by the production of thick and sticky mucus, which can severely affect various organs and systems in the body.
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:24
Cystic Fibrosis: Management
Cystic fibrosis (CF) is an autosomal recessive disorder that predominantly affects individuals of Northern European descent, occurring at a rate of 1 in 3500. It is caused by a genetic mutation in a gene on chromosome 7, most commonly the ΔF508 mutation, that codes for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. This results in thicker mucus secretions and obstruction pathologies in multiple organs, including the lungs and sinuses.
Sinus disease and chronic sinusitis...
Sinus disease and chronic sinusitis...