
(57) 血红蛋白模型系统的Fe Mössbauer异构体转移:电子结构计算
Yong Zhang1, Junhong Mao, Eric Oldfield
1Department of Chemistry, University of Illinois at Urbana-Champaign, 600 South Mathews Avenue, Urbana, IL 61801, USA.
Journal of the American Chemical Society
|June 27, 2002

概括
密度函数理论 (DFT) 的计算准确地预测了铁-57 Mössbauer 异构体在各种化学系统中的移位. 这一进步使无机,有机金属和金属蛋白化合物在各种旋转状态下能够精确地表征.
科学领域:
- 计算化学的计算化学
- 固态化学 固态化学
- 生物有机化学 生物有机化学
背景情况:
- 铁-57 Mössbauer光谱对于表征含铁化合物至关重要.
- 准确预测Mössbauer同位素移位 (delta(Fe)) 对于理解电子结构至关重要.
- 之前的计算方法在预测不同铁系统和旋转状态的delta ((Fe)) 方面存在局限性.
研究的目的:
- 为了报告密度函数理论 (DFT) 对铁-57的计算,Mössbauer 异构体移位 (delta(Fe)).
- 评估DFT方法对广泛的无机,有机金属和金属蛋白/金属氨酸系统的准确性.
- 在各种自旋状态中研究计算和实验德尔塔值之间的相关性.
主要方法:
- 对24个模型系统进行了密度函数理论 (DFT) 计算.
- 计算包括S = 0, 1/2, 1, 3/2, 2, 和 5/2旋转状态.
- 使用混合交换相关函数B3LYP,并与其他函数进行比较.
主要成果:
- 在计算和实验同位素转移之间达成了很好的一致性 (R(2) = 0.973-0.981).
- 平方根均值偏差很小 (0.07-0.08 mm s(-1)),占整个 delta (Fe) 范围的3-4%.
- 对于正常和中间旋转状态,DFT准确地复制了异构体移位,涵盖了从-0.90到1.44毫米秒秒-1的广泛范围.
结论:
- DFT方法,特别是使用B3LYP函数的方法,可以准确预测铁-57同位素的移位.
- 这项研究证明了DFT对各种铁化学环境和旋转状态的可靠性.
- 分子轨道分析揭示了铁核对电荷密度的关键贡献,与同位素移位良好相关.
相关概念视频
 01:18
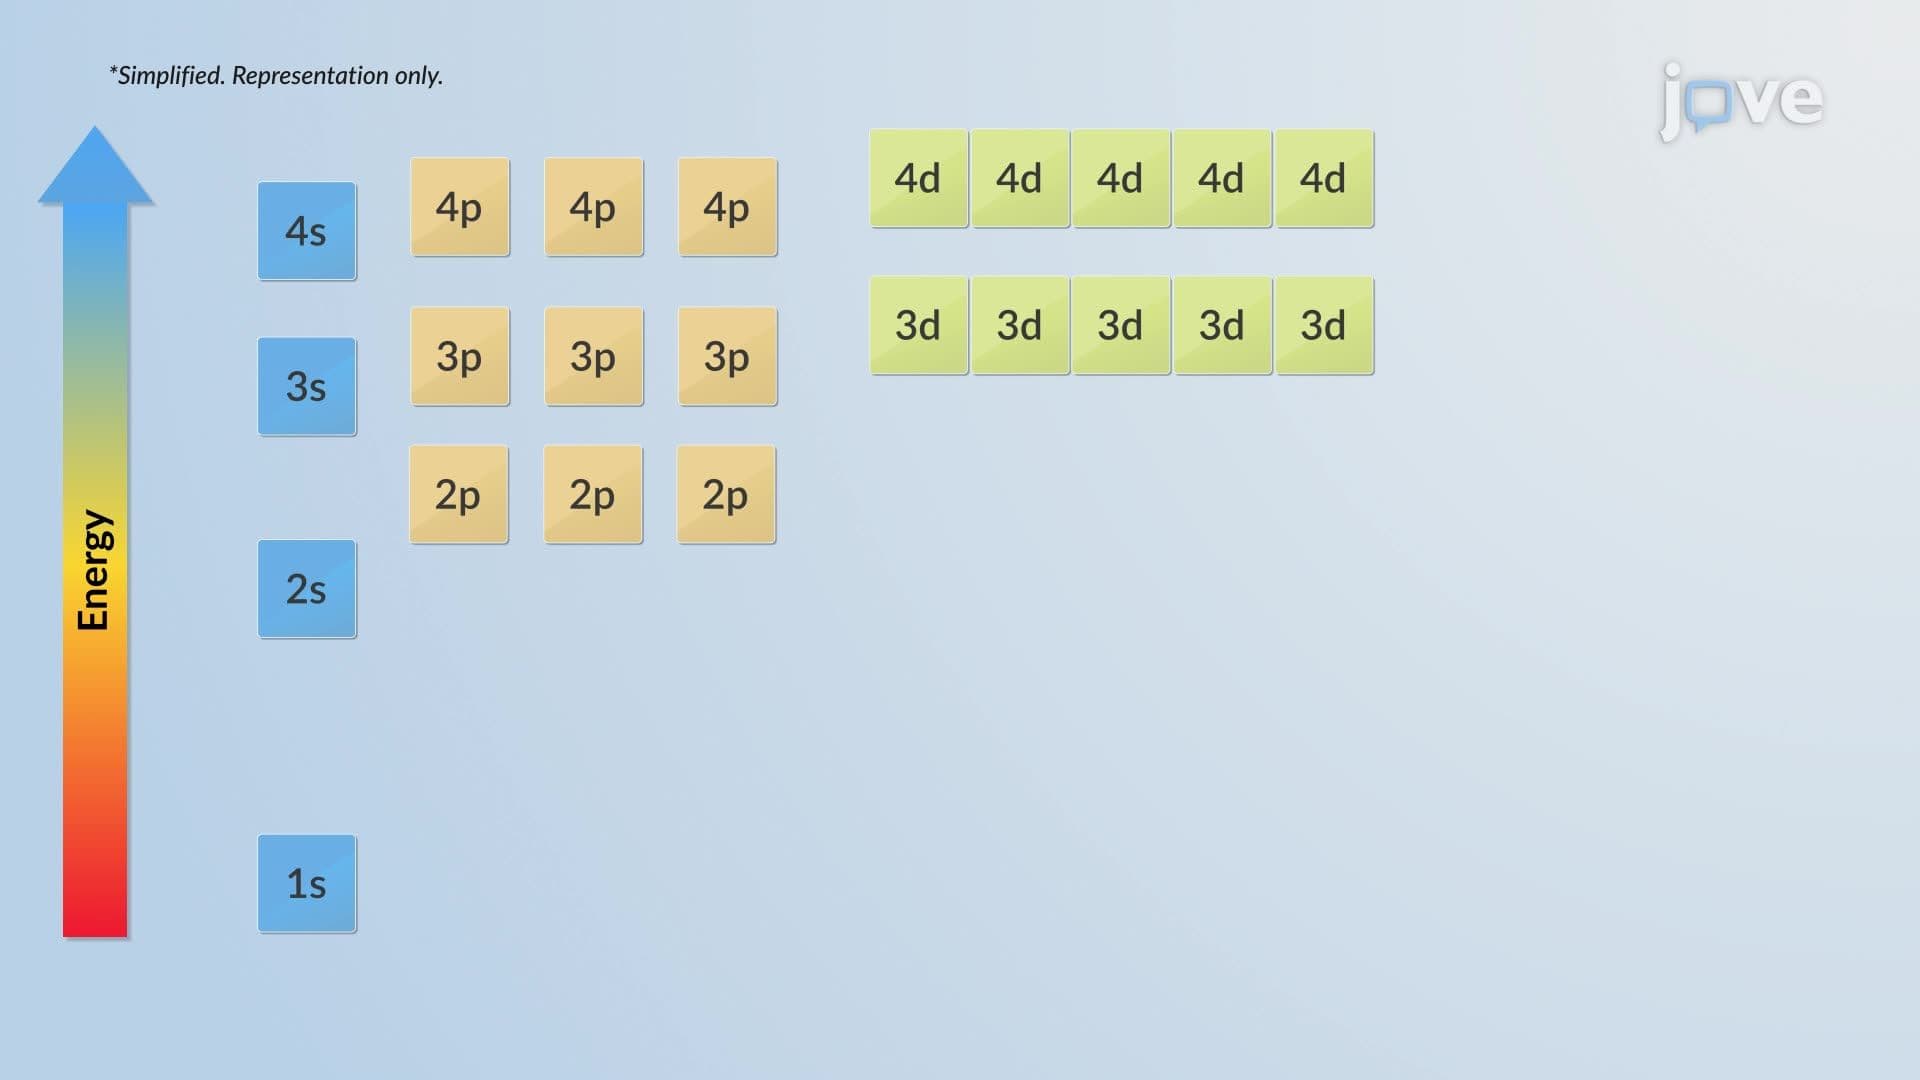
Electron Orbital Model
Orbitals are the areas outside of the atomic nucleus where electrons are most likely to reside. They are characterized by different energy levels, shapes, and three-dimensional orientations. The location of electrons is described most generally by a shell or principal energy level, then by a subshell within each shell, and finally, by individual orbitals found within the subshells.The first shell is closest to the nucleus, and it has only one subshell with a single spherical orbital called the...
 02:18
The Bohr Model
Following the work of Ernest Rutherford and his colleagues in the early twentieth century, the picture of atoms consisting of tiny dense nuclei surrounded by lighter and even tinier electrons continually moving about the nucleus was well established. This picture was called the planetary model since it pictured the atom as a miniature “solar system” with the electrons orbiting the nucleus like planets orbiting the sun. The simplest atom is hydrogen, consisting of a single proton as the nucleus...
 03:26
Electron Configuration of Multielectron Atoms
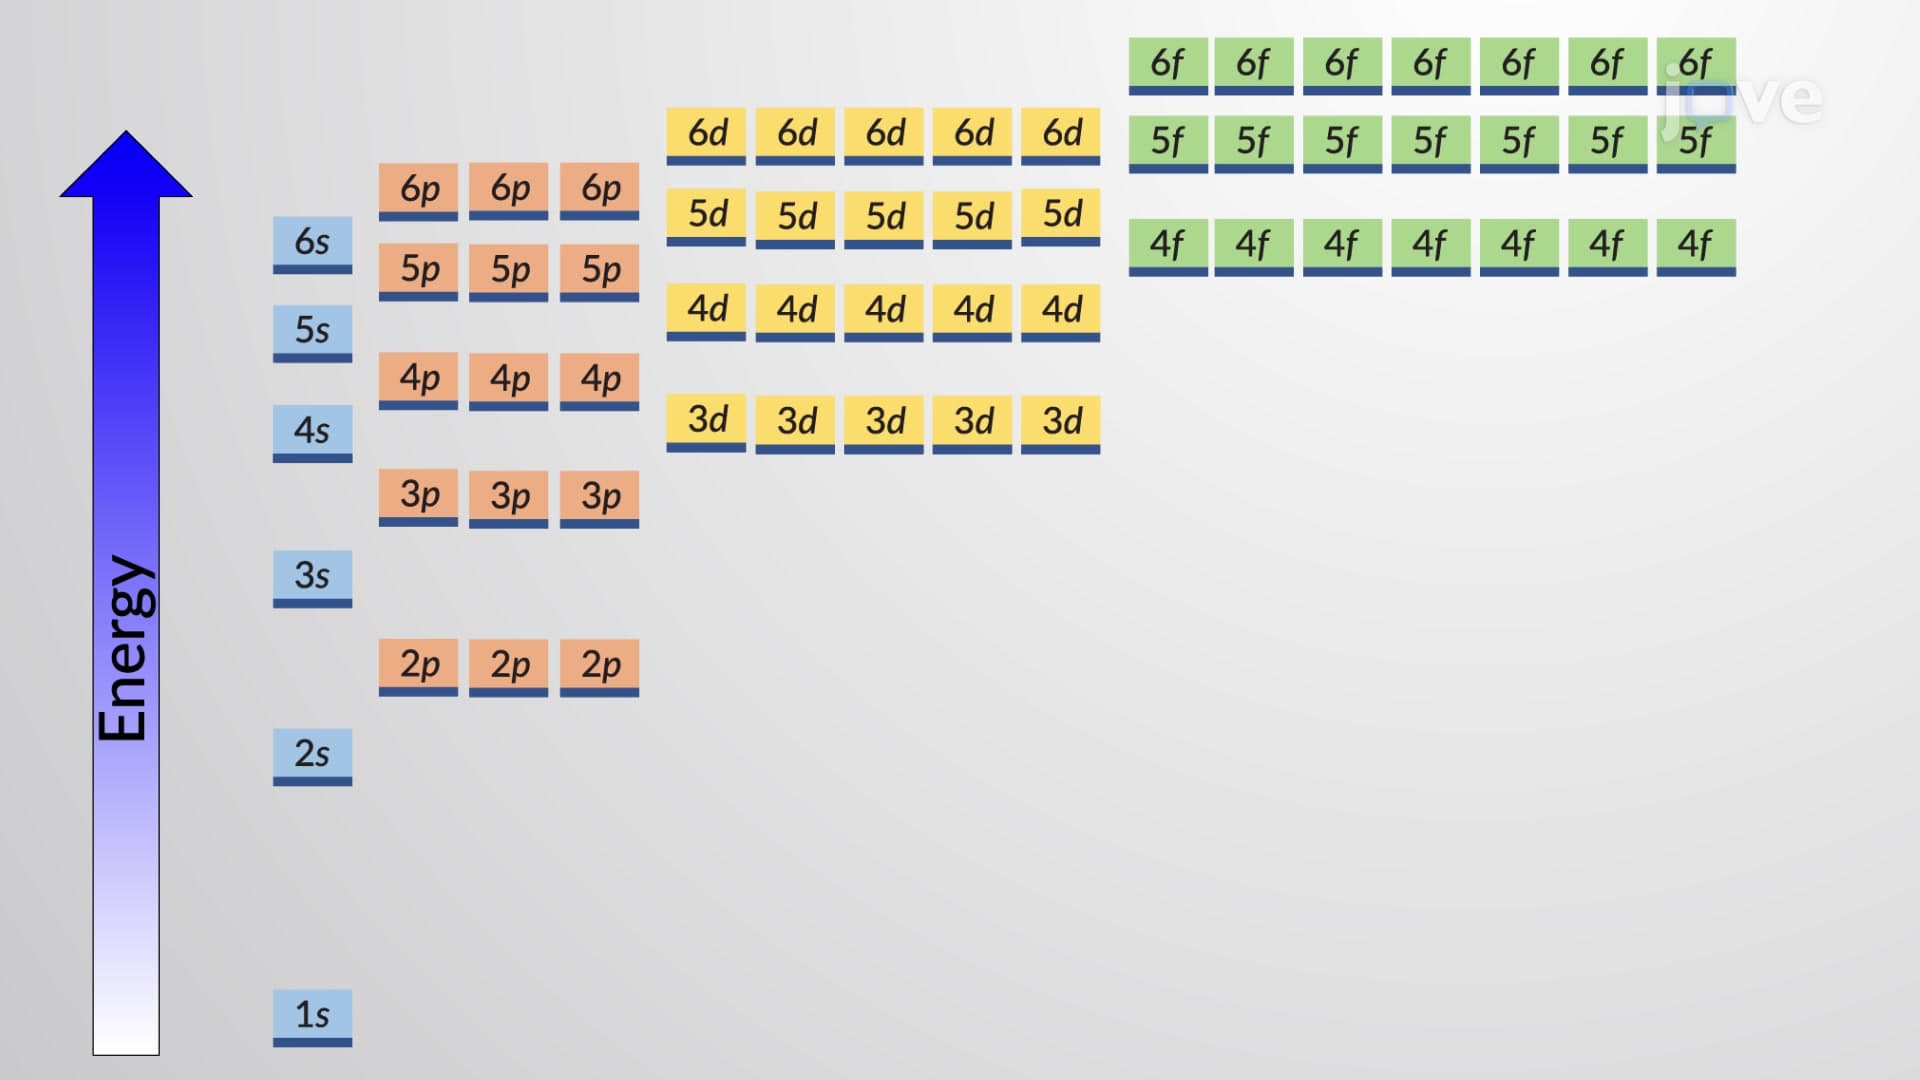
The alkali metal sodium (atomic number 11) has one more electron than the neon atom. This electron must go into the lowest-energy subshell available, the 3s orbital, giving a 1s22s22p63s1 configuration. The electrons occupying the outermost shell orbital(s) (highest value of n) are called valence electrons, and those occupying the inner shell orbitals are called core electrons. Since the core electron shells correspond to noble gas electron configurations, we can abbreviate electron...
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum numbers: n, l, ml, and...
 01:08
Hückel's Rule Diagram of π MOs: Frost Circle
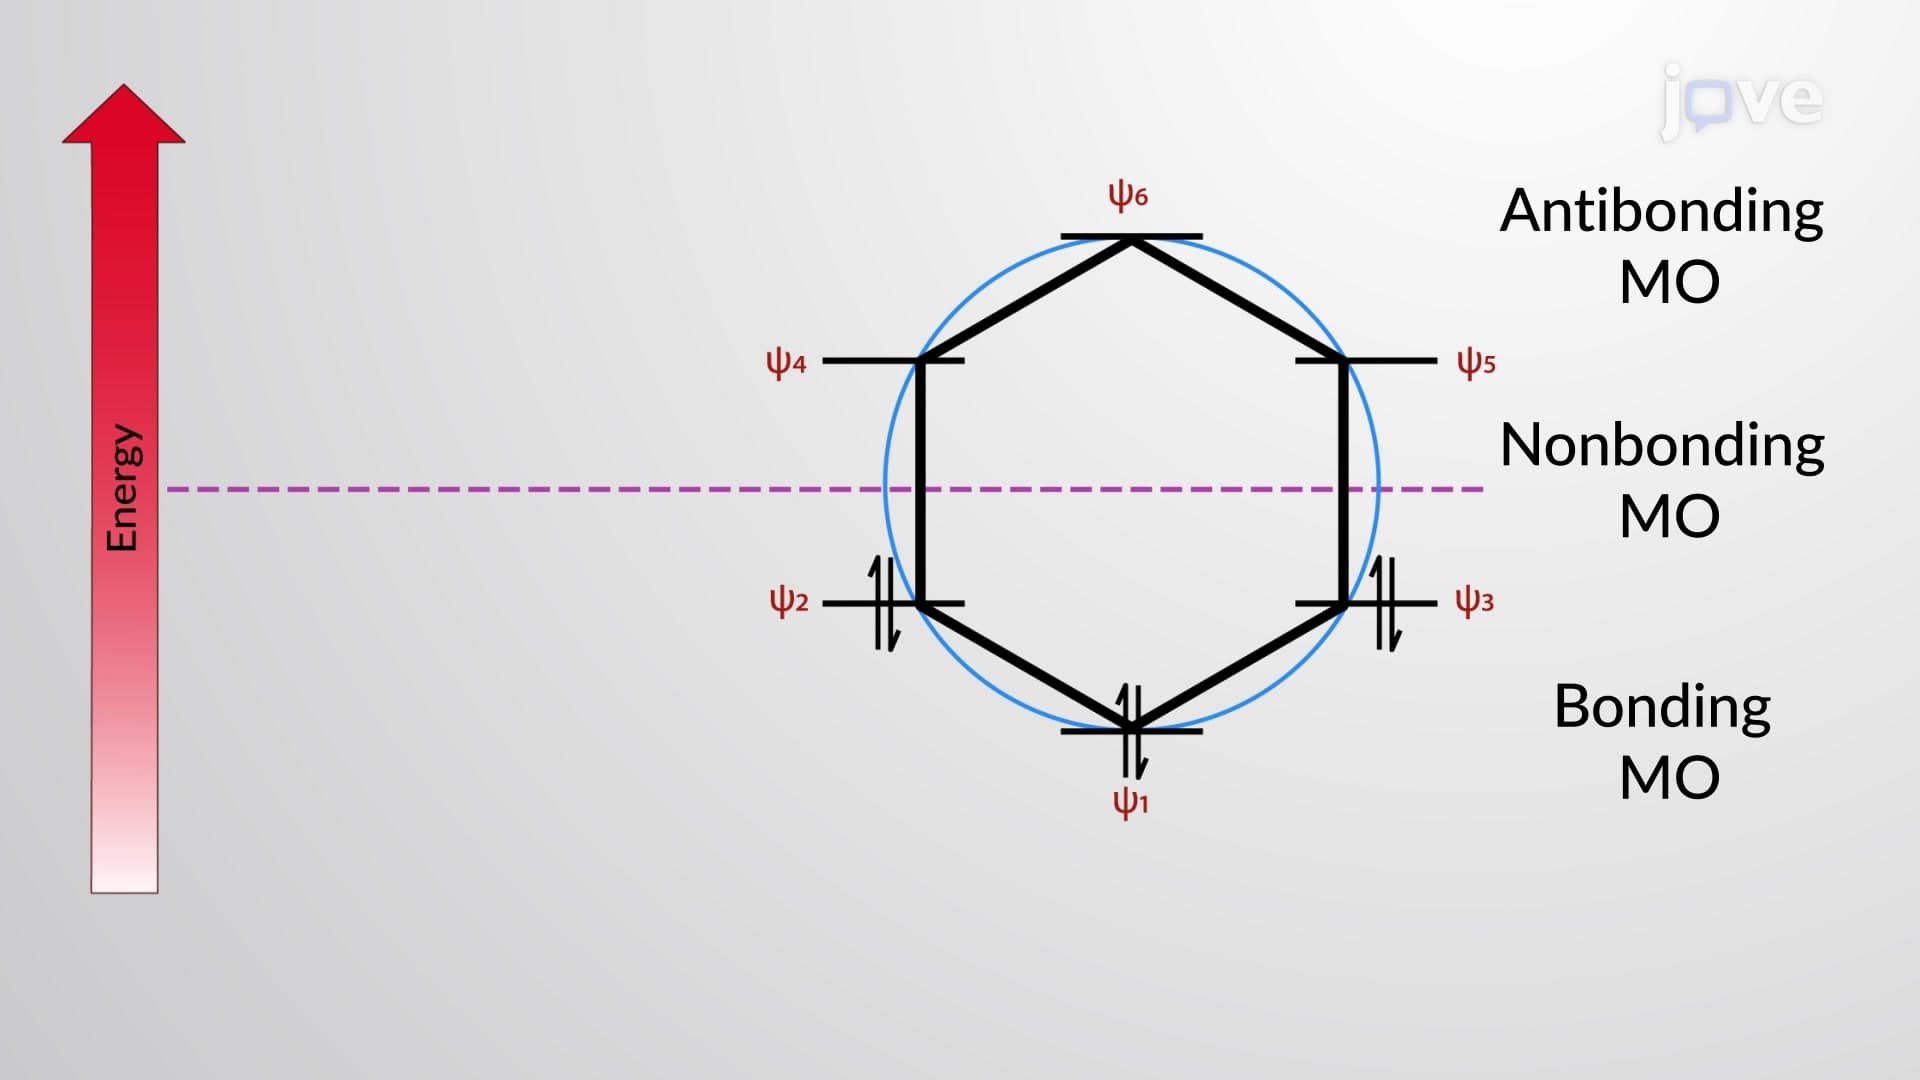
The Frost circle or the inscribed polygon method is a graphical method for determining the relative energies of π molecular orbitals (MOs) for planar, fully conjugated, and monocyclic compounds. This method was first described by A. A. Frost and Boris Musulin in 1953.
A Frost circle is constructed by drawing a polygon whose number of edges is equal to the number of carbons of the given cyclic system, with one of the vertices pointing down. Then, a circle is drawn enclosing the polygon so that...
A Frost circle is constructed by drawing a polygon whose number of edges is equal to the number of carbons of the given cyclic system, with one of the vertices pointing down. Then, a circle is drawn enclosing the polygon so that...
 01:27
π Electron Effects on Chemical Shift: Overview
An applied magnetic field causes loosely bound π-electrons in organic molecules to circulate, producing a local or induced diamagnetic field over a large spatial volume. As the molecules tumble in solution, the field generated by π-electrons in spherical substituents results in a zero net field. However, the net field generated by π-electrons in non-spherical substituents is not zero. The effect of this induced field depends on the orientation of the molecule with respect to B0, resulting in...