固态67ZnNMR光谱研究和ab initio分子轨道计算在合成模拟的碳酸 anhydrase的合成模拟
Andrew S Lipton1, Catherine Bergquist, Gerard Parkin
1Macromolecular Structure & Dynamics Directorate, WR Wiley Environmental Molecular Sciences Laboratory, Richland, Washington 99352, USA.
Journal of the American Chemical Society
|March 27, 2003

概括
低温固态67ZnNMR光谱显示,复合体中的四极合常数随着替代物电子负性的增加而增加. 这一发现有助于理解的活性部位,比如在碳酸中.
科学领域:
- 无机化学 无机化学 有机化学
- 固态NMR光谱学 固态NMR光谱学
- 有机金属化学 有机金属化学
背景情况:
- 三 ((pyrazolyl) 酸联体在稳定金属中心方面具有多功能.
- 复合物在生物系统中起着至关重要的作用,特别是在像碳酸无水酶这样的酶中.
- 固态NMR光谱学提供了四极核的电子环境的详细信息.
研究的目的:
- 用固态NMR研究复合物的电子性质,使用不同的替代剂.
- 为了将光谱数据与替代物电子负性相关联.
- 使用计算方法建模碳酸酸酶的活性部位.
主要方法:
- 使用了低温固态67ZnNMR光谱学.
- 三 (pyrazolyl) 酸盐复合物的合成和特征 [Tp (but,Me) ] ZnX (X = Br, Cl, OH).
- 对模型复合体的电场梯度张量进行计算计算.
主要成果:
- 的四极合常量 (Cq) 显示,随着替代物X (Br < Cl << OH) 的电子负性,的单调性增加.
- 对[Tp(But,Me) ]ZnOH的实验性NMR数据与计算结果一致.
- 对甲基利米达氧化复合物的计算,模拟碳酸无酶,与实验和计算值有很好的一致性.
结论:
- 中心的电子环境对结合替代剂的性质敏感.
- 固态67Zn核磁共振是检测有机金属复合体中的电子结构的宝贵工具.
- 该研究提供了有关生物系统的协调化学的见解.
相关概念视频
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
At room temperature, the chair conformer of cyclohexane undergoes rapid ring flipping between two equivalent chair conformers at a rate of approximately 105 times per second. These two chair conformers are in equilibrium. The rapid ring flipping results in the interconversion of the axial proton to an equatorial proton and an equatorial to the axial proton. Such interconversions are too rapid and cannot be detected on the NMR timescale. Hence, the NMR spectrometer cannot distinguish between the...
 01:15
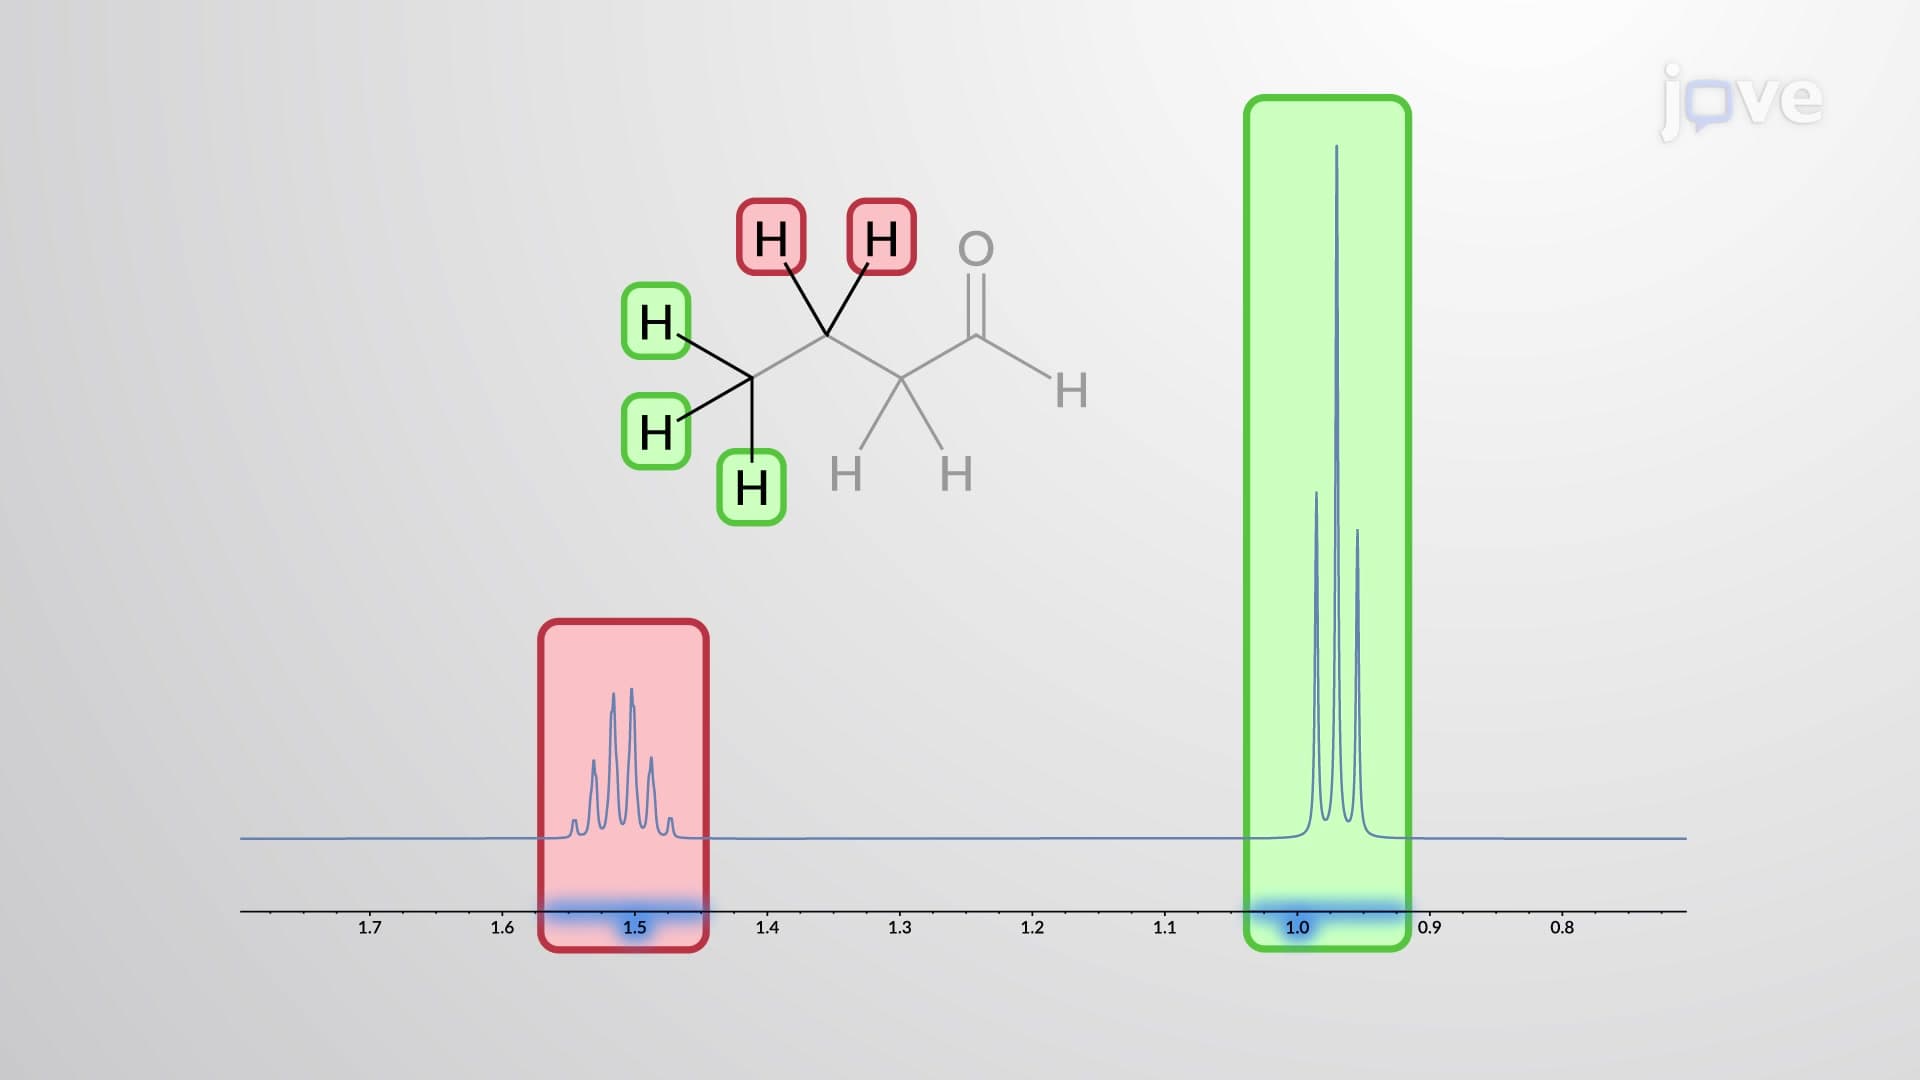
NMR Spectroscopy and Mass Spectrometry of Aldehydes and Ketones
In aldehydes, the hydrogen atom connected to the carbonyl carbon helps distinguish aldehydes from other carbonyl compounds using ¹H NMR spectroscopy. The closeness of aldehydic hydrogen to the electrophilic carbonyl carbon highly deshields the hydrogen atom causing its signal to appear around 10 ppm in the ¹H NMR spectra. α hydrogens split the aldehydic proton signal, which helps identify the number of α hydrogens in the molecule. For instance, one α hydrogen creates a doublet for an aldehydic...
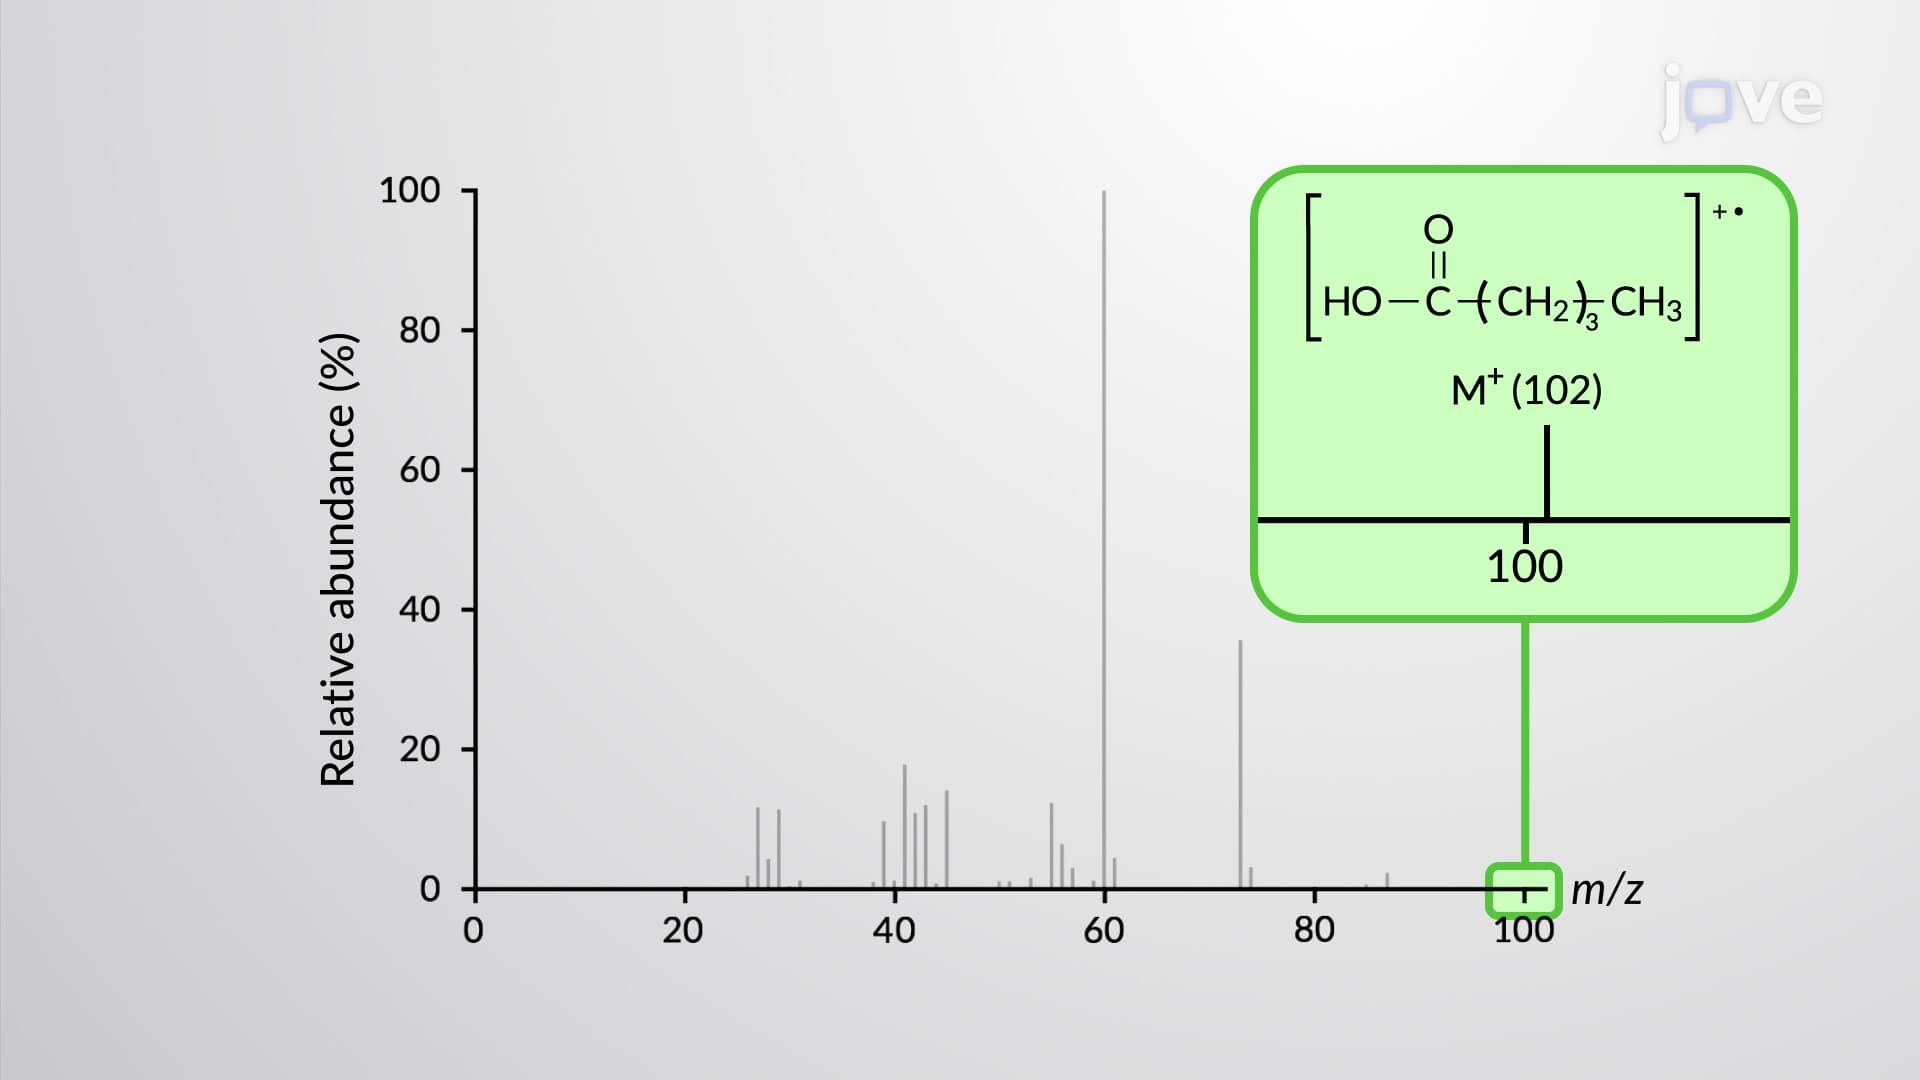 01:30
NMR and Mass Spectroscopy of Carboxylic Acids
In ¹H NMR spectroscopy, acidic protons (–COOH) of carboxylic acids are highly deshielded and absorb far downfield, at around 9–12 ppm. The chemical shift value depends on the concentration and solvent used.
While α protons of carboxylic acids absorb at 2–2.5 ppm, β protons absorb further upfield.
Carboxylic acids are easily identified by dissolving them in deuterium oxide, which results in a rapid exchange of the acidic protons with deuterium. This leads to the disappearance of the acidic...
While α protons of carboxylic acids absorb at 2–2.5 ppm, β protons absorb further upfield.
Carboxylic acids are easily identified by dissolving them in deuterium oxide, which results in a rapid exchange of the acidic protons with deuterium. This leads to the disappearance of the acidic...
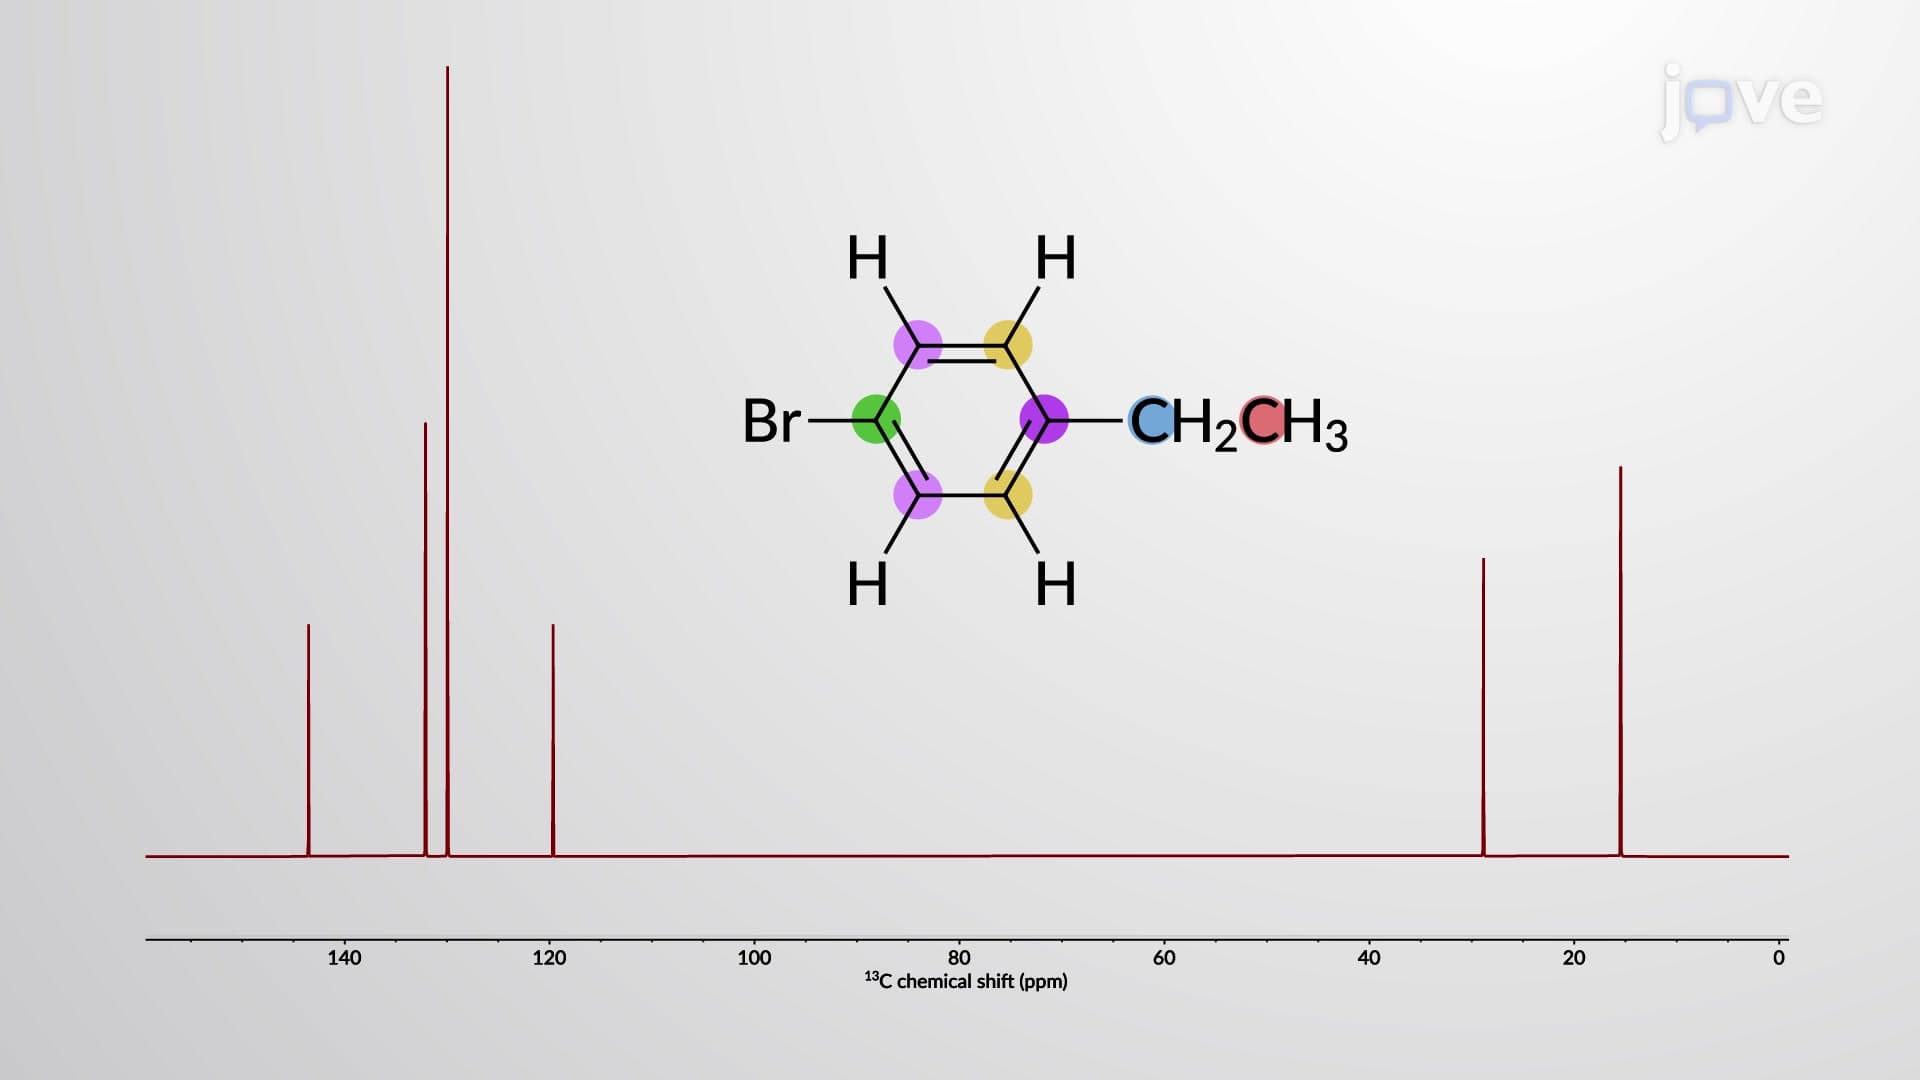 01:37
NMR Spectroscopy of Benzene Derivatives
Simple unsubstituted benzene has six aromatic protons, all chemically equivalent. Therefore, benzene exhibits only a singlet peak at δ 7.3 ppm in the 1H NMR spectrum. The observed shift is far downfield because the aromatic ring current strongly deshields the protons. Any substitution on the benzene ring makes the aromatic protons nonequivalent, and the protons split each other. The peak is, therefore, no longer a singlet and the splitting pattern and their associated coupling constants depend...
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
The axial and equatorial protons in cyclohexane can be distinguished by performing a variable-temperature NMR experiment. In this process, except for one proton, the remaining eleven protons are replaced by deuterium. The deuterium substitution avoids the possible peak splitting caused by the spin-spin coupling between the adjacent protons. The remaining proton flips between the axial and equatorial positions.
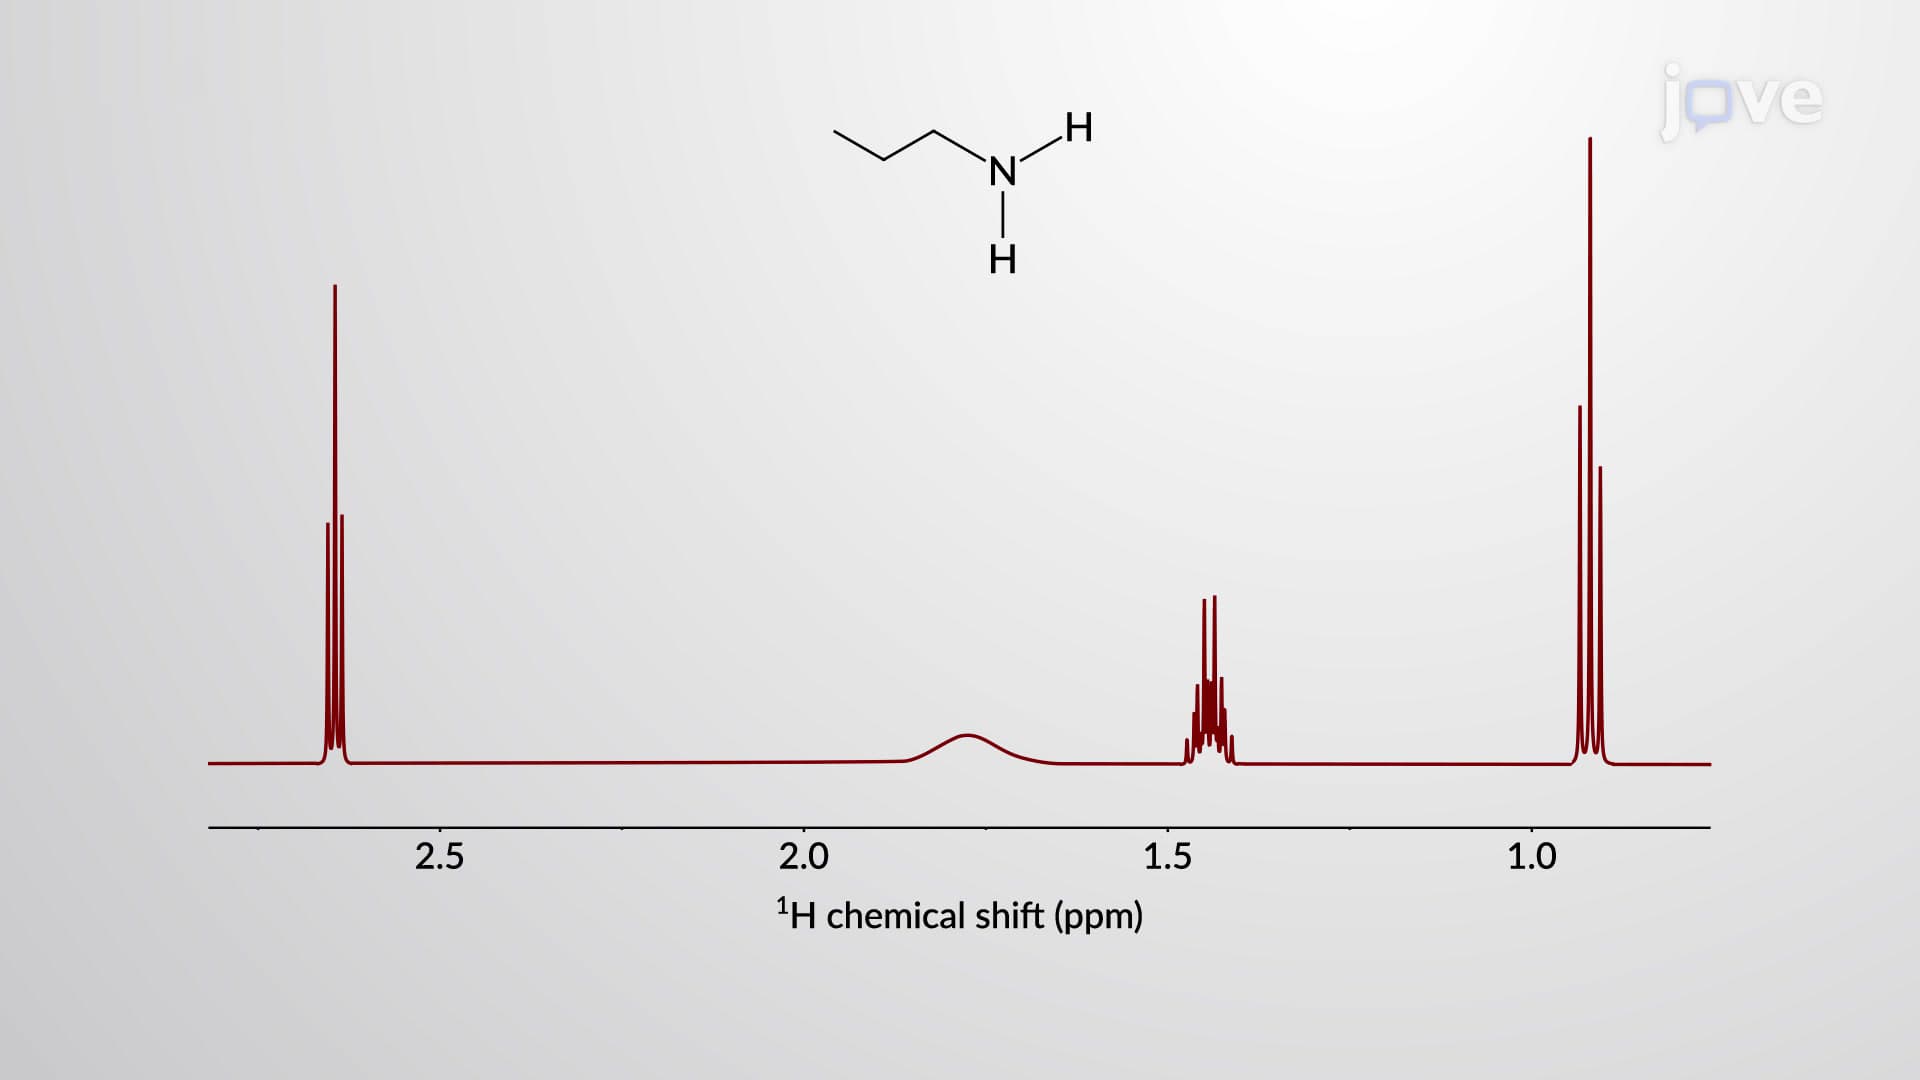 01:19
NMR Spectroscopy Of Amines
In proton NMR spectroscopy, primary amines and secondary amines showcase their N–H protons as a broad signal in the chemical shift range between δ 0.5 and 5 ppm. The exact position in this range depends on several factors, including sample concentration, hydrogen bonding, and the type of solvent used. Since amine protons undergo fast proton exchange in solution, the protons are labile and therefore do not participate in any splitting with adjacent protons. Thus, the observed peak is broad and...