
原子的平面网由新的二元抗氧化物Ti2Sbb中的方形和组成
Shahab Derakhshan1, Abdeljalil Assoud, Katja M Kleinke
1Department of Chemistry, University of Waterloo, Waterloo, ON, Canada N2L 3G1.
Journal of the American Chemical Society
|July 1, 2004

概括
研究人员发现了一种新的抗氧化物,Ti(2) Sb,具有独特的晶体结构. 这种结构增强了-结合,降低了材料的总能量,提高了稳定性.
科学领域:
- 固态化学 固态化学
- 晶体学 晶体学是指结晶学.
- 材料科学是一种材料科学.
背景情况:
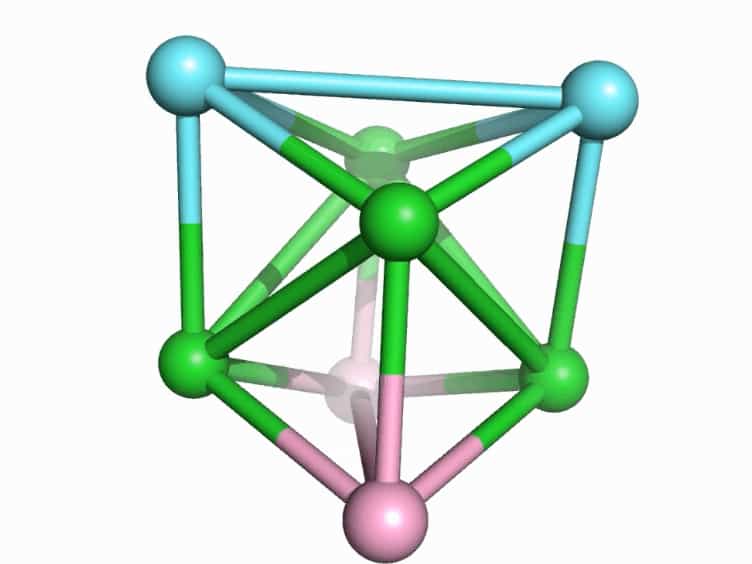
- 2Sb晶体类型具有带有短拉拉键的正方形平面网.
- 了解二元抗氧化物中的结构变异对于开发新材料至关重要.
研究的目的:
- 为了确定二元抗氧化物Ti(2) Sb. 的晶体结构.
- 研究Ti(2) Sb. 的结合特性和电子结构.
- 了解其结构形成背后的驱动力.
主要方法:
- 单晶X射线衍射. 单晶X射线衍射.
- 现实空间对分布函数 (PDF) 分析使用X射线和中子粉的衍射数据.
- 电子结构计算.
主要成果:
- Ti(2) Sb 结晶成一个扭曲的 La(2) Sb 型结构.
- 网被变形成正方形和方形,增强了Ti-Ti的结合.
- 电子结构计算证实,由于加强的Ti-Ti相互作用,乱降低了总能量.
结论:
- Ti(2) Sb 独特的扭曲结构优化了 Ti-Ti 的结合.
- 增强的Ti-Ti相互作用是驱动观察到的结构和电子特性的主要因素.
- 这一发现为相关的二元抗氧化物材料的设计原则提供了洞察力.
相关概念视频
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
49.1K
Tetrahedral Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
49.1K
 02:42
Valence Bond Theory
11.4K
Coordination compounds and complexes exhibit different colors, geometries, and magnetic behavior, depending on the metal atom/ion and ligands from which they are composed. In an attempt to explain the bonding and structure of coordination complexes, Linus Pauling proposed the valence bond theory, or VBT, using the concepts of hybridization and the overlapping of the atomic orbitals. According to VBT, the central metal atom or ion (Lewis acid) hybridizes to provide empty orbitals of suitable...
11.4K
 02:57
Coordination Number and Geometry
19.3K
For transition metal complexes, the coordination number determines the geometry around the central metal ion. Table 1 compares coordination numbers to molecular geometry. The most common structures of the complexes in coordination compounds are octahedral, tetrahedral, and square planar.
19.3K
 02:42
Ionic Crystal Structures
19.0K
Ionic crystals consist of two or more different kinds of ions that usually have different sizes. The packing of these ions into a crystal structure is more complex than the packing of metal atoms that are the same size.
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
19.0K
 02:37
Metallic Solids
21.1K
Metallic solids such as crystals of copper, aluminum, and iron are formed by metal atoms. The structure of metallic crystals is often described as a uniform distribution of atomic nuclei within a “sea” of delocalized electrons. The atoms within such a metallic solid are held together by a unique force known as metallic bonding that gives rise to many useful and varied bulk properties.
All metallic solids exhibit high thermal and electrical conductivity, metallic luster, and malleability....
All metallic solids exhibit high thermal and electrical conductivity, metallic luster, and malleability....
21.1K
 02:27
Predicting Molecular Geometry
46.4K
VSEPR Theory for Determination of Electron Pair Geometries
46.4K