
1,5-的催化异构化为双环[3.1.0]素
Michael R Luzung1, Jordan P Markham, F Dean Toste
1Center for New Directions in Organic Synthesis, Department of Chemistry, University of California-Berkeley, Berkeley, CA 94720, USA.
Journal of the American Chemical Society
|September 2, 2004

概括
黄金(I) 催化剂促进1,5-的循环异构,形成双循环[3.1.0]素. 这种反应可以容忍多种不同的替代,并实现出色的立体特异性和奇拉性转移,使复杂分子合成.
科学领域:
- 有机化学 有机化学
- 催化剂是一种催化剂.
- 合成方法论 合成方法论
背景情况:
- 1,5-类是有机合成中的多功能基质.
- 黄金的催化已经成为激活不和债券的强大工具.
研究的目的:
- 为了研究1,5-enynes.的金(I) 催化循环异构化.
- 探索这种转型的范围和局限性.
- 为了实现双环[3.1.0]烯衍生物的立体选择性和反选择性合成.
主要方法:
- 使用阴离子三四金 (((I) 复合物作为催化剂.
- 采用各种替代的1,5-因作为基质.
- 使用光谱方法 (例如,NMR) 和手术色谱学分析反应产物.
主要成果:
- 从1,5-enynes中有效合成双环[3.1.0]素.
- 不同替代模式的耐受性,包括四级碳.
- 1,2-非替代烯的立体特异性循环异构化.
- 从丰富的原料中转移高的选择性和性.
- 通过并联反应成功制备三环系统.
结论:
- 金(I) 催化提供了一种强大的合成bicyclo[3.1.0]hexenes的方法.
- 这种反应对立体化学和酶选择性有很好的控制.
- 这种方法扩大了对复杂的碳循环框架的访问.
相关概念视频
 02:43
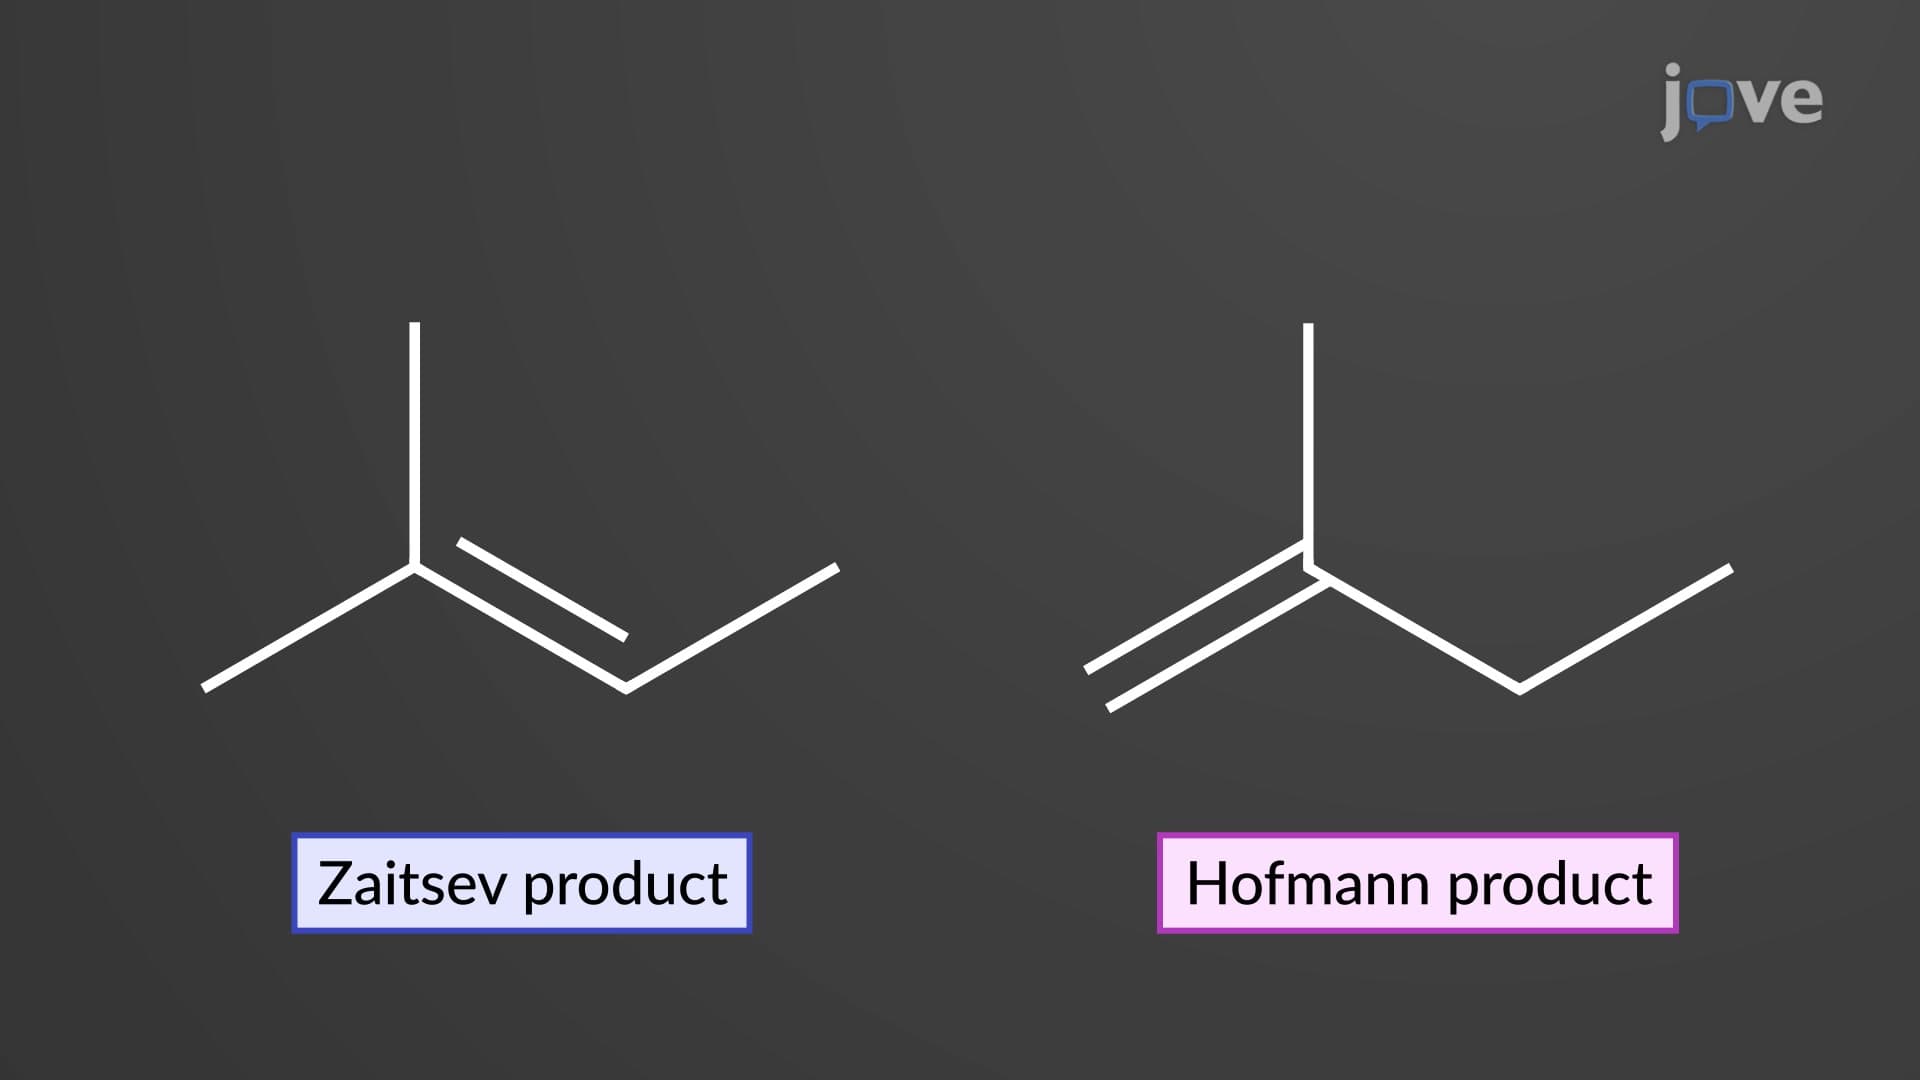
E1 Reaction: Stereochemistry and Regiochemistry
One of the critical aspects of the E1 reaction mechanism, as also observed in E2, is the regiochemistry, with multiple regioisomers obtained as products. In the example discussed, the presence of water as a weak base favors elimination over substitution to generate two alkenes. Given that alkenes’ stability increases with the number of alkyl groups across the double bond, typically, E1 reactions lead to the Zaitsev product, for this is more substituted and stable than the Hofmann product.
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration
The rate of acid-catalyzed hydration of alkenes depends on the alkene's structure, as the presence of alkyl substituents at the double bond can significantly influence the rate.
 01:21
Acid-Catalyzed α-Halogenation of Aldehydes and Ketones
By replacing an α-hydrogen with a halogen, acid-catalyzed α-halogenation of aldehydes or ketones yields a monohalogenated product
In the first step of the mechanism, the acid protonates the carbonyl oxygen resulting in a resonance-stabilized cation, which subsequently loses an α-hydrogen to form an enol tautomer. The C=C bond in an enol is highly nucleophilic because of the electron-donating nature of the –OH group. Consequently, the double bond attacks an electrophilic halogen to form a...
In the first step of the mechanism, the acid protonates the carbonyl oxygen resulting in a resonance-stabilized cation, which subsequently loses an α-hydrogen to form an enol tautomer. The C=C bond in an enol is highly nucleophilic because of the electron-donating nature of the –OH group. Consequently, the double bond attacks an electrophilic halogen to form a...
 01:17
Thermal Electrocyclic Reactions: Stereochemistry
The stereochemistry of electrocyclic reactions is strongly influenced by the orbital symmetry of the polyene HOMO. Under thermal conditions, the reaction proceeds via the ground-state HOMO.
Selection Rules: Thermal Activation
Conjugated systems containing an even number of π-electron pairs undergo a conrotatory ring closure. For example, thermal electrocyclization of (2E,4E)-2,4-hexadiene, a conjugated diene containing two π-electron pairs, gives trans-3,4-dimethylcyclobutene.
Selection Rules: Thermal Activation
Conjugated systems containing an even number of π-electron pairs undergo a conrotatory ring closure. For example, thermal electrocyclization of (2E,4E)-2,4-hexadiene, a conjugated diene containing two π-electron pairs, gives trans-3,4-dimethylcyclobutene.
 01:26
Photochemical Electrocyclic Reactions: Stereochemistry
The absorption of UV–visible light by conjugated systems causes the promotion of an electron from the ground state to the excited state. Consequently, photochemical electrocyclic reactions proceed via the excited-state HOMO rather than the ground-state HOMO. Since the ground- and excited-state HOMOs have different symmetries, the stereochemical outcome of electrocyclic reactions depends on the mode of activation; i.e., thermal or photochemical.
Selection Rules: Photochemical Activation
Selection Rules: Photochemical Activation
 01:21
[3,3] Sigmatropic Rearrangement of 1,5-Dienes: Cope Rearrangement
The Cope rearrangement is classified as a [3,3] sigmatropic shift in 1,5-dienes, leading to a more stable, isomeric 1,5-diene. The reaction involves a concerted movement of six electrons, four from two π bonds and two from a σ bond, via an energetically favorable chair-like transition state.