在和-表面上对二氧化碳吸附点的振动识别
Ismaila Dabo1, Andrzej Wieckowski, Nicola Marzari
1Department of Materials Science and Engineering, Massachusetts Institute of Technology, 77 Massachusetts Avenue, Cambridge, Massachusetts 02139, USA. dabo_is@mit.edu

密度函数理论准确地预测了和-表面上的一氧化碳 (CO) 振动频率. 这种方法正确地模拟了由于的频率变化,即使其他方法在吸附点稳定性方面失败.
科学领域:
- 表面科学是一门学科.
- 计算化学的计算化学
- 材料科学 材料科学 材料科学
背景情况:
- 了解吸附分子的振动特性对于催化和表面化学至关重要.
- 一氧化碳 (CO) 在等过渡金属上的吸附是一种基本的模型系统.
- 准确预测吸附点稳定性和振动频率仍然是计算方法的挑战.
研究的目的:
- 为了研究在和-表面上被吸附的CO的振动特性.
- 评估密度函数理论 (DFT) 方法,特别是Perdew-Burke-Ernzerhof (PBE) 通用梯度近似 (GGA) 的准确性,用于预测C-O拉伸频率和频率转移.
- 为了阐明C-O频率在共吸附时发生的变化的化学起源.
主要方法:
- 在PBE-GGA近似中使用了密度功能扰动理论 (DFPT).
- 对和-表面上吸附的二氧化碳进行了计算.
- 使用轨道解析分析状态的力和频率密度,以及GGA+U方法来了解频率转移的化学起源.
主要成果:
- 计算的C-O拉伸频率与实验光谱测量结果非常一致.
- 在单层吸附时的频率变化得到了准确的预测.
- 该研究强调了PBE-GGA在预测振动频率方面的成功,与其在预测吸附点稳定性方面的局限性形成鲜明对比.
结论:
- DFT计算,特别是PBE-GGA,对于预测吸附CO的绝对振动频率非常准确.
- 该研究确定了DFT在强化学吸收模式下对振动性质预测的可靠性.
- 轨道解析分析提供了对控制C-O振动频率转移的电子因素的见解.
更多相关视频
 11:38
In situ FTIR Spectroscopy as a Tool for Investigation of Gas/Solid Interaction: Water-Enhanced CO2 Adsorption in UiO-66 Metal-Organic Framework
Published on: February 1, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
相关概念视频
 01:25
Adsorption Isotherms II
 01:29
Adsorption Isotherms I
 01:28
Adsorption of Gases on Solids
 01:22
Heterogeneous Catalysis
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
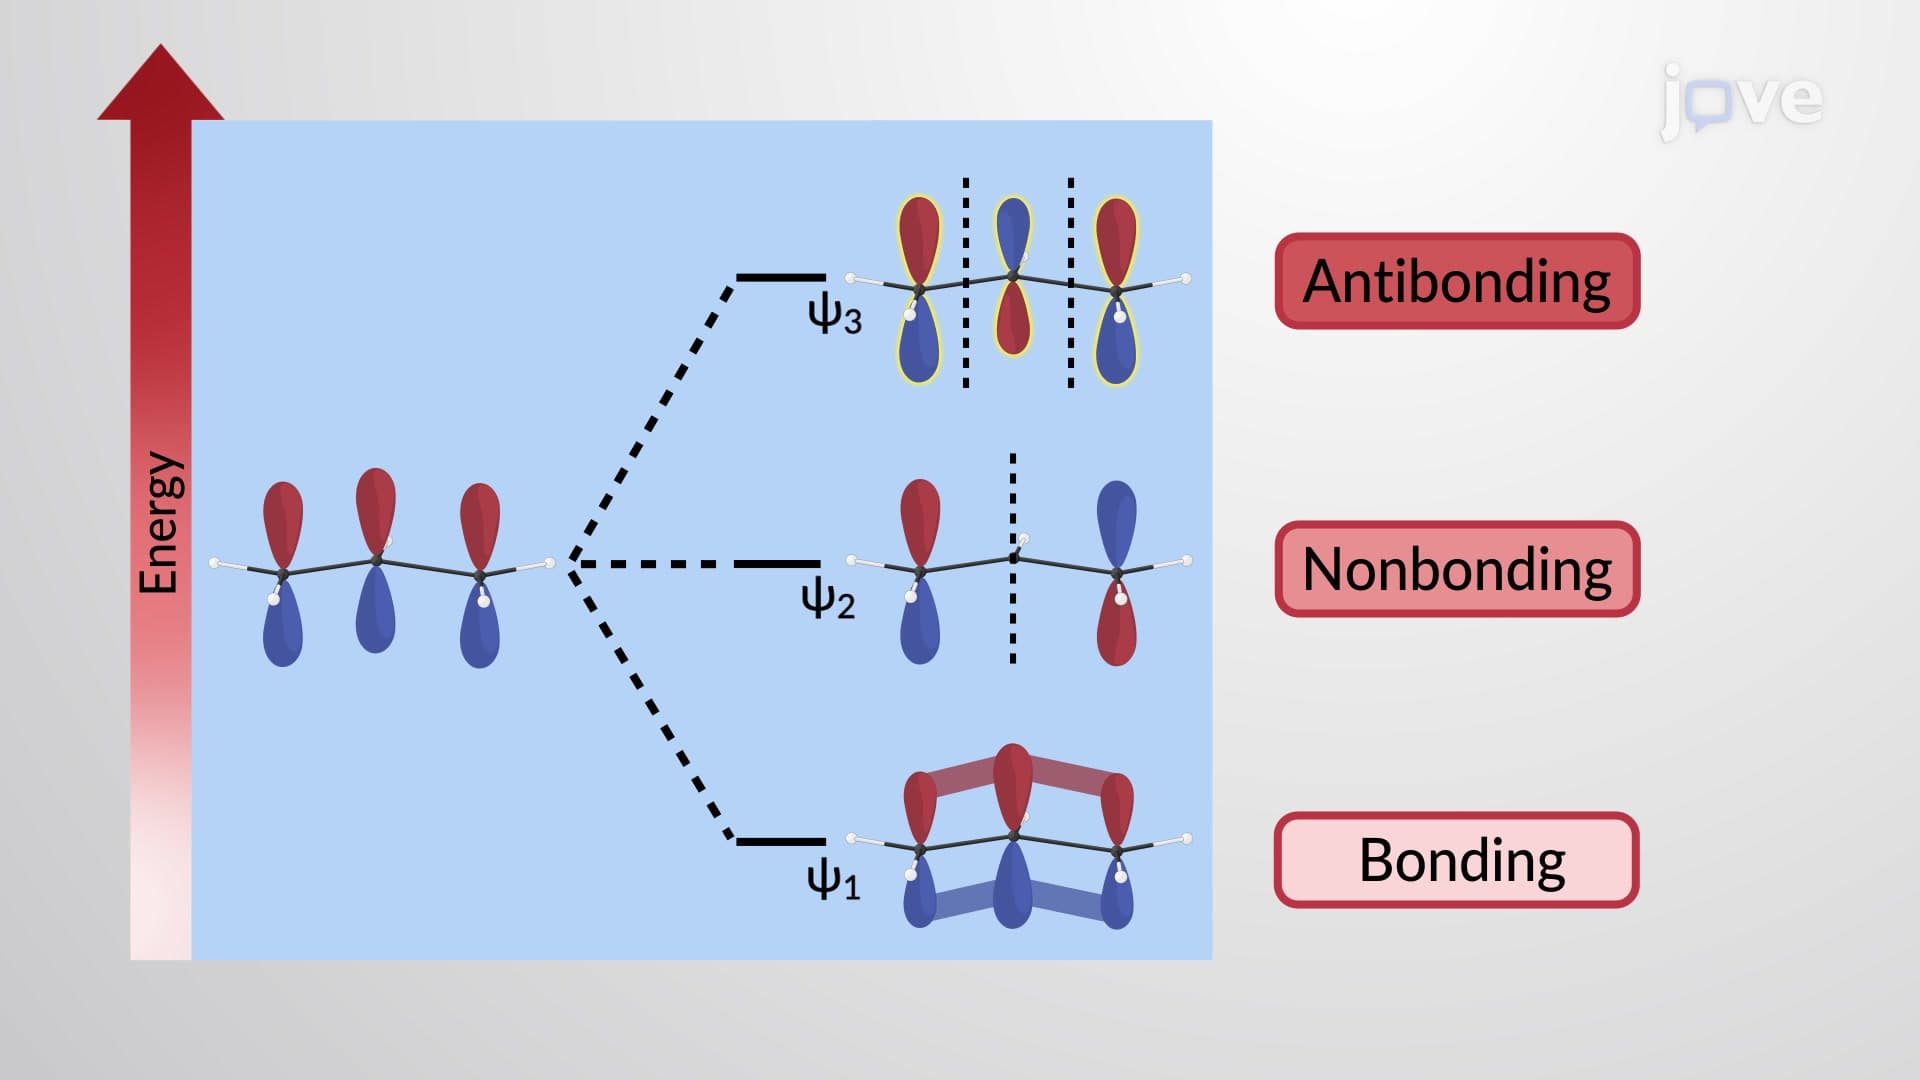 01:18
π Molecular Orbitals of the Allyl Cation and Anion