
H(2) 从多环芳中喷射:对质子化1,2-二二甲进行红外多光子解离研究
Martin Vala1, Jan Szczepanski, Jos Oomens
1Department of Chemistry and Center for Chemical Physics, P.O. Box 117200, University of Florida, Gainesville, Florida 32611-7200, USA. mvala@chem.ufl.edu
Journal of the American Chemical Society
|April 8, 2009

概括
这项研究使用光谱学和理论来分析质子化1,2-二甲 (DHN),揭示其结构和解离途径. 这些发现支持了从多环芳 (PAH) 离子中形成星际 (H2) 的新模型.
科学领域:
- 天体化学是天体化学.
- 物理化学 物理化学
- 频谱学是一种光谱学.
背景情况:
- 了解星际环境中分子 (H2) 的形成对天体化学至关重要.
- 多环芳 (PAH) 在太空中很丰富,可以被电离,可能在H2形成中发挥作用.
- 1,2-二甲 (DHN) 作为研究与星际化学相关的化和光解离过程的模型系统.
研究的目的:
- 为了研究质子化1,2-二二甲 (DHN) 的结构和光解离机制.
- 将实验光谱数据与理论计算进行比较,以验证结构赋值.
- 提供关于星际H2形成的拟议模型的见解,其中涉及PAH离子.
主要方法:
- 使用矩阵隔离红外吸收光谱来研究中性DHN.
- 采用多光子红外光解离 (IRMPD) 动作光谱技术,使用里埃变换离子循环子共振质谱仪和自由电子激光器对质子化DHN进行了测试.
- 进行密度函数理论 (DFT) 计算 (B3LYP/6-31++G(d,p)) 来确定结构,能量和光谱.
主要成果:
- DHN的质子化主要发生在3位,形成1,2,3-三纳烯 (1,2,3-THN+).
- 实验IRMPD对质子化DHN的频谱与1,2,3-THN+的计算频谱密切匹配.
- 在IR辐射下观察到1,2,3-THN+的两个解离通道,产物离子在m/z 129和m/z 91处,提供了机械洞察力.
结论:
- 该研究证实了质子化DHN的结构,并阐明了其光解离路径.
- 实验和理论数据支持星际H2形成的新模型.
- 这种模型涉及PAH离子的化,其次是光溶性H2喷射,DHN作为相关的模拟物.
相关概念视频
 01:13
¹H NMR: Complex Splitting
A proton M that is coupled to a proton X results in doublet signals for M. However, NMR-active nuclei can be simultaneously coupled to more than one nonequivalent nucleus. When M is coupled to a second proton A, such as in styrene oxide, each peak in the doublet is split into another doublet.
Splitting diagrams or splitting tree diagrams are routinely used to depict such complex couplings. While drawing splitting diagrams, the splitting with the larger coupling constant is usually applied first.
Splitting diagrams or splitting tree diagrams are routinely used to depict such complex couplings. While drawing splitting diagrams, the splitting with the larger coupling constant is usually applied first.
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Cycloheptatriene is a neutral monocyclic unsaturated hydrocarbon that consists of an odd number of carbon atoms and an intervening sp3 carbon in the ring. The three double bonds in the ring correspond to 6 π electrons, which is a Huckel number, and therefore satisfies the criteria of 4n + 2 π electrons. However, the intervening sp3 carbon disrupts the continuous overlap of p orbitals. As a result, cycloheptatriene is not aromatic.
Removing one hydrogen from the intervening CH2 group with both...
Removing one hydrogen from the intervening CH2 group with both...
 00:48
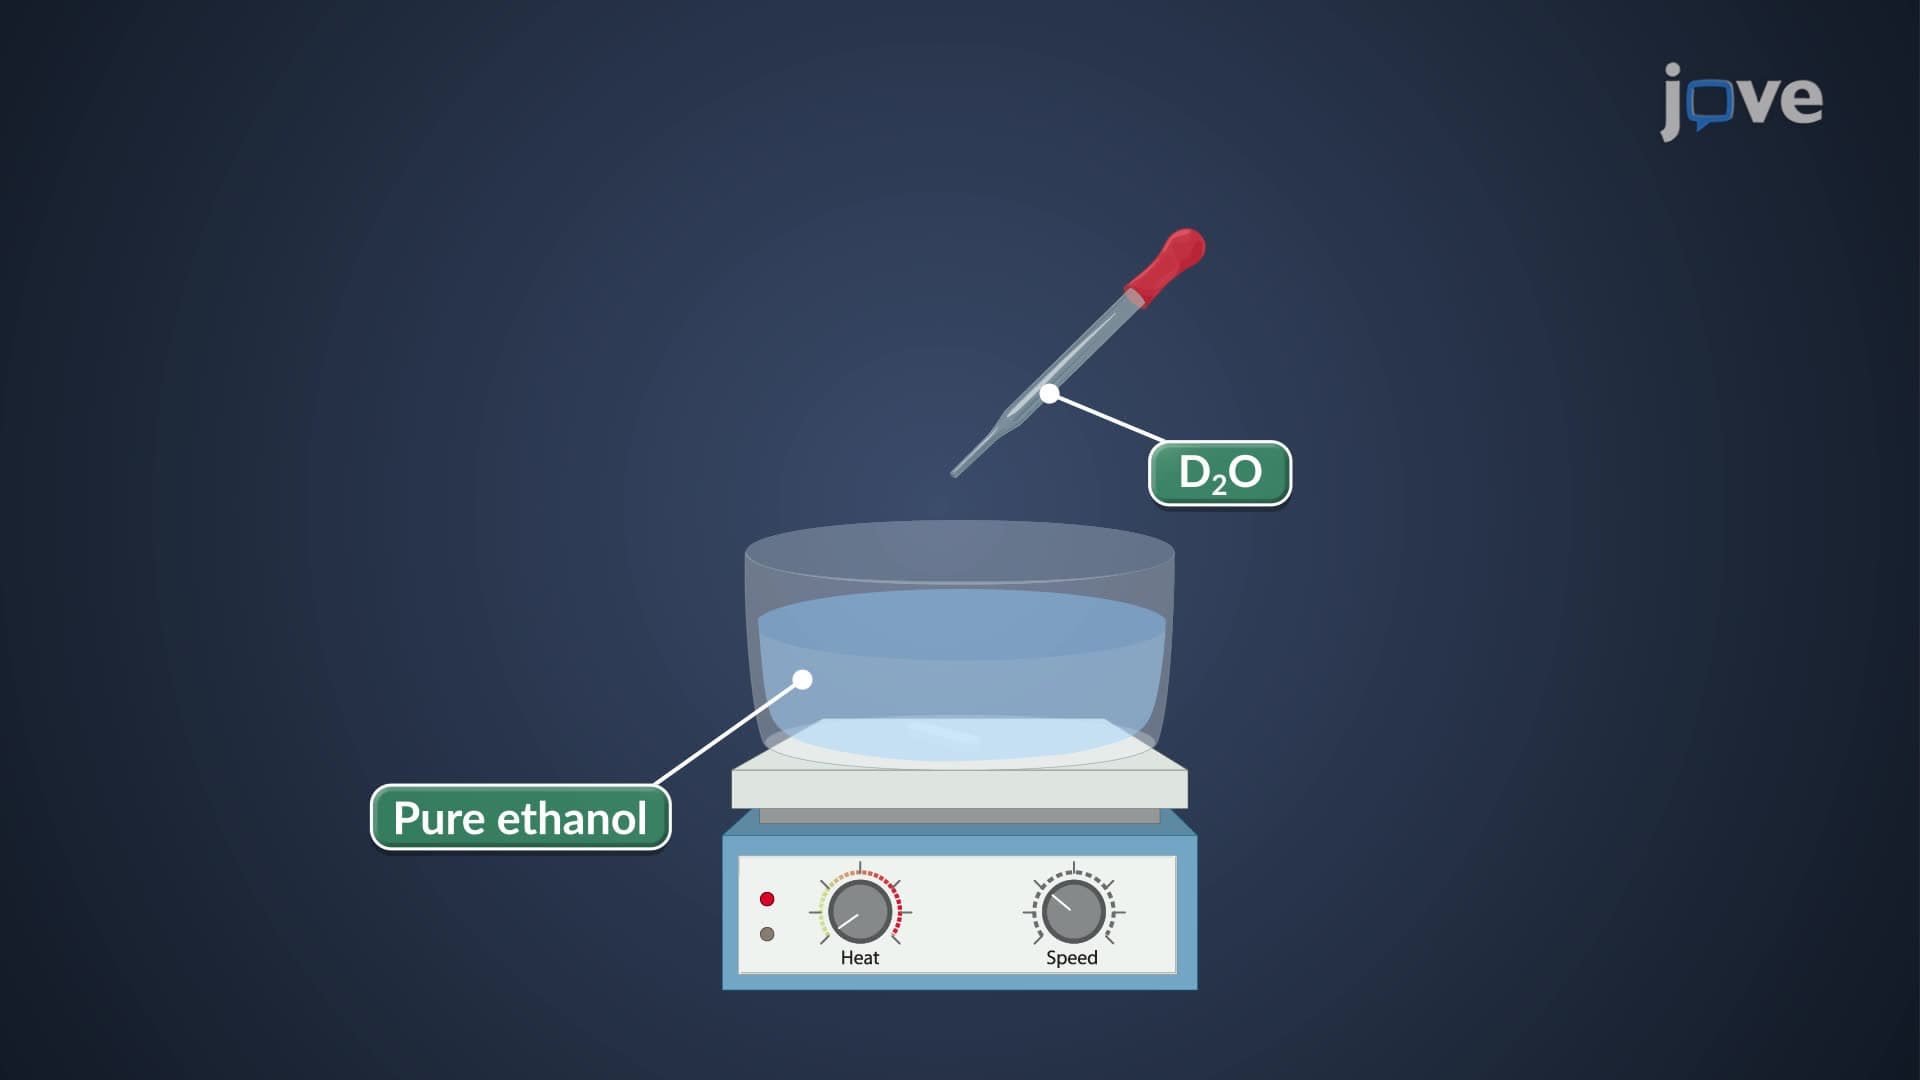
¹H NMR of Labile Protons: Deuterium (²H) Substitution
This lesson illustrates the role of deuterium substitution in simplifying the NMR spectrum of compounds comprising labile protons. One method employed is the use of deuterium. Amongst the three isotopes of hydrogen, deuterium (2H) has a nucleus composed of one proton and one neutron. When the D2O solvent is added to a pure dry ethanol solution, its labile proton is substituted with deuterium.
 01:14
NMR Spectroscopy of Aromatic Compounds
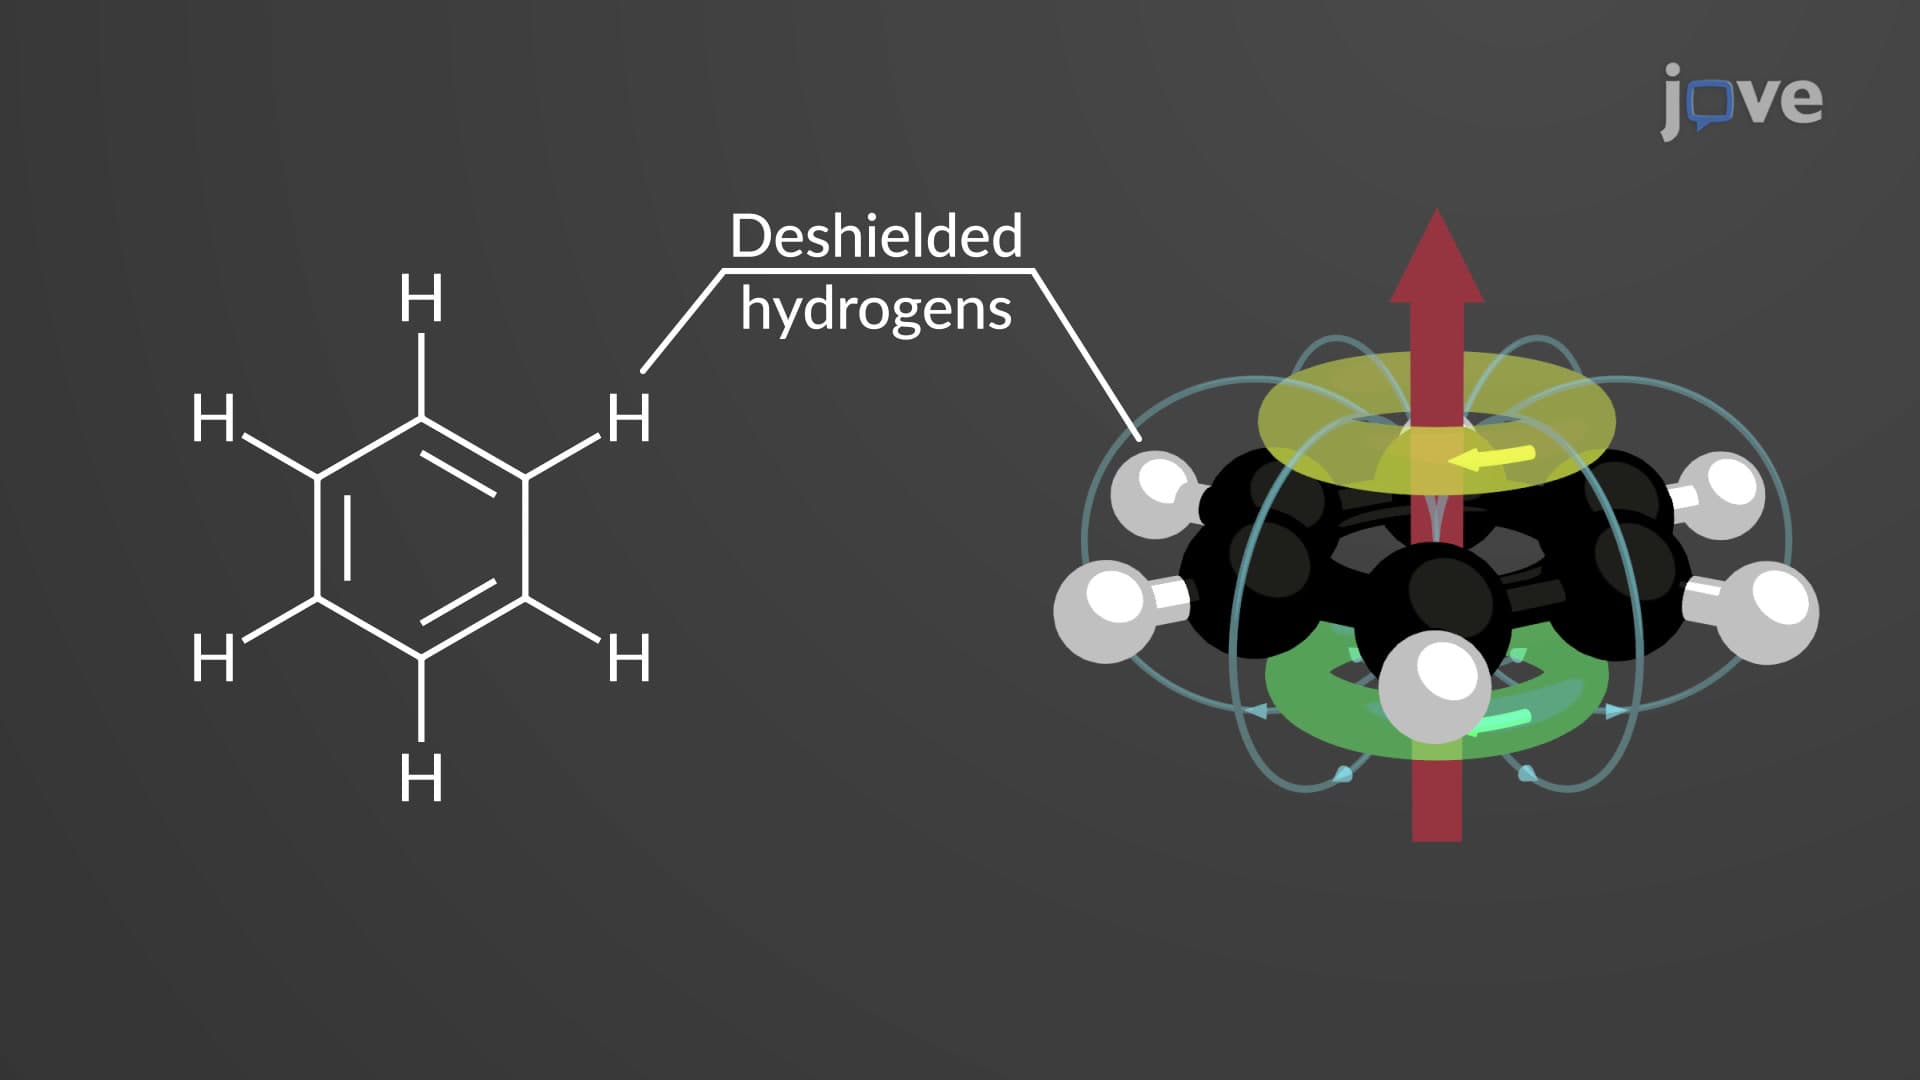
Aromatic compounds can be identified or analyzed using proton NMR and carbon‐13 NMR. Typically, aromatic hydrogens or hydrogens directly bonded to the aromatic rings are strongly deshielded by the aromatic ring current. Therefore, they absorb in the range of 6.5–8.0 ppm in proton NMR spectra. For instance, aromatic hydrogens directly bonded to the benzene ring absorb at 7.3 ppm. However, aromatic hydrogens of larger rings absorb farther upfield or downfield than the ideal range. Consider...
 01:29
UV–Vis Spectroscopy: Woodward–Fieser Rules
UV–Visible absorption spectra of conjugated dienes arise from the lowest energy π → π* transitions. The light-absorbing part of the molecule is called the chromophore, and the substituents directly attached to the chromophore are called auxochromes. A strong correlation exists between the absorption maxima, λmax, and the structure of a conjugated π system. The Woodward–Fieser rules predict the value of λmax for a given structure by adding the contributions...
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
The axial and equatorial protons in cyclohexane can be distinguished by performing a variable-temperature NMR experiment. In this process, except for one proton, the remaining eleven protons are replaced by deuterium. The deuterium substitution avoids the possible peak splitting caused by the spin-spin coupling between the adjacent protons. The remaining proton flips between the axial and equatorial positions.