在H-ZSM-5上进行乙化量的量子化学建模接近化学精度:混合MP2:DFT研究
Niels Hansen1, Torsten Kerber, Joachim Sauer
1Department of Chemical Engineering, Hamburg University of Technology, D-21073 Hamburg, Germany. n.hansen@tuhh.de
Journal of the American Chemical Society
|August 4, 2010

概括
这项研究使用混合MP2:DFT方法精确计算基化在H-ZSM-5上,为工业应用提供精确的吸附度和激活能量障碍.
科学领域:
- 计算化学计算化学
- 催化科学 催化科学
- 材料科学 材料科学 材料科学
背景情况:
- 烯与乙烯的化在H-ZSM-5等化物上是一个关键的工业过程.
- 准确的反应能量的理论预测对于催化剂设计和优化至关重要.
研究的目的:
- 使用先进的计算方法,准确确定 H-ZSM-5 上基化的吸附能量和反应障碍.
- 为了比较不同理论方法的性能,包括MP2:DFT和PBE+D,与高级计算.
主要方法:
- 混合MP2:DFT方案将周期密度函数理论 (DFT) 与MP2能量计算相结合.
- 通过对MP2能量的完整基础设定极限进行推断,并通过CCSD (T) 计算包含更高阶的相关性效应.
- 对潜在能量表面的分散贡献和自我相互作用校正错误的分析.
主要成果:
- 乙烯,和乙基在298K的吸附的最终度分别为-46,-78和-110kJ/mol.
- 在653K的内部阻被确定为117kJ/mol (单步) 和119/94kJ/mol (双步化).
- 模拟的明显激活能量 (66kJ/mol) 与实验数据 (58-76kJ/mol) 非常接近.
结论:
- 混合MP2:DFT方案为H-ZSM-5的基化提供了准确的能量.
- 散射力显著影响潜在能量表面,需要仔细处理.
- 与高水平方法相比,PBE+D方法对吸附能量的一致性很好,但对内在能量障碍的一致性较低.


相关概念视频
 01:18
Structure of Benzene: Molecular Orbital Model
According to the molecular orbital (MO) model, benzene has a planar structure with a regular hexagon of six sp2 hybridized carbons. As shown in Figure 1, each carbon is bonded to three other atoms with C–C–C and H–C–C bond angles of 120°. The C–H bond length is 109 pm, and the C–C bond length is 139 pm which is midway between the single bond length of sp3 hybridized carbons (154 pm) and sp2 hybridized carbons (133 pm).
 01:07
Structure of Benzene: Kekulé Model
In 1865, August Kekule suggested the structure of benzene according to the structural theory of organic chemistry based on the three assertions—formula of benzene is C6H6, all the hydrogens of benzene are equivalent, and each carbon must have four bonds due to its tetravalency.
He proposed that benzene has a cyclic structure of six carbon atoms attached to one hydrogen atom each, with three alternating pi bonds.
He proposed that benzene has a cyclic structure of six carbon atoms attached to one hydrogen atom each, with three alternating pi bonds.
 01:37
NMR Spectroscopy of Benzene Derivatives
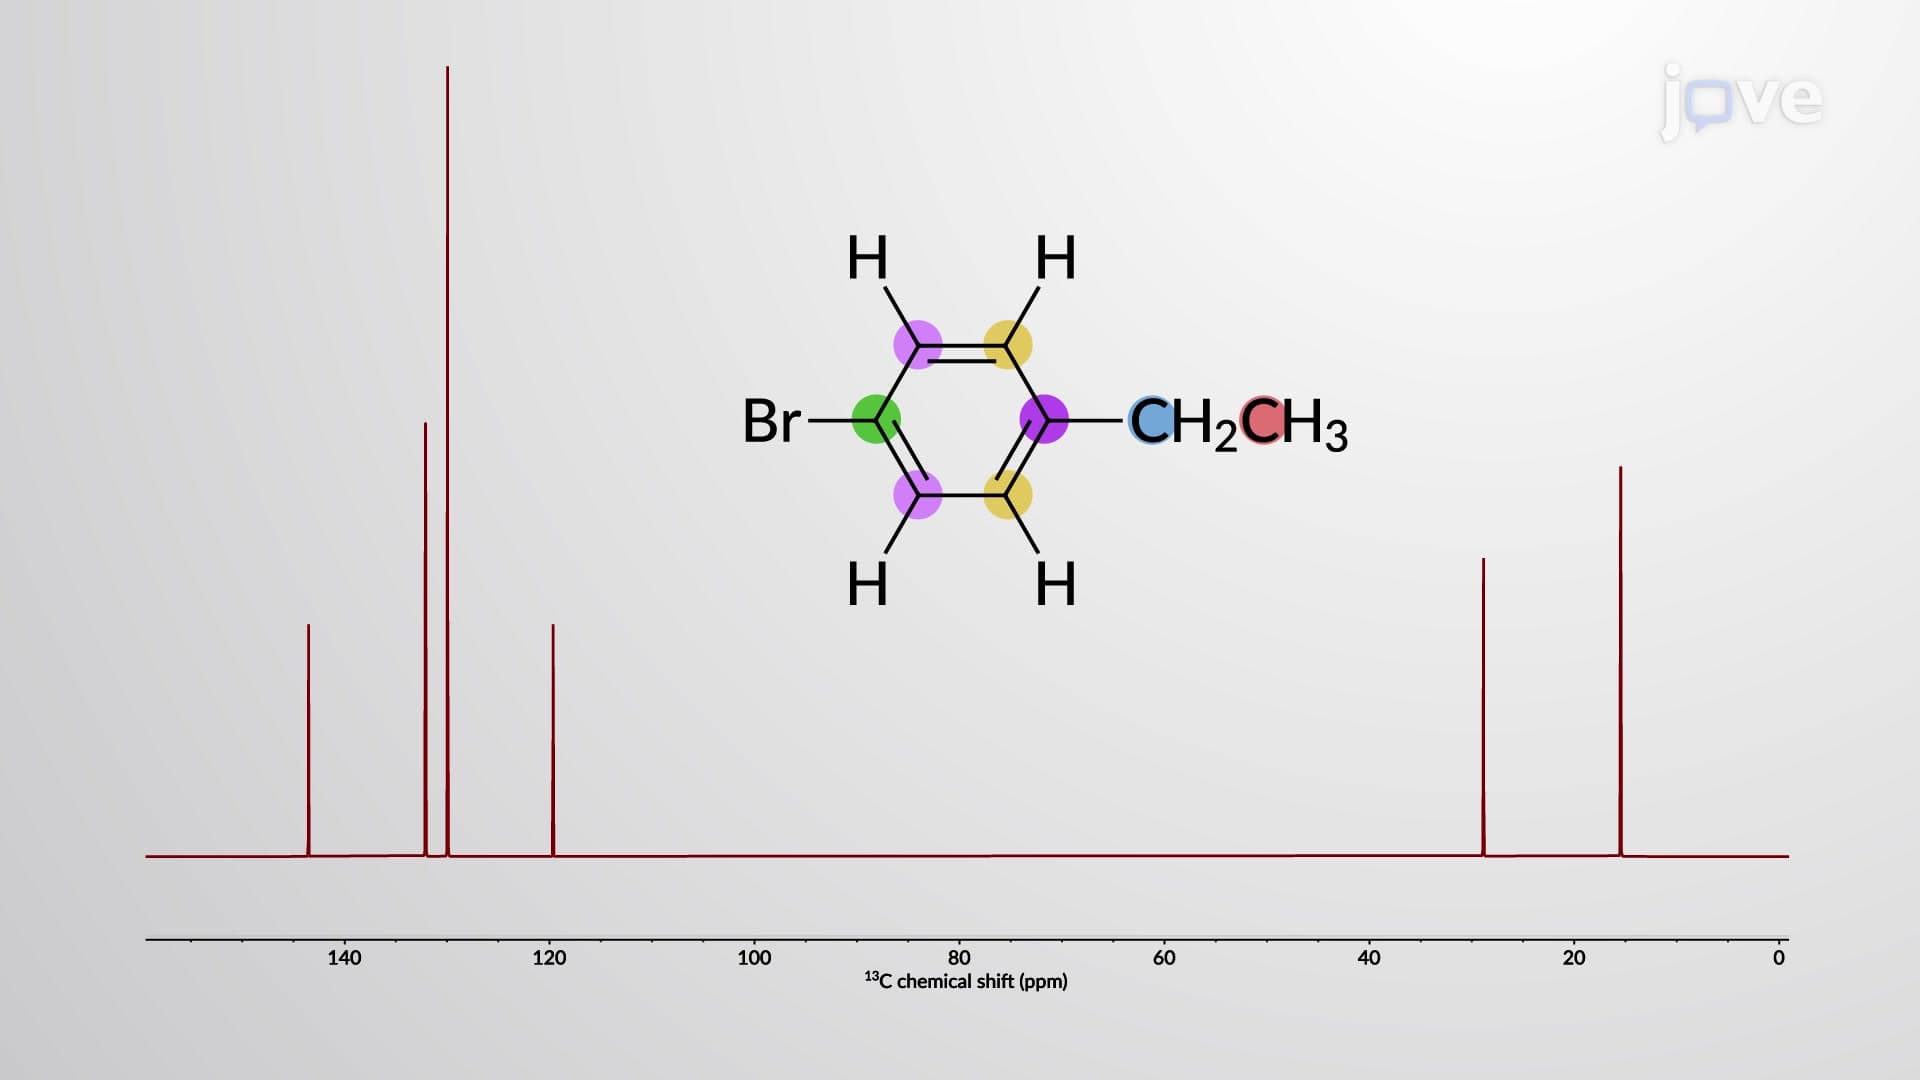
Simple unsubstituted benzene has six aromatic protons, all chemically equivalent. Therefore, benzene exhibits only a singlet peak at δ 7.3 ppm in the 1H NMR spectrum. The observed shift is far downfield because the aromatic ring current strongly deshields the protons. Any substitution on the benzene ring makes the aromatic protons nonequivalent, and the protons split each other. The peak is, therefore, no longer a singlet and the splitting pattern and their associated coupling constants depend...
 03:51
Molecular Orbital Theory II
Molecular Orbital Energy Diagrams
 01:18
Directing and Steric Effects in Disubstituted Benzene Derivatives
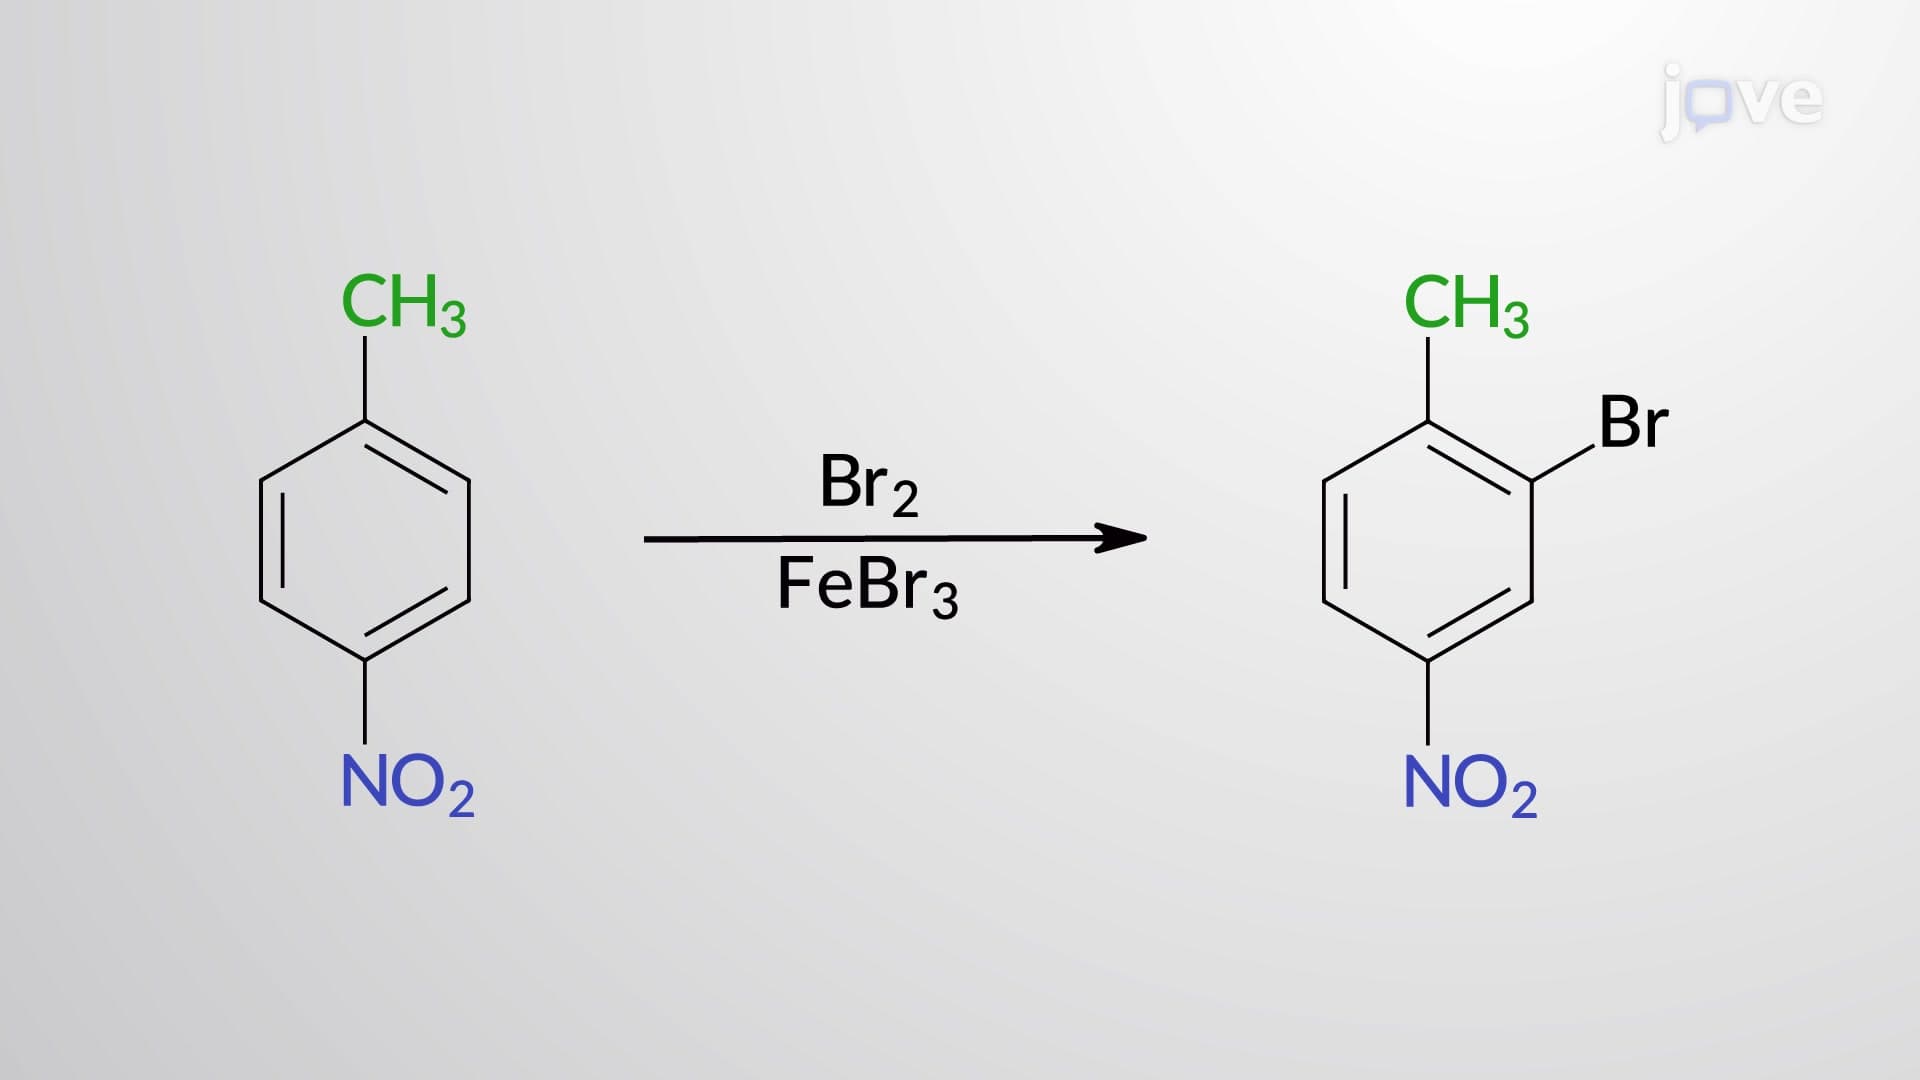
When disubstituted benzenes undergo electrophilic substitution, the product distribution depends on the directing effect of both substituents. When the directing effects of both substituents reinforce each other, a single product is obtained. For example, bromination of p-nitrotoluene occurs ortho to the methyl group and meta to the nitro group, which is the same position, resulting in a single product. However, if the directing effects of the two groups oppose each other, the more strongly...
 01:24
π Molecular Orbitals of 1,3-Butadiene
Conjugated dienes have lower heats of hydrogenation than cumulated and isolated dienes, making them more stable. The enhanced stabilization of conjugated systems can be understood from their π molecular orbitals.
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...