甲在初始潜在能量表面上的光解离动态的三态轨迹表面跳跃研究
Bina Fu1, Benjamin C Shepler, Joel M Bowman
1Department of Chemistry and Cherry L. Emerson Center for Scientific Computation, Emory University, Atlanta, Georgia 30322, USA.

本研究详细介绍了使用高级表面跳跃计算的甲光解离动态. 它揭示了一种新的异构化途径,并提供了关于激素和分子产物形成的见解.
科学领域:
- 化学动力学 化学动力学
- 摄影化学的使用.
- 理论化学 理论化学
背景情况:
- 甲光解离是大气化学中的一个关键过程.
- 了解激发电子状态的动态对于预测反应路径至关重要.
研究的目的:
- 为了研究甲的全维,三态光解离动力学.
- 探索电子状态S(1),T(1) 和S(0) 的初始潜在能量表面 (PES) 和旋转轨道合.
- 阐明反应机制,产品分布和能量分割.
主要方法:
- 全维,三态表面跳跃计算.
- 在S(0),S(1和T(1) 状态中使用初始潜在能量表面 (PES).
- 在S(1) 发起的研究中首次包含了ab initio计算的旋转轨道合.
主要成果:
- 计算了取决于时间的电子状态群体和根基和分子产品的分支比.
- 在T(1) 状态上通过S(0) 状态确定了一种新的异构化途径.
- 报告了H + HCO产品的转换能量分布和H (2) 产品的波动能量分布.
- 观察到一个漫游路径,导致低旋转CO和高振动H2种群.
结论:
- 该研究提供了对甲光解离动态的全面了解.
- 新的异构化路径和观察到的漫游动态为反应机制提供了新的见解.
- 结果与之前的轨迹计算一致,验证了采用的方法.
更多相关视频
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
相关概念视频
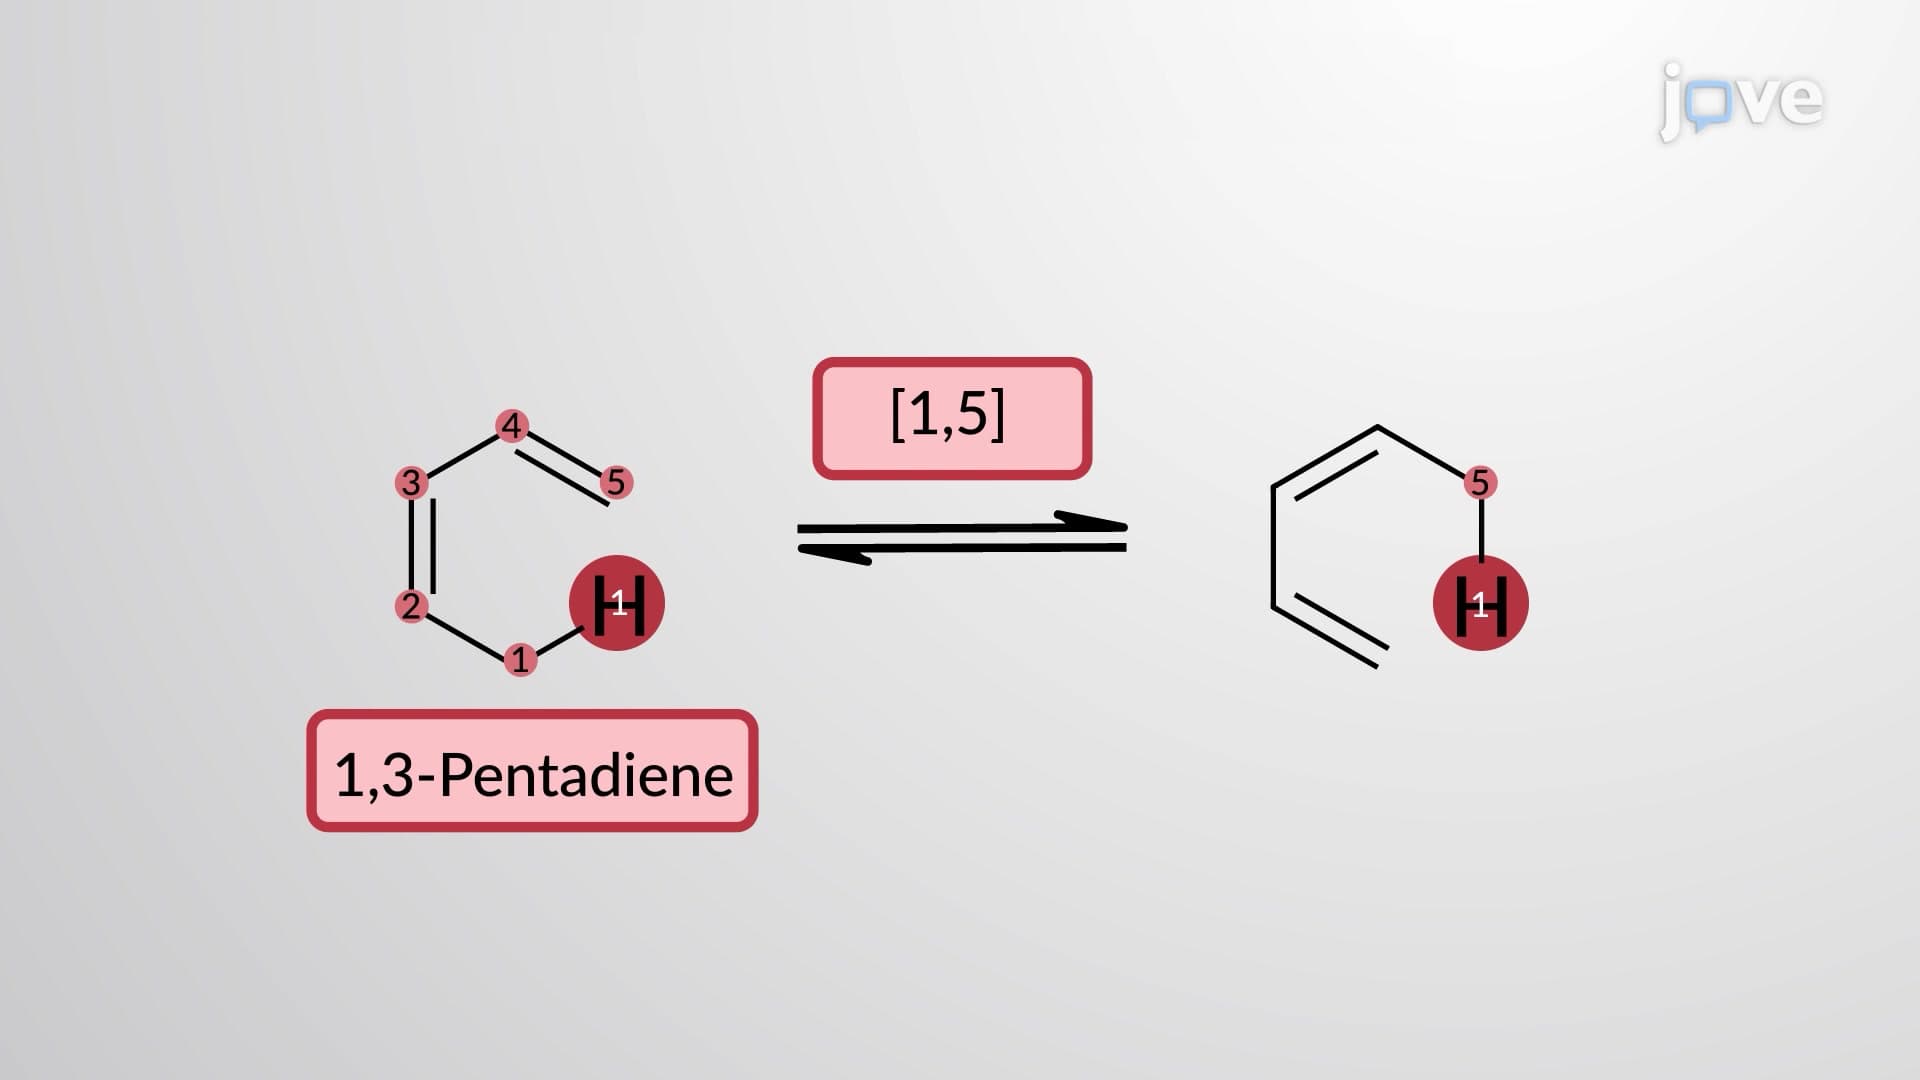 01:16
Thermal Sigmatropic Reactions: Overview
Sigmatropic shifts are classified based on an order term [i, j ], where i and j indicate the number of atoms across which each end of the σ bond migrates. Below are examples of a [3,3] sigmatropic shift in 1,5-hexadiene, referred to as...
 02:13
Energy Diagrams, Transition States, and Intermediates
 01:25
Deactivation Processes: Jablonski Diagram
 01:14
Molecular Spectroscopy: Absorption and Emission
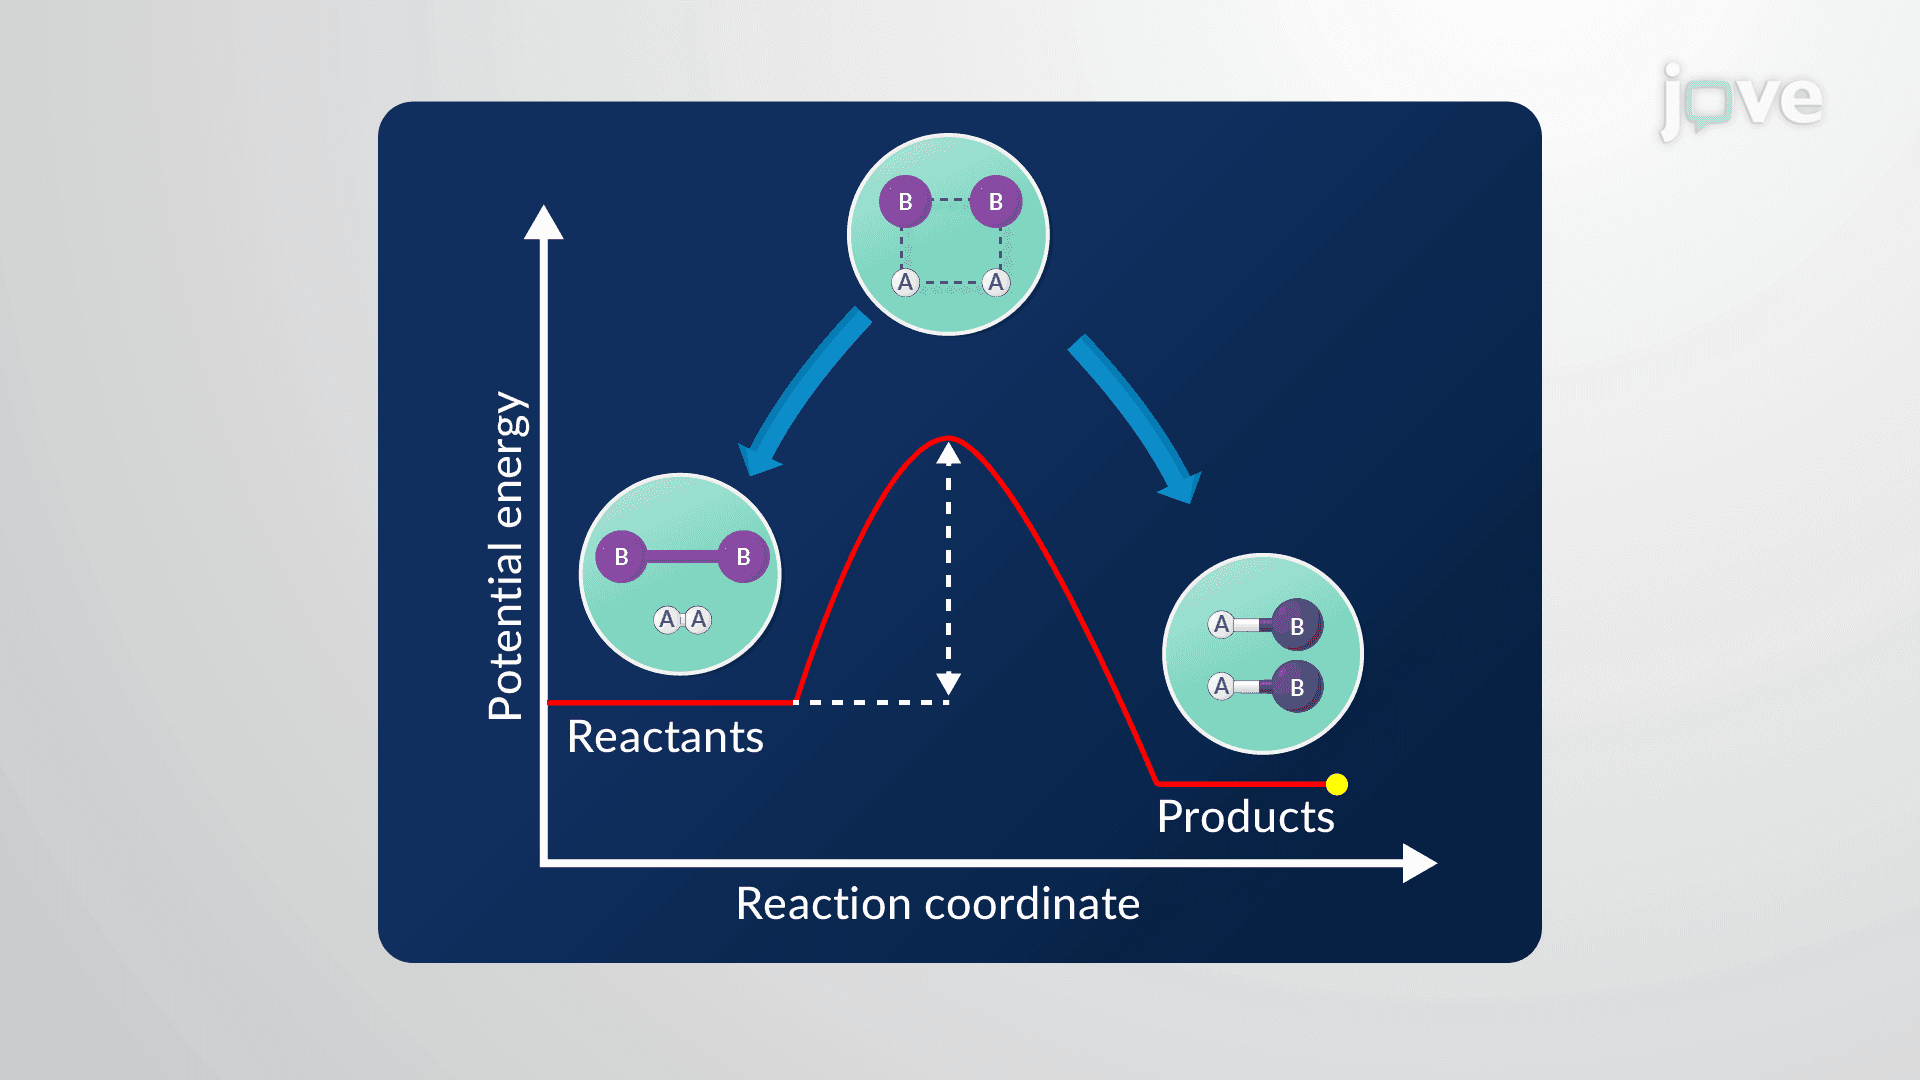 01:25
Transition State Theory
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions