在M2质子通道中的两个药物结合点上的自由能量计算
Ruo-Xu Gu1, Limin Angela Liu, Dong-Qing Wei
1State Key Laboratory of Microbial Metabolism, Luc Montagnier Biomedical Research Institute, and College of Life Sciences and Biotechnology, Shanghai Jiao Tong University, Shanghai Minhang District, China 200240.
Journal of the American Chemical Society
|June 30, 2011

概括
针对流感A M2通道的阿达曼坦类药物在孔隙内结合得更稳定,但需要更高的能量. C端表面为这些抗病毒药物提供了动力学上有利的结合点.
科学领域:
- 生物物理学的生物物理.
- 计算化学计算化学
- 病毒学 病毒学
背景情况:
- 流感A M2通道是阿达曼坦类药物的目标.
- 建议有两个药物结合点:毛孔和C端表面.
- 了解结合机制对于药物耐药性和疗效至关重要.
研究的目的:
- 研究阿达曼坦类药物与M2通道结合的热力学和动力学方面.
- 确定药物相互作用的首选结合点和能量景观.
- 提供一个框架来协调相互矛盾的实验和计算发现.
主要方法:
- 分子动力学模拟.分子动力学模拟.
- 平均力计算的潜力.
- 在生物pH值下分析M2-脂质双层系统.
主要成果:
- 孔隙结合部位在热力学上更有利 (~7 kcal/mol) 于稳定的药物结合和通道抑制.
- 与C端表面相比,与孔隙结合需要显著更高的能量屏障 (~10kcal/mol).
- 在动力学上,C端表面结合部位更有利.
结论:
- M2通道孔提供了一个更稳定的结合点,解释了药物的疗效.
- 动力学上有利的表面位置可能在药物耐药性中起作用.
- 这项研究提供了M2药物相互作用的统一观点,有助于未来的药物设计.
相关概念视频
 01:22
Protein-Drug Binding: Determination Methods
Determining protein-drug binding can be achieved through indirect and direct methods, each providing valuable insights into the interaction between proteins and drugs.
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 02:18
The Equilibrium Binding Constant and Binding Strength
The equilibrium binding constant (Kb) quantifies the strength of a protein-ligand interaction. Kb can be calculated as follows when the reaction is at equilibrium:
 01:16
Protein-Drug Binding: Mechanism and Kinetics
Protein-drug binding refers to the interaction between drugs and proteins within the body. This binding process can occur intracellularly, involving drug interactions with enzymes or receptors within cells, or extracellularly, involving plasma proteins in the blood.
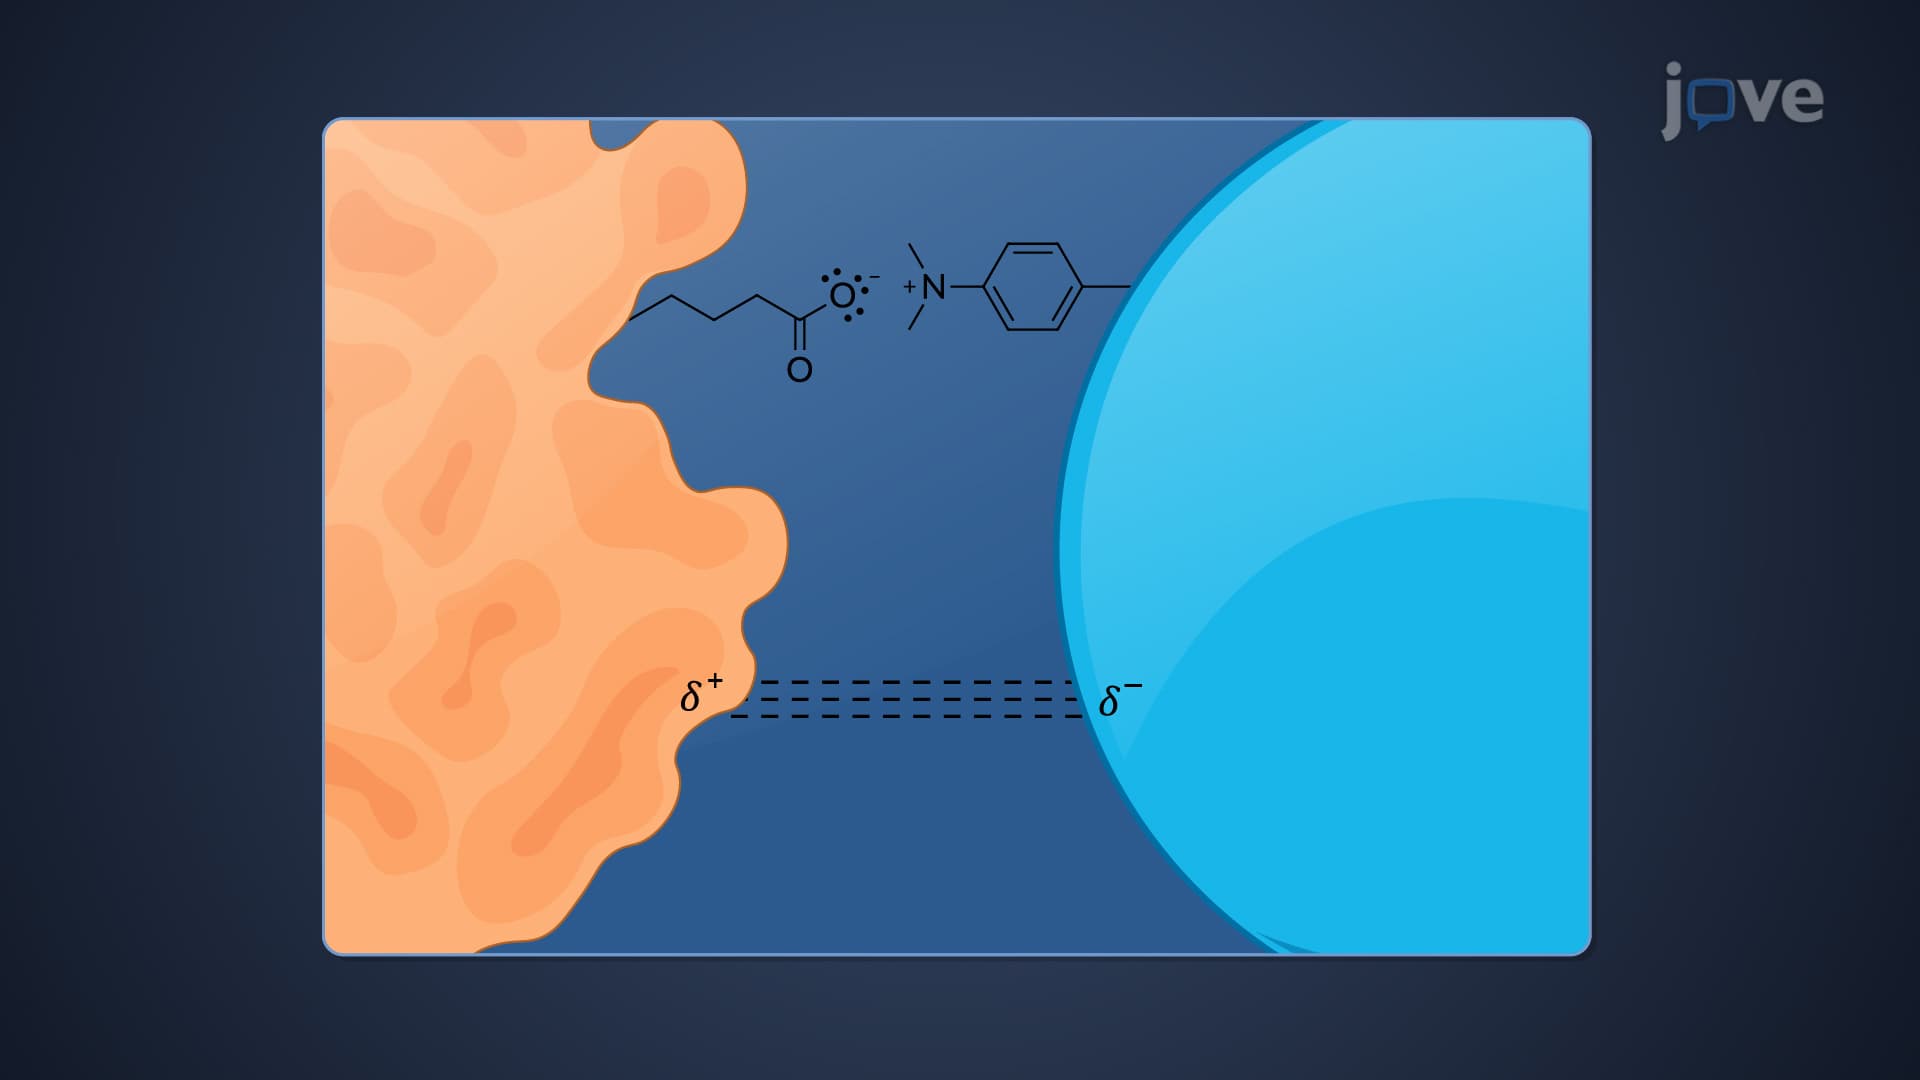
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:29
The Two-State Receptor Model
The two-state receptor model explains a drug's interaction with receptors, such as G protein-coupled receptors and ligand-gated ion channels, to induce or inhibit a biological response. When no natural ligands are present, a receptor exists in an equilibrium of inactive (Ri) and active (Ra) conformations. The inactive form does not produce a response, while the active form generates a basal effect known as constitutive activity.
The binding affinity of a drug determines its interaction with one...
The binding affinity of a drug determines its interaction with one...
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
Physiological models with protein binding in pharmacokinetics offer a sophisticated approach to understanding drug disposition. These models consider drug-protein interactions, enabling them to effectively predict drug concentrations in different organs and tissues. This precision aids in accurate drug dosing, providing a significant advantage over conventional models. A key process within these models is equilibration, which ensures that drug concentrations achieve a steady state within the...
 01:30
Quantitative Aspects of Drug-Receptor Interaction
The receptor occupancy theory connects a drug's response to the number of occupied receptors. With higher drug concentrations, more receptors are occupied, leading to increased responses. The formation of drug-receptor complexes involves association and dissociation rates, which reach equilibrium when the forward and backward reactions are equal. The equilibrium association constant (Ka) and its inverse, the equilibrium dissociation constant (Kd), indicate drug affinity. Higher Ka and lower Kd...