相关实验视频
Updated: Apr 21, 2026
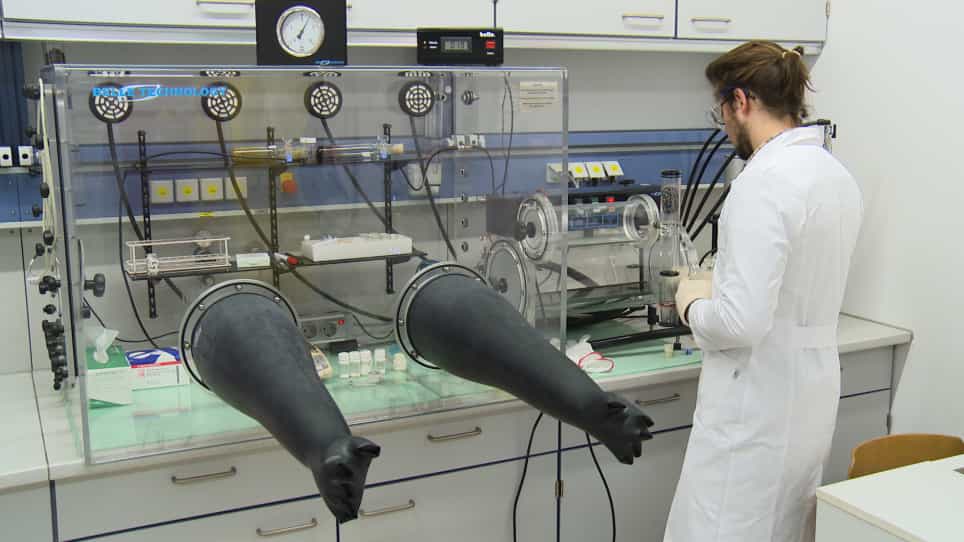

Isolating Free Carbenes, their Mixed Dimers and Organic Radicals
Published on: April 19, 2019
作为主要组元素结合的硬币金属:碳化合物复合物的结构和结合[TM(cAAC) ]和[TM(cAAC) ] (TM = Cu, Ag, Au) (TM = Cu, Ag, Au)
Paul Jerabek1, Herbert W Roesky, Guy Bertrand
1Fachbereich Chemie, Philipps-Universität Marburg , Hans-Meerwein-Strasse, D-35032 Marburg, Germany.

量子化学计算显示,周期性 (基) 氨基) 碳 (cAAC) 连接物在中形成更强的金属-连接物键[TM(cAAC) ]+) 比中性复合物[TM(cAAC) ]中形成更强的金属-连接物键. 相对论效应显著影响债券强度,黄金复合体在非相对论水平上显示较弱的债券.
科学领域:
- 无机化学 无机化学
- 量子化学 是一个量子化学.
- 计算化学的计算化学
背景情况:
- 循环 (基) 氨基) 碳素 (cAACs) 是有机金属化学中的多功能联体.
- 11组金属 (Cu,Ag,Au) 形成具有不同电子性质的复合物.
- 了解金属-连接体结合对于设计新材料和催化剂至关重要.
研究的目的:
- 为了研究中性[TM(cAAC) 2]和阴离子[TM(cAAC) 2] (TM = Cu,Ag,Au) 复合体中的金属连接.
- 阐明相对论对这些复合体电子结构和结合作用的作用.
- 为了比较中性和充电金属cAAC系统之间的粘合特性.
主要方法:
- 使用密度函数理论 (DFT) 的计算.
- 使用化学价值的自然轨道 (EDA-NOCV) 进行电荷和能量分解分析.
- 进行非相对论和相对论计算以进行比较.
主要成果:
- 在[TM(cAAC) 2+) 中的金属碳键较长,但具有比[TM(cAAC) 2更高的键解离能.
- 在中性复合体中,由于π回捐,cAAC连接体作为比σ捐赠体更强的π接受体而不是σ捐赠体.
- 相对论效应削弱TM-C债券,特别是黄金,改变债券强度顺序.
- 非相对论计算高估了保利排斥,低估了轨道相互作用.
结论:
- 在[TM(cAAC) 2和[TM(cAAC) 2+) 中的结合有显著差异,其中体表现出更强的内在相互作用.
- 相对论效应对于准确描述11组重金属复合体中的结合至关重要.
- 该EDA-NOCV方法提供了比传统的NBO分析更深入地了解金属连接体相互作用.
更多相关视频
 09:45
Accessing Valuable Ligand Supports for Transition Metals: A Modified, Intermediate Scale Preparation of 1,2,3,4,5-Pentamethylcyclopentadiene
Published on: March 20, 2017
 19:58
Palladium N-Heterocyclic Carbene Complexes: Synthesis from Benzimidazolium Salts and Catalytic Activity in Carbon-carbon Bond-forming Reactions
Published on: July 30, 2017
相关概念视频
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group...
 02:10
Carbocations
 01:07
Radicals: Electronic Structure and Geometry
Accordingly, the structure of a trivalent radical lies between the geometries of carbocations and carbanions. An sp2-hybridized carbocation is trigonal planar, while an sp3-hybridized carbanion is trigonal pyramidal. Here, the difference in geometry is...
 02:42
Valence Bond Theory
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than...
 01:23
Properties of Organometallic Compounds