

用光解密过渡状态:光异构化作为布伦斯特酸催化中的机制工具
Polyssena Renzi1, Johnny Hioe1, Ruth M Gschwind1
1Institut für Organische Chemie, Universität Regensburg , D-93040 Regensburg, Germany.
Journal of the American Chemical Society
|May 6, 2017

概括
这项研究引入了一种基于光的新方法 (DTS-hν),用于实验性地探测酸催化中的过渡状态. 这种方法揭示了核友性添加伊米因的不同反应途径和反选择性指纹.
科学领域:
- 有机化学
- 催化剂
- 反应机制
背景情况:
- 选择性布伦斯特酸催化被广泛使用,但缺乏实验性过渡状态的洞察力.
- 机械学的理解往往严重依赖于理论计算.
研究的目的:
- 开发一种实验方法,DTS-hν,用于阐明酸催化反应中的过渡状态.
- 调查相互竞争的反应途径及其对酶选择性的影响.
主要方法:
- 使用光 (365 nm LED) 作为机械探测器的二次链的光异构化.
- 分析反应速率和酶选择性的特征变化,以确定活性过渡状态.
- 将实验结果与量子化学计算和非共价相互作用分析相关联.
主要成果:
- 在催化过程中证明了因明双键的光诱导同质化,产生了独特的反应动态指纹.
- 鉴定了I型Z和II型Z途径在ketimine不对称转移化中占主导地位.
- 已发现I型E和II型E途径在酸乙与阿尔迪明的核友添加中是活跃的.
- 在胺降解中观察到反应速度显著加快,高达177%.
结论:
- DTS-hν方法为过渡状态结构和竞争途径提供了直接的实验证据.
- 光诱导的异构化是解剖复杂的催化机制的强大工具.
- 这种方法提供了一个对理论计算的补充实验策略,以了解反选择性催化.
相关概念视频
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration
9.8K
The rate of acid-catalyzed hydration of alkenes depends on the alkene's structure, as the presence of alkyl substituents at the double bond can significantly influence the rate.
9.8K
 02:36
Regioselectivity and Stereochemistry of Hydroboration
9.6K
A significant aspect of hydroboration–oxidation is the regio- and stereochemical outcome of the reaction.
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn stereochemistry.
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn stereochemistry.
9.6K
 01:26
Photochemical Electrocyclic Reactions: Stereochemistry
2.3K
The absorption of UV–visible light by conjugated systems causes the promotion of an electron from the ground state to the excited state. Consequently, photochemical electrocyclic reactions proceed via the excited-state HOMO rather than the ground-state HOMO. Since the ground- and excited-state HOMOs have different symmetries, the stereochemical outcome of electrocyclic reactions depends on the mode of activation; i.e., thermal or photochemical.
Selection Rules: Photochemical Activation
Selection Rules: Photochemical Activation
2.3K
 02:35
Regioselectivity of Electrophilic Additions-Peroxide Effect
11.3K
In the presence of organic peroxides, the addition of hydrogen bromide to an alkene yields the isomer that is not predicted by Markovnikov’s rule. For example, the addition of hydrogen bromide to 2-methylpropene in the presence of peroxides gives 1-bromo-2-methylpropane. This addition reaction proceeds via a free radical mechanism, which reverses the regioselectivity. The free radical reaction mechanism involves three stages: initiation, propagation, and termination.
11.3K
 01:14
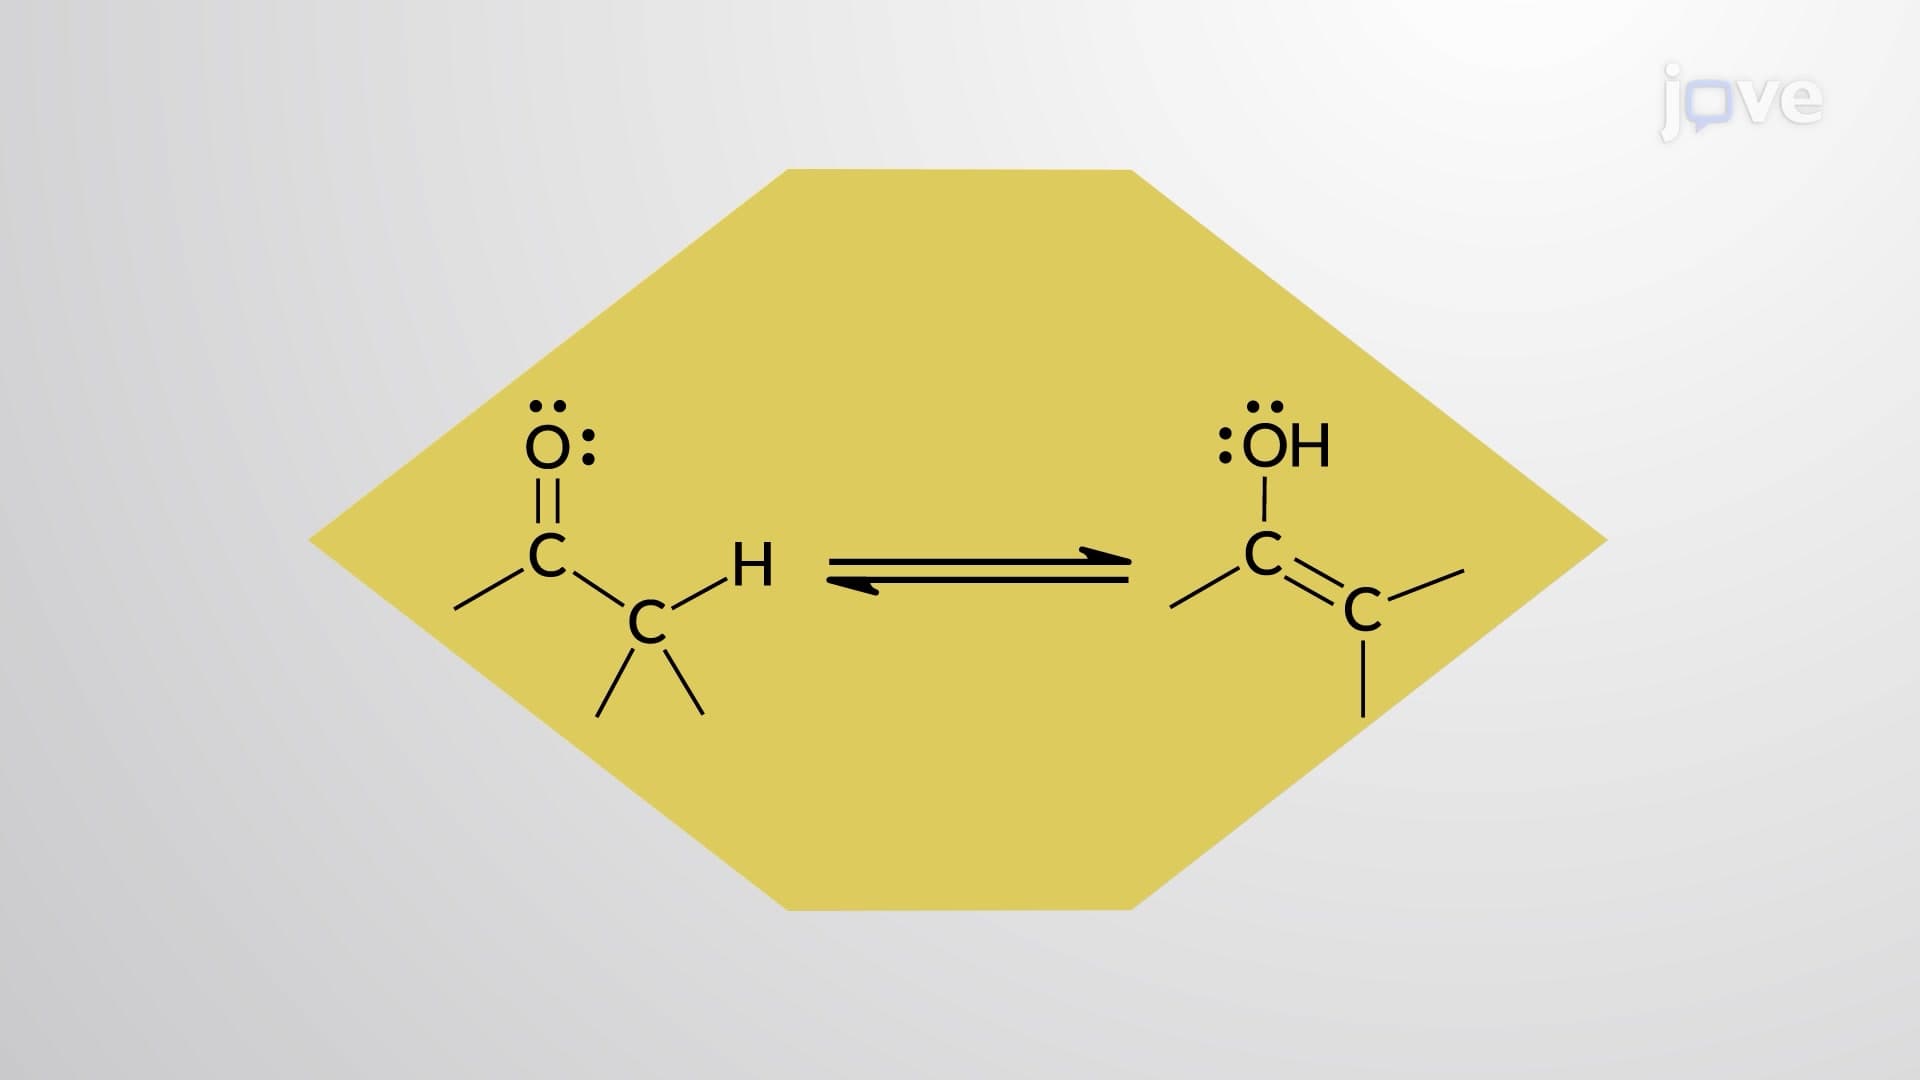
Keto–Enol Tautomerism: Mechanism
8.0K
The keto and enol forms are known as tautomers and they constantly interconvert (or tautomerize) between the two forms under acid or base catalyzed conditions. Both the reactions involve the same steps—protonation and deprotonation— although in the reverse order.
8.0K
 02:17
Reduction of Alkenes: Asymmetric Catalytic Hydrogenation
3.9K
Catalytic hydrogenation of alkenes is a transition-metal catalyzed reduction of the double bond using molecular hydrogen to give alkanes. The mode of hydrogen addition follows syn stereochemistry.
The metal catalyst used can be either heterogeneous or homogeneous. When hydrogenation of an alkene generates a chiral center, a pair of enantiomeric products is expected to form. However, an enantiomeric excess of one of the products can be facilitated using an enantioselective reaction or an...
The metal catalyst used can be either heterogeneous or homogeneous. When hydrogenation of an alkene generates a chiral center, a pair of enantiomeric products is expected to form. However, an enantiomeric excess of one of the products can be facilitated using an enantioselective reaction or an...
3.9K