

6D-QSAR用于预测人类阿尔多减少酶抑制剂的生物活性,使用类星受体表面建模
Babak Sokouti1, Maryam Hamzeh-Mivehroud2,3
1Biotechnology Research Center, Tabriz University of Medical Sciences, Tabriz, Iran.
BMC chemistry
|June 22, 2023

概括
多维定量结构-活性关系 (QSAR) 模型有助于药物设计. 应用在人类阿尔多缩酶抑制剂上的更高维度6D-QSAR模型,与3D-QSAR模型相比,显示出更高的预测性能.
科学领域:
- 药用化学 医学化学
- 计算化学的计算化学
- 药物设计 药物设计
背景情况:
- 定量结构-活性关系 (QSAR) 分析是合理药物设计的基石,其历史跨越了五十年.
- 多维QSAR建模为开发用于设计新型化学化合物的预测模型提供了强大的方法.
研究的目的:
- 开发和比较人类阿尔多缩酶 (AR) 抑制剂的多维QSAR模型 (3D-QSAR和6D-QSAR).
- 通过使用解离常数 (Kd) 值来评估不同维的QSAR模型的预测性能.
主要方法:
- 使用Pentacle和Quasar软件生成3D和6D-QSAR模型.
- 使用解离常数 (Kd) 值作为模型开发的终点.
- 使用内部和外部验证指标评估模型性能.
主要成果:
- 三维和六维QSAR模型都显示了可比的内部验证统计数据.
- 外部验证显示,6D-QSAR模型在预测终点值方面明显优于3D-QSAR模型.
- 一个趋势表明,增加的QSAR模型维度与增强的预测性能相关.
结论:
- 与3D-QSAR相比,6D-QSAR模型对人类阿尔多缩酶抑制剂的预测精度更高.
- QSAR模型的维度似乎是影响预测性能的关键因素.
- 需要进一步的研究来验证这些发现在各种化学数据集和生物标之间.
更多相关视频
 00:05
In Silico Modeling Method for Computational Aquatic Toxicology of Endocrine Disruptors: A Software-Based Approach Using QSAR Toolbox
Published on: August 28, 2019
14.0K
 08:31
Biosensor-based High Throughput Biopanning and Bioinformatics Analysis Strategy for the Global Validation of Drug-protein Interactions
Published on: December 1, 2020
5.1K
相关概念视频
 01:28
Structure-Activity Relationships and Drug Design
794
Drug design is a dynamic field that involves discovering and developing new medications based on specific biological targets. This process heavily relies on structure-activity relationships (SAR) and quantitative structure-activity relationships (QSAR) to guide the design and optimization of efficient drugs.
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
794
 01:30
Quantitative Aspects of Drug-Receptor Interaction
1.1K
The receptor occupancy theory connects a drug's response to the number of occupied receptors. With higher drug concentrations, more receptors are occupied, leading to increased responses. The formation of drug-receptor complexes involves association and dissociation rates, which reach equilibrium when the forward and backward reactions are equal. The equilibrium association constant (Ka) and its inverse, the equilibrium dissociation constant (Kd), indicate drug affinity. Higher Ka and lower...
1.1K
 01:15
G Protein-coupled Receptors
12.3K
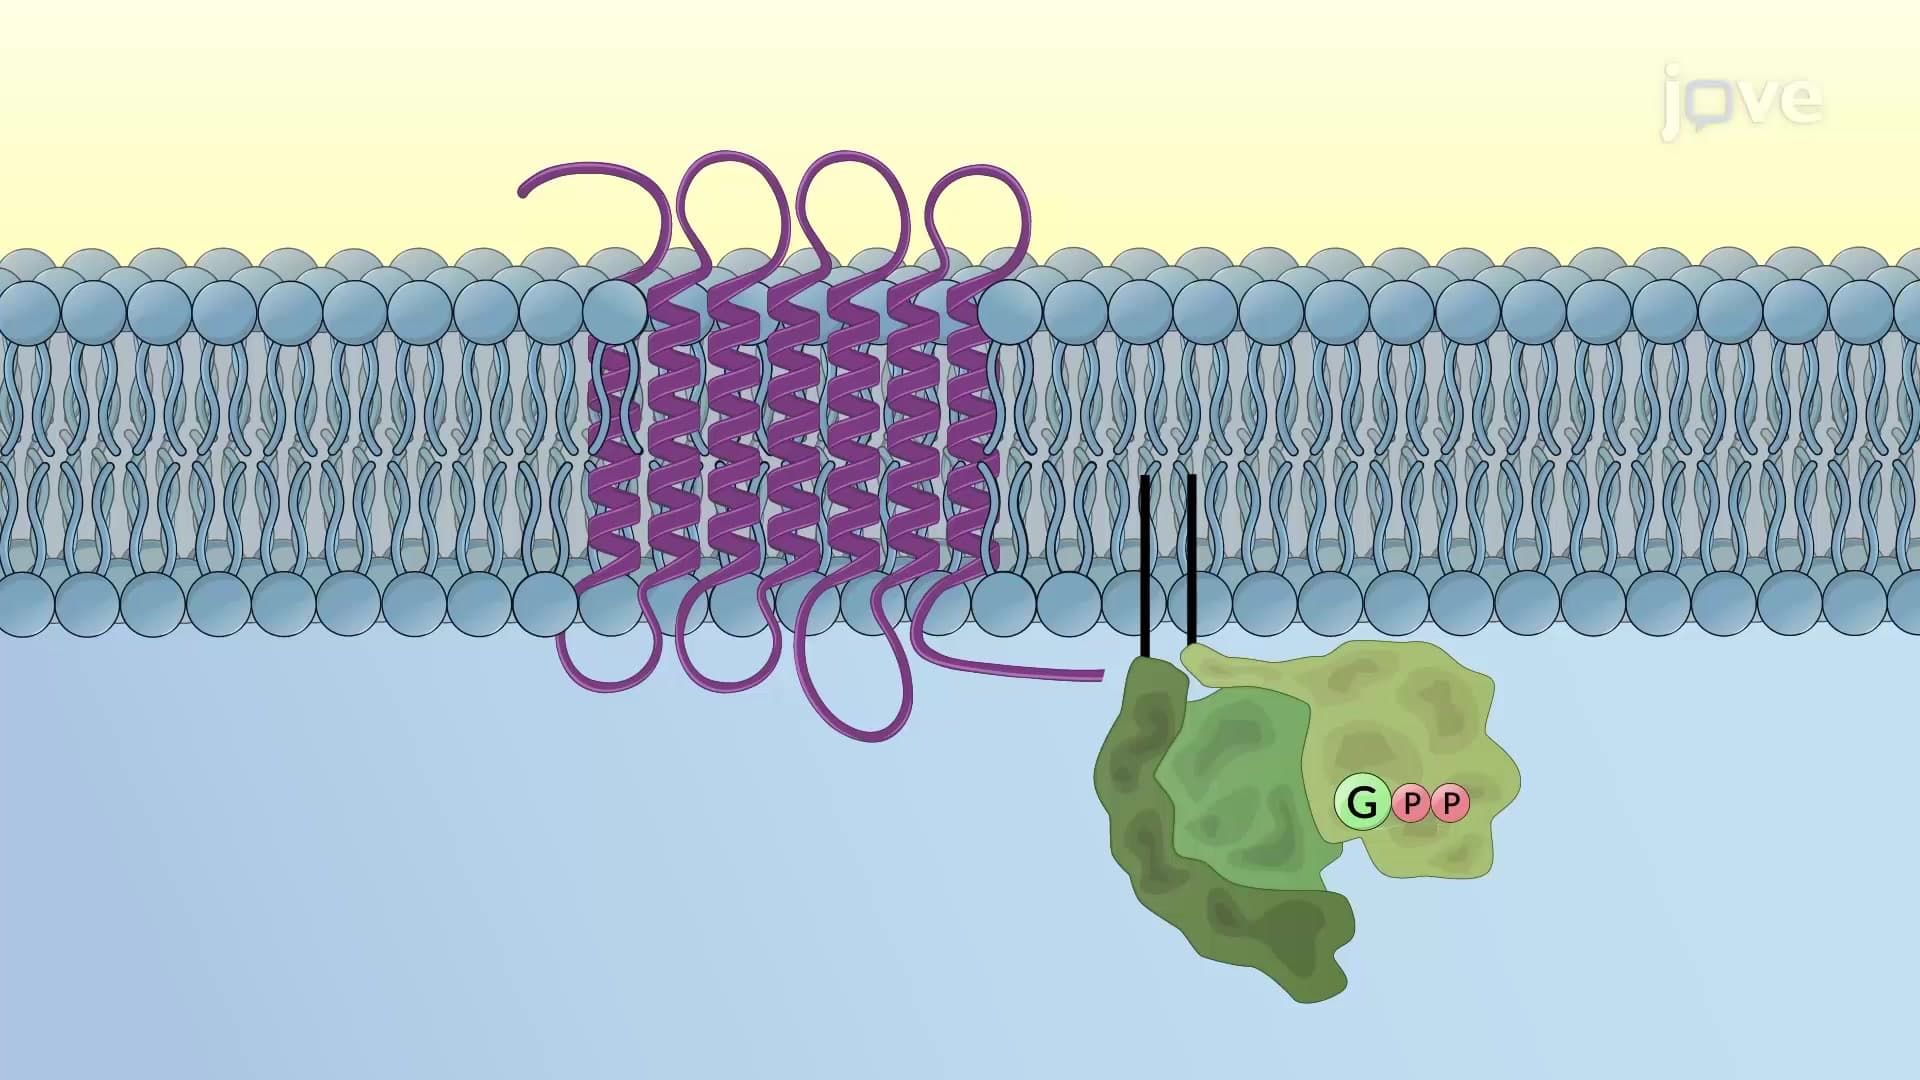
G Protein-Coupled Receptors or GPCRs are membrane-bound receptors that transiently associate with heterotrimeric G proteins and induce an appropriate response to sensory stimuli such as light, odors, hormones, cytokines, or neurotransmitters.
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...
12.3K