
在数据依赖性获取中对质谱参数的优化,用于基于假定赋值的非目标代谢学
Hailemariam Abrha Assress1,2, Mario G Ferruzzi1,2, Renny S Lan1,2
1Arkansas Children's Nutrition Center, Little Rock, Arkansas 72202, United States.

概括
优化质谱学参数,如MS/MS事件和信号值,显著提高了在非目标代谢学中的代谢物识别. 本研究提供了改善复杂生物样本中代谢物覆盖率和光谱质量的策略.
科学领域:
- 分析化学 分析化学
- 生物化学 生化学
- 质谱测量质量谱测量
背景情况:
- 非定位代谢学依赖于全面的代谢物识别.
- 数据依赖采集 (DDA) 质谱需要仔细的参数优化,以获得最大的MS/MS覆盖范围.
- 准确的代谢物注释对于理解生物途径至关重要.
研究的目的:
- 研究各种质谱参数对非目标代谢学中的代谢物注释的影响.
- 确定Exploris 480-Orbitrap质谱仪的最佳设置,以改善代谢物识别.
- 提出提高代谢物覆盖率和光谱质量的策略.
主要方法:
- 系统地探索质谱参数,包括质分辨率,射频水平,信号强度值,MS/MS事件数量,周期时间,碰撞能量,最大离子注入时间 (MIT),动态排除和自动增强控制 (AGC) 目标值.
- 在数据依赖采集 (DDA) 实验中使用了Exploris 480-Orbitrap质谱仪.
- 评估了参数变化对代谢物注释率的影响.
主要成果:
- 通过十次MS/MS扫描,2.0m/z隔离窗口,1x10^4信号强度值,180,000个MS分辨率和30,000个MS/MS分辨率在70%的射频水平上,实现了最佳的注释.
- 将5x10^6 (MS) 和1x10^5 (MS/MS) 的AGC目标值与100 ms (MS) 和50 ms (MS/MS) 的MIT相结合,改善了代谢物注释.
- 一个10秒的排除时间和两阶段碰撞能量优化了光谱质量.
结论:
- 质谱参数显著影响代谢学结果,影响代谢物覆盖率和识别信心.
- 该研究为DDA实验提供了优化的参数,以增强在非目标代谢学中的代谢物识别.
- 建议使用真实标准进行进一步验证,并在不同的协议上进行测试.
更多相关视频
 08:23
2 in 1: One-step Affinity Purification for the Parallel Analysis of Protein-Protein and Protein-Metabolite Complexes
Published on: August 6, 2018
11.4K
 11:00
Untargeted Metabolomics from Biological Sources Using Ultraperformance Liquid Chromatography-High Resolution Mass Spectrometry UPLC-HRMS
Published on: May 20, 2013
22.6K
相关概念视频
 01:33
Peptide Identification Using Tandem Mass Spectrometry
6.6K
Tandem mass spectrometry, also known as MS/MS or MS2, is an analytical technique that employs two mass analyzers. Essentially it is a series of mass spectrometers that helps isolate a particular biomolecule and then helps study its chemical properties.
This technique helps gather information regarding the protein from which the peptide was obtained and to study the peptides’ amino acid sequence. Identifying peptides from a complex mixture is an important component of the growing field of...
This technique helps gather information regarding the protein from which the peptide was obtained and to study the peptides’ amino acid sequence. Identifying peptides from a complex mixture is an important component of the growing field of...
6.6K
 01:19
Mass Spectrometry: Overview
5.4K
Mass spectrometry is an analytical technique used to determine the molecular mass and molecular formula of a compound. The basic principle of mass spectrometry is to generate ions from the analyte molecule and measure these ion abundances against their molecular mass. One common type of ionization, known as electrospray ionization or EI, bombards the analyte molecules in the gas phase with high-energy electron beams. The electron beams displace an electron from the molecule and leave...
5.4K
 01:21
Mass Spectrometry: Complex Analysis
829
Mass spectrometry is an important technique for the identification of pure compounds. However, it has some limitations for the analysis of complex mixtures, often due to excessive fragmentation making the spectrum too complicated to decipher. Mass spectrometry can be combined with suitable separation methods in sequence, forming hyphenated methods, which are useful in the analysis of complex mixtures.
GC–MS is a powerful hyphenated method commonly used in forensics and environmental...
GC–MS is a powerful hyphenated method commonly used in forensics and environmental...
829
 01:15
High-Resolution Mass Spectrometry (HRMS)
1.4K
The resolution of a mass spectrometer depends on the efficiency of separating ions with different ion masses. The mass of an atom is approximated to the sum of the masses of protons and neutrons inside, considering the masses of protons and neutrons as equal. However, the masses of the proton (1.6726 × 10⁻²⁴ g) and neutron (1.6749 × 10⁻²⁴ g) are not truly equal. There is a minor error in the expression of atomic masses relative to the simplest atom of hydrogen. For...
1.4K
 01:16
Mass Spectrometers
5.7K
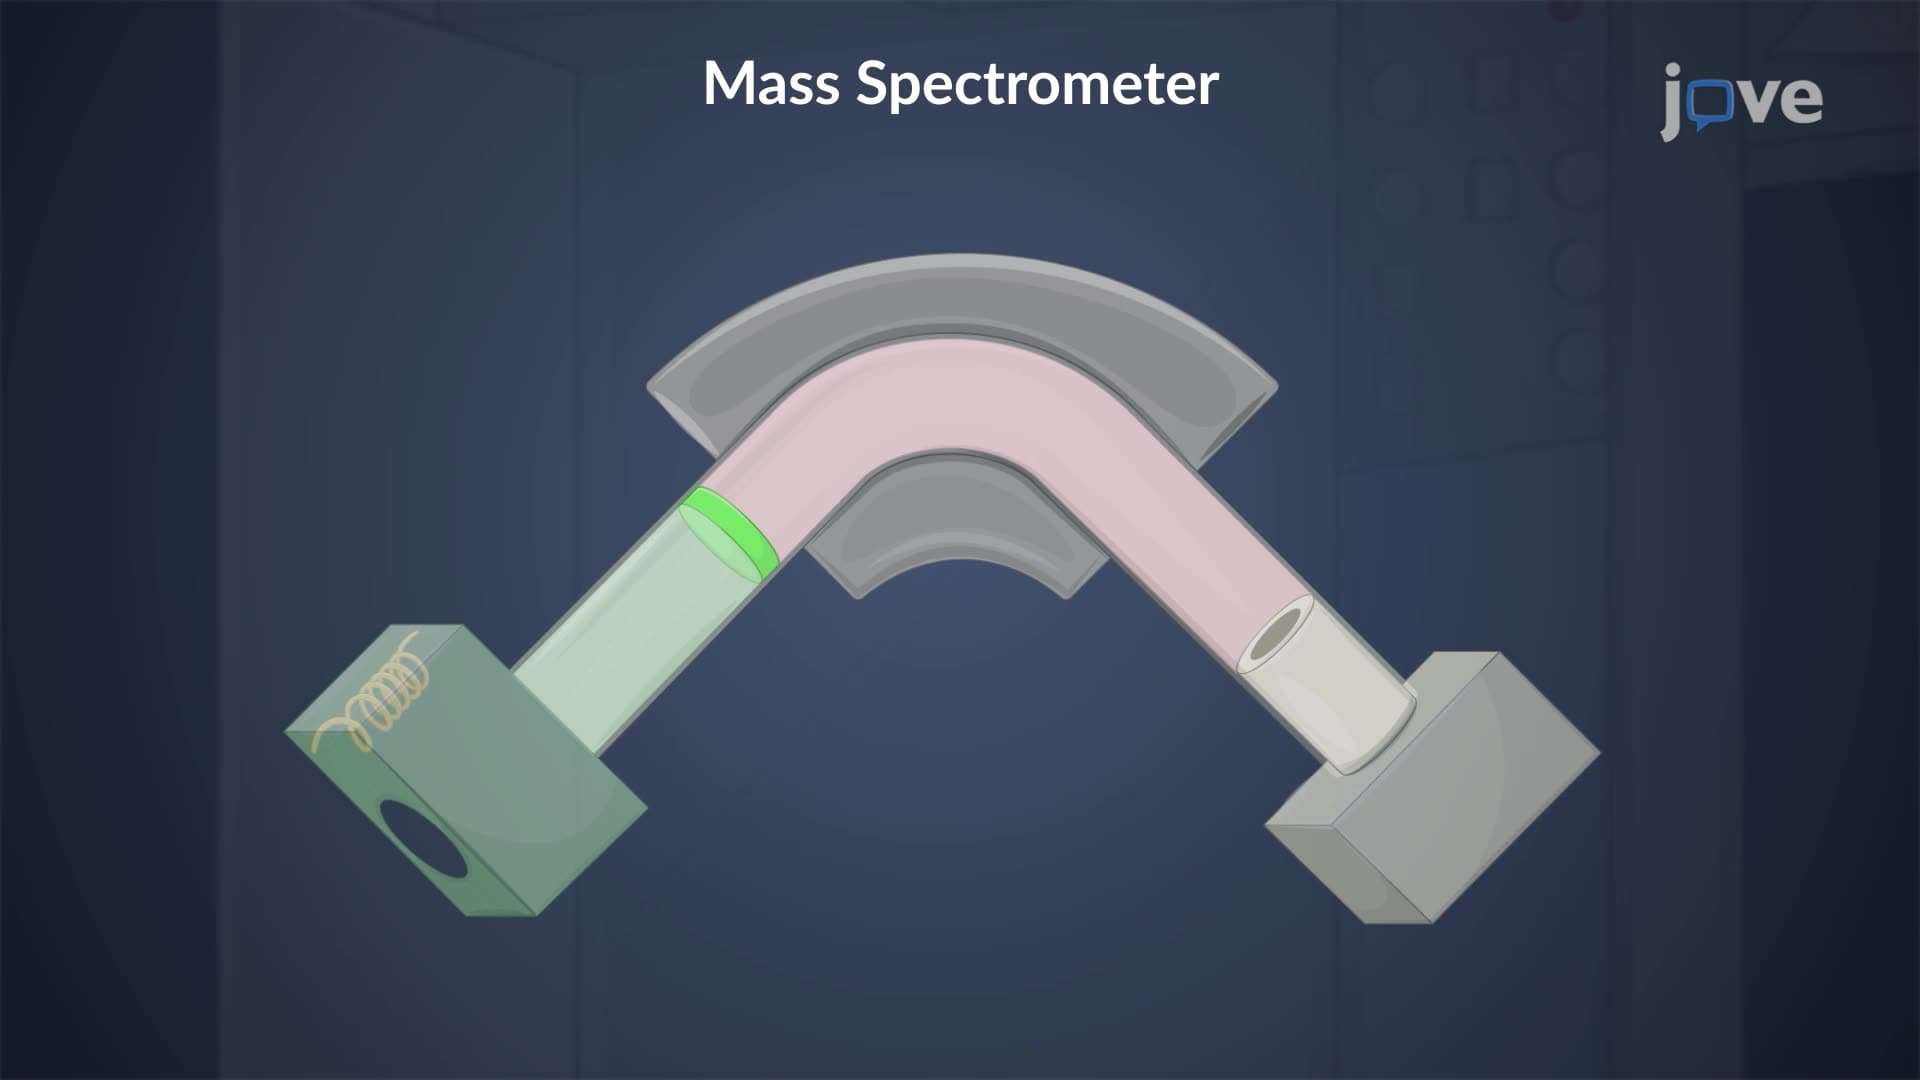
This lesson details the instrumentation of a mass spectrometer—a physical instrument to perform mass spectrometry on analyte molecules and record the characteristic mass spectra. This is achieved via three chief functions:
5.7K