
CodingDiv:分析SNP水平的微生物多样性,以区分病毒基因组中的编码和非编码区域
Eric Olo Ndela1, François Enault1
1Université Clermont Auvergne, CNRS, LMGE, F-63000 Clermont-Ferrand, France.
Bioinformatics (Oxford, England)
|July 14, 2023

概括
预测病毒基因是一个挑战. CodingDiv通过检测潜在编码区域的SNP级微生物多样性来识别蛋白质编码区域,帮助精确的病毒基因预测.
科学领域:
- 基因组学就是基因组学.
- 生物信息学是一种生物信息学.
- 病毒学 病毒学
背景情况:
- 病毒基因预测是复杂的,因为基因大小小小和重叠.
- 准确识别所有病毒基因仍然是基因组学的一个重大挑战.
研究的目的:
- 介绍CodingDiv,这是一个用于预测病毒基因的新工具.
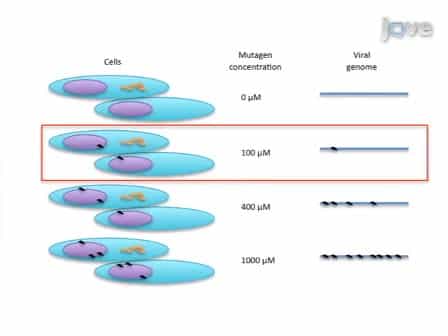
- 在病毒基因组的潜在编码区域中检测单核酸多态 (SNP) 级微生物多样性.
主要方法:
- 编码Div使用元基因组读取和外部序列数据库.
- 它分析SNP模式,区分同义词与非同义词替代.
- 蛋白质编码区域通过同义和非同义SNP的较高比例来识别.
主要成果:
- 在SNP层面上,CodingDiv有效地检测到微型多样性.
- 该工具有助于识别病毒基因组内的蛋白质编码区域.
- 这种方法提高了病毒基因预测的准确性.
结论:
- CodingDiv为准确预测病毒基因提供了一个强大的解决方案.
- 该工具利用SNP微多样性分析来增强基因识别.
- 这种方法有助于更好地了解病毒基因组组织.

相关概念视频
 02:26
Comparing Copy Number Variations and SNPs
17.8K
Sequencing of the human genome has opened up several best-kept secrets of the genome. Scientists have identified thousands of genome variations that exist within a population. These variations can be a single nucleotide or a larger chromosomal variation.
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...
17.8K
 01:05
Single Nucleotide Polymorphisms-SNPs
15.3K
A single nucleotide polymorphism or SNP is a single nucleotide variation at a specific genomic position in a large population. It is the most prevalent type of sequence variation found in the human genome. Point mutations that occur in more than 1% of the population qualify as SNPs. These are present once every 1000 nucleotides on an average in the human genome. Replacement of a purine with another purine (A/G) or a pyrimidine with another pyrimidine (C/T) is known as a transition. In contrast,...
15.3K
 01:26
Size and Structure of Viral Genomes
65
Viral genomes exhibit remarkable diversity in size, structure, and composition, influencing their replication strategies and interactions with host cells. These genomes consist of either DNA or RNA and may be linear or circular. Additionally, they can be single-stranded or double-stranded, with each configuration affecting how the virus propagates within a host. RNA viruses, for instance, generally have smaller genomes than DNA viruses, a factor that contributes to their high mutation rates and...
65
 02:54
Evolutionary Relationships through Genome Comparisons
5.8K
Genome comparison is one of the excellent ways to interpret the evolutionary relationships between organisms. The basic principle of genome comparison is that if two species share a common feature, it is likely encoded by the DNA sequence conserved between both species. The advent of genome sequencing technologies in the late 20th century enabled scientists to understand the concept of conservation of domains between species and helped them to deduce evolutionary relationships across diverse...
5.8K
 01:29
Modern Molecular Taxonomy
53
Advancements in molecular biology have revolutionized the identification and characterization of bacteria, with multiple methods leveraging DNA sequencing for enhanced precision. As sequencing technologies improve and costs decline, these approaches are increasingly used in clinical, environmental, and evolutionary studies.Multilocus Sequence Typing (MLST) examines several housekeeping genes, essential chromosomal genes encoding cellular functions, to distinguish strains. Approximately...
53
 01:11
Genome-wide Association Studies-GWAS
13.6K
Genome-wide association studies or GWAS are used to identify whether common SNPs are associated with certain diseases. Suppose specific SNPs are more frequently observed in individuals with a particular disease than those without the disease. In that case, those SNPs are said to be associated with the disease. Chi-square analysis is performed to check the probability of the allele likely to be associated with the disease.
GWAS does not require the identification of the target gene involved in...
GWAS does not require the identification of the target gene involved in...
13.6K