通过轴 Fe4 C 实现的 d-轨道电子移位 原子集群提供高性能 Fe-N-C 催化剂,用于氧降解反应
Long-Ji Yuan1, Bo Liu2, Li-Xiao Shen3
1Shenzhen Key Laboratory of Special Functional Materials, Shenzhen Engineering Laboratory for Advance Technology of Ceramics, Guangdong Research Center for Interfacial Engineering of Functional Materials, College of Materials Science and Engineering, Shenzhen University, Shenzhen, 518060, P. R. China.
Advanced materials (Deerfield Beach, Fla.)
|July 14, 2023

概括
一种新型的-Fe-N-C催化剂稳定了使用Fe4C集群的活性Fe-pyrrolic N4位点,用于燃料电池中高效的氧降解反应.
科学领域:
- 电化学 电化学 电化学
- 材料科学 材料科学 材料科学
- 催化剂是一种催化剂.
背景情况:
- 铁--碳 (Fe-N-C) 催化剂在质子交换膜燃料电池 (PEMFC) 中的氧降解反应 (ORR) 中是的有希望的替代品.
- 铁烯酸N4结构被确定为ORR高度活跃的部位,但其稳定性仍然是一个重大挑战.
- 开发稳定高效的非贵金属催化剂对于推进燃料电池技术至关重要.
研究的目的:
- 构建一个稳定的Fe-N-C催化剂与Fe-pyrrolic N4活性中心,以提高ORR性能.
- 调查轴 Fe4C 原子集群在稳定 Fe-pyrrolic N4 结构和优化催化活性中的作用.
- 通过原子集群相互作用来探索调整单原子催化剂电子结构的新策略.
主要方法:
- 合成一种含有Fe-pyrrolic N4活性中心和轴性Fe4C原子集群的-Fe-N-C催化剂.
- 在酸性环境中对催化剂的ORR活性进行电化学评估,包括半波电位测量.
- 在H2-O2和H2-Air燃料电池中测试催化剂的性能,以确定功率密度.
- 理论计算 (例如,DFT) 和现场光谱分析以分析电子结构和结合相互作用.
主要成果:
- 合成的Cyan-Fe-N-C催化剂在酸性介质中实现了0.836V (相对于RHE) 的ORR高半波潜力.
- 催化剂在1.0bar时提供了令人印象深刻的870mWcm-2 (H2-O2) 和478mWcm-2 (H2-Air) 的功率密度.
- 理论和光谱分析证实,轴性Fe4C集群通过扰乱其电子特性和优化OH*吸附来稳定Fe-pyrrolic N4结构.
结论:
- 轴性Fe4C原子集群有效地稳定了Fe-pyrrolic N4活性位点,克服了Fe-N-C催化剂开发中的一个主要挑战.
- 单个原子和原子集群之间的强烈相互作用为设计具有定制电子结构的先进单原子催化剂提供了可行的策略.
- 这项工作为为燃料电池应用开发高效和稳定的非贵金属催化剂铺平了道路.
相关概念视频
 02:58
Crystal Field Theory - Octahedral Complexes
26.9K
Crystal Field Theory
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
26.9K
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
43.1K
Tetrahedral Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
43.1K
 02:42
Valence Bond Theory
8.8K
Coordination compounds and complexes exhibit different colors, geometries, and magnetic behavior, depending on the metal atom/ion and ligands from which they are composed. In an attempt to explain the bonding and structure of coordination complexes, Linus Pauling proposed the valence bond theory, or VBT, using the concepts of hybridization and the overlapping of the atomic orbitals. According to VBT, the central metal atom or ion (Lewis acid) hybridizes to provide empty orbitals of suitable...
8.8K
 01:18
Electron Orbital Model
68.0K
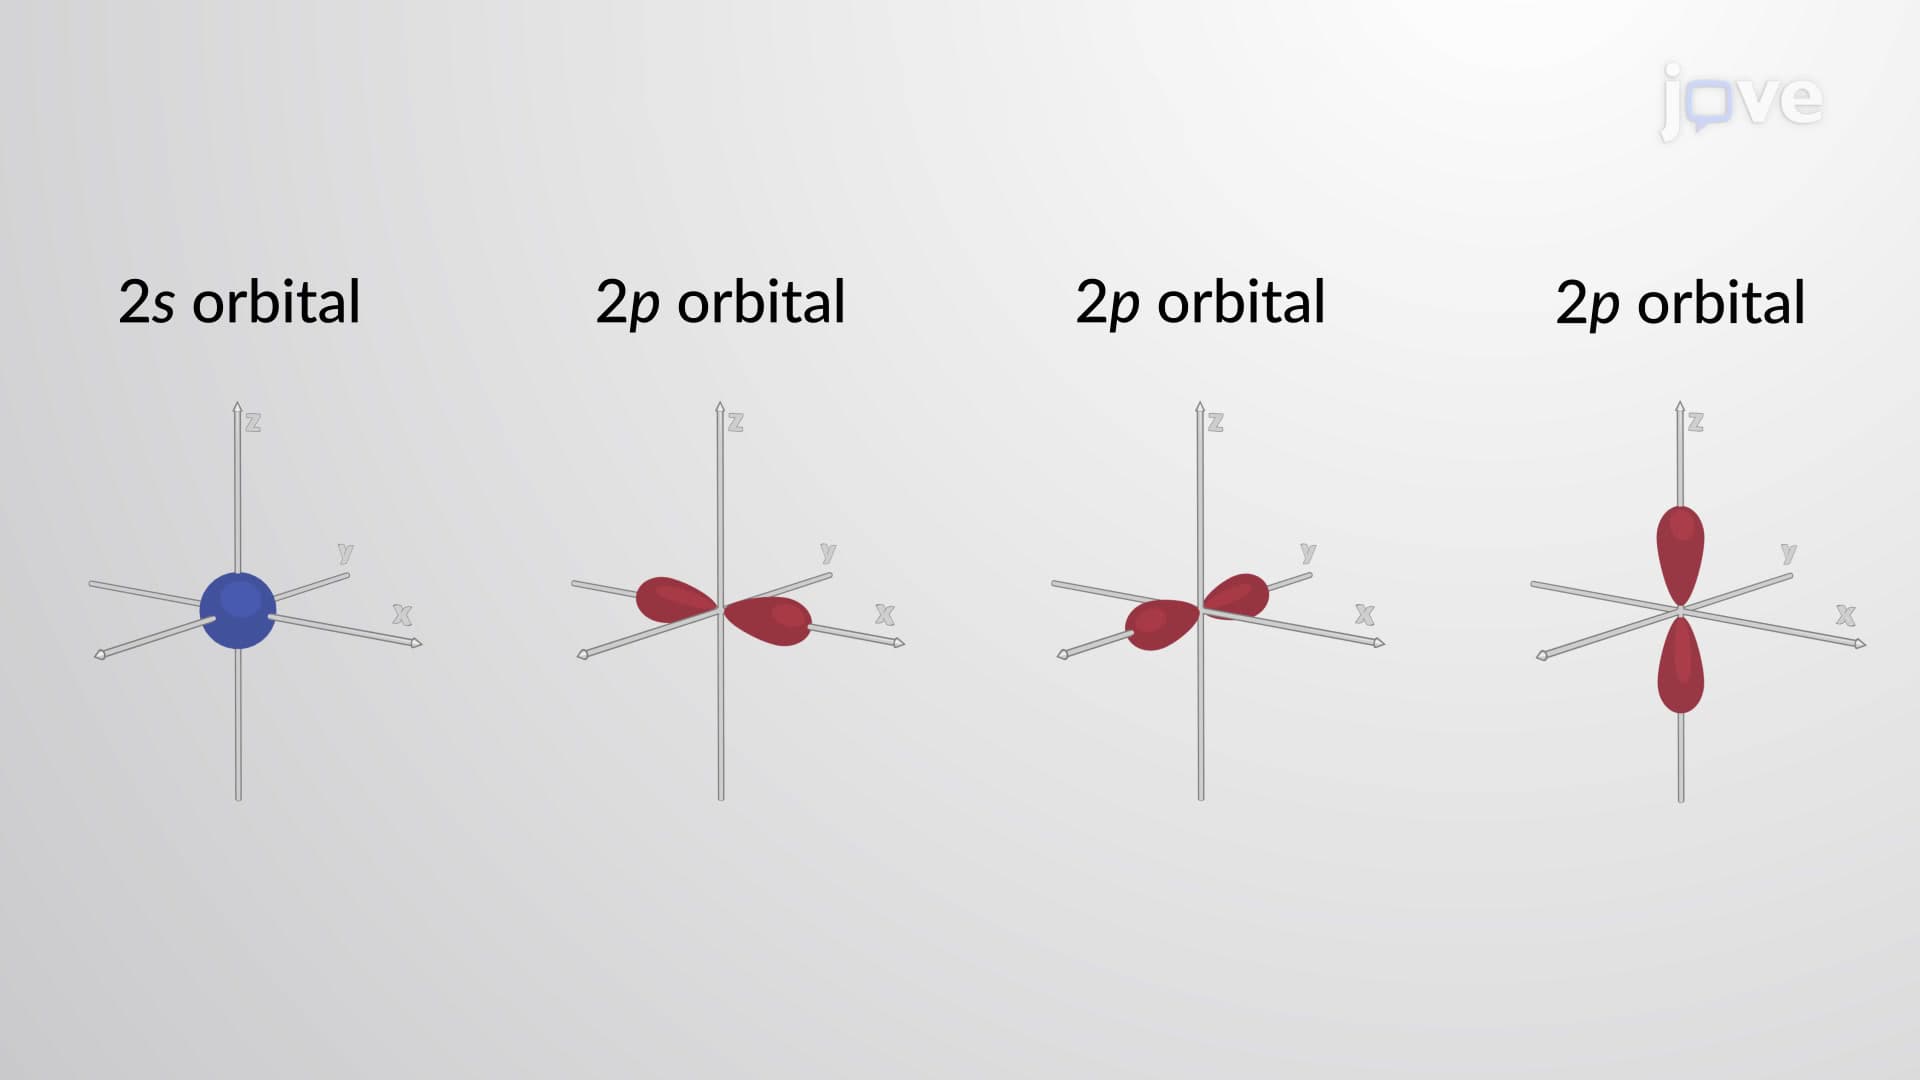
Orbitals are the areas outside of the atomic nucleus where electrons are most likely to reside. They are characterized by different energy levels, shapes, and three-dimensional orientations. The location of electrons is described most generally by a shell or principal energy level, then by a subshell within each shell, and finally, by individual orbitals found within the subshells.
The first shell is closest to the nucleus, and it has only one subshell with a single spherical orbital called the...
The first shell is closest to the nucleus, and it has only one subshell with a single spherical orbital called the...
68.0K
 02:44
Atomic Orbitals
33.8K
An atomic orbital represents the three-dimensional regions in an atom where an electron has the highest probability to reside. The radial distribution function indicates the total probability of finding an electron within the thin shell at a distance r from the nucleus. The atomic orbitals have distinct shapes which are determined by l, the angular momentum quantum number. The orbitals are often drawn with a boundary surface, enclosing densest regions of the cloud.
33.8K
 03:21
The Energies of Atomic Orbitals
24.1K
In an atom, the negatively charged electrons are attracted to the positively charged nucleus. In a multielectron atom, electron-electron repulsions are also observed. The attractive and repulsive forces are dependent on the distance between the particles, as well as the sign and magnitude of the charges on the individual particles. When the charges on the particles are opposite, they attract each other. If both particles have the same charge, they repel each other.
24.1K