
多切片空间解析的转录学数据的部分对齐
Xinhao Liu1, Ron Zeira2, Benjamin J Raphael3
1Department of Computer Science, Princeton University, Princeton, New Jersey 08540, USA.
Genome research
|August 8, 2023

概括
通过部分对齐多个2D组织切片,PASTE2可以实现空间基因表达的3D重建. 这种方法克服了现有方法的局限性,改善了空间转录学数据集成和分析.
科学领域:
- 计算生物学是一种计算生物学.
- 基因组学就是基因组学.
- 生物信息学是一种生物信息学.
背景情况:
- 空间解析转录组学 (SRT) 在组织片内提供基因表达数据.
- 当前的SRT方法分析2D切片,丢失3D空间信息.
- 整合来自多个切片的数据是具有挑战性的,因为对齐问题和组织形态变化.
研究的目的:
- 开发一种用于部分对齐和3D重建多切片SRT数据的新方法.
- 为了使空间转录学数据在多个组织切片中准确整合.
- 克服现有方法的局限性,这些方法假设完整的切片重叠或保存形态.
主要方法:
- 介绍PASTE2,一个用于部分对齐和3D重建多片段SRT数据集的方法.
- 制定一个新的部分化格罗莫夫-瓦瑟斯坦最佳运输问题,通过条件梯度算法解决.
- 包括一个模型选择程序来估计切片重叠分数和可选的组织学图像数据的使用.
主要成果:
- 与现有方法相比,PASTE2在模拟和真实数据上实现了更准确的对齐.
- 在使用16个立体隔离切片*Drosophila*胚胎中的3D基因表达图的成功重建.
- 证明了来自各种SRT技术的多切片数据集的准确对齐.
结论:
- PASTE2有效地集成了多切片空间转录学数据,恢复丢失的3D信息.
- 该方法促进了对空间基因表达模式的详细研究.
- PASTE2 增强了 SRT 技术在各种生物应用中的实用性.
相关概念视频
 03:21
RNA-seq
10.1K
RNA sequencing, or RNA-Seq, is a high-throughput sequencing technology used to study the transcriptome of a cell. Transcriptomics helps to interpret the functional elements of a genome and identify the molecular constituents of an organism. Additionally, it also helps in understanding the development of an organism and the occurrence of diseases.
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
10.1K
 02:34
DNA Microarrays
17.5K
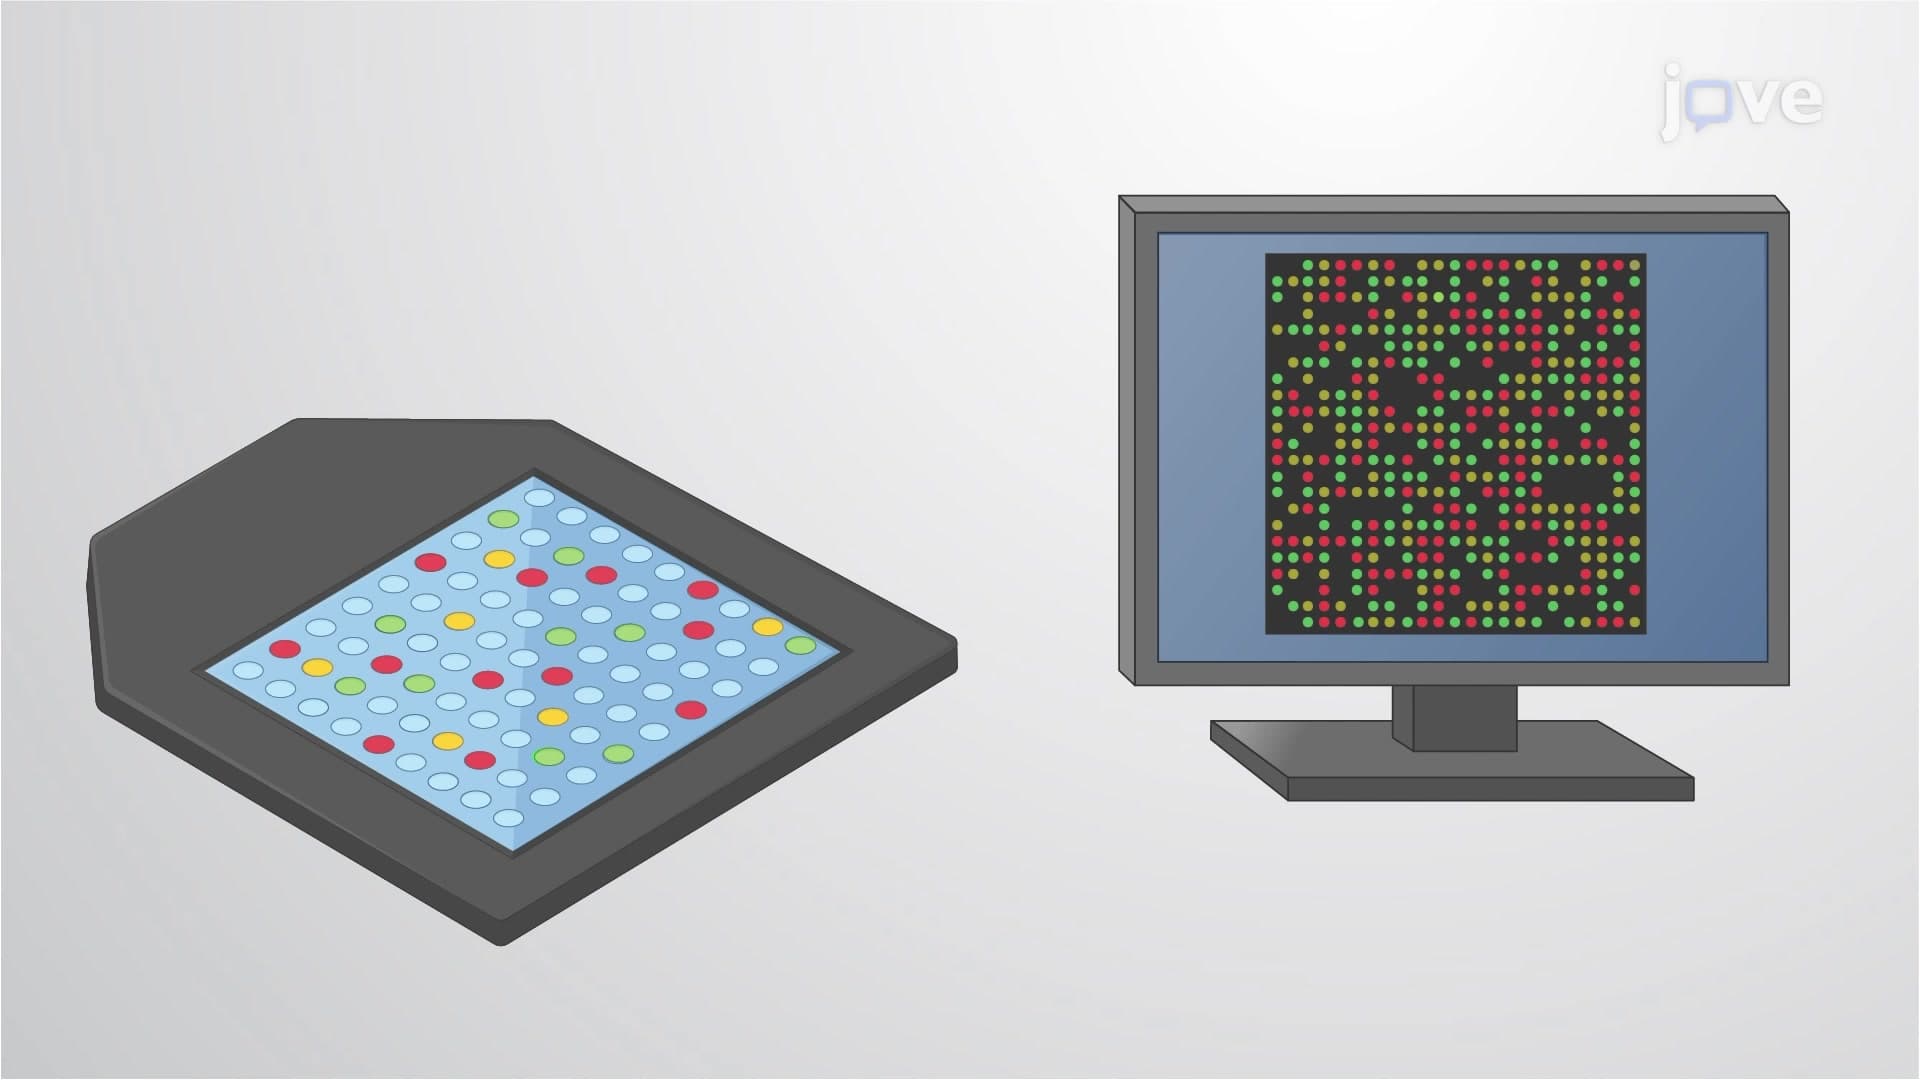
Microarrays are high-throughput and relatively inexpensive assays that can be automated to analyze large quantities of data at a time. They are used in genome-wide studies to compare gene or protein expression under two varied conditions, such as healthy and diseased states. Microarrays consist of glass or silica slides on which probe molecules are covalently attached through surface functionalization. Most commonly, the slides are prepared through the chemisorption of silanes to silica...
17.5K