MT-MAG:准确和可解释的机器学习,用于对元基因组组装基因组的完全或部分分类学赋值
Wanxin Li1, Lila Kari1, Yaoliang Yu1
1School of Computer Science, University of Waterloo, Waterloo, Ontario, Canada.
PloS one
|August 18, 2023

概括
我们介绍了MT-MAG,这是一种用于分类元基因组组装基因组 (MAG) 的机器学习工具. 这种无对齐的软件准确地对大型数据集进行分类,性能优于现有的工具,并提供部分分类以改善分类学赋值.
科学领域:
- 基因组学就是基因组学.
- 生物信息学是一种生物信息学.
- 机器学习 机器学习
背景情况:
- 大基因组组装基因组 (MAGs) 对于理解微生物群落至关重要.
- 准确地分类MAG的分类学是必不可少的,但具有挑战性,特别是对于大型和多样化的数据集.
- 现有的工具通常依赖于对齐或遗传标记,限制了它们的适用性.
研究的目的:
- 开发和评估MT-MAG,一种基于机器学习的新型软件,用于MAG的层次分类.
- 为分类完整和部分MAG提供一个强大的和有效的工具.
- 为分类学分配提供无对齐和无标记的方法.
主要方法:
- MT-MAG使用k-mer频率 (k=7) 作为分类的唯一特征,避免了序列对齐.
- 该工具在广泛的元基因组数据集 (245.68 Gbp) 上进行了训练,并在各种数据集 (9.6 Gbp) 上进行了测试.
- 开发了一种"加权分类准确性"指标,以计算不同分类学等级的部分分类的信息性.
主要成果:
- 在基准数据集上,MT-MAG的平均加权分类准确率为87.32%.
- 它的表现明显超过了DeepMicrobes,在物种层面的加权分类准确度高出34.79%.
- MT-MAG在物种层面完全分类了67.70%的序列,而深层微生物的分类率为47.45%,并为95.13%的基因组提供了部分或完整的分类.
结论:
- MT-MAG是MAG的分类学分类的高度准确和高效的工具,包括部分分类.
- 它的无对齐和无标记方法比现有方法具有优势.
- MT-MAG通过提供可靠的分类学赋值和保密来增强对元基因组数据的分析.
相关概念视频
 01:29
Modern Molecular Taxonomy
52
Advancements in molecular biology have revolutionized the identification and characterization of bacteria, with multiple methods leveraging DNA sequencing for enhanced precision. As sequencing technologies improve and costs decline, these approaches are increasingly used in clinical, environmental, and evolutionary studies.Multilocus Sequence Typing (MLST) examines several housekeeping genes, essential chromosomal genes encoding cellular functions, to distinguish strains. Approximately...
52
 01:20
Applications of Molecular Taxonomy
41
Molecular taxonomy has revolutionized the understanding and classification of bacteria, providing precise insights into their diversity, evolutionary relationships, and ecological roles. By utilizing molecular techniques such as DNA sequencing and fingerprinting, researchers have made significant strides in various fields related to bacterial studies.Resolving Taxonomic AmbiguitiesMolecular taxonomy has been instrumental in distinguishing closely related bacterial species initially thought to...
41
 02:54
Evolutionary Relationships through Genome Comparisons
5.8K
Genome comparison is one of the excellent ways to interpret the evolutionary relationships between organisms. The basic principle of genome comparison is that if two species share a common feature, it is likely encoded by the DNA sequence conserved between both species. The advent of genome sequencing technologies in the late 20th century enabled scientists to understand the concept of conservation of domains between species and helped them to deduce evolutionary relationships across diverse...
5.8K
 03:36
Genome Annotation and Assembly
18.9K
The genome refers to all of the genetic material in an organism. It can range from a few million base pairs in microbial cells to several billion base pairs in many eukaryotic organisms. Genome assembly refers to the process of taking the DNA sequencing data and putting it all back together in a correct order to create a close representation of the original genome. This is followed by the identification of functional elements on the newly assembled genome, a process called genome annotation.
18.9K
 01:24
Microbial Classification System
45
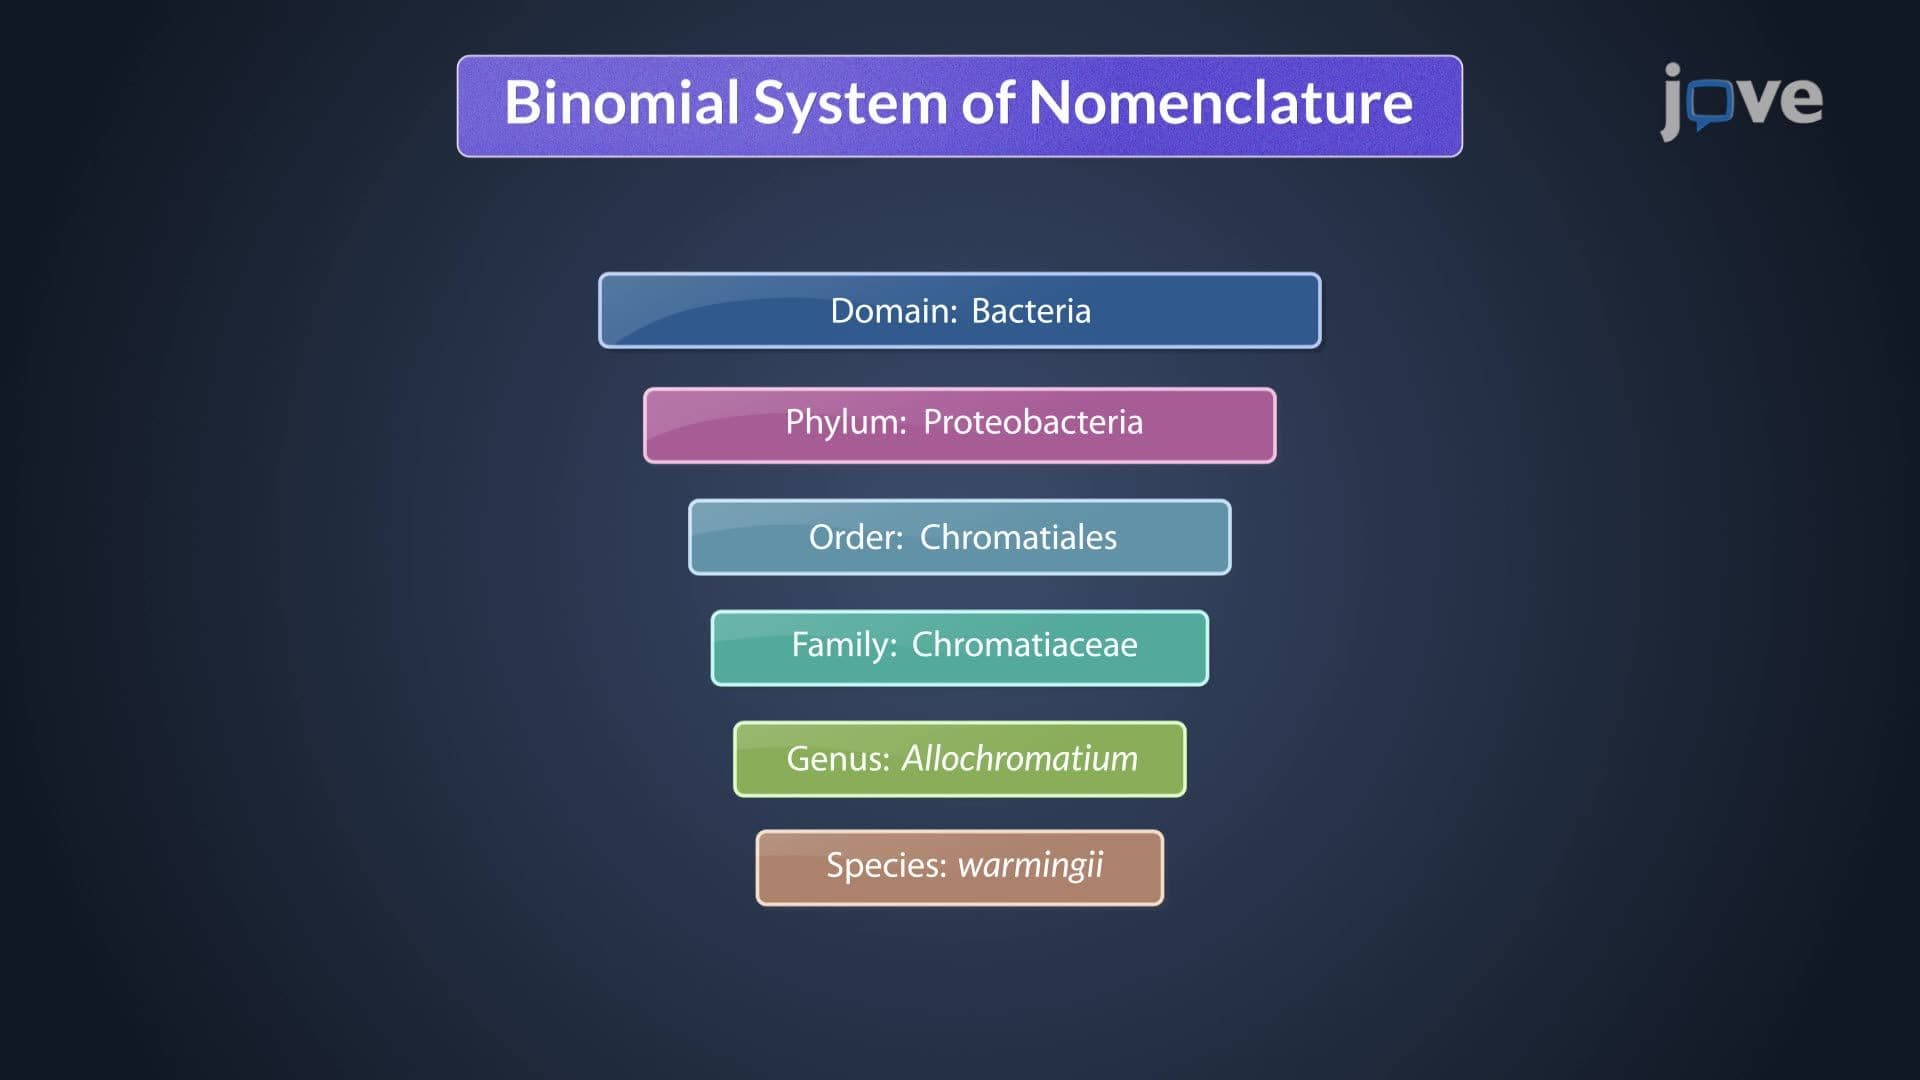
Classification is the process of organizing organisms into hierarchically inclusive groups based on their phenotypic similarities or evolutionary relationships. A species comprises one or more strains, and closely related species are grouped into genera. Genera are further classified into families, families into orders, orders into classes, and so forth, up to the domain level, which is the broadest taxonomic rank derived from a combination of phenotypic and genotypic data.The nomenclature of...
45