
在中链乙烯-CoA脱酶患者之间的临床异质性的个性化建模
Christoff Odendaal1, Emmalie A Jager1,2, Anne-Claire M F Martines1
1Laboratory of Paediatrics, University of Groningen, University Medical Centre Groningen, Groningen, the Netherlands.
BMC biology
|September 4, 2023

概括
计算模型显示,保持较高的自由线粒体CoASH和通路流量可以区分无症状和症状的中链甲酸脱酶缺乏症 (MCADD) 患者,为代谢疾病异质性提供了新的见解.
科学领域:
- 生物化学 生化学
- 系统生物学 系统生物学
- 代谢障碍 代谢障碍 代谢障碍
背景情况:
- 单基因代谢的先天性错误表现出显著的表型异质性,即使在具有相同基因变异的个体中.
- 中链乙-CoA脱酶缺乏症 (MCADD) 是线粒体脂肪酸氧化 (mFAO) 的常见先天性错误,临床结果不可预测.
- 从无症状到严重脱补偿的MCADD患者表型的变化原因尚不清楚.
研究的目的:
- 开发和验证人类肝脏mFAO的动力模型,以调查MCADD患者间异质性的来源.
- 测试维持增加的自由线粒体CoASH和通路流量的假设,使无症状患者与症状的MCADD患者有所区别.
主要方法:
- 构建和实验验证人类肝脏mFAO的动力模型,包括基于水溶性的代谢物分区.
- 在代谢去补偿过程中对患者尿液代谢物的分析.
- 对MCADD患者衍生纤维细胞的蛋白质组分析.
- 使用患者特异性蛋白质组学数据开发个性化代谢模型.
主要成果:
- 经过验证的动态模型准确地预测了MCADD中减少的mFAO流量和CoASH水平,其中CoA被分离为乙烯-CoA.
- 患者尿液中中短链甲酸的积累反映了该模型预测的甲酸池.
- 个性化模型在无症状的MCADD患者中显示出更高的通路流量和CoASH,这与短链乙-CoA脱酶 (SCAD) 水平升高有关.
结论:
- 一个经过验证的人类肝脏mFAO的动态模型与可溶性依赖的代谢物分区为MCADD的表型异质性提供了新的解释.
- 个性化代谢建模为改善个人风险评估和代谢先天错误管理提供了一个有希望的方法.
- 升高的SCAD水平代表了在无症状的MCADD患者中潜在的,以前未被探索的补偿机制.

相关概念视频
 01:28
Overview of Fatty Acid Metabolism
30.8K
Lipids also are sources of energy that power cellular processes. Like carbohydrates, lipids are composed of carbon, hydrogen, and oxygen, but these atoms are arranged differently. Most lipids are nonpolar and hydrophobic. Major types include fats and oils, waxes, phospholipids, and steroids.
Fatty acids are catabolized in a process called beta-oxidation, which takes place in the matrix of the mitochondria and converts their fatty acid chains into two-carbon units of acetyl groups. The acetyl...
Fatty acids are catabolized in a process called beta-oxidation, which takes place in the matrix of the mitochondria and converts their fatty acid chains into two-carbon units of acetyl groups. The acetyl...
30.8K
 02:59
Animal Mitochondrial Genetics
7.6K
Among all the organelles in an animal cell, only mitochondria have their own independent genomes. Animal mitochondrial DNA is a double-stranded, closed-circular molecule with around 20,000 base pairs. Mitochondrial DNA is unique in that one of its two strands, the heavy, or H, -strand is guanine rich, whereas the complementary strand is cytosine rich and called the light, or L, -strand. Compared to nuclear DNA, mitochondrial DNA has a very low percentage of non-coding regions and is marked by...
7.6K
 01:20
Inborn Errors of Metabolism
194
Phenylketonuria (PKU) is a protein metabolism disorder characterized by high blood levels of the amino acid phenylalanine. This results from a mutation in the gene responsible for phenylalanine hydroxylase, an enzyme that converts phenylalanine into tyrosine. When this enzyme is deficient, phenylalanine builds up in the blood, leading to symptoms such as vomiting, rashes, seizures, growth deficiency, and severe mental retardation. An early diagnosis and a diet restricting phenylalanine intake...
194
 01:25
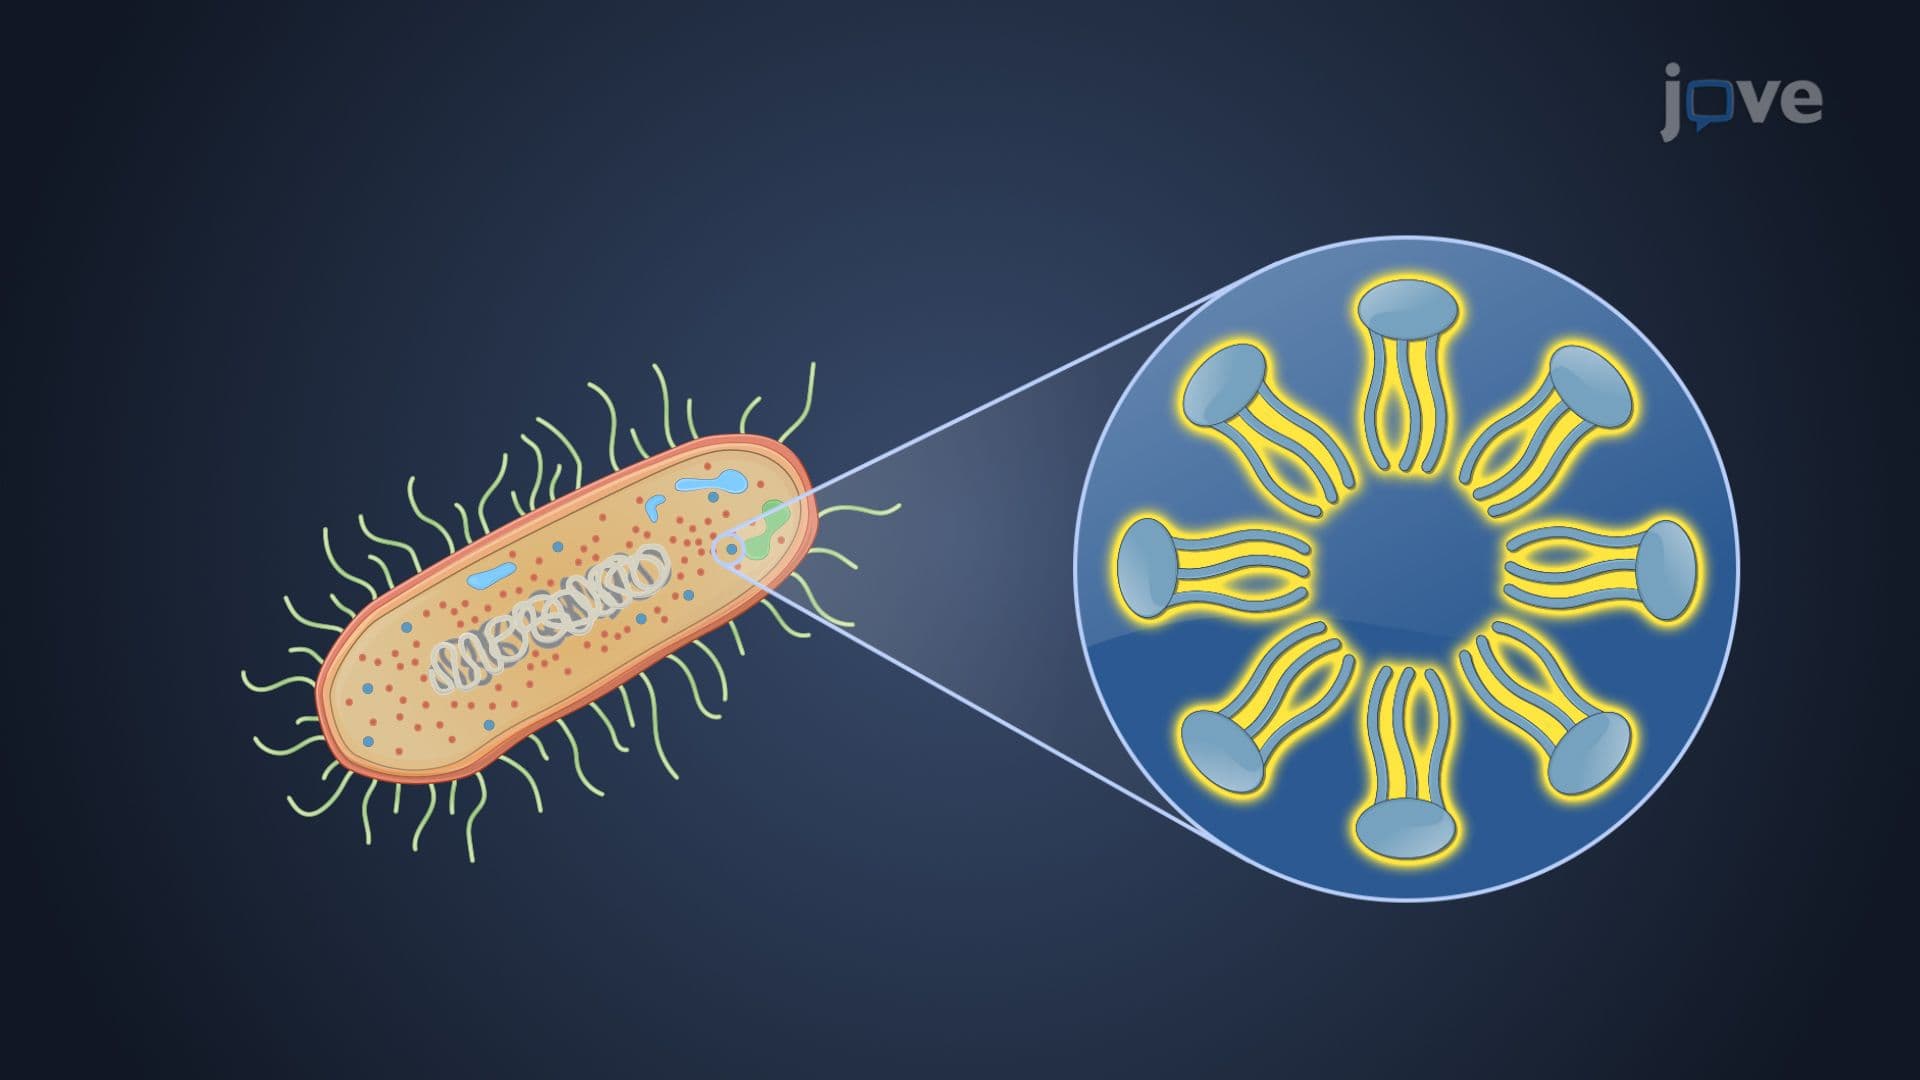
Lipid Catabolism
48
Triglycerides serve as crucial long-term energy storage molecules in microorganisms, providing a dense source of metabolic energy. Their breakdown is mediated by lipases, which hydrolyze triglycerides into glycerol and free fatty acids. Each of these components follows distinct metabolic pathways, ultimately contributing to ATP synthesis and cellular energy homeostasis.Glycerol MetabolismGlycerol, released from triglyceride hydrolysis, is phosphorylated by glycerol kinase to form...
48