
整合内部碎片在解释更大的寡核酸的自上而下的测序数据
Thomas Kenderdine1, William McIntyre1, Ghazaleh Yassaghi1
1University of Connecticut, Storrs, Connecticut 06269, United States.
Journal of the American Society for Mass Spectrometry
|September 20, 2023

概括
随着寡核酸大小的增加,碎片化从末端转移到内部部分. 分析这些内部碎片,以及终端碎片,显著提高了核酸测序的精度和覆盖范围.
科学领域:
- 质谱测量质量谱测量
- 分析化学是一种分析化学.
- 生物化学 生物化学
背景情况:
- 对核酸的直接分析面临着更大分子的挑战.
- 随着寡核酸大小的增加,终端序列的碎片产量会减少.
- 这限制了较大的寡核酸的气相测序.
研究的目的:
- 通过使用碰撞诱导解离 (CID) 来研究寡合核酸中的碎片化模式.
- 评估内部碎片对于核酸测序的有用性.
- 开发综合的数据解释策略,以改善序列化.
主要方法:
- 聚氨基核酸的碰撞诱导解离 (CID) (20-364 nt).
- 系统地评估实验因素 (碰撞能量,前体电荷,温度,杂热剂).
- 碎片阶梯形成和对齐的计算模拟.
主要成果:
- 碎片化从终端转移到内部碎片,随着寡核酸大小的增加.
- 内部碎片形成自相一致的梯子,有助于序列对齐和识别.
- 结合内部和终端碎片数据,可以增强序列和核酸覆盖范围.
- 来自内部碎片的赋值模两可,与终端碎片相似.
结论:
- 内部碎片提供有价值的序列信息,补充终端碎片.
- 综合数据解释策略对于测序较大的寡核酸至关重要.
- 这种方法克服了当前气相测序方法的局限性.
相关概念视频
 03:00
Next-generation Sequencing
89.9K
The first human genome sequencing project cost $2.7 billion and was declared complete in 2003, after 15 years of international cooperation and collaboration between several research teams and funding agencies. Today, with the advent of next-generation sequencing technologies, the cost and time of sequencing a human genome have dropped over 100 fold.
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
89.9K
 03:21
RNA-seq
10.0K
RNA sequencing, or RNA-Seq, is a high-throughput sequencing technology used to study the transcriptome of a cell. Transcriptomics helps to interpret the functional elements of a genome and identify the molecular constituents of an organism. Additionally, it also helps in understanding the development of an organism and the occurrence of diseases.
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
10.0K
 03:36
Genome Annotation and Assembly
18.9K
The genome refers to all of the genetic material in an organism. It can range from a few million base pairs in microbial cells to several billion base pairs in many eukaryotic organisms. Genome assembly refers to the process of taking the DNA sequencing data and putting it all back together in a correct order to create a close representation of the original genome. This is followed by the identification of functional elements on the newly assembled genome, a process called genome annotation.
18.9K
 01:57
Sanger Sequencing
754.7K
DNA sequencing is a fundamental technique that is routinely used in the biological sciences. This method can be applied to a range of questions at different scales - from the sequencing of a cloned DNA fragment or the study of a mutation in a gene up to whole-genome sequencing. However, despite the widespread use of sequencing today, it was not until 1977 that Fredrick Sanger and his collaborators developed the chain-termination method to decode DNA sequences. It relies on the separation of a...
754.7K
 01:05
Maxam-Gilbert Sequencing
11.2K
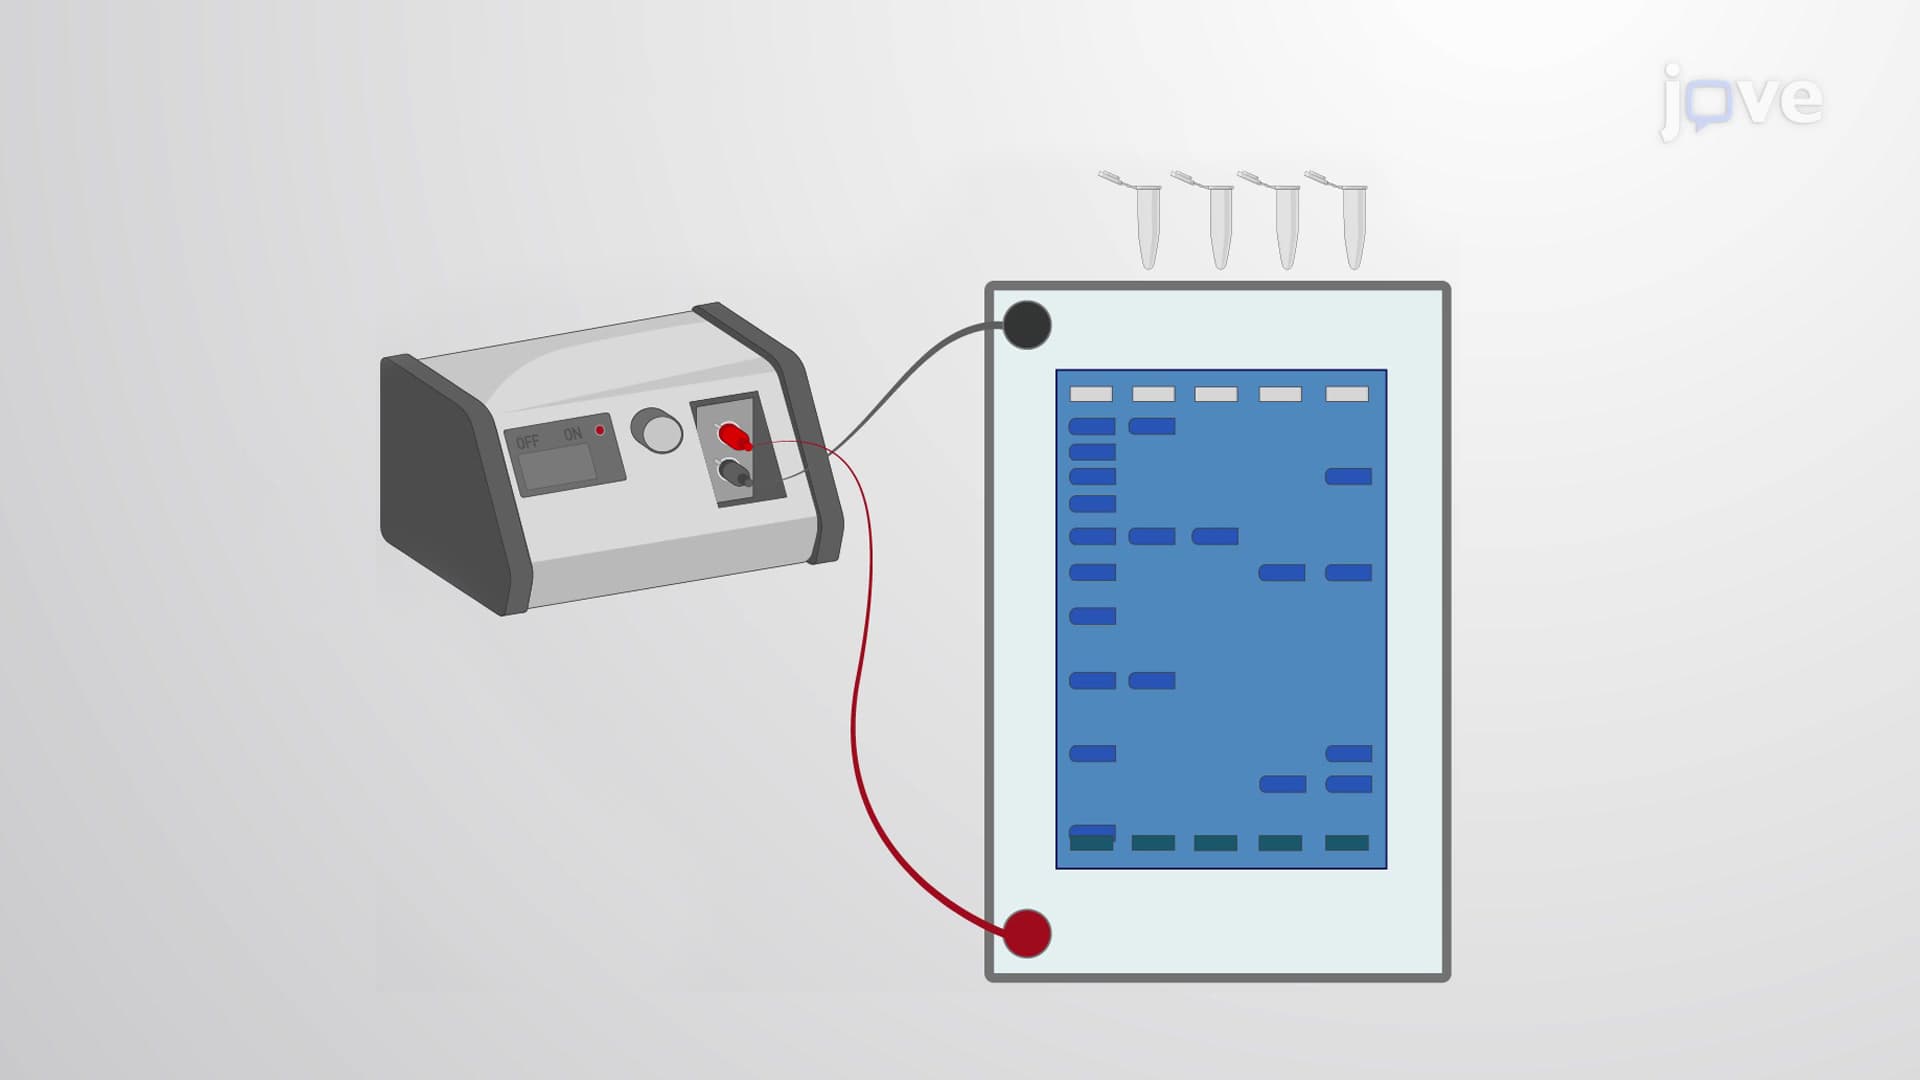
In the same year as the discovery of the Sanger sequencing method, another group of scientists, Allan Maxam and Walter Gilbert, demonstrated their chemical-cleavage method for DNA sequencing. The Maxam-Gilbert method relies on using different chemicals that can cleave the DNA sequence at specific sites, the separation of resulting DNA fragments of variable size using electrophoresis, and deciphering the DNA sequence from the resulting gel bands.
Challenges of the Maxam-Gilbert Method
The...
Challenges of the Maxam-Gilbert Method
The...
11.2K