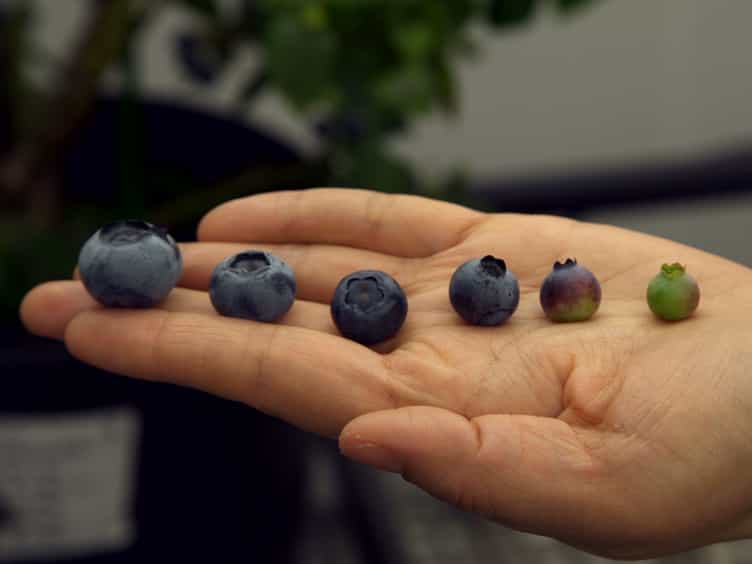

基于近红外光谱的PE包装蓝的可溶性固体含量建模与反向传播神经网络和部分最小方形 (BP-PLS) 算法
Ya Chen1, Yaoxiang Li1, Roger A Williams2
1College of Mechanical and Electrical Engineering, Northeast Forestry University, Harbin, China.
Journal of food science
|September 27, 2023

概括
近红外 (NIR) 谱学与反向传播 (BP) 神经网络和部分最小方形 (PLS) 算法 (BP-PLS) 结合,提供了一种快速,非破坏性的方法来确定包装蓝中的可溶性固体含量 (SSC). 这种先进的技术提高了整个蓝供应链质量控制的准确性.
科学领域:
- 农业科学 农业科学
- 频谱学是一种光谱学.
- 数据科学数据科学数据科学
背景情况:
- 蓝的质量属性,如溶性固体含量 (SSC),在整个供应链中发生变化.
- 准确和快速的质量评估对于尽量减少收获后损失和满足市场需求至关重要.
- 现有的近红外 (NIR) 光谱研究通常使用未包装的样本,限制了对现实世界供应链条件的适用性.
研究的目的:
- 开发一种使用NIR光谱的快速,非破坏性的方法来评估聚乙烯 (PE) 包装蓝的SSC.
- 为了比较NIR模型对未包装和PE包装蓝的预测性能.
- 通过减轻包装材料的干扰,提高PE包装蓝SSC预测的准确性.
主要方法:
- 近红外 (NIR) 谱学被用于蓝中SSC的定量分析.
- 光谱预处理技术包括多倍散射校正,标准正常变量,萨维茨基-戈莱和规范化.
- 波长优化利用了非信息变量消除,竞争性适应性重权采样和代保留信息变量.
- 部分最小平方 (PLS) 回归被用来构建初始的SSC预测模型.
- 开发了一种混合BP-PLS算法,结合了后传神经网络和基于残留学习的PLS,以解决包装干扰.
主要成果:
- 对未包装蓝的PLS模型实现了高预测准确性 (RP2 = 0.953,RMSEP = 0.611).
- 由于包装效应,PE包装蓝的初始PLS模型显示精度降低 (RP2 = 0.876,RMSEP = 0.632).
- 开发的BP-PLS模型显著提高了PE包装蓝的SSC预测准确度 (RP2=0.947,RMSEP=0.414).
结论:
- 与NIR光谱学相结合的BP-PLS方法提供了一个非常准确和快速的方法来确定PE包装蓝的SSC.
- 这种技术对于在供应链的各个阶段,从收获到销售,对蓝的质量监测有价值.
- 该研究展示了一种有希望的非破坏性实时质量评估方法,有助于满足市场需求和减少损失.
相关概念视频
 01:32
UV–Vis Spectroscopy of Conjugated Systems
7.1K
Organic compounds with conjugated double bonds show strong absorption features in the UV–visible region of the electromagnetic spectrum attributed to π → π* electronic excitations. Generally, a UV–vis absorption spectrum is recorded as a plot of absorbance vs wavelength. The wavelength of maximum absorbance, which manifests as a peak in the absorption spectrum, is denoted as λmax.
One of the factors influencing λmax is the extent...
One of the factors influencing λmax is the extent...
7.1K
 01:29
IR and UV–Vis Spectroscopy of Aldehydes and Ketones
5.8K
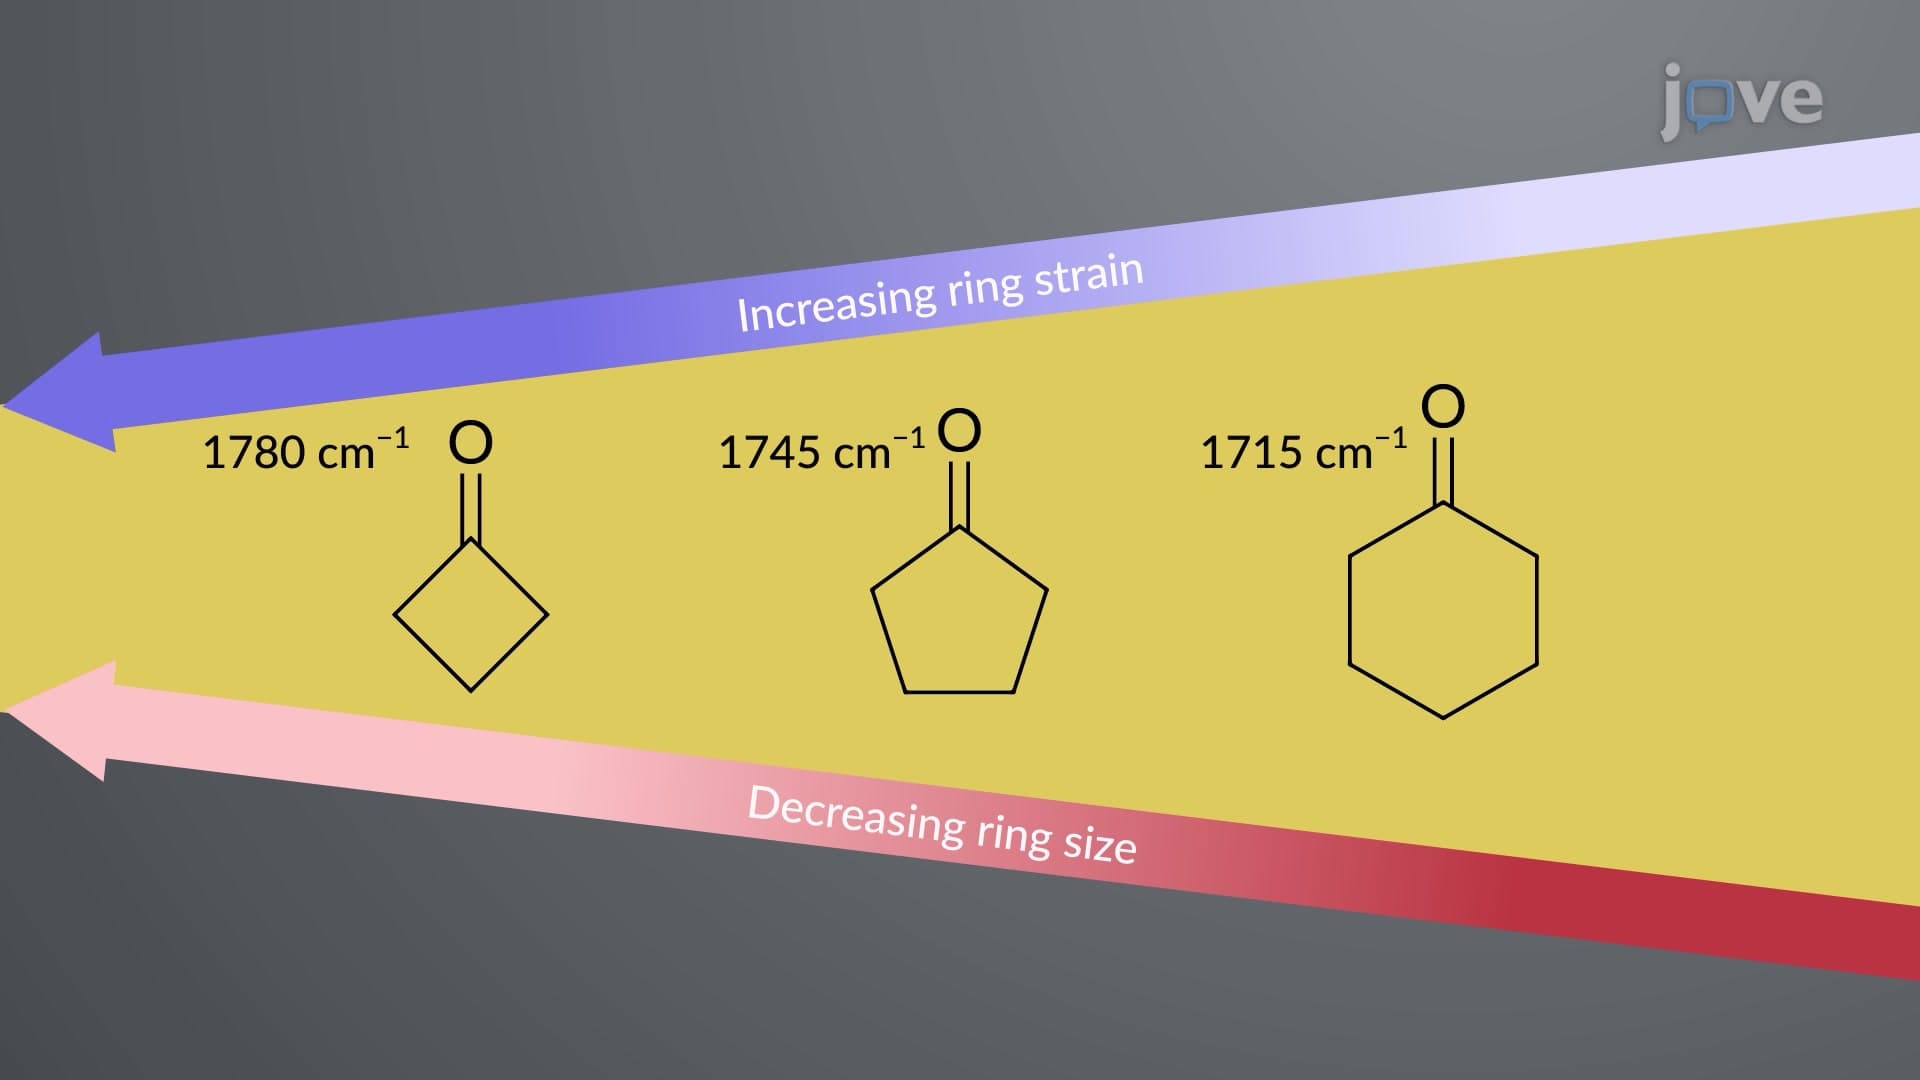
Infrared spectroscopy, also known as vibrational spectroscopy, is mainly used to determine the types of bonds and functional groups in molecules. In aldehydes and ketones, the carbonyl (C=O) bond shows an absorption around 1710 cm-1. The C=O bond vibration of an aldehyde occurs at lower frequencies than that of a ketone. In addition to the C=O absorption in an aldehyde, the aldehydic C–H bond also gives two peaks in the 2700–2800 cm-1 range. This absorption, coupled with the...
5.8K
 01:28
IR and UV–Vis Spectroscopy of Carboxylic Acids
4.2K
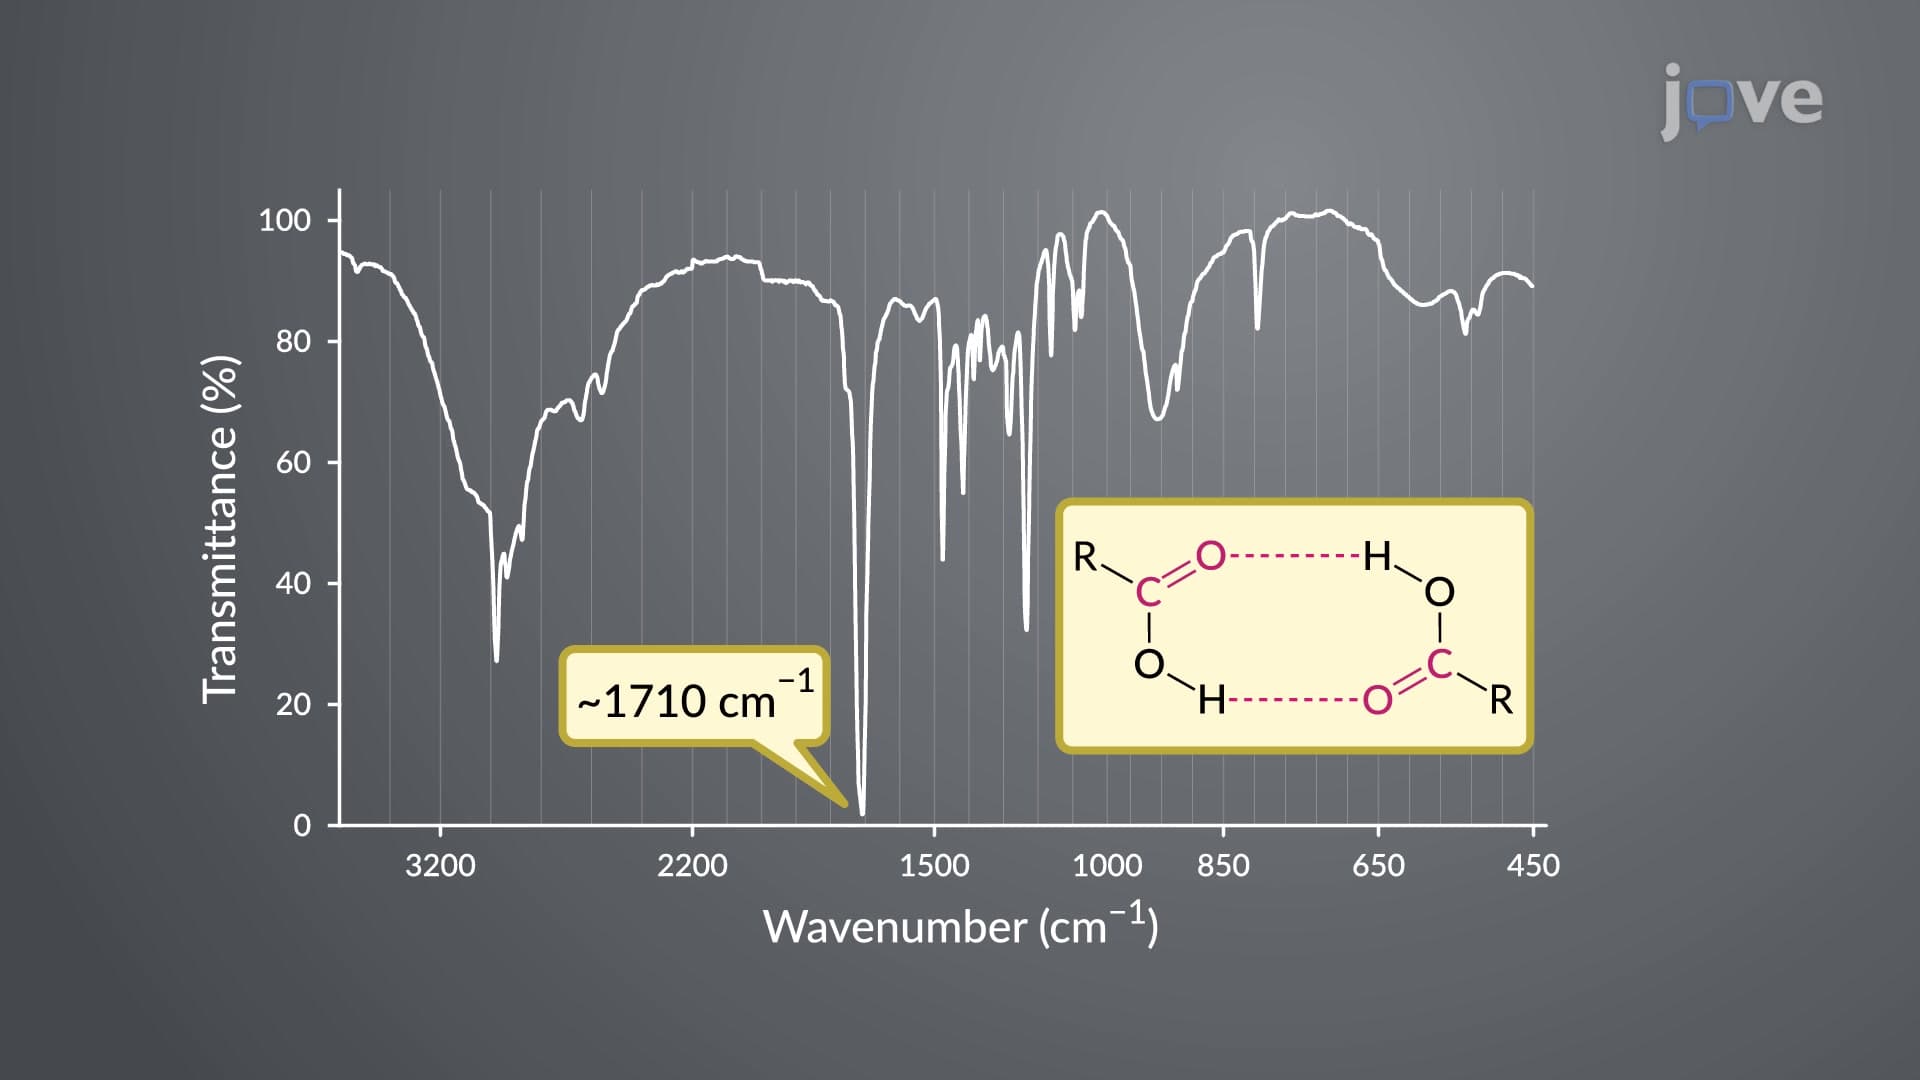
In IR spectroscopy of carboxylic acids, the C=O bond shows a characteristic band between 1710 and 1760 cm⁻¹, and the O–H bond exhibits a broad band between 2500 and 3300 cm⁻¹.
However, the stretching absorptions for the C=O bond vary depending on the structure of carboxylic acids. The C=O bond of the free carboxylic acids shows a higher stretching frequency, 1760 cm−1, while H-bonded carboxylic acids (dimers) exhibit stretching absorptions at a lower frequency,...
However, the stretching absorptions for the C=O bond vary depending on the structure of carboxylic acids. The C=O bond of the free carboxylic acids shows a higher stretching frequency, 1760 cm−1, while H-bonded carboxylic acids (dimers) exhibit stretching absorptions at a lower frequency,...
4.2K
 01:26
Spectroscopy of Carboxylic Acid Derivatives
2.4K
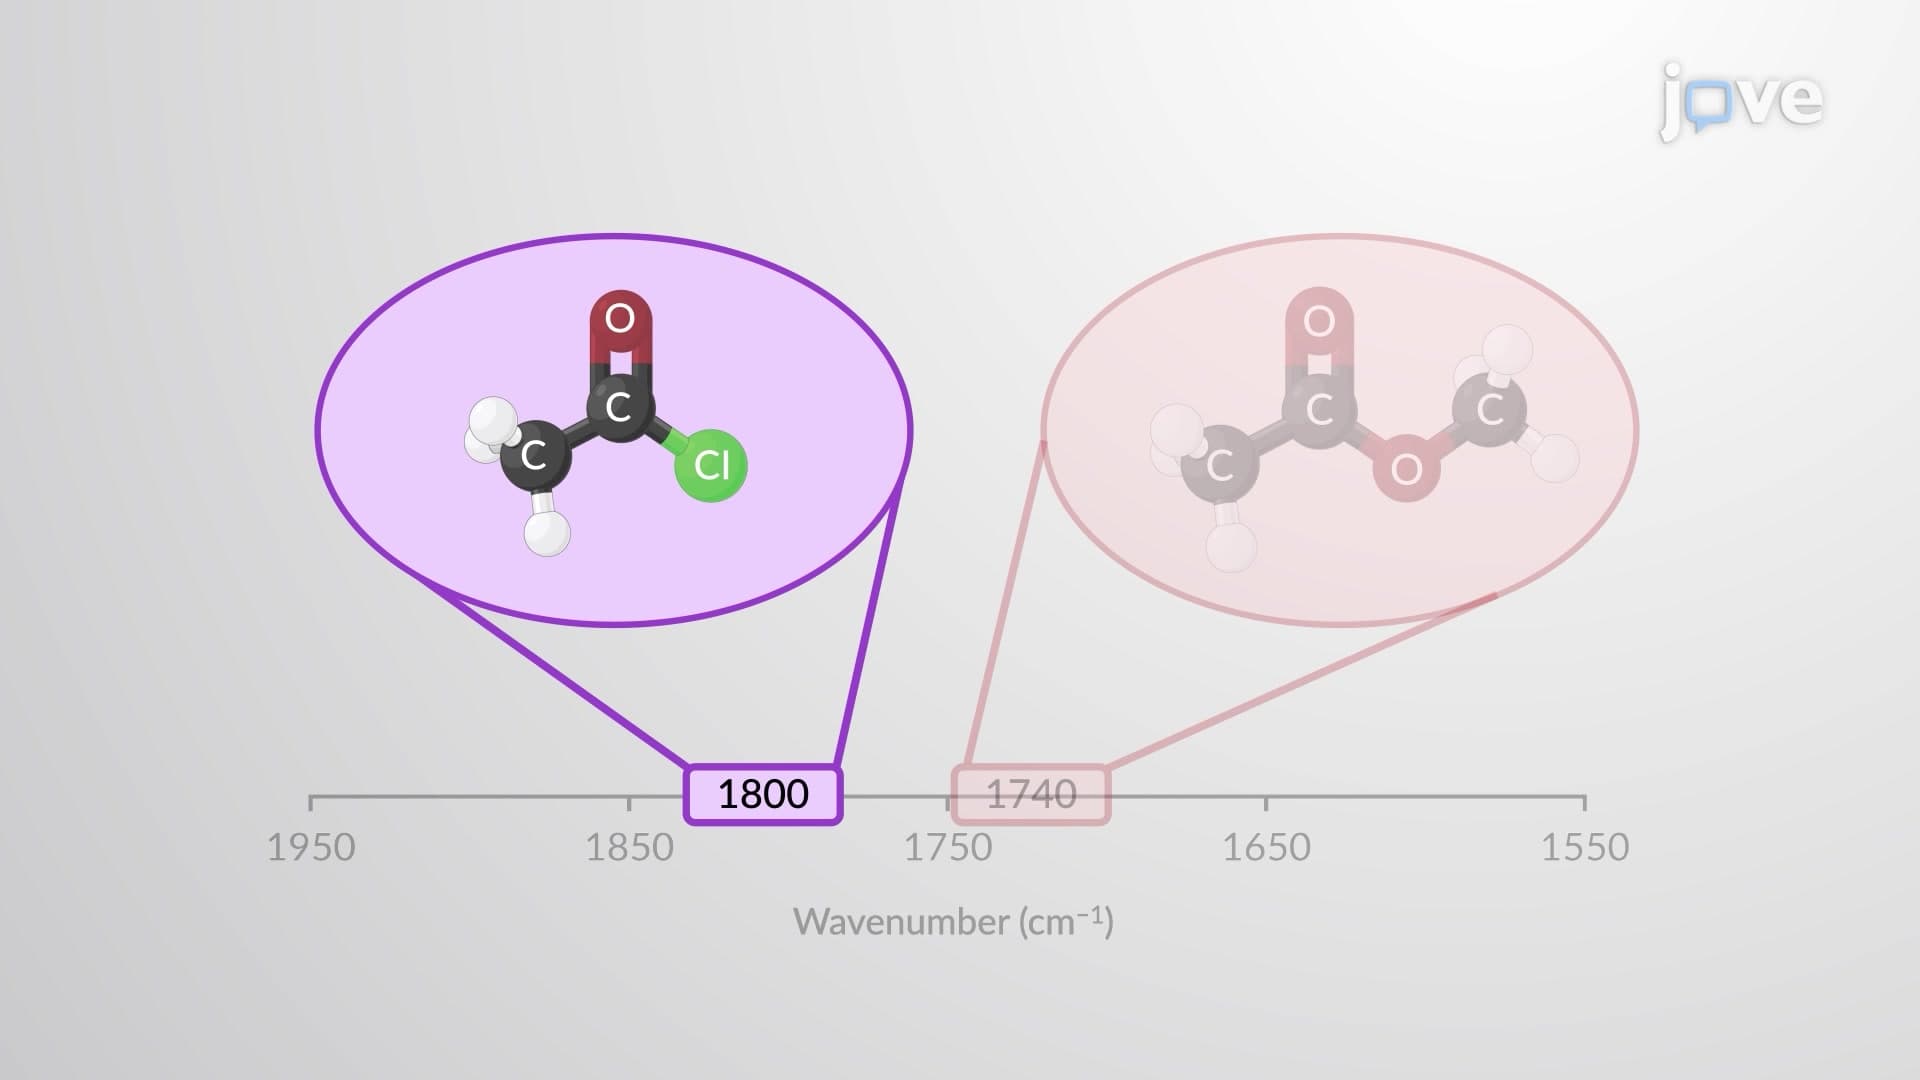
Infrared spectroscopy is primarily used to determine the types of bonds and functional groups. In carboxylic acid derivatives, a typical carbonyl bond absorption is observed around 1650–1850 cm−1. For esters, the absorption is recorded at around 1740 cm−1, while acid halides show the absorption at about 1800 cm−1. Another acid derivative, the acid anhydrides, exhibit two carbonyl absorption around 1760 cm−1 and 1820 cm−1, arising from the symmetrical and...
2.4K
 01:14
NMR Spectroscopy of Aromatic Compounds
4.8K
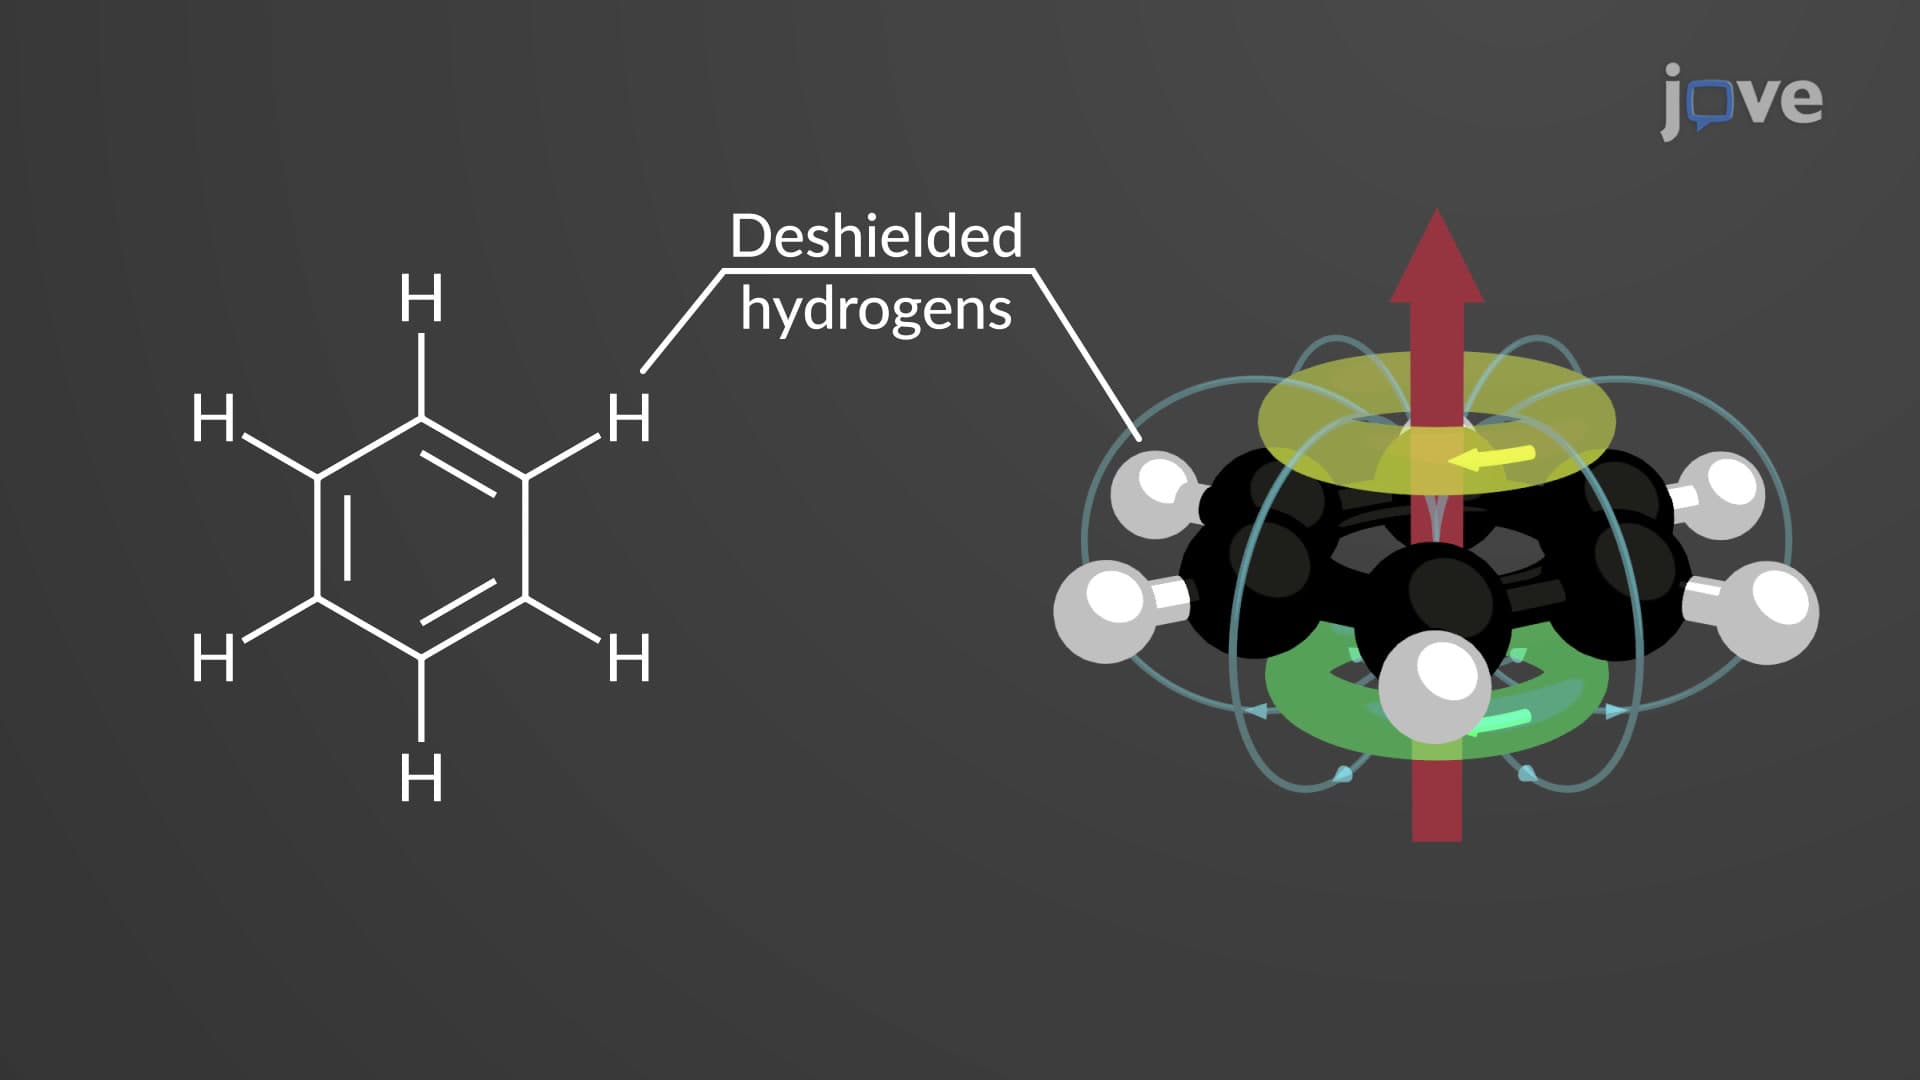
Aromatic compounds can be identified or analyzed using proton NMR and carbon‐13 NMR. Typically, aromatic hydrogens or hydrogens directly bonded to the aromatic rings are strongly deshielded by the aromatic ring current. Therefore, they absorb in the range of 6.5–8.0 ppm in proton NMR spectra. For instance, aromatic hydrogens directly bonded to the benzene ring absorb at 7.3 ppm. However, aromatic hydrogens of larger rings absorb farther upfield or downfield than the ideal range.
4.8K
 01:29
UV–Vis Spectroscopy: Woodward–Fieser Rules
24.6K
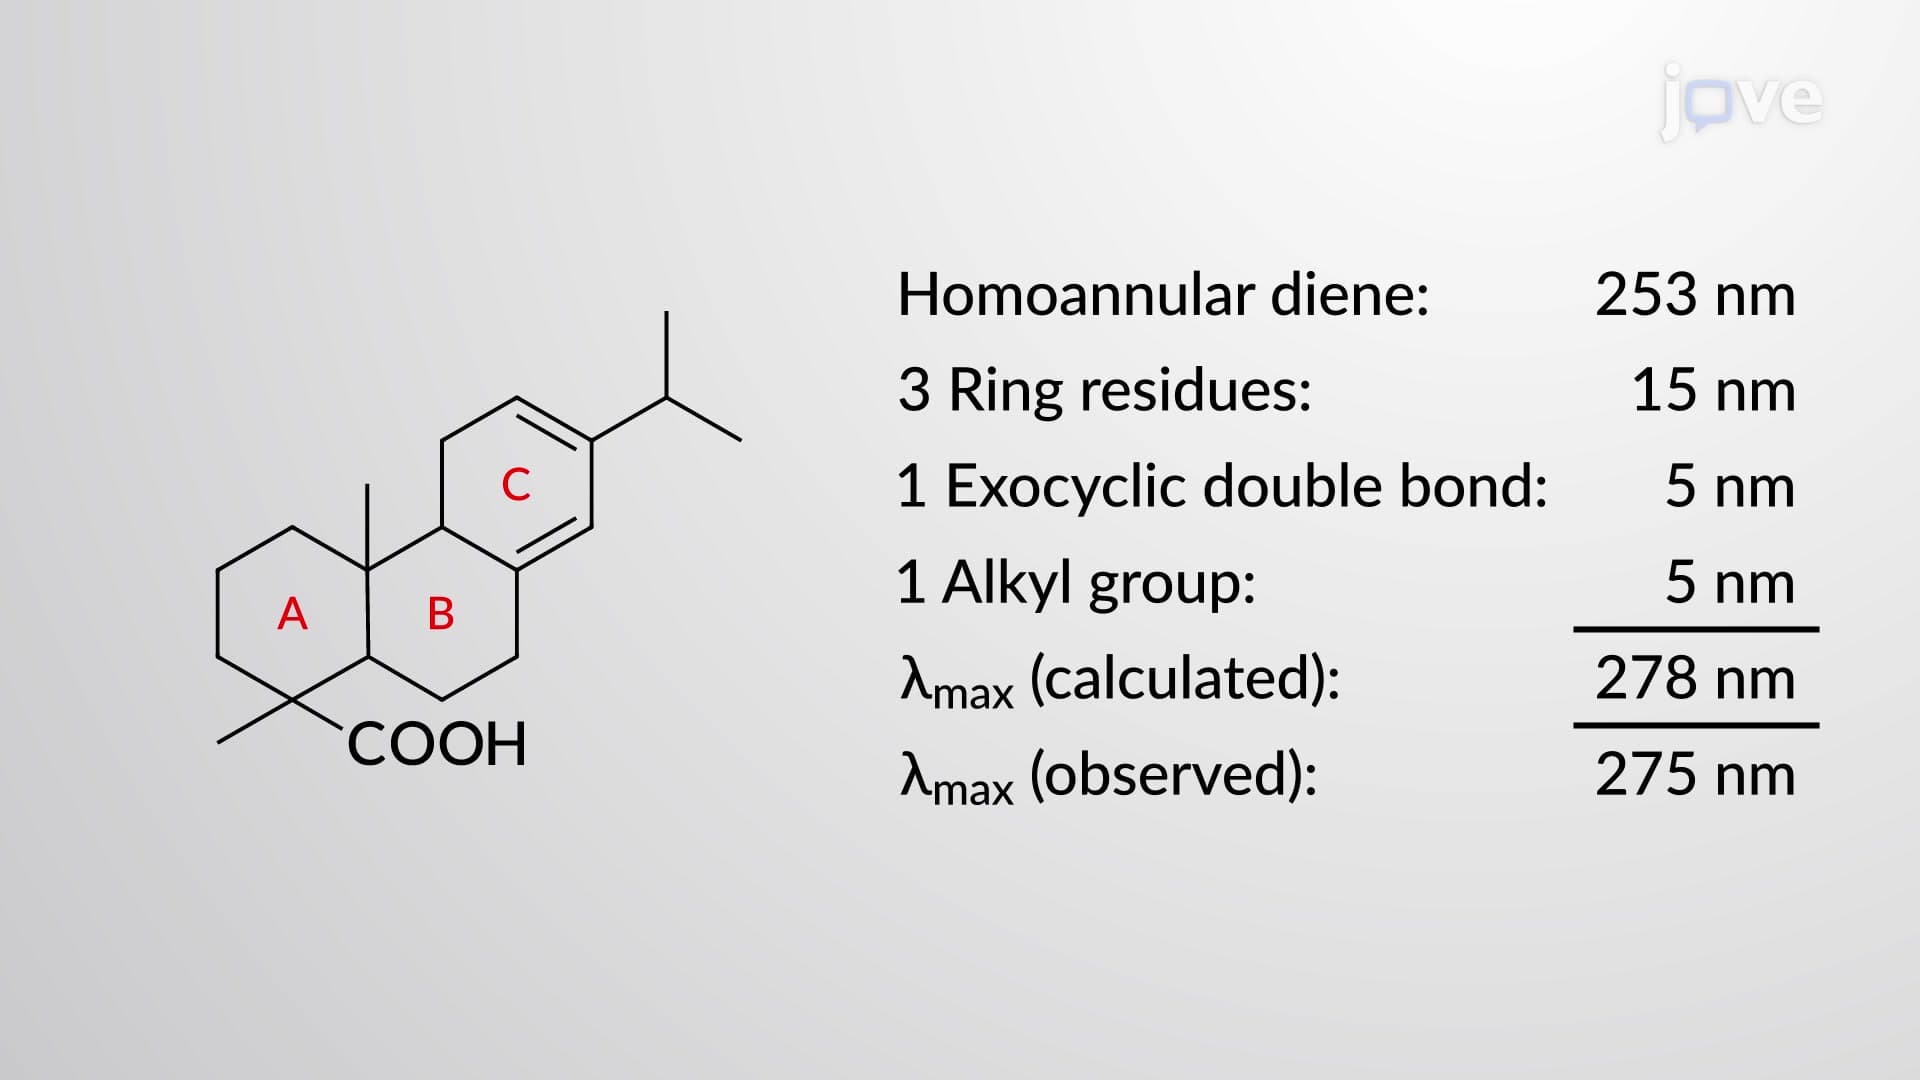
UV–Visible absorption spectra of conjugated dienes arise from the lowest energy π → π* transitions. The light-absorbing part of the molecule is called the chromophore, and the substituents directly attached to the chromophore are called auxochromes. A strong correlation exists between the absorption maxima, λmax, and the structure of a conjugated π system. The Woodward–Fieser rules predict the value of λmax for a given...
24.6K