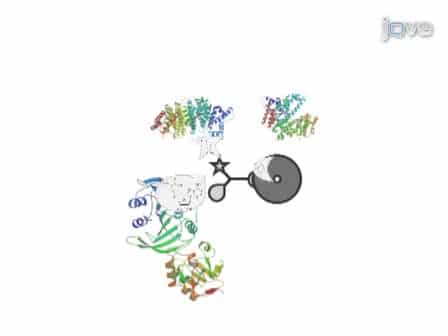

蛋白3D:通过molSimplify从蛋白质数据库中分析和提取含有金属的部位
Freya Edholm1, Aditya Nandy1,2, Clorice R Reinhardt1
1Department of Chemical Engineering, Massachusetts Institute of Technology, Cambridge, Massachusetts, USA.
Journal of computational chemistry
|October 24, 2023

概括
研究人员开发了一种新方法,使用molSimplify程序分析金属酶活性位点. 这种方法简化了蛋白质中金属结合点的研究,使得高通量计算分析成为可能.
科学领域:
- 生物化学 生物化学
- 计算化学计算化学
- 结构生物学 结构生物学
背景情况:
- 金属酶是各种化学反应的关键催化剂.
- 研究金属酶活性位点是具有挑战性的,因为它们融入更大的蛋白质结构.
- 了解这些网站的电子和几何性质是控制反应性和选择性的关键.
研究的目的:
- 开发一种用于分析金属酶活性位点的计算方法.
- 为了促进对金属酶结构的量子力学计算.
- 扩大molSimplify程序研究含金属蛋白质的能力.
主要方法:
- 在 molSimplify 程序中实现了一个新的 protein3D 类.
- 利用蛋白质数据库来获取晶体结构.
- 在非血红素铁酶周围提取的局部环境,具有两个histidine图案.
- 为量子力学计算准备了372种蛋白质结构.
主要成果:
- 成功提取并准备了372个金属酶结构用于计算分析.
- 证明了蛋白3D类对分析特定酶动机的有用性.
- 扩大了molSimplify的范围,包括复杂的蛋白质金属活性位点.
结论:
- 新的蛋白3D类增强了molSimplify分析金属酶的能力.
- 这种方法可以对含有金属的活性部位进行高通量计算研究.
- 有助于更深入地了解金属酶的功能和设计.
相关概念视频
 01:33
Proteomics
7.4K
A proteome is the entire set of proteins that a cell type produces. We can study proteomes using the knowledge of genomes because genes code for mRNAs, and the mRNAs encode proteins. Although mRNA analysis is a step in the right direction, not all mRNAs are translated into proteins.
Proteomics is the study of proteomes' function. It involves the large-scale systematic study of the proteome to denote the protein complement expressed by a genome. Scientist Mark Wilkins coined the term...
Proteomics is the study of proteomes' function. It involves the large-scale systematic study of the proteome to denote the protein complement expressed by a genome. Scientist Mark Wilkins coined the term...
7.4K
 01:49
Conserved Binding Sites
4.2K
Many proteins’ biological role depends on their interactions with their ligands, small molecules that bind to specific locations on the protein known as ligand-binding sites. Ligand-binding sites are often conserved among homologous proteins as these sites are critical for protein function.
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
4.2K