
对n → π*相互作用的分子轨道和拓电子密度研究:胺和胺的情况
Flor María Briceño-Vargas1, Mariana Quesadas-Rojas2, Gumersindo Mirón-López3
1Laboratory of Theoretical Chemistry, Faculty of Chemistry, Autonomous University of Yucatan Merida Yucatan 97069 Mexico ramiro.quijano@correo.uady.mx.

分散力显著加强胺和胺的n → π*相互作用. 胺相互作用比胺更强,尽管拓学方法并没有完全捕捉到这一趋势.
科学领域:
- 计算化学是一种计算化学.
- 量子化学是一种量子化学.
- 分子相互作用分子相互作用.
背景情况:
- 非共价相互作用在分子系统中至关重要.
- 了解n → π*相互作用对于各种化学和生物过程很重要.
研究的目的:
- 调查分散力在胺和胺的n → π*相互作用中的作用.
- 使用各种计算方法,比较胺和胺之间的n → π*相互作用的强度.
主要方法:
- 密度函数理论 (DFT) 使用不同的函数来评估分散效应.
- 自然结合轨道 (NBO) 分析.
- 独立梯度模型 (IGM) 和减少密度梯度 (RDG) 分析.
主要成果:
- 分散力对n → π*相互作用的强度有显著的贡献.
- 预测胺 n → π* 相互作用比胺更强.
- IGM分析显示,n → π*相互作用的dg高度值低于0.025.
结论:
- 分散力是调节 n → π* 相互作用强度的一个关键因素.
- 虽然计算方法通常同意相对强度,电子密度拓方法没有完全复制实验和NBO发现的胺与胺相互作用.
更多相关视频
 07:20
Amide Coupling Reaction for the Synthesis of Bispyridine-based Ligands and Their Complexation to Platinum as Dinuclear Anticancer Agents
Published on: May 28, 2014
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
相关概念视频
 02:35
Molecular Orbital Theory I
 02:40
MO Theory and Covalent Bonding
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:18
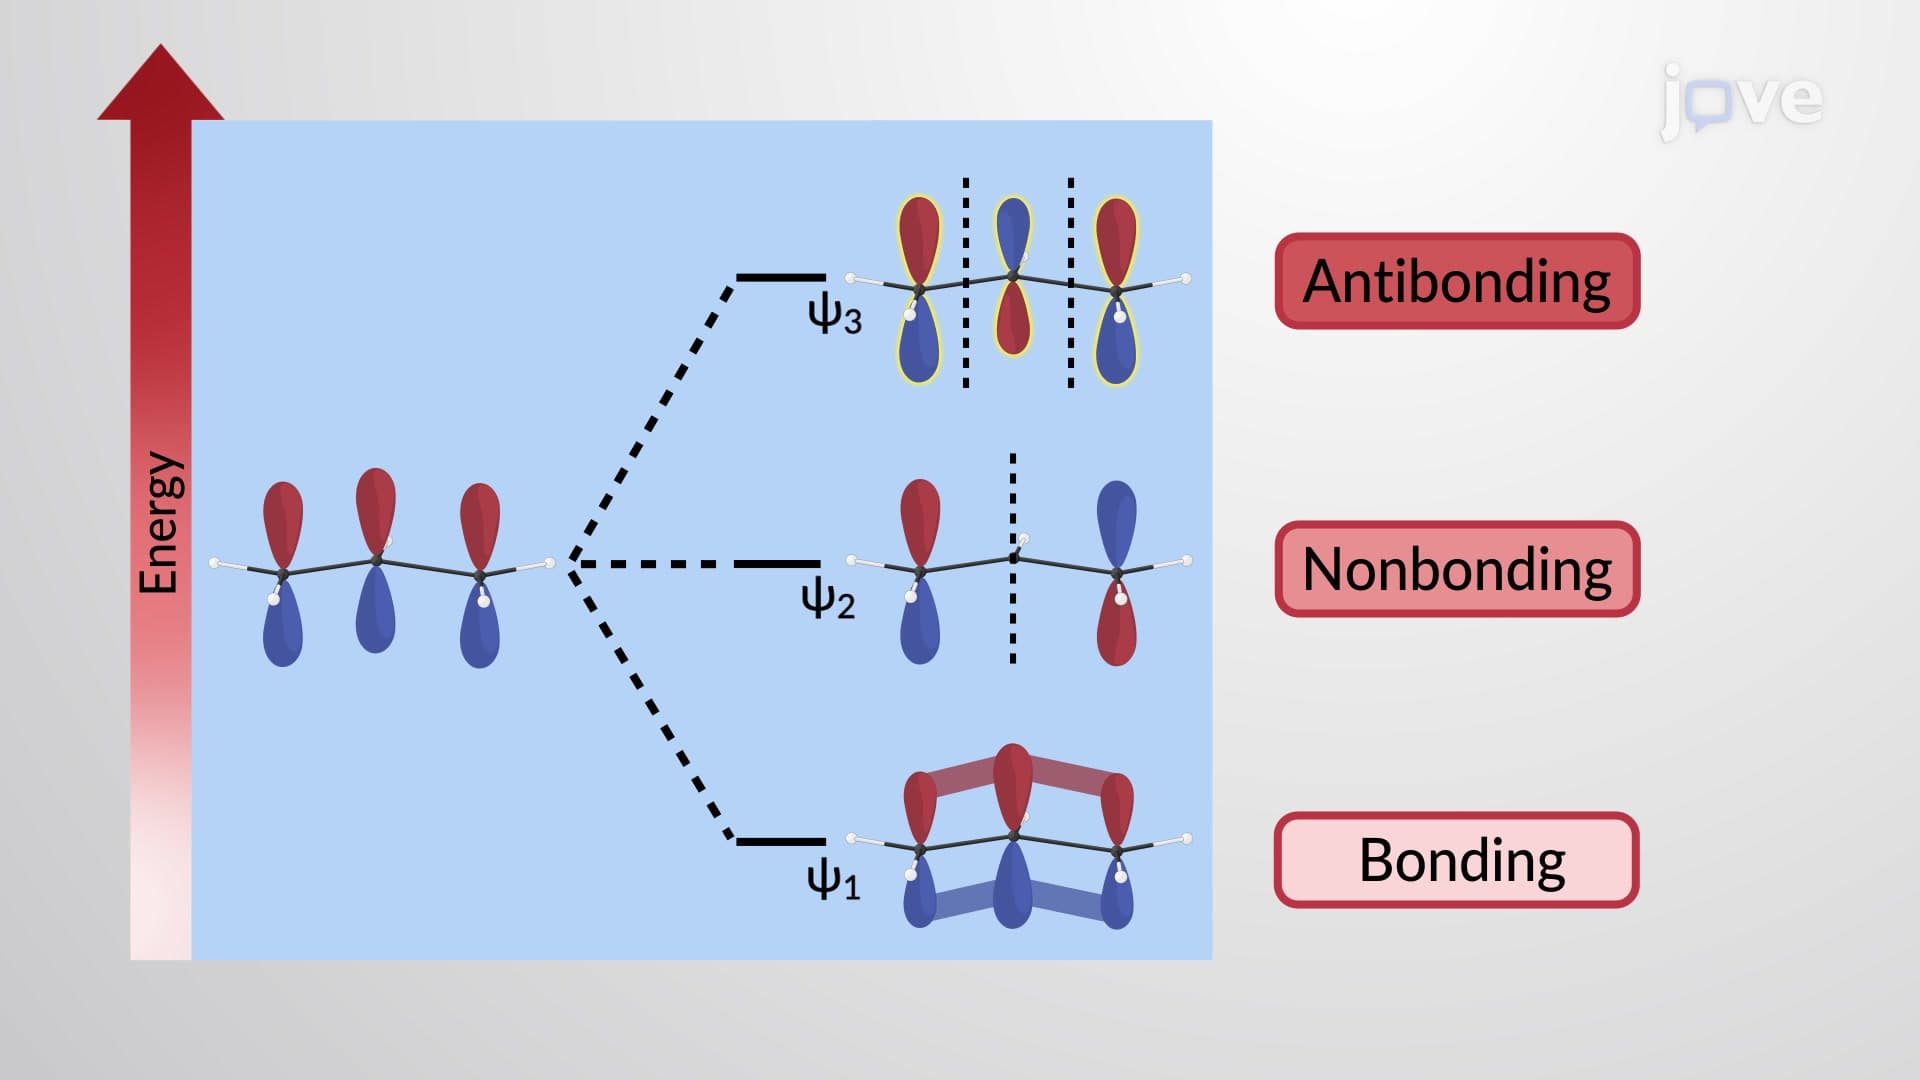
π Molecular Orbitals of the Allyl Cation and Anion
 01:27
π Electron Effects on Chemical Shift: Overview
 03:51
Molecular Orbital Theory II