比较密度功能理论金属-连接体键解离合体与试验溶液阶段激活对键解离合体的解离合体
Edward N Brothers1, Ashfaq A Bengali2, Giovanni Scalmani1
1Gaussian, Inc., 340 Quinnipiac Street, Wallingford, Connecticut 06492, United States.

密度函数理论 (DFT) 对金属-连接体键解离的预测在43种方法中进行了比较. M11plus表现出色,而M06,r2SCAN和revM11也与实验数据有很好的一致性.
科学领域:
- 计算化学计算化学
- 量子化学 是一个量子化学.
- 材料科学 材料科学 材料科学
背景情况:
- 密度函数理论 (DFT) 对于预测化学反应结果至关重要.
- 之前的研究比较了DFT在溶液相金属-连接体键解离中的激活和反应度的预测.
- 准确的理论预测对于推进化学应用和材料设计至关重要.
研究的目的:
- 为了评估43种不同密度的功能方法的预测准确度,用于金属-连接体键解离度.
- 为了识别DFT函数,准确地模拟金属复合体中的联体解离度.
- 评估分散校正对各种DFT函数的性能的影响.
主要方法:
- 对30个金属-联结体复合物的联结体解离度进行计算分析.
- 系统地比较43种不同的密度函数理论 (DFT) 方法的结果.
- 用和没有经验分散校正的函数的评估.
主要成果:
- 在预测连接体解离度方面,对一些DFT函数发现了显著的不足.
- 针对某些函数的分散校正的应用和有效性,发现了一些挑战.
- M11plus功能显示出出色的性能,准确地预测了解离度.
- 像M06,r2SCAN,M06-L和revM11这样的函数与实验数据很好地一致,没有经验分散校正.
- 当应用分散校正时,包括 ωB97X-D,BLYP,B3LYP,CAM-B3LYP 和 PBE0 在内的功能性表现良好.
结论:
- 选择 DFT 函数对预测金属结合体键解离的准确性产生重大影响.
- 对于这些类型的计算,M11plus成为一个非常准确的函数.
- 其他一些函数提供可靠的预测,有或没有分散校正,这取决于具体的方法.
- 这项研究为在计算化学研究中选择适当的DFT方法提供了有价值的指导.
更多相关视频
 16:11
Thermochemical Studies of NiII and ZnII Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
相关概念视频
 02:13
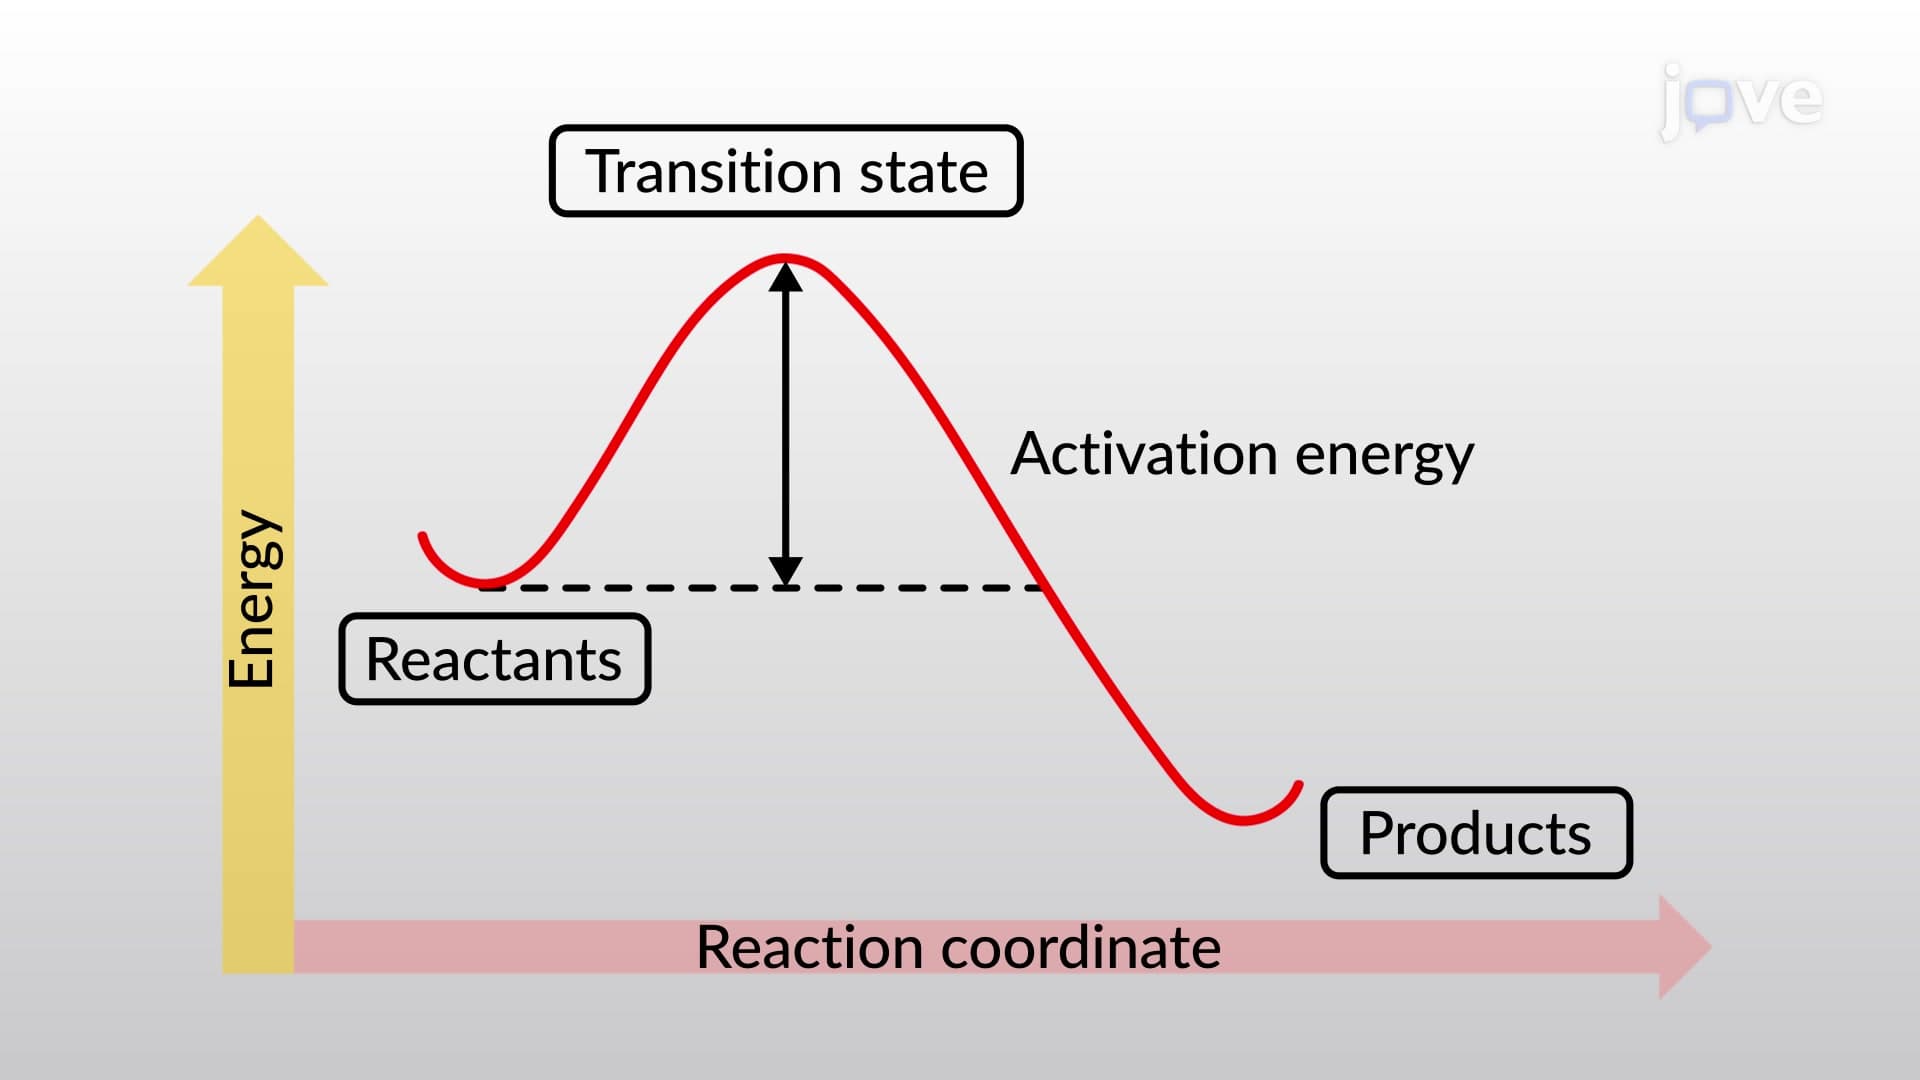
Bond Dissociation Energy and Activation Energy
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:07
Ladder Diagrams: Complexation Equilibria
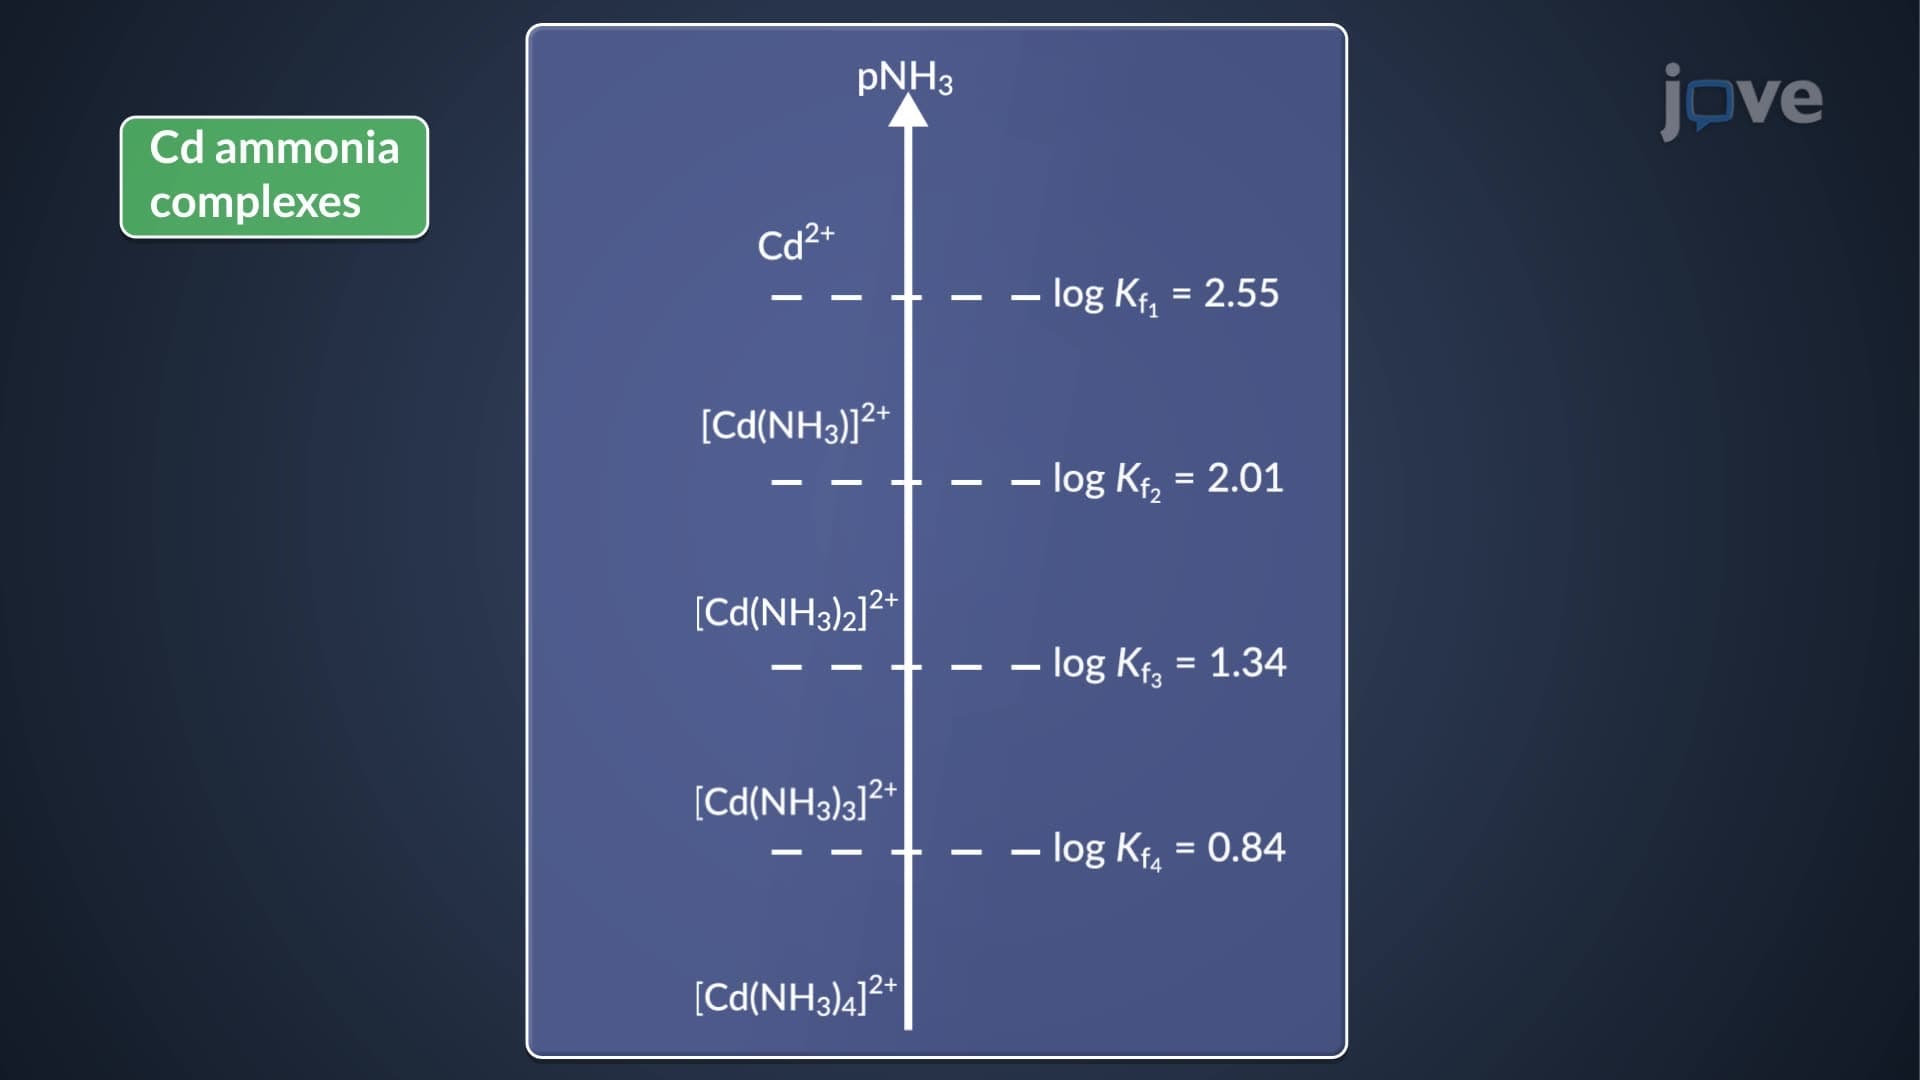
The formation constant, K1, for the formation of Cd(NH3)2+ complex from cadmium and ammonia is 3.55 × 10². Log K1 (i.e. pNH3) is 2.55, and...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:23
Complexation Equilibria: Overview
The equilibrium constant of the complexation reaction is represented as the formation constant...
 02:49
Bond Energies and Bond Lengths