
天然起源的糖和聚醇作为可溶性和溶解增强的载体
Madan Sai Poka1, Marnus Milne1, Anita Wessels2
1Department of Pharmaceutical Sciences, School of Pharmacy, Sefako Makgatho Health Sciences University, Molotlegi Street, Pretoria 0208, South Africa.
Pharmaceutics
|November 25, 2023

概括
糖和多醇可提高药物的溶解性和溶解率在固体分散 (SDs). 这些晶体载体比合成聚合物具有优势,可以避免再结晶和体积问题,从而改善药物输送.
科学领域:
- 制药科学 制药科学
- 药物输送系统 药物输送系统
- 材料科学 材料科学 材料科学
背景情况:
- 合成聚合物通常用于药物:辅助剂固体分散 (SD),但在储存过程中经常导致再结晶问题.
- 高分子量聚合物可以导致最终剂型的粘度和体积增加.
- 一个理想的固体分散载体应该是水友性,非湿透性,具有高结合能力,高玻璃过渡温度 (Tg),并安全使用.
研究的目的:
- 审查在固体分散 (SD) 中使用糖和多作为载体,以提高难溶性药物的可溶性和溶解率.
- 讨论糖和聚醇在SD配方中的优势,而不是合成聚合物.
- 探索过去的成功,最近的进步,以及糖和聚醇在药物输送中的未来潜力.
主要方法:
- 关于在固体分散配方中使用糖和多的研究的文献综述.
- 对各种晶体载体的物理化学性质的分析,包括乳糖,糖糖,银糖,曼尼托尔,索尔比托尔和异.
- 基于糖/聚合物的SD与基于聚合物的SD的性能比较.
主要成果:
- 像德克斯特,糖和曼尼托尔这样的晶体载体可以显著提高SDs中的药物溶解性和溶解率.
- 糖和多醇避免了在聚合物基SD中观察到的再结晶的常见缺点.
- 这些载体为高分子量合成聚合物提供了更安全,更小的替代品.
结论:
- 糖和多醇是开发固体分散 (SD) 的有希望,安全和有效的载体,以提高水溶性较差的药物的可溶性.
- 它们有利的物理化学性质使它们成为合成聚合物的理想替代品,解决了再结晶和配方体积等问题.
- 对糖和聚醇的进一步研究为推进药物输送技术和改善治疗结果提供了重大潜力.
相关概念视频
 01:28
Factors Influencing Drug Absorption: Pharmaceutical Parameters
134
Solid dosage forms such as tablets and capsules undergo rigorous manufacturing processes to ensure stability and effectiveness. Their dissolution and absorption properties are influenced significantly by the choice of excipients (inactive ingredients that serve various roles in the formulation), and the methodology applied during production. The manufacturing parameters, such as compression force and granulation techniques, significantly affect dissolution rates. Elevated compression forces...
134
 01:23
Factors Affecting Dissolution: Particle Size and Effective Surface Area
855
Dissolution kinetics, an essential aspect of oral drug delivery, is significantly influenced by the drug's particle size. According to the Noyes-Whitney dissolution model, the dissolution rate correlates directly with the drug's surface area. The larger the surface area, the higher the drug's solubility in water, leading to a faster drug dissolution rate. Reducing particle size increases the effective surface area, enhancing the dissolution process. Micronization and nanosizing are...
855
 01:21
Factors Affecting Dissolution: Polymorphism, Amorphism and Pseudopolymorphism
311
Polymorphism refers to the existence of a drug substance in multiple crystalline forms, known as polymorphs. Recently, this term has been expanded to include solvates (forms containing a solvent), amorphous forms (non-crystalline forms), and desolvated solvates (forms from which the solvent has been removed).
Some polymorphic crystals possess lower aqueous solubility than their amorphous counterparts, leading to incomplete absorption. For instance, the oral suspension of Chloramphenicol, which...
Some polymorphic crystals possess lower aqueous solubility than their amorphous counterparts, leading to incomplete absorption. For instance, the oral suspension of Chloramphenicol, which...
311
 02:16
Solution Formation
31.7K
There is no one solvent that can dissolve every type of solute. Some substances that readily dissolve in a certain solvent might be insoluble in a different solvent. A simple way to predict which substances dissolve in which solvent is the phrase "like dissolves like". This means that polar substances, such as salt and sugar, dissolve in a polar substance like water. In contrast, non-polar substances are more soluble in non-polar solvents such as carbon tetrachloride.
This selective...
This selective...
31.7K
 01:27
Factors Influencing Drug Absorption: Drug Dissolution
521
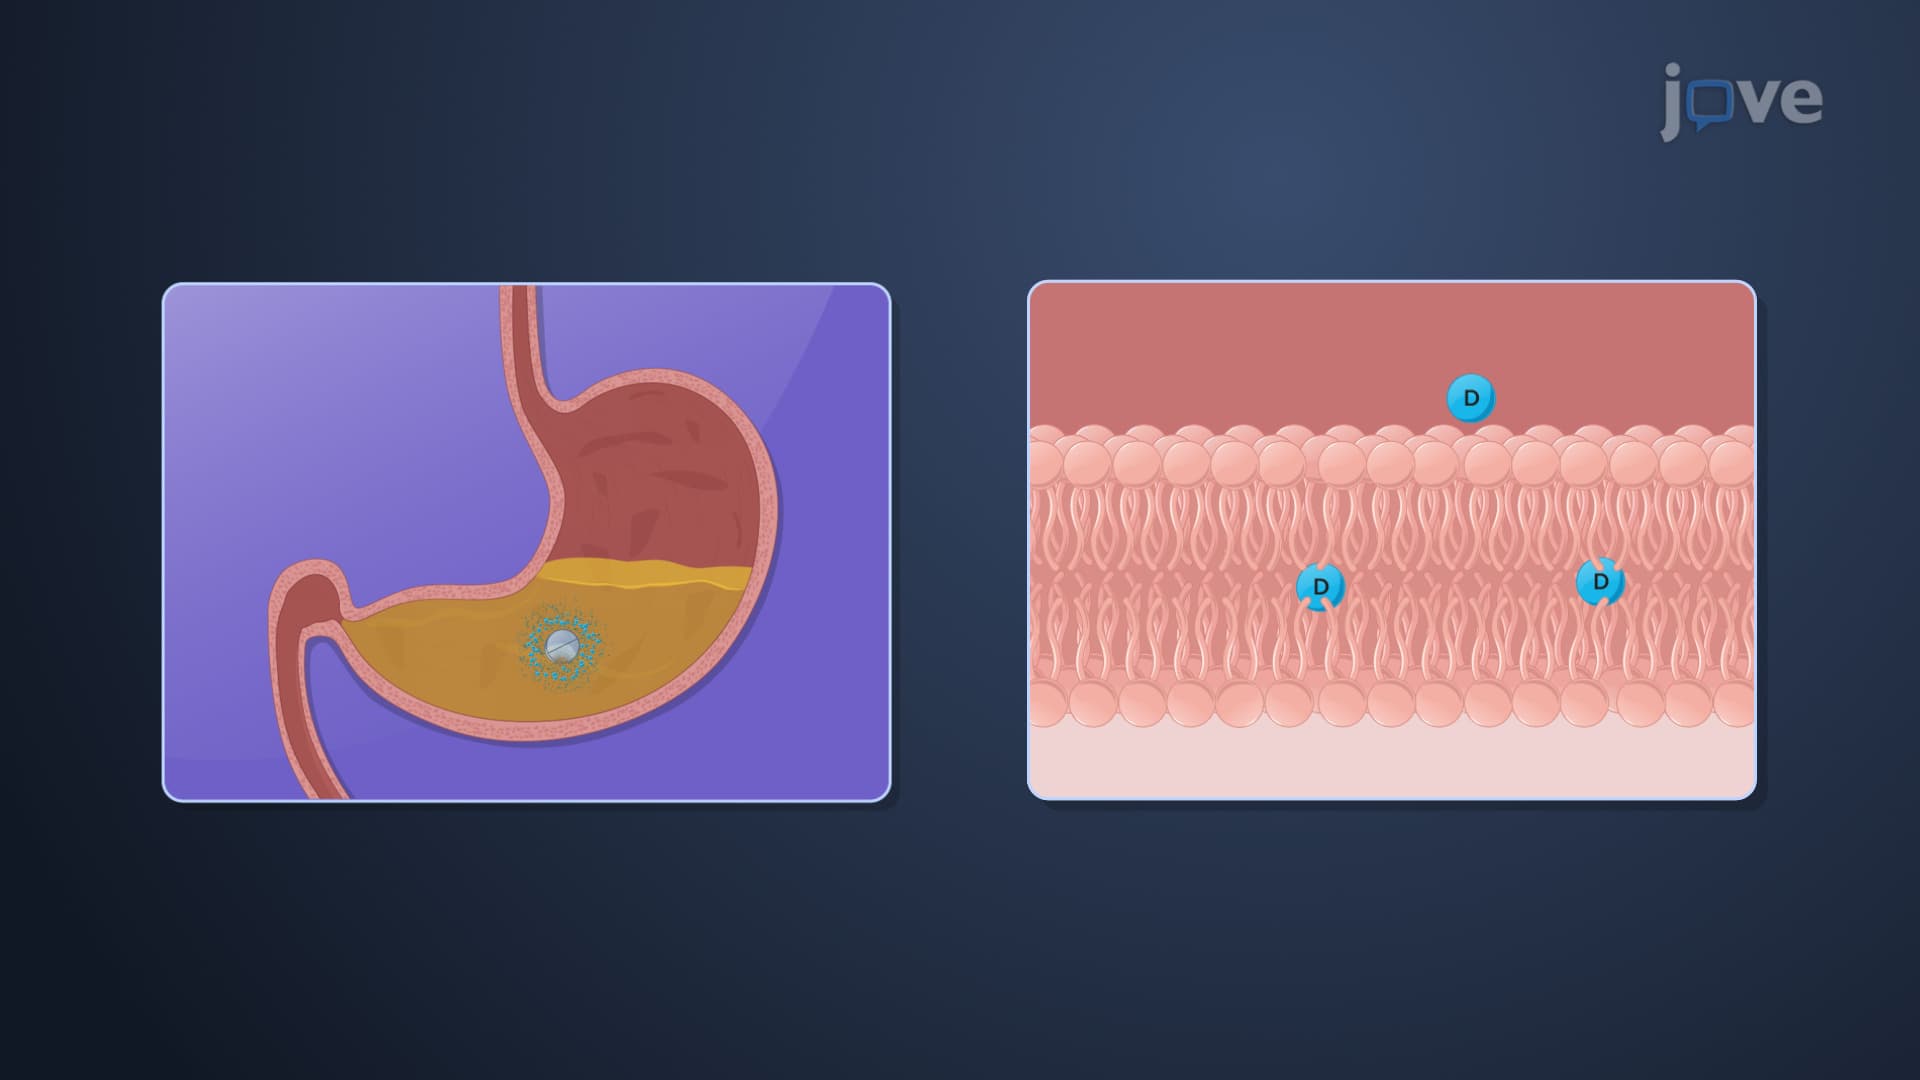
The pharmacokinetic journey of drugs from solid oral dosage forms into systemic circulation is multifaceted. It begins with disintegration, a prerequisite ensuring a solid dosage form's subdivision into minute particles. Dissolution occurs next as these granulated entities solubilize in gastrointestinal fluids. This solubilization is crucial for the succeeding stage, permeation, which describes the traversal of the drug across the intestinal membrane and its subsequent entry into the blood...
521
 01:20
Factors Affecting Dissolution: Drug Permeability, Stability and Stereochemistry
208
Orally administered drugs primarily enter the systemic circulation via passive diffusion through the intestinal membranes. The drug's absorption is influenced by drug stability in the gastrointestinal GI tract, membrane permeability, the surface area available for absorption, luminal drug concentration, and residence time in the lumen. Drug permeability can be enhanced by adjusting the lipophilicity, polarity, or molecular size of the drug, promoting its passive transport across intestinal...
208