
改进了使用完整参考基因组和升空对照的序列映射
Nae-Chyun Chen1, Luis F Paulin2, Fritz J Sedlazeck2,3
1Department of Computer Science, Johns Hopkins University, Baltimore, MD, USA. cnaechy1@jhu.edu.
Nature methods
|November 30, 2023

概括
levioSAM2使得基因组组件在参考之间能够快速准确地进行提升. 将读数对准到像T2T-CHM13这样的高质量组合,并将读数提升到较旧的引用,可以提高变体调用精度,特别是对于医学上相关的基因.
科学领域:
- 基因组学就是基因组学.
- 生物信息学是一种生物信息学.
背景情况:
- 完整的基因组组合,如端粒到端粒 (T2T),提供了增强的分析和变异发现.
- 基本的基因组资源通常与较旧的参考基因组联系在一起,因此需要交叉引用兼容性的方法.
研究的目的:
- 引入levioSAM2,一种新的方法,用于高效精确地提升基因组特征,并读取不同基因组组件之间的对齐.
- 通过利用高质量的T2T引用来证明levioSAM2在提高变量调用准确度方面的实用性.
主要方法:
- 开发和应用levioSAM2,一个基于全基因组图的升空工具.
- 短序和长序的调整可以读取高质量的T2T-CHM13和较旧的GRC引用.
- 在levioSAM2处理的数据和基于标准GRC的映射之间对变异调用准确性的比较分析.
主要成果:
- levioSAM2提供了不同的基因组组件之间的快速和准确的升空.
- 与直接的GRC映射相比,对齐读取到T2T-CHM13和提升到GRC引用可以提高变量调用精度.
- 观察到小和结构变异调用错误的显著减少,特别是对于具有较低质量的GRC引用的复杂,医学相关的基因.
结论:
- levioSAM2 便于基因组数据在参考组件之间进行翻译.
- 使用levioSAM2的高质量的T2T引用可以提高对较旧,广泛使用的引用的变体检测的准确性.
- 该方法为基因组分析提供了实质性的改进,特别是在具有生物和医学意义的区域.
相关概念视频
 03:36
Genome Annotation and Assembly
18.9K
The genome refers to all of the genetic material in an organism. It can range from a few million base pairs in microbial cells to several billion base pairs in many eukaryotic organisms. Genome assembly refers to the process of taking the DNA sequencing data and putting it all back together in a correct order to create a close representation of the original genome. This is followed by the identification of functional elements on the newly assembled genome, a process called genome annotation.
18.9K
 03:21
RNA-seq
10.0K
RNA sequencing, or RNA-Seq, is a high-throughput sequencing technology used to study the transcriptome of a cell. Transcriptomics helps to interpret the functional elements of a genome and identify the molecular constituents of an organism. Additionally, it also helps in understanding the development of an organism and the occurrence of diseases.
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
10.0K
 03:00
Next-generation Sequencing
89.0K
The first human genome sequencing project cost $2.7 billion and was declared complete in 2003, after 15 years of international cooperation and collaboration between several research teams and funding agencies. Today, with the advent of next-generation sequencing technologies, the cost and time of sequencing a human genome have dropped over 100 fold.
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
89.0K
 02:54
Evolutionary Relationships through Genome Comparisons
5.8K
Genome comparison is one of the excellent ways to interpret the evolutionary relationships between organisms. The basic principle of genome comparison is that if two species share a common feature, it is likely encoded by the DNA sequence conserved between both species. The advent of genome sequencing technologies in the late 20th century enabled scientists to understand the concept of conservation of domains between species and helped them to deduce evolutionary relationships across diverse...
5.8K
 01:05
Maxam-Gilbert Sequencing
11.2K
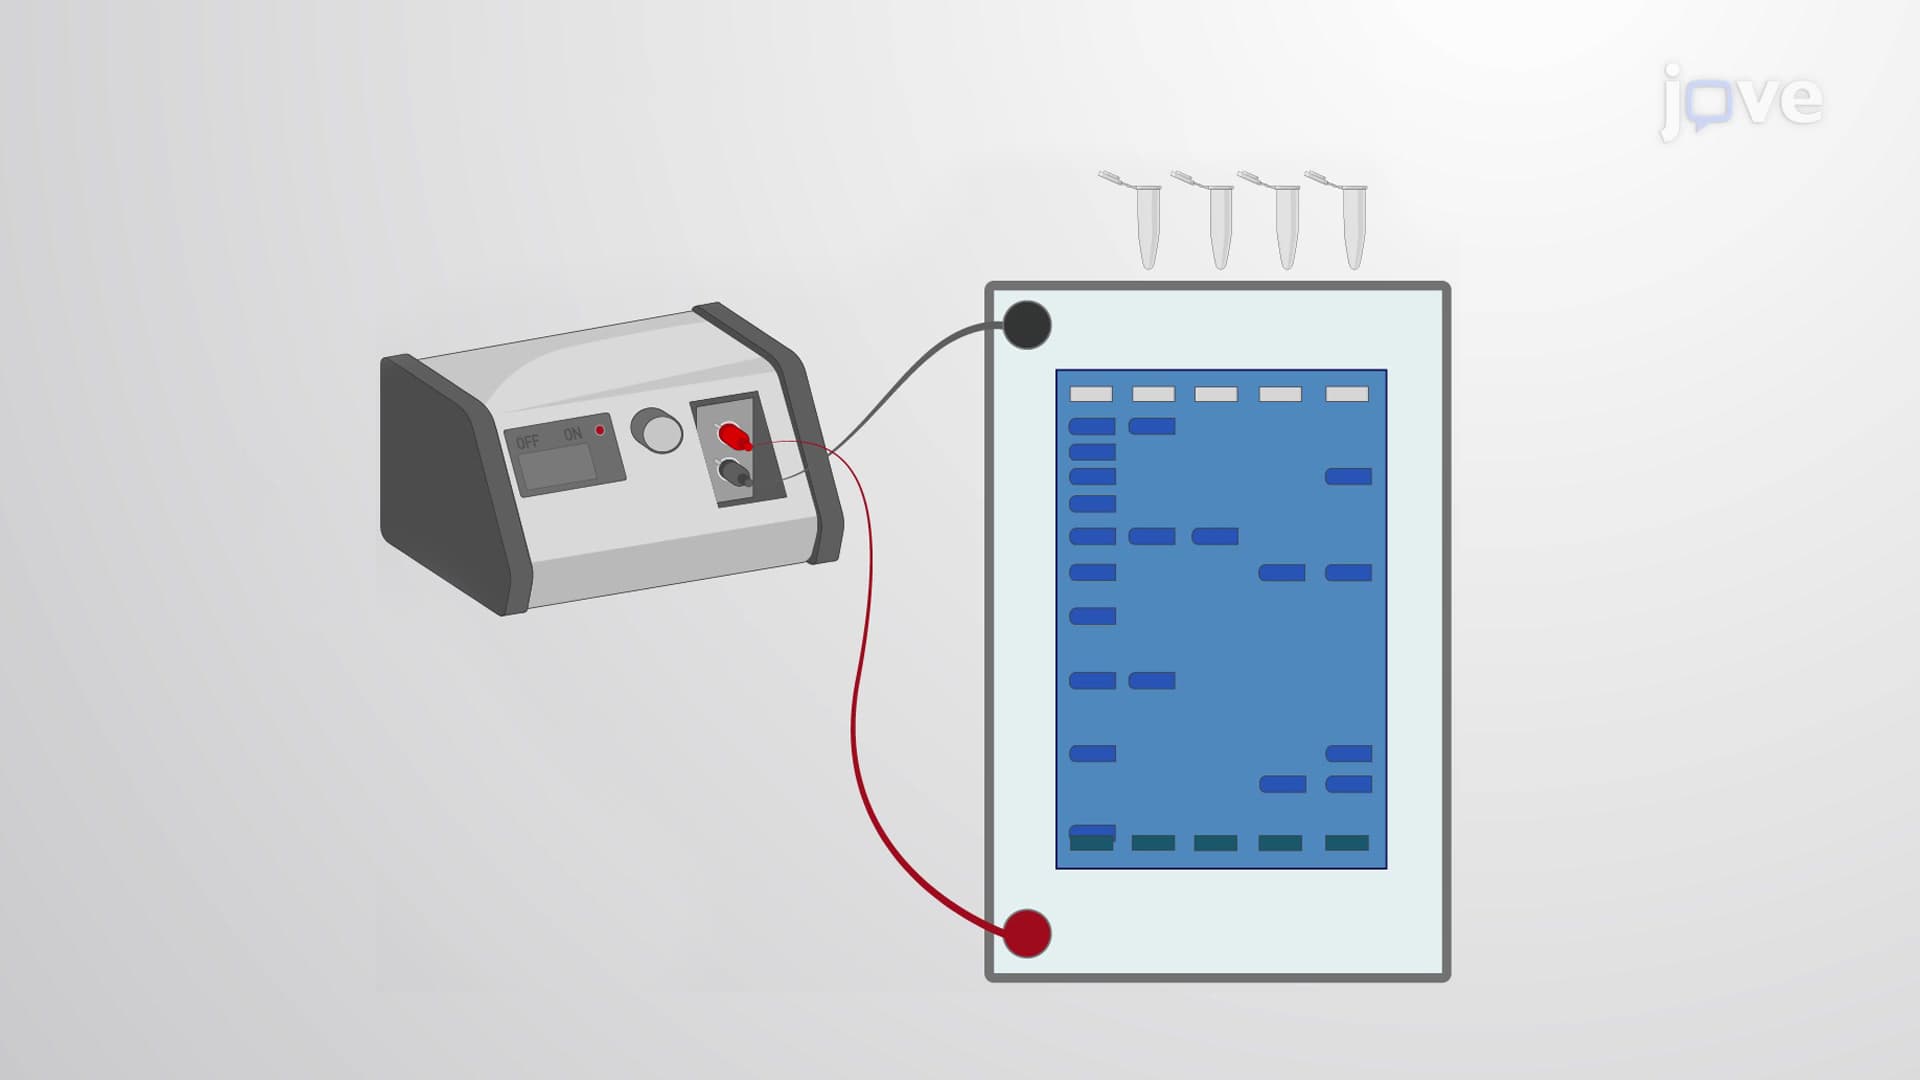
In the same year as the discovery of the Sanger sequencing method, another group of scientists, Allan Maxam and Walter Gilbert, demonstrated their chemical-cleavage method for DNA sequencing. The Maxam-Gilbert method relies on using different chemicals that can cleave the DNA sequence at specific sites, the separation of resulting DNA fragments of variable size using electrophoresis, and deciphering the DNA sequence from the resulting gel bands.
Challenges of the Maxam-Gilbert Method
The...
Challenges of the Maxam-Gilbert Method
The...
11.2K