

灵活的基于DMRG的框架,用于无和的振动计算
Nina Glaser1, Alberto Baiardi1, Markus Reiher1
1Department of Chemistry and Applied Biosciences, ETH Zurich, Vladimir-Prelog-Weg 2, 8093 Zurich, Switzerland.
Journal of chemical theory and computation
|December 7, 2023

概括
我们开发了一种新的n-模式振动密度矩阵重规范化组 (vDMRG) 方法,用于研究无和的分子. 这种方法可以有效地计算像甲基洛西兰这样复杂的系统的振动频率.
科学领域:
- 计算化学的计算化学
- 量子力学就是量子力学.
- 分子光谱学 分子光谱学
背景情况:
- 准确预测分子振动频率对于理解分子特性和反应性至关重要.
- 强烈不和的分子和高维的潜在能量表面 (PES) 对传统的计算方法构成重大挑战.
- 振动密度矩阵重规范化组 (vDMRG) 是一个强大的工具,但它对一般无调系统的应用有限.
研究的目的:
- 引入振动密度矩阵重规范化组 (vDMRG) 算法的新配方.
- 扩展vDMRG框架以处理一般的,高维的潜在能量表面 (PES) 和无振动的哈密尔顿.
- 为了能够有效地计算无和的过渡频率,包括激发状态.
主要方法:
- 开发了振动哈密尔顿的n-模式第二量子化形式主义.
- 实施了一种n-mode vDMRG方法,在 PES 功能形式和单粒子基础集中提供灵活性.
- 通过振动自相一致场 (vSCF) 算法优化的一组无和的模态基础组合的n模式vDMRG.
- 嵌入激发状态准算法用于过渡频率计算.
主要成果:
- 在n-模式的vDMRG方法成功地处理一般的,高维的不和的潜在能量表面.
- 首次应用vDMRG使用优化和的模态基础设置在飞机上构建的PES.
- 证明了该方法对甲基洛克西兰 (methyloxirane) 的能力,甲基洛克西兰是一种具有24种合振动模式的分子,计算了无和的过渡频率.
结论:
- 新的n-mode vDMRG框架为研究强烈不和的分子振动提供了灵活和高效的方法.
- 这种方法克服了以前的vDMRG配方的局限性,使复杂分子系统的准确计算成为可能.
- 对甲基洛克西兰的证明应用突显了n模式vDMRG在推进分子光谱学和计算化学方面的潜力.


相关概念视频
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
1.3K
A covalently bonded heteronuclear diatomic molecule can be modeled as two vibrating masses connected by a spring. The vibrational frequency of the bond can be expressed using an equation derived from Hooke's law, which describes how the force applied to stretch or compress a spring is proportional to the displacement of the spring. In this case, the atoms behave like masses, and the bond acts like a spring.
According to Hooke's law, the vibrational frequency is directly proportional to...
According to Hooke's law, the vibrational frequency is directly proportional to...
1.3K
 01:24
IR Spectroscopy: Molecular Vibration Overview
2.3K
When Infrared (IR) radiation passes through a covalently bonded molecule, the bonds transition from lower to higher vibrational levels. The fundamental vibrational motions that result in infrared absorption can be classified as stretching or bending vibrations.
Stretching vibrations are vibrational motions that occur along the bond line, changing the bond length or distance between two bonded atoms. They are further distinguished as symmetric or asymmetric. In symmetric stretching, the...
Stretching vibrations are vibrational motions that occur along the bond line, changing the bond length or distance between two bonded atoms. They are further distinguished as symmetric or asymmetric. In symmetric stretching, the...
2.3K
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
1.1K
The axial and equatorial protons in cyclohexane can be distinguished by performing a variable-temperature NMR experiment. In this process, except for one proton, the remaining eleven protons are replaced by deuterium. The deuterium substitution avoids the possible peak splitting caused by the spin-spin coupling between the adjacent protons. The remaining proton flips between the axial and equatorial positions.
1.1K
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
852
At room temperature, the chair conformer of cyclohexane undergoes rapid ring flipping between two equivalent chair conformers at a rate of approximately 105 times per second. These two chair conformers are in equilibrium. The rapid ring flipping results in the interconversion of the axial proton to an equatorial proton and an equatorial to the axial proton. Such interconversions are too rapid and cannot be detected on the NMR timescale. Hence, the NMR spectrometer cannot distinguish between the...
852
 01:08
IR Spectrum Peak Splitting: Symmetric vs Asymmetric Vibrations
1.0K
Identical bonds within a polyatomic group can stretch symmetrically (in-phase) or asymmetrically (out-of-phase). Similar to hydrogen bonding, these vibrations also influence the shape of the IR peak. Generally, asymmetric stretching frequencies are higher than symmetric stretching frequencies. For example, primary amines exhibit two distinct IR peaks between 3300–3500 cm−1 corresponding to the symmetric and asymmetric N-H stretching, while secondary amines exhibit a single...
1.0K
 01:10
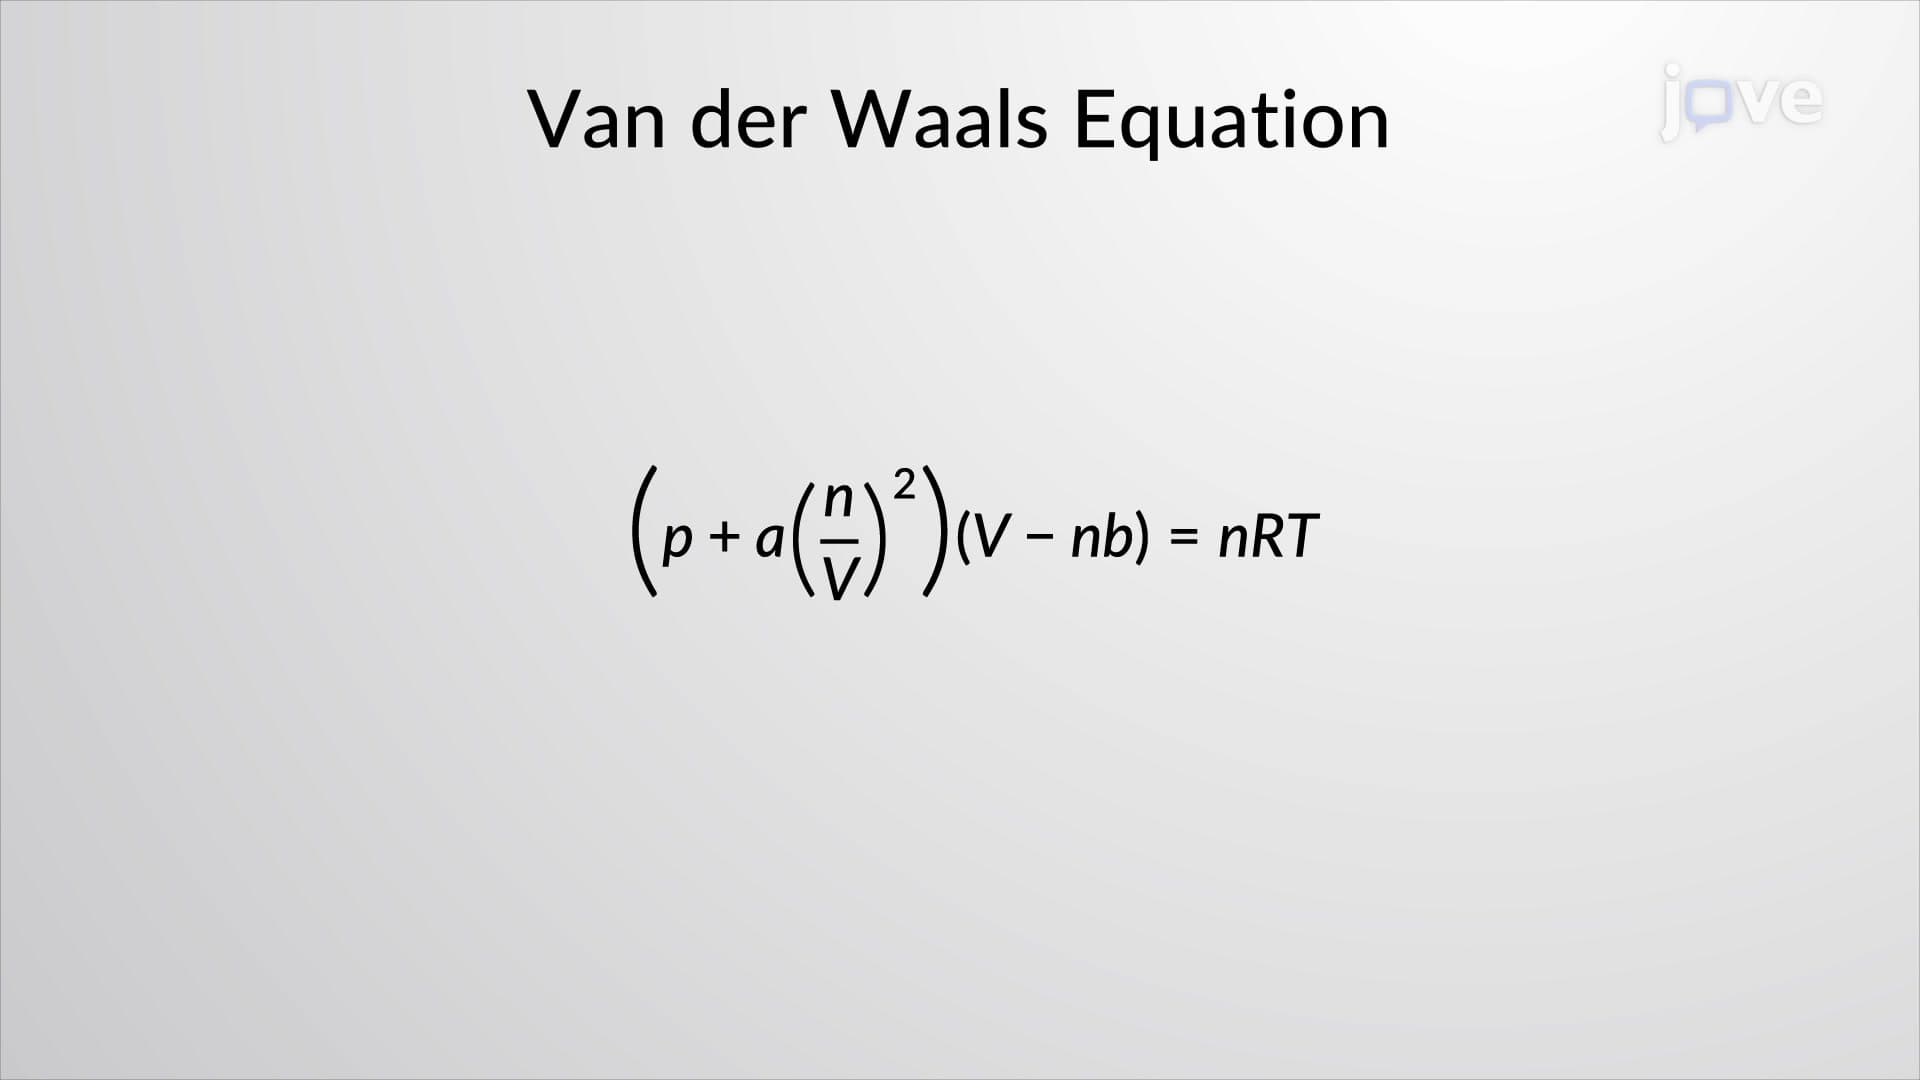
Van der Waals Equation
4.1K
The ideal gas law is an approximation that works well at high temperatures and low pressures. The van der Waals equation of state (named after the Dutch physicist Johannes van der Waals, 1837−1923) improves it by considering two factors.
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
4.1K