

对于NO A2Σ+ + CO2X1Σg+碰撞复合物的兴奋状态范德瓦尔斯潜在能量表面
Luca Craciunescu1, Eirik M Liane2, Adam Kirrander2
1Institute of Chemical Sciences, School of Engineering and Physical Sciences, Heriot-Watt University, Edinburgh, Scotland EH14 4AS, United Kingdom.

这项研究使用先进的计算方法揭示了激发NO + CO2相互作用中的更深的范德瓦尔斯井. 这些发现支持了分子系统中能量转移的实验观测.
科学领域:
- 化学物理 化学物理
- 计算化学计算化学
- 频谱学是一种光谱学.
背景情况:
- 了解分子间力量对于预测分子相互作用和能量转移至关重要.
- NO A2Σ+ + CO2 系统是研究激发状态动态和范德瓦尔斯相互作用的关键模型.
- 准确的潜在能量表面 (PES) 对于解释实验数据至关重要.
研究的目的:
- 计算和分析NO A2Σ+ + CO2 X1Σg+系统的兴奋状态范德瓦尔斯潜在能量表面 (PES).
- 为了比较对自然轨道合集群单双和双双与扰动三重 (PNO-CCSD(T)) 与传统合集群方法的准确性和效率.
- 为了研究多参考效应的作用,使用完整的第二阶活性空间扰动理论 (CASPT2).
主要方法:
- 使用合集群理论,特别是PNO-CCSD (T),用于准确计算分子特性和范德瓦尔斯最小值.
- 采用高度扩散的基础集和对权衡校正来减轻基础集叠加错误.
- 执行CASPT2计算,以评估多引用对PES的影响.
主要成果:
- PNO-CCSD (T) 提供与CCSD (T) 相同的准确性,计算成本降低.
- 在激发状态的PES上发现了许多范德瓦尔斯井,其深度高达830厘米-1,比地面状态深得多.
- 计算的远程vdW表面表现出异质性,支持关于旋转能量转移的实验发现.
结论:
- 开发的计算方法提供了激发状态vdW-PESs的准确和高效的表征.
- 较深的和异型的vdW井在NO + CO2系统的动态和能量转移中发挥着重要作用.
- 这些结果增强了对激发电子状态中的分子间相互作用的理解.
更多相关视频
 07:53
Analysis of Complex Molecules and Their Reactions on Surfaces by Means of Cluster-Induced Desorption/Ionization Mass Spectrometry
Published on: March 1, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
相关概念视频
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 01:10
Van der Waals Equation
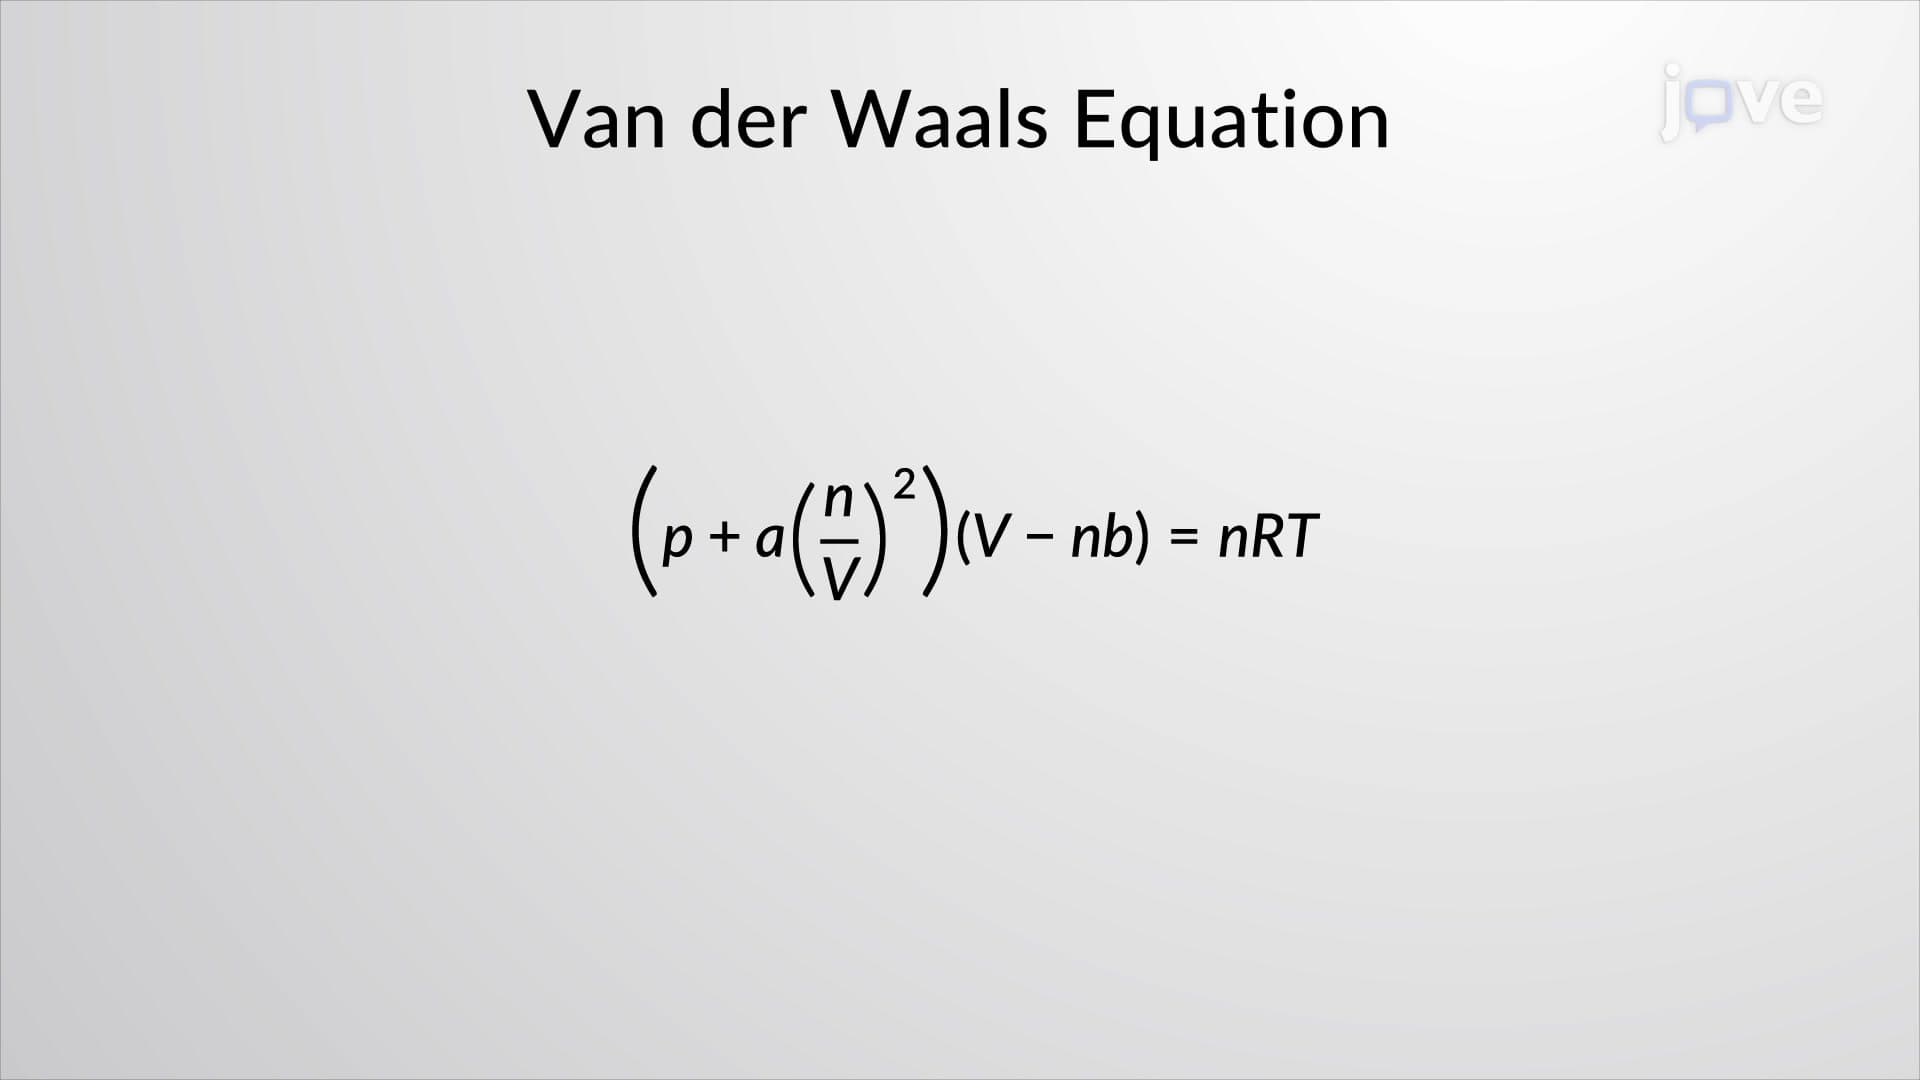
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:24
Van der Waals Interactions
 02:42
Valence Bond Theory
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 03:51
Molecular Orbital Theory II