
优化符合人群的陷,以最大限度地提高预测和实验性手术光谱之间的相似性
Cody L Covington1,2, Andrew R Puente2, Prasad L Polavarapu2
1Department of Chemistry, Austin Peay State University, Clarksville, Tennessee 37044, United States.
The journal of physical chemistry. A
|December 28, 2023

概括
对泛托拉克和其他分子的手术光谱的相似性分析揭示了局限性. 从光谱相似性获得的符合性群体并不比基于能量的方法更准确.
科学领域:
- 计算化学的计算化学
- 频谱学是一种光谱学.
- 分子形状分析 分子形状分析
背景情况:
- 确定精确的分子构造对于理解化学性质至关重要.
- 石眼光谱学,包括振动圆形二重化 (VCD) 和拉曼光学活性 (ROA),为构造分析提供实验数据.
- 计算方法通常用于预测光谱并将其与实验结果进行比较.
研究的目的:
- 调查计算和实验手术光谱之间的相似性分析的可靠性,以确定形状群体.
- 通过光谱相似性优化与基于能量的博尔茨曼群体获得的构造群体进行比较.
- 评估不同分子和手术技术之间的相似性分析的稳定性.
主要方法:
- 计算的手术光谱 (VCD和ROA) 对于潘托拉克,长水素和N-乙-三甲基.
- 对相同分子的实验性手术光谱.
- 类似性分析比较计算和实验光谱,以确定形状群体.
主要成果:
- 对于泛托拉克顿,从VCD和ROA相似性分析中得出的最佳符合组不一致,这表明光谱优化存在局限性.
- 通过优化光谱相似性获得的符合性群体没有比基于标准能量的博尔兹曼群体更准确.
- 删除单个对应器通常只会导致光谱相似性的微小变化,这表明多个对应器集可以产生可接受的匹配.
结论:
- 石眼光谱的相似性分析不是准确确定分子形状群体的最终方法.
- 光谱配合的模糊性意味着多个构造分布可以重现实验数据,挑战独特的确定.
- 与光谱相似性优化相比,基于能源的方法仍然是估计构型群体的更可靠方法.
相关概念视频
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
847
At room temperature, the chair conformer of cyclohexane undergoes rapid ring flipping between two equivalent chair conformers at a rate of approximately 105 times per second. These two chair conformers are in equilibrium. The rapid ring flipping results in the interconversion of the axial proton to an equatorial proton and an equatorial to the axial proton. Such interconversions are too rapid and cannot be detected on the NMR timescale. Hence, the NMR spectrometer cannot distinguish between the...
847
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
1.0K
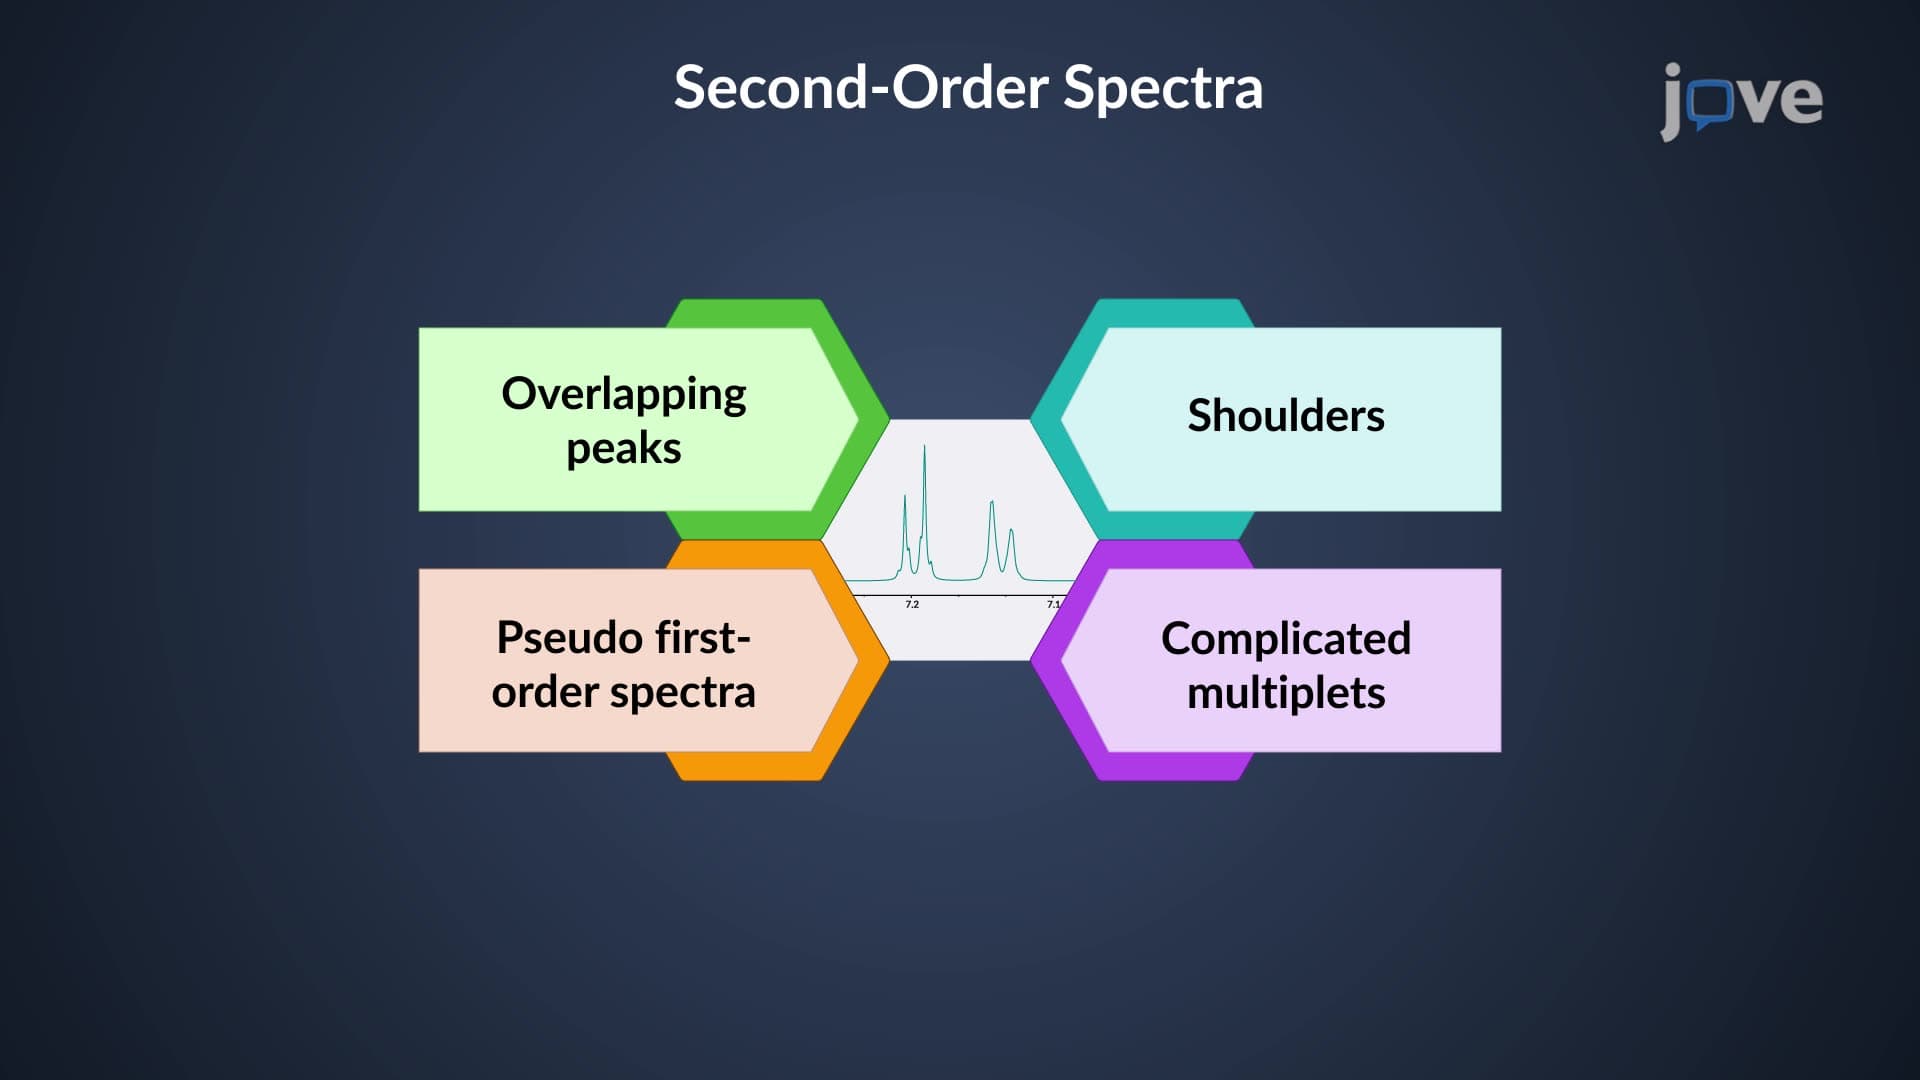
Spin systems where the difference in chemical shifts of the coupled nuclei is greater than ten times J are called first-order spin systems. These nuclei are weakly coupled, and their chemical shifts and coupling constant can generally be estimated from the well-separated signals in the spectrum.
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are...
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are...
1.0K
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
1.1K
The axial and equatorial protons in cyclohexane can be distinguished by performing a variable-temperature NMR experiment. In this process, except for one proton, the remaining eleven protons are replaced by deuterium. The deuterium substitution avoids the possible peak splitting caused by the spin-spin coupling between the adjacent protons. The remaining proton flips between the axial and equatorial positions.
1.1K
 01:03
¹H NMR Chemical Shift Equivalence: Homotopic and Heterotopic Protons
2.4K
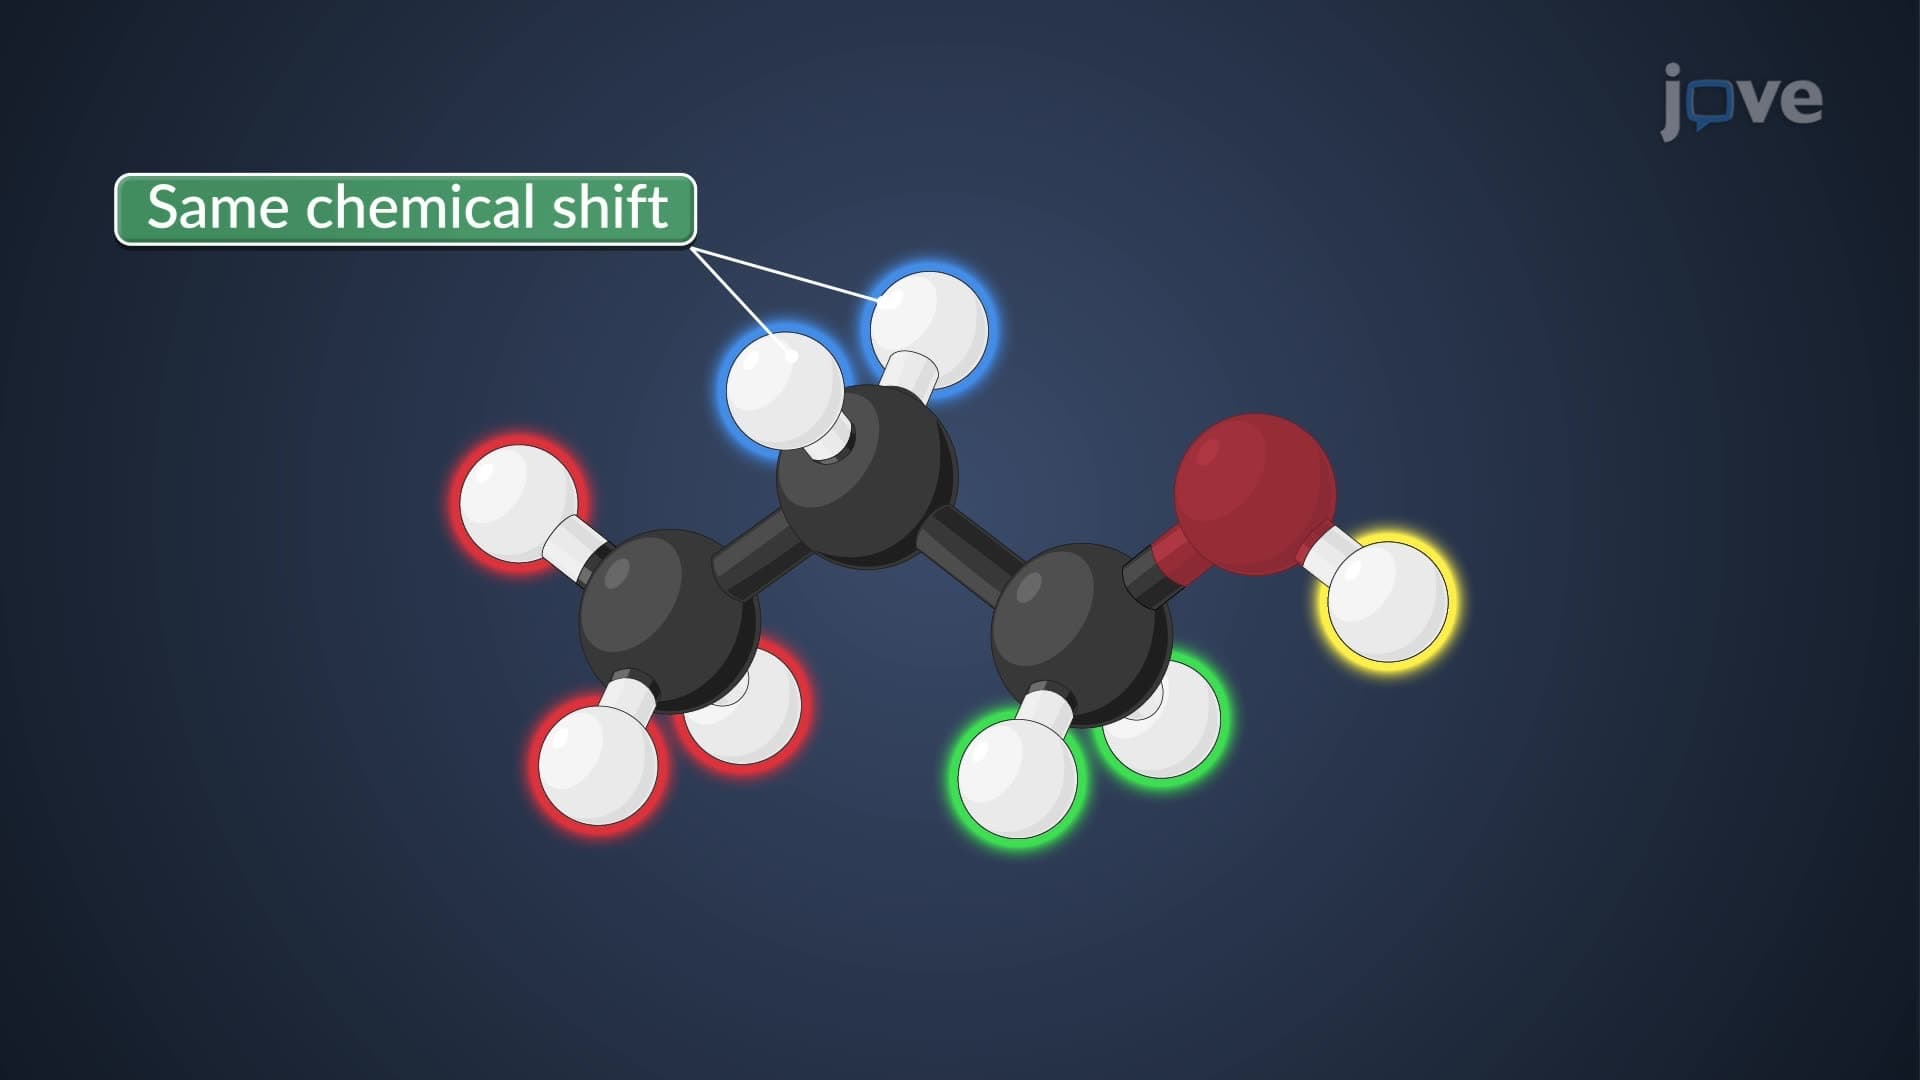
Protons in identical electronic environments within a molecule are chemically equivalent and have the same chemical shift. The replacement test is a useful tool to identify chemical equivalence and predict NMR spectra. A substituent replaces each of the protons being examined and the resulting molecules are compared. If the same molecule is obtained, the protons are equivalent or homotopic. Replacement of any hydrogens in ethane by chlorine yields chloroethane because all six protons are...
2.4K
 02:11
Conformations of Cyclohexane
12.5K
Cyclohexane does not exist in a planar form due to the high angle and torsional strain it would experience in the planar structure. Instead, it adopts non-planar chair and boat conformations.
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal...
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal...
12.5K
 02:29
Conformations of Cycloalkanes
11.8K
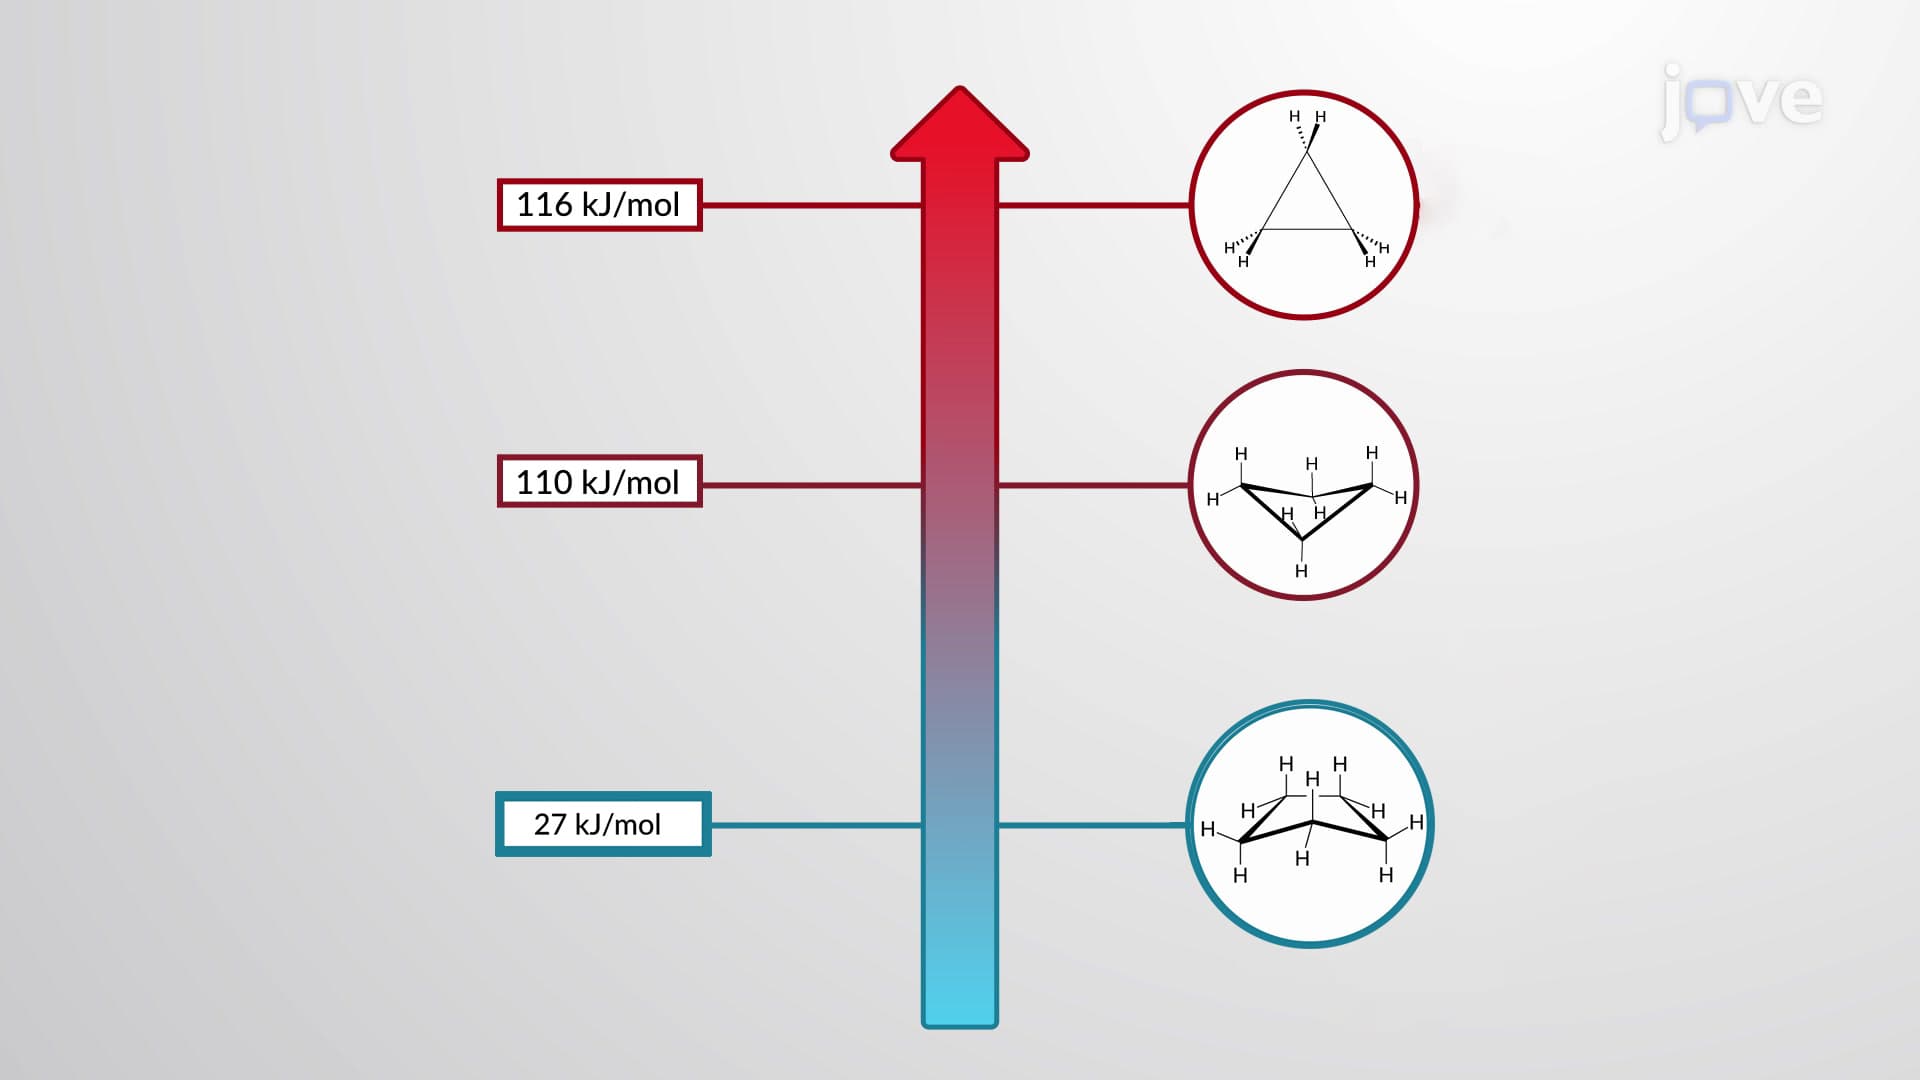
Adolf von Baeyer attempted to explain the instabilities of small and large cycloalkane rings using the concept of angle strain — the strain caused by the deviation of bond angles from the ideal 109.5° tetrahedral value for sp3 hybridized carbons. However, while cyclopropane and cyclobutane are strained, as expected from their highly compressed bond angles, cyclopentane is more strained than predicted, and cyclohexane is virtually strain-free. Hence, Baeyer’s theory that...
11.8K