

模拟使用刚性体的水的MD描述需要一个小的时间步骤,以确保Equipartition
Dilipkumar N Asthagiri1, Thomas L Beck1
1Oak Ridge National Laboratory, One Bethel Valley Road, Oak Ridge, Tennessee 37830-6012, United States.

在水模拟中使用更长的时间步骤会破坏分子运动类型之间的能量平衡. 较短的时间步骤对于准确捕捉水系统中的旋转动力学和热力学特性至关重要.
科学领域:
- 计算化学和物理计算化学和物理
- 分子动力学模拟的模拟.
- 水性系统是水性系统.
背景情况:
- 水系统的模拟通常会结水中的结合振动和角度曲.
- 这允许更长的集成时间步骤 (δt),通常使用δt = 2 fs用于刚性水模型.
- 假设是结振动简化了计算,没有显著的物理影响.
研究的目的:
- 为了研究变化的时间步骤 (δt) 对水分子动力学模拟中的能量均等分配的影响.
- 了解转换和旋转模式之间没有均等分配的原因.
- 评估时间步骤对旋转放松动态和热力学特性的影响.
主要方法:
- 模拟了水的SPC/E模型,使用从0.5 fs到3.0 fs的时间步骤范围,并使用质量再分区进行高达4 fs的时间步骤.
- 分析了转换质心速度和角速度的自相关性.
- 检查了波动-散流关系及其对能量分配和热力学属性的影响.
主要成果:
- 对于 δt ≥ 0.5 fs,转换模式和旋转模式之间的均分并未实现,旋转模式显示较低的温度.
- 旋转放松发生在与振动周期相似的时间尺度上,挑战了结振动的逻辑.
- 时间步骤 δt ≥ 1 fs 不准确地捕捉快速旋转放松,有效地减慢它,并影响热力学特性,如潜在能量和过量的水合.
结论:
- 时间步骤的选择显著影响了对水的分子动力学模拟的准确性,特别是关于旋转动力学和能量分配的精确性.
- 较短的时间步骤 (例如,0.5 fs) 是必要的,以准确地捕捉快速旋转放松,并确保可靠的热力学属性计算.
- 该研究强调了考虑波动的时间演变的重要性,以及在平衡分子动力学中时间步骤选择所带来的局限性.
更多相关视频
 07:31
Author Spotlight: Advancing Cell Membrane Biophysics - Exploring Interactions and Challenges Through Experimental and Computational Approaches
Published on: September 1, 2023
 06:37
Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
相关概念视频
 01:08
Molecular Weight of Step-Growth Polymers
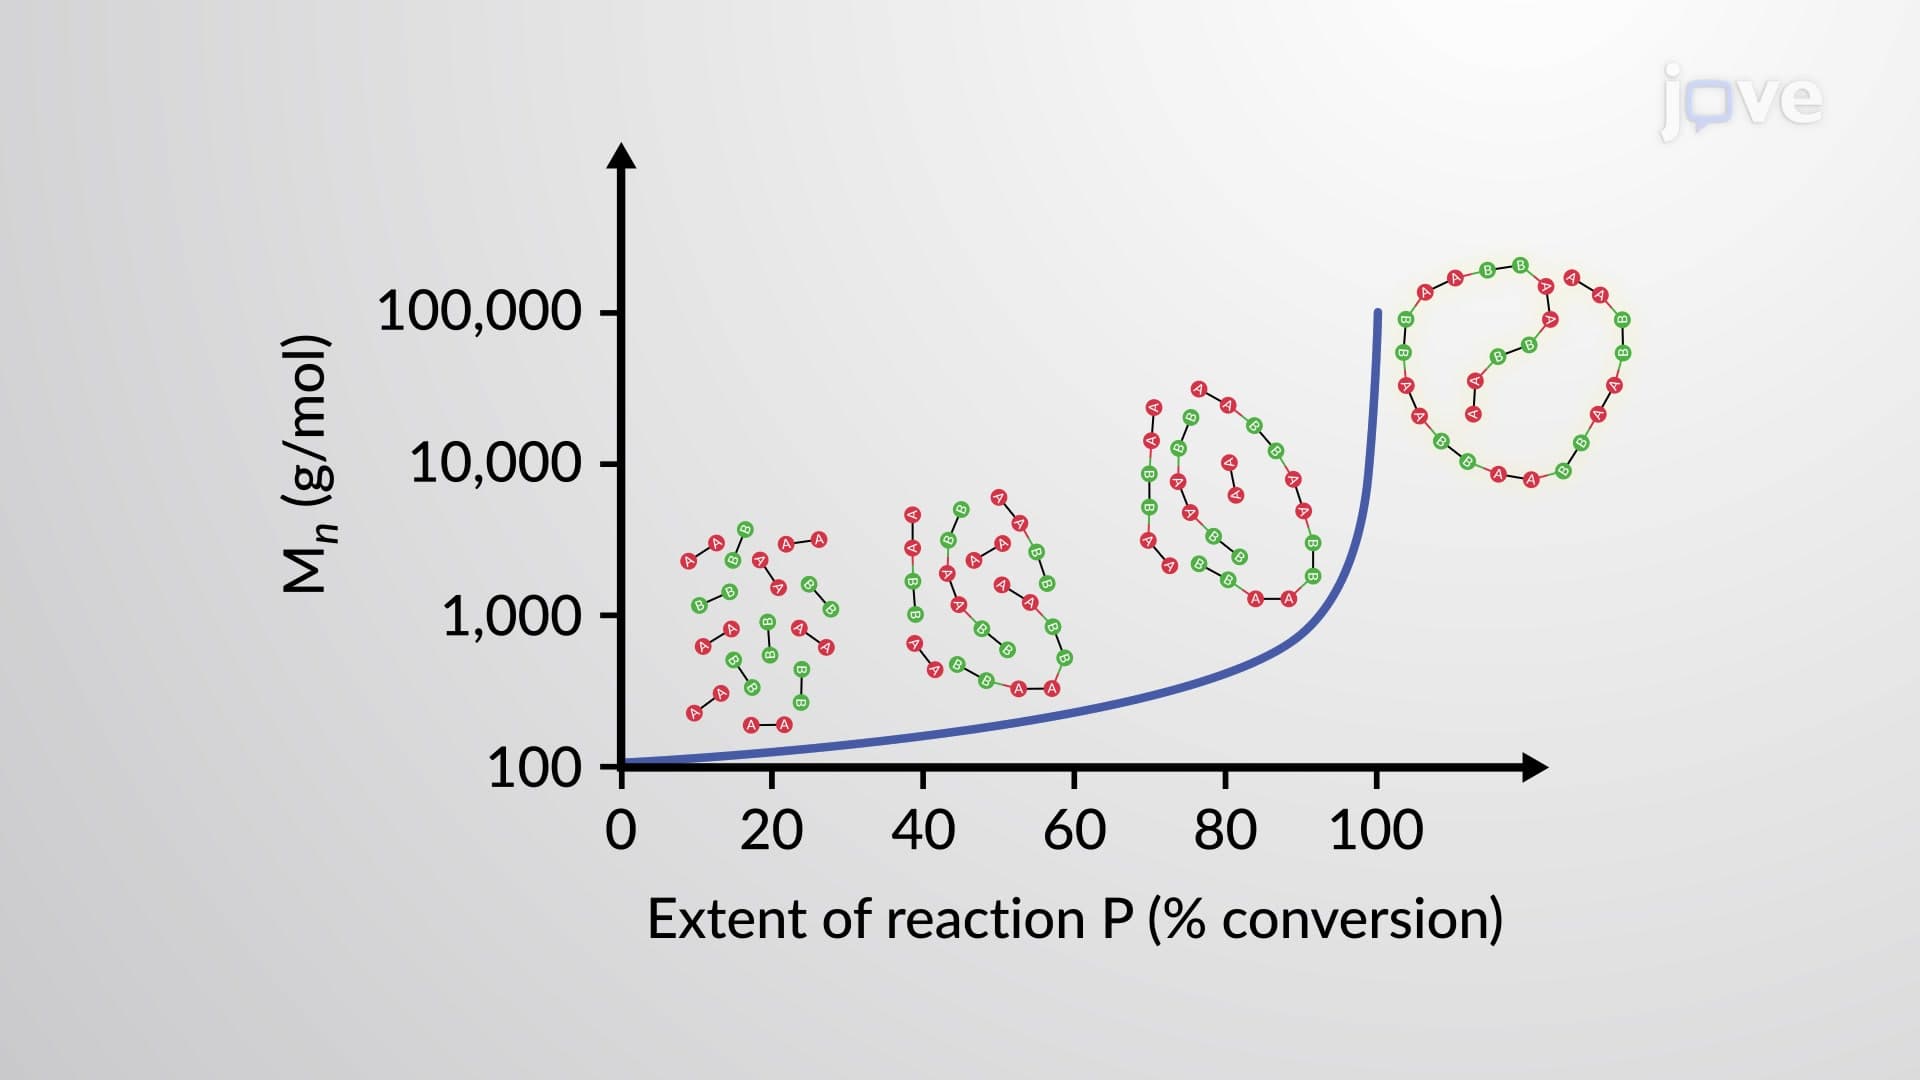
As the step-growth polymerization involves step-wise condensation of monomers, the molecular weight also builds up eventually. Consequently, high molecular weight polymers are obtained at the late stages of the polymerization, where 99% of monomers have been consumed.
The extent of the...
 01:22
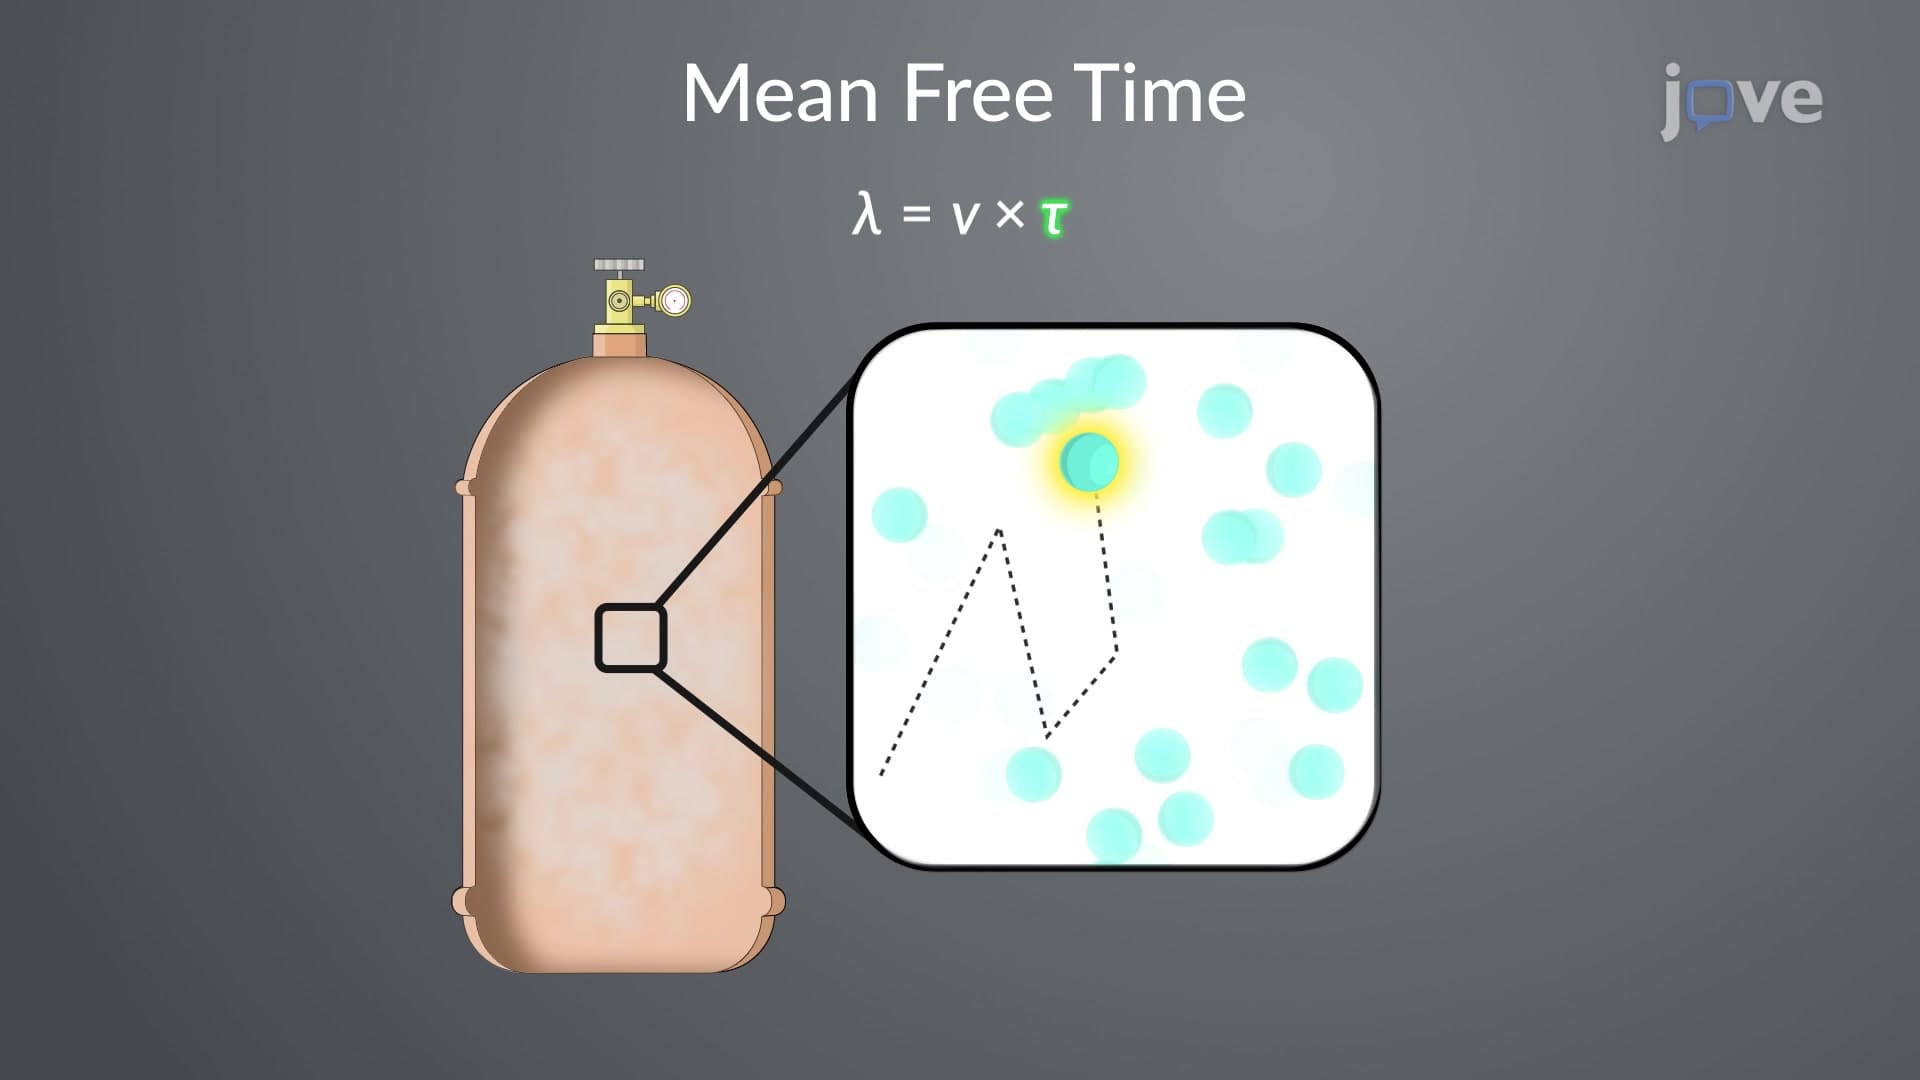
Mean free path and Mean free time
 01:27
Distribution of Molecular Speeds
 01:30
Typical Model Studies
 01:20
Maxwell-Boltzmann Distribution: Problem Solving
This distribution function f(v) is defined by saying that the expected number N (v1,v2) of particles with speeds between v1 and v2 is given by
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution