在氧化α-SiO2(001) 表面上多核U(VI) 物种的结构:从DFT计算中的见解
Pengyuan Gao1,2,3, Qiang Jin1,2, Zongyuan Chen1,2
1Radiochemistry Laboratory, School of Nuclear Science and Technology, Lanzhou University, 730000 Lanzhou, China. guozhj@lzu.edu.cn.
Physical chemistry chemical physics : PCCP
|January 16, 2024

概括
这项研究揭示了多核 (VI) 物种如何与二氧化表面结合. 密度函数理论计算显示静电相互作用和化学键驱动吸附,影响.
科学领域:
- 地质化学 地质化学
- 环境科学 环境科学
- 计算化学的计算化学
背景情况:
- 多核 (VI) 种在水性环境中普遍存在.
- 了解它们与矿物表面的相互作用对于环境修复和核废物管理至关重要.
- 这些物种在矿物表面的特异性结构和结合是不确定的.
研究的目的:
- 为了研究多核的结构和结合性质的多核 (VI) 吸附在氧化α-SiO2 (001) 表面的物种.
- 阐明主要的吸附配置和复杂形成背后的驱动力.
- 为了提供原子层面的洞察力,对在二氧化上的封存.
主要方法:
- 使用密度函数理论 (DFT) 的计算.
- 模拟集中在三个特定的多核 (VI) 物种上: (UO2) 2 (OH) 3+, (UO2) 2 (OH) 22+,和 (UO2) 3 (O) (OH) 3+.
- 分析包括结合能,轨道杂交和键相互作用.
主要成果:
- 该研究确定了不同 (VI) 种在特定的表面部位上具有不同的吸附模式 (端对与侧对).
- (UO2) 2 ((OH) 22+和 (UO2) 3 ((O) ((OH) 3+更喜欢侧向结构,而 (UO2) 2 ((OH) 3+更喜欢侧向配置.
- 静电相互作用和部分共价键,特别是涉及U-5f和U-6d轨道与表面氧 (O_s)-2p轨道,是关键的结合力. 结合强度在类似的位点下降的顺序是 (UO2) 2 (((OH) 3+ > (UO2) 2 ((OH) 22+ > (UO2) 3 (((O) ((OH) 3+).
结论:
- 多核 (VI) 物种在吸附到二氧化表面时表现出特定的结构偏好.
- 吸附是由静电力,化学键和键的组合控制的.
- DFT计算为在矿物表面的复杂化行为提供了宝贵的见解,有助于预测其环境命运.
更多相关视频
 12:05
U2O5 Film Preparation via UO2 Deposition by Direct Current Sputtering and Successive Oxidation and Reduction with Atomic Oxygen and Atomic Hydrogen
Published on: February 21, 2019
8.1K
 08:18
Microscopic Visualization of Porous Nanographenes Synthesized through a Combination of Solution and On-Surface Chemistry
Published on: March 4, 2021
1.8K
相关概念视频
 02:42
Valence Bond Theory
8.6K
Coordination compounds and complexes exhibit different colors, geometries, and magnetic behavior, depending on the metal atom/ion and ligands from which they are composed. In an attempt to explain the bonding and structure of coordination complexes, Linus Pauling proposed the valence bond theory, or VBT, using the concepts of hybridization and the overlapping of the atomic orbitals. According to VBT, the central metal atom or ion (Lewis acid) hybridizes to provide empty orbitals of suitable...
8.6K
 02:42
Ionic Crystal Structures
14.3K
Ionic crystals consist of two or more different kinds of ions that usually have different sizes. The packing of these ions into a crystal structure is more complex than the packing of metal atoms that are the same size.
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
14.3K
 01:29
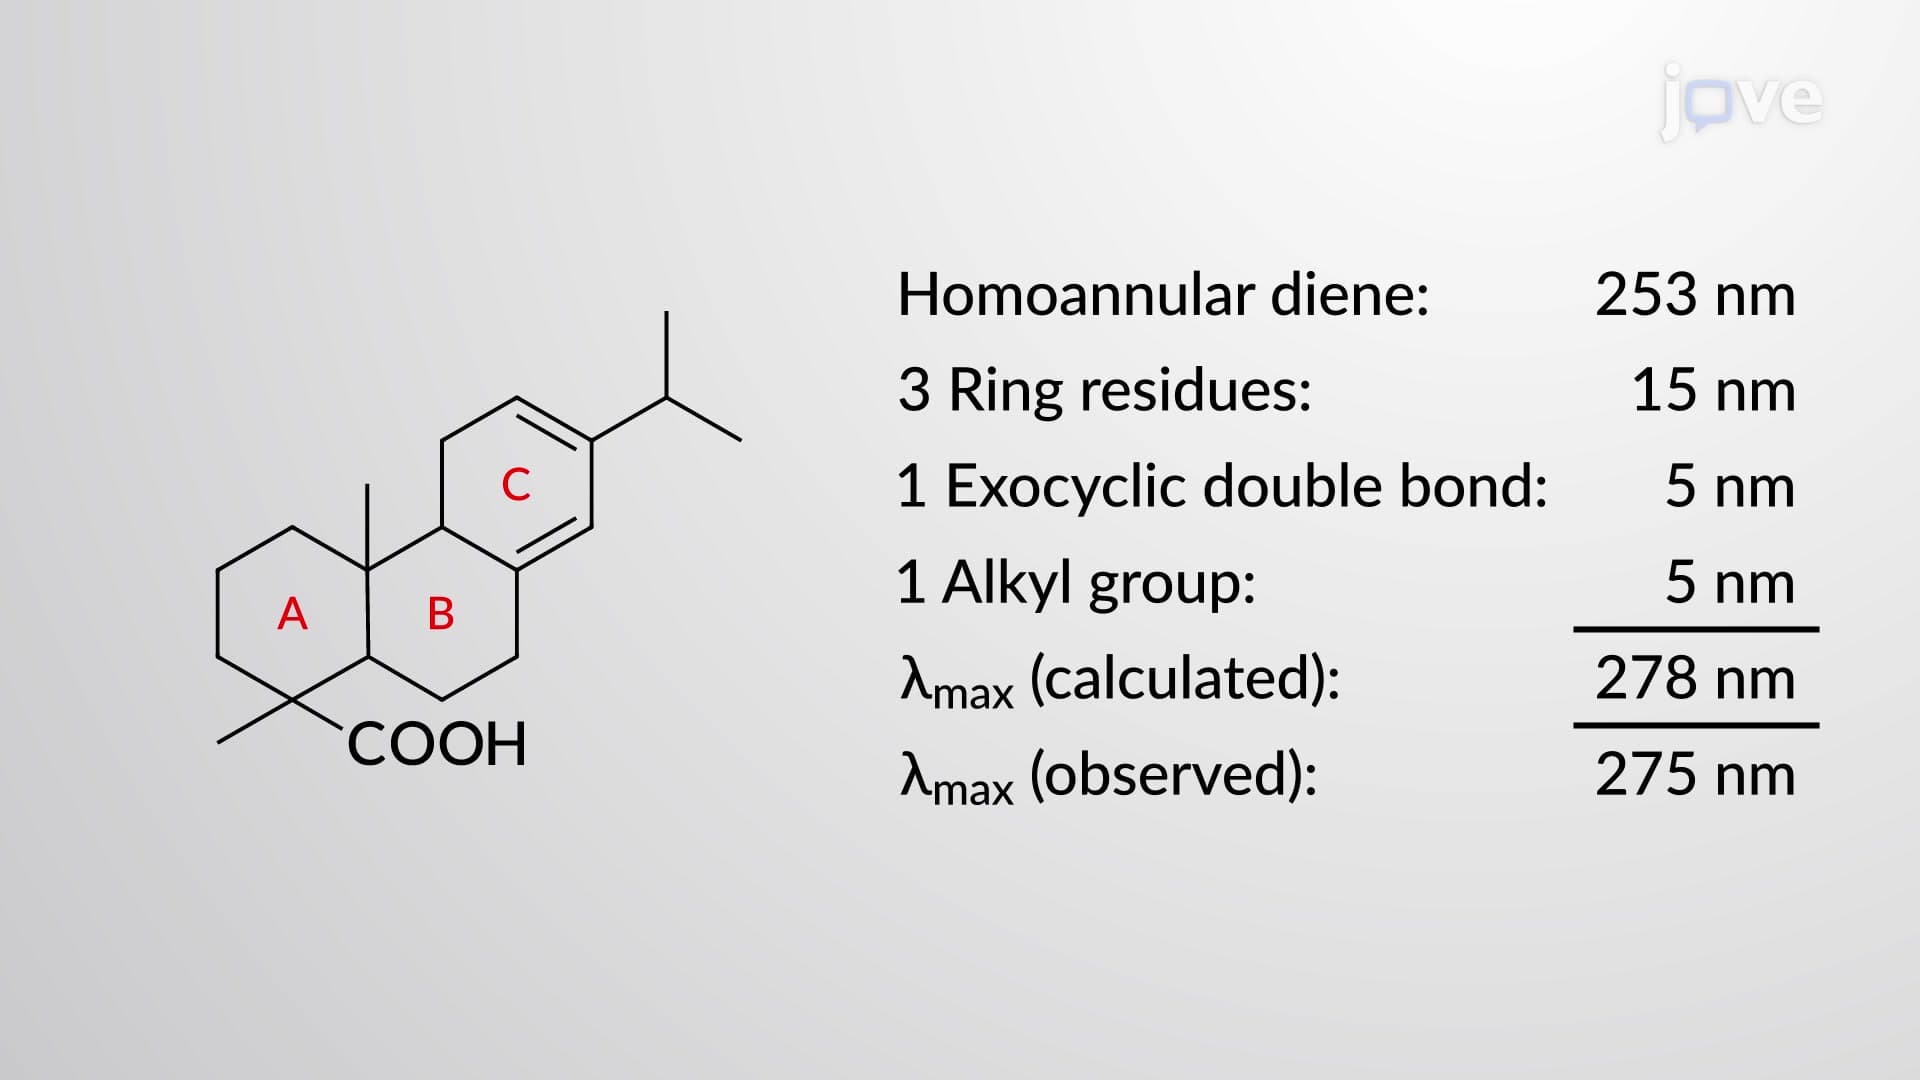
UV–Vis Spectroscopy: Woodward–Fieser Rules
24.4K
UV–Visible absorption spectra of conjugated dienes arise from the lowest energy π → π* transitions. The light-absorbing part of the molecule is called the chromophore, and the substituents directly attached to the chromophore are called auxochromes. A strong correlation exists between the absorption maxima, λmax, and the structure of a conjugated π system. The Woodward–Fieser rules predict the value of λmax for a given...
24.4K
 02:58
Crystal Field Theory - Octahedral Complexes
26.5K
Crystal Field Theory
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
26.5K
 02:46
Electron Configurations
16.7K
Electron configurations and orbital diagrams can be determined by applying the Aufbau principle (each added electron occupies the subshell of lowest energy available), Pauli exclusion principle (no two electrons can have the same set of four quantum numbers), and Hund’s rule of maximum multiplicity (whenever possible, electrons retain unpaired spins in degenerate orbitals).
The relative energies of the subshells determine the order in which atomic orbitals are filled (1s, 2s, 2p, 3s, 3p,...
The relative energies of the subshells determine the order in which atomic orbitals are filled (1s, 2s, 2p, 3s, 3p,...
16.7K