利用等级交互来进行蛋白质表面学习
IEEE journal of biomedical and health informatics
|January 19, 2024

概括
本研究介绍了HCGNet,这是一种用于蛋白质表面分析的深度学习模型. HCGNet有效地整合了化学和几何特征,提高了蛋白质-蛋白质相互作用预测的准确性.
科学领域:
- 结构生物信息学 结构生物信息学
- 计算生物学是一种计算生物学.
- 深度学习应用程序深度学习应用程序
背景情况:
- 预测蛋白质与蛋白质相互作用至关重要,但具有挑战性.
- 现有的方法通常会单独分析几何和化学特征.
- 需要模拟原子关系和层次特征相互作用.
研究的目的:
- 开发一种用于蛋白质表面分析的新型深度学习框架.
- 有效地整合化学和几何特征,以改善预测.
- 为了提高蛋白质-蛋白质相互作用部位预测的准确性.
主要方法:
- 开发了分层化学和几何特征交互网络 (HCGNet).
- 采用深度学习来建模原子之间的关系和层次特征相互作用.
- 桥梁化学和几何特征学习用于蛋白质表面分析.
主要成果:
- 在蛋白位预测方面,HCGNet表现出卓越的性能.
- 在现场预测方面,与最先进的方法相比,实现了2.3%的改进.
- 该框架成功地整合了层次化的特征交互.
结论:
- HCGNet提供了一种基于原则的方法来学习蛋白质表面.
- 模型捕捉特征交互的能力提高了预测准确度.
- 这项工作推动了结构生物信息学和蛋白质相互作用预测领域的发展.
更多相关视频
 07:08
Optimization of Synthetic Proteins: Identification of Interpositional Dependencies Indicating Structurally and/or Functionally Linked Residues
Published on: July 14, 2015
7.3K
 08:38
Genome-wide Protein-protein Interaction Screening by Protein-fragment Complementation Assay PCA in Living Cells
Published on: March 3, 2015
13.4K
相关概念视频
 02:04
Protein-protein Interfaces
12.5K
Many proteins form complexes to carry out their functions, making protein-protein interactions (PPIs) essential for an organism's survival. Most PPIs are stabilized by numerous weak noncovalent chemical forces. The physical shape of the interfaces determines the way two proteins interact. Many globular proteins have closely-matching shapes on their surfaces, which form a large number of weak bonds. Additionally, many PPIs occur between two helices or between a surface cleft and a...
12.5K
 01:30
Assembly of Signaling Complexes
5.8K
Multiprotein signaling complexes are formed in a dynamic process involving protein-protein interactions at the cytoplasmic domain of transmembrane receptors or enzymatic and non-enzymatic proteins associated with the receptor. These complexes ensure the activation and propagation of intracellular signals that regulate cell functions.
Interaction domains in cell signaling
Interaction domains recognize exposed features of their binding partners containing post-translationally modified sequences,...
Interaction domains in cell signaling
Interaction domains recognize exposed features of their binding partners containing post-translationally modified sequences,...
5.8K
 02:26
Protein Networks
4.0K
An organism can have thousands of different proteins, and these proteins must cooperate to ensure the health of an organism. Proteins bind to other proteins and form complexes to carry out their functions. Many proteins interact with multiple other proteins creating a complex network of protein interactions.
These interactions can be represented through maps depicting protein-protein interaction networks, represented as nodes and edges. Nodes are circles that are representative of a protein,...
These interactions can be represented through maps depicting protein-protein interaction networks, represented as nodes and edges. Nodes are circles that are representative of a protein,...
4.0K
02:04Protein-Protein Interfaces
3.8K
3.8K
 01:24
Protein Organization
6.5K
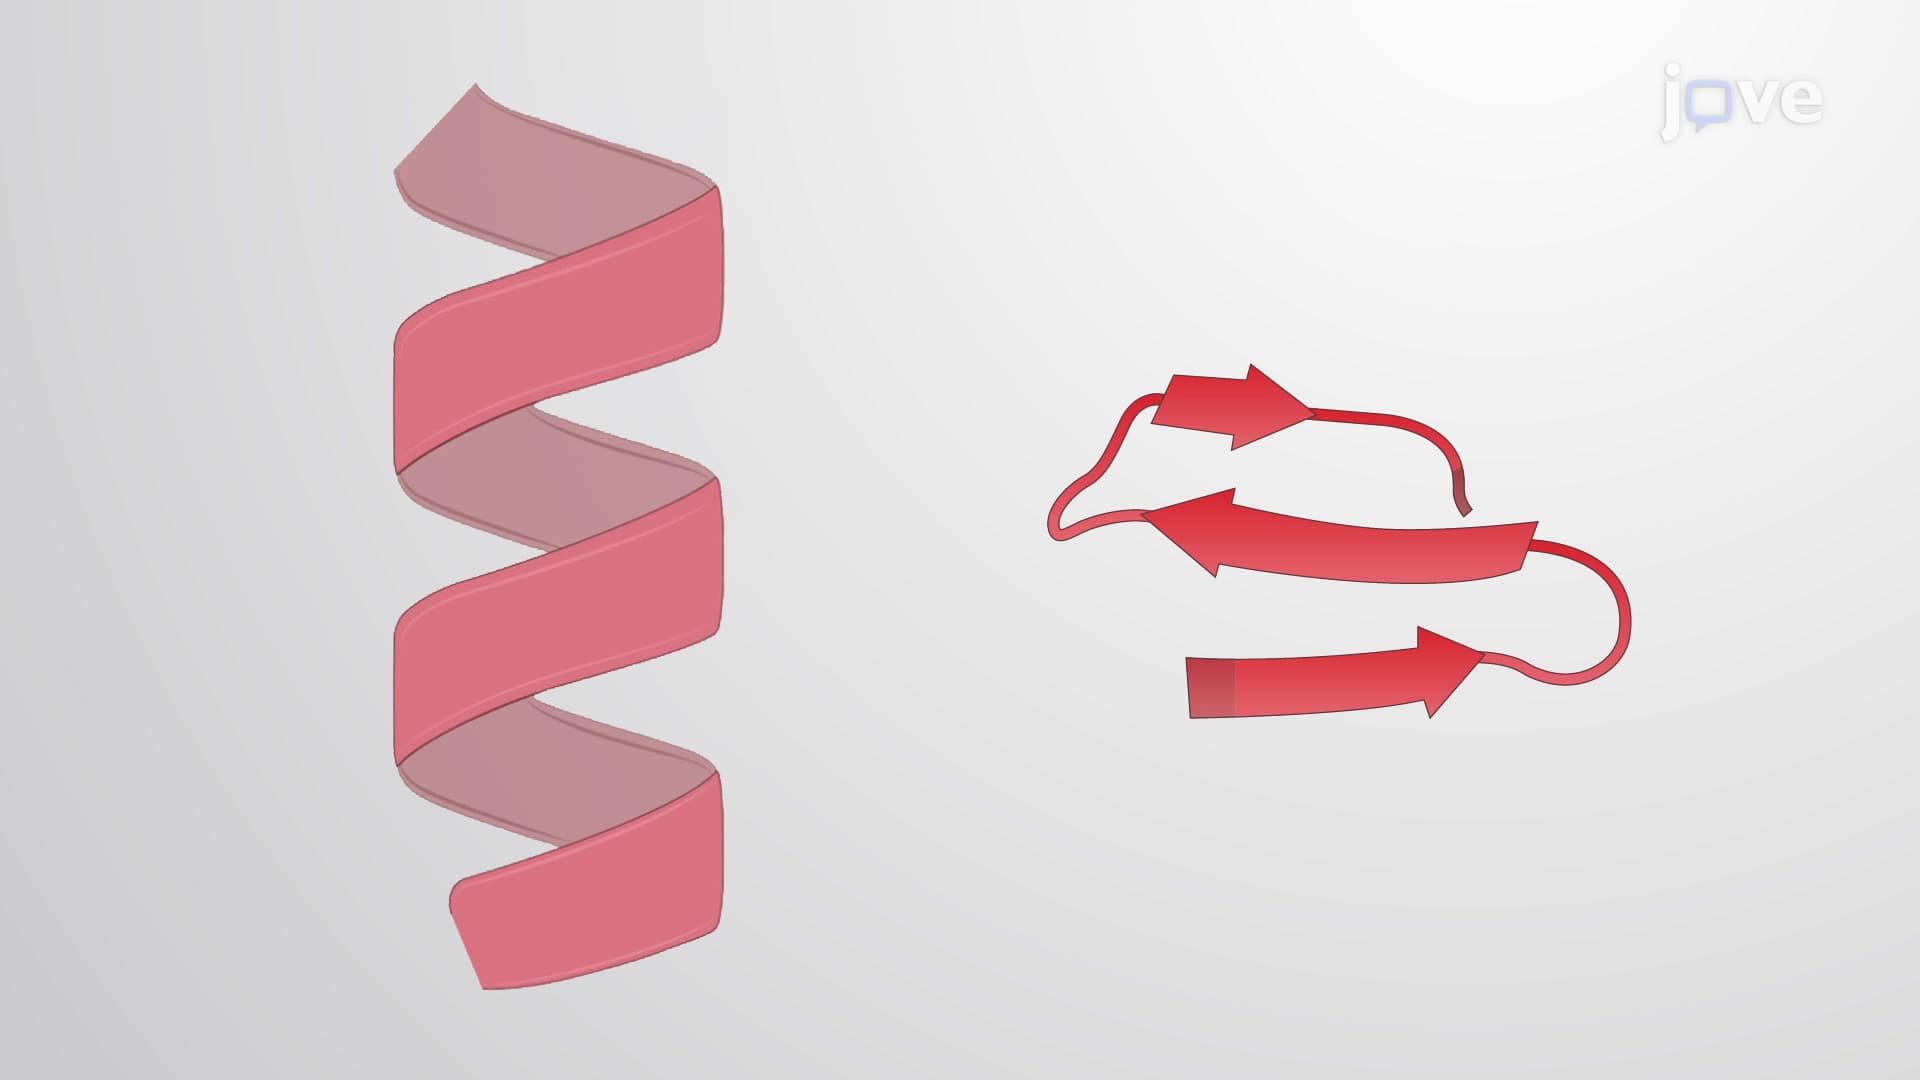
Proteins are polymers of amino acid residues. They are versatile and responsible for different cellular functions, including DNA replication, molecular transport, catalysis, and structural support. Proteins have a hierarchical structure comprising at least three levels of organization: primary, secondary, and tertiary structure. Some large proteins have a quaternary structure where individual protein subunits are linked together.
The primary structure of a protein is its amino acid sequence....
The primary structure of a protein is its amino acid sequence....
6.5K
 01:49
Conserved Binding Sites
4.2K
Many proteins’ biological role depends on their interactions with their ligands, small molecules that bind to specific locations on the protein known as ligand-binding sites. Ligand-binding sites are often conserved among homologous proteins as these sites are critical for protein function.
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
4.2K