
低密度脂蛋白非瑞斯对万科米辛血清度的最小影响:一个案例报告
Yoko Hiyama1, Takashi Tomita1, Hiroaki Matsuo2
1Department of Pharmaceutical Services, Hiroshima University Hospital, 1-2-3 Kasumi, Minami-ku, Hiroshima, 734-8551, Japan.

概括
低密度脂蛋白非瑞斯 (LDL-A) 在接受透析的患者中对万科米辛水平的影响最小. 在一个案例研究中,这种血液净化疗法没有显著改变血清中万科米辛度.
科学领域:
- 腎臟病學 (nephrology) 是一種醫學專業.
- 药理学 药理学是指药理学的学科.
- 心血管医学 心血管医学
背景情况:
- 低密度脂蛋白非瑞斯 (LDL-A) 是一种至关重要的血液净化疗法,用于消化动脉样硬化的耐火性.
- 患有糖尿病病的患者往往需要复杂的治疗,包括透析和其他治疗.
相关概念视频
 01:09
Hepatic Drug Clearance: Effect of Protein Binding
198
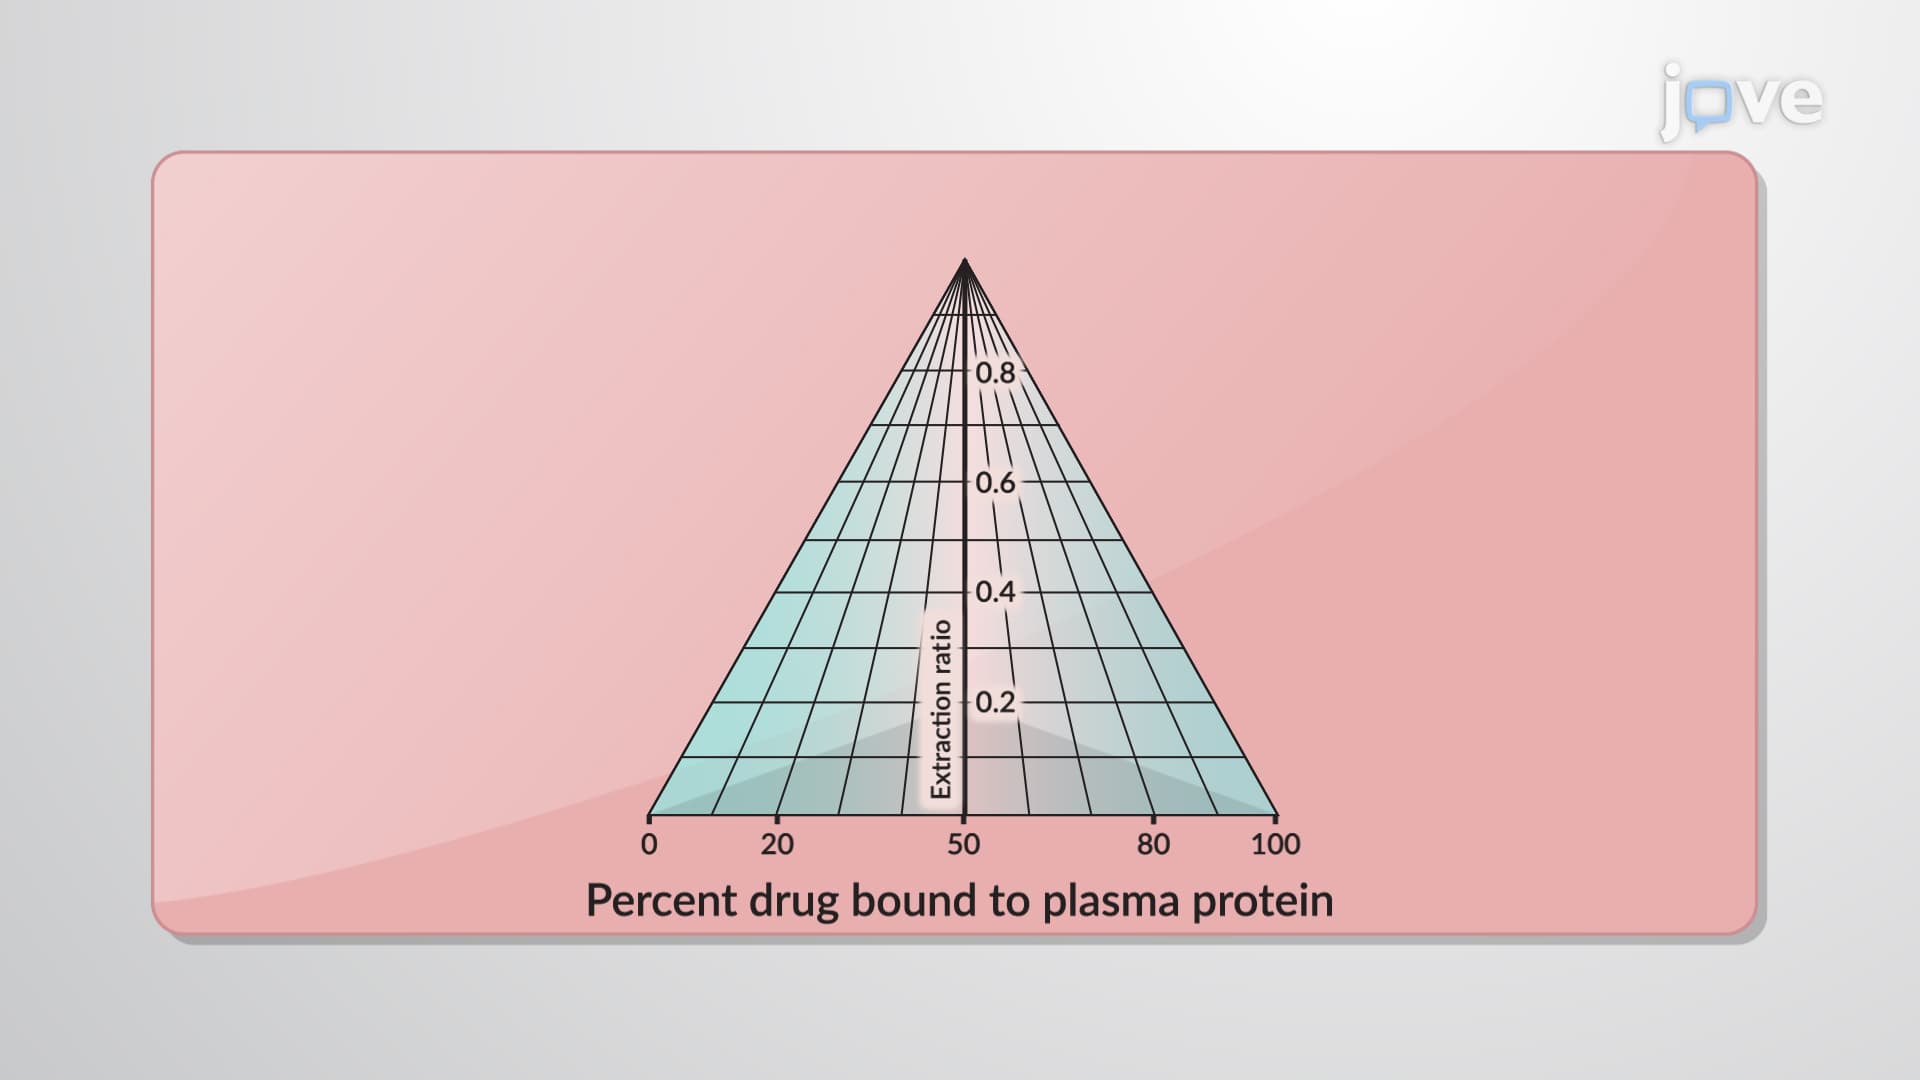
Hepatic clearance is influenced by protein binding based on the drug's extraction ratio. Drugs with high extraction ratios are considered flow-limited and remain unaffected by protein binding during hepatic clearance. On the other hand, drugs with low extraction ratios may be impacted by plasma protein binding, although the extent of this influence depends on the fraction of the drug bound.
For low-extraction-ratio drugs that are less than 80% protein-bound, minor changes in protein binding...
For low-extraction-ratio drugs that are less than 80% protein-bound, minor changes in protein binding...
198
 01:18
Factors Affecting Protein-Drug Binding: Drug-Related Factors
107
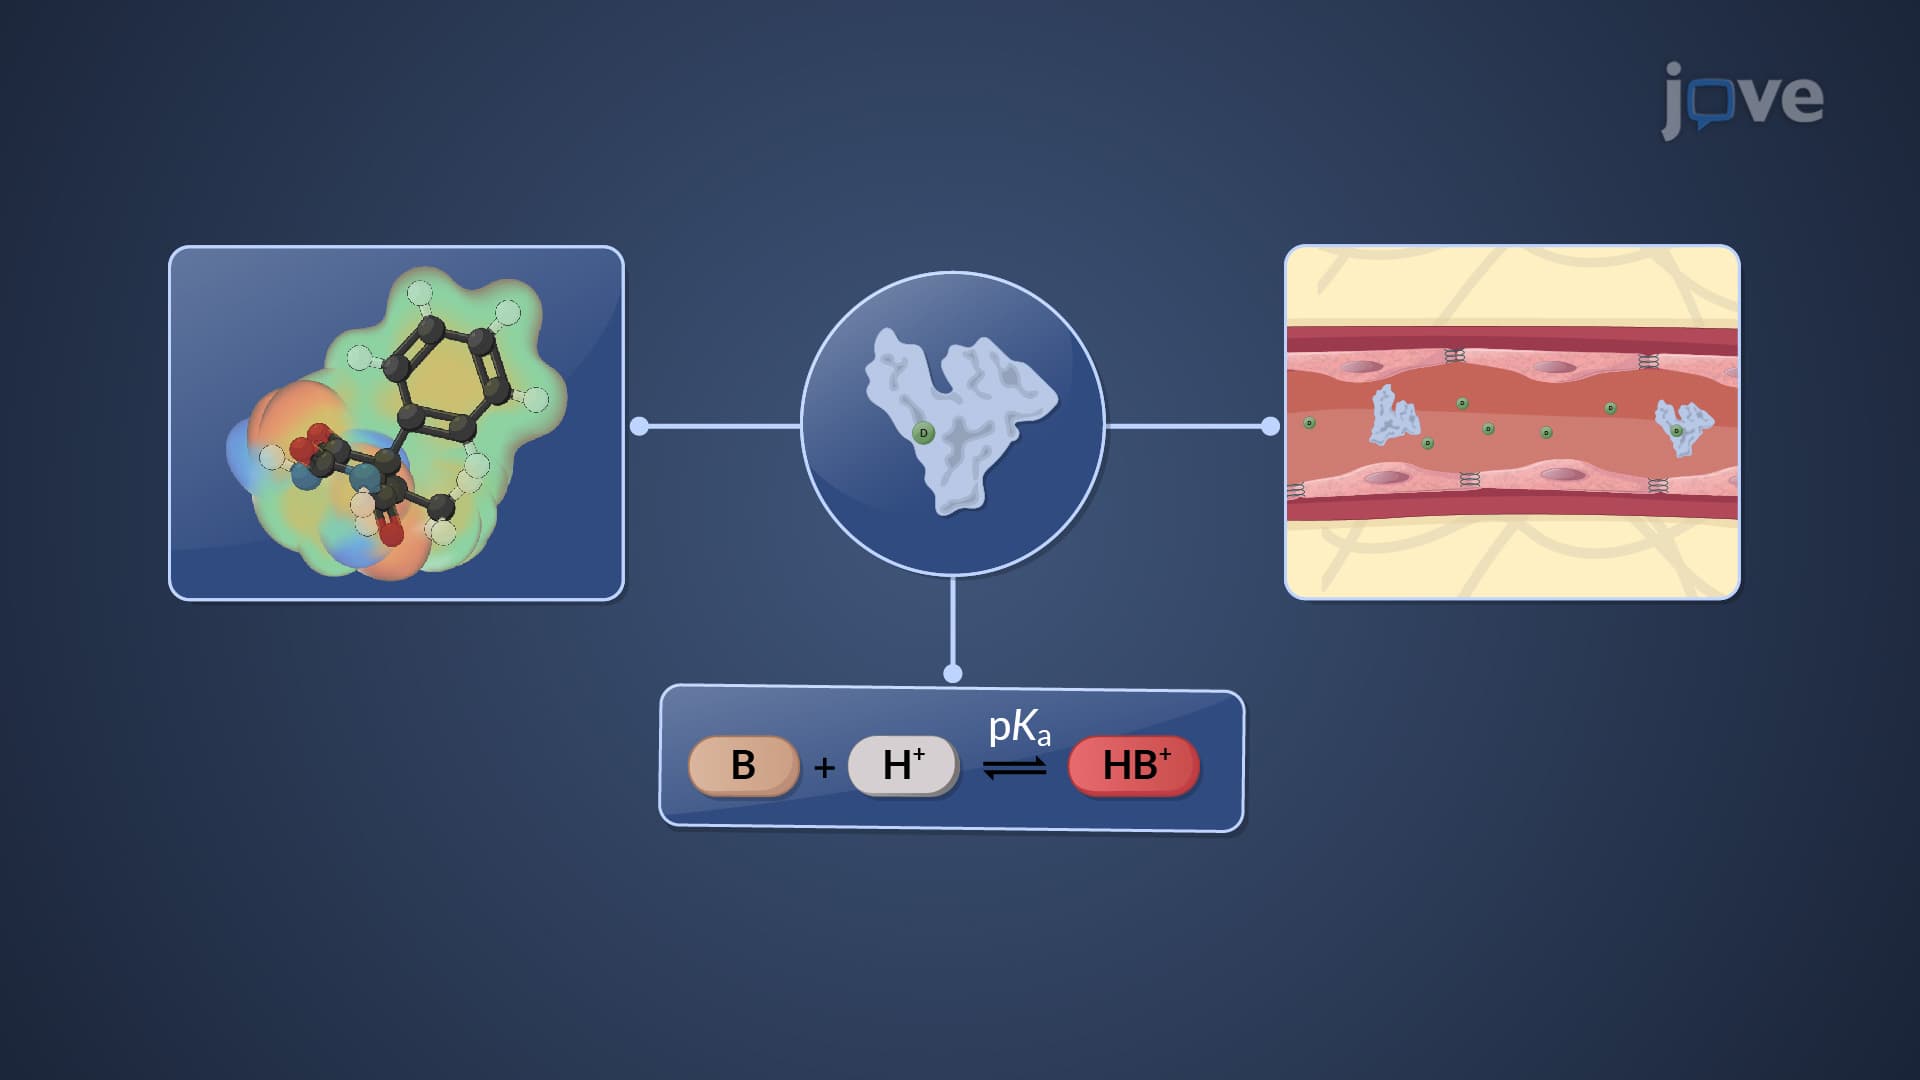
Drug binding to proteins is a complex phenomenon influenced by various drug-related factors, each playing a significant role in the interaction between drugs and proteins within the body.
One crucial factor in drug-protein binding is the drug's lipophilicity or its affinity for fat. More lipophilic drugs tend to have higher binding extents. For example, highly lipophilic drugs like cloxacillin exhibit substantial protein binding, with as much as 95% of the drug binding to proteins. In...
One crucial factor in drug-protein binding is the drug's lipophilicity or its affinity for fat. More lipophilic drugs tend to have higher binding extents. For example, highly lipophilic drugs like cloxacillin exhibit substantial protein binding, with as much as 95% of the drug binding to proteins. In...
107
 01:25
Drug Distribution: Volume of Distribution
5.1K
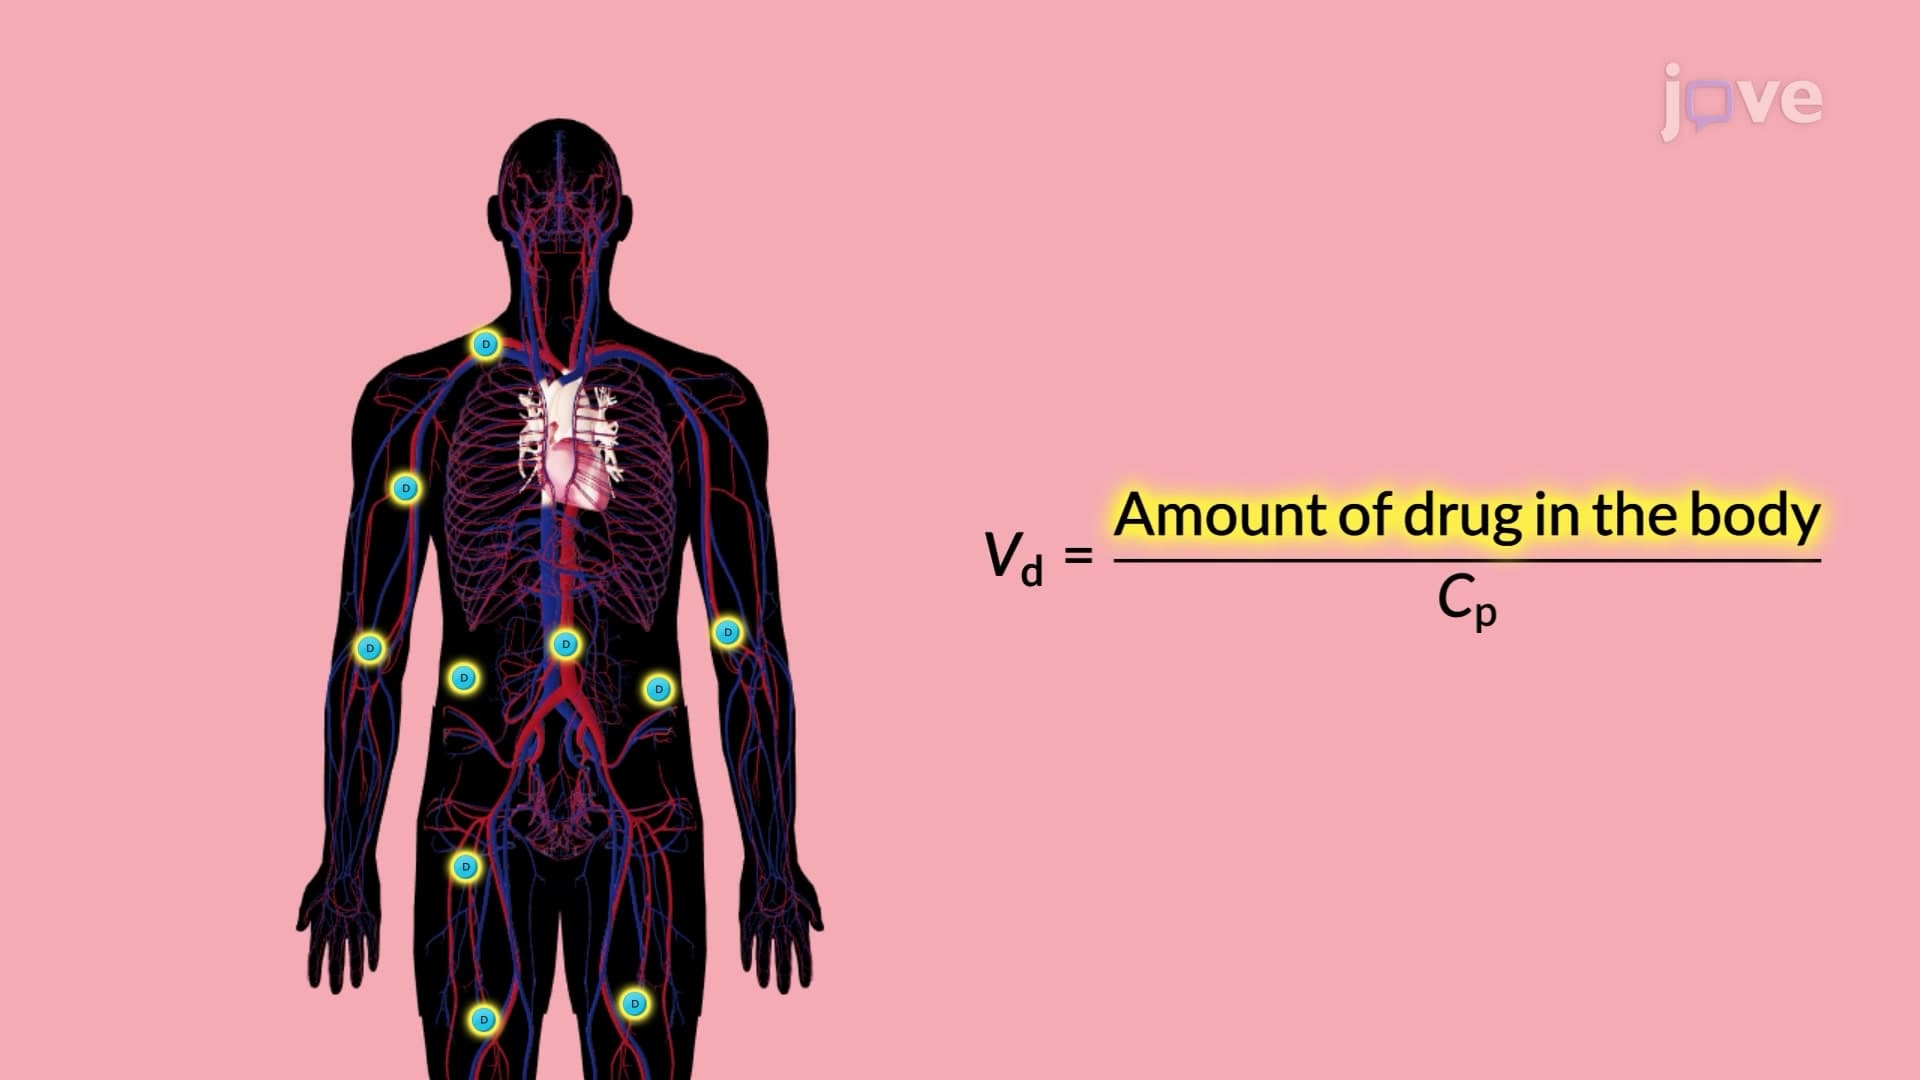
The volume of distribution refers to the theoretical volume necessary to contain the entire amount of an administered drug at the same concentration observed in the blood plasma. The body's intracellular fluid compartment, which makes up two-thirds of the total body water, is contrasted with the extracellular fluid compartment—comprising plasma and interstitial fluid—that accounts for one-third. The volume of distribution can vary depending on the characteristics of the drug.
5.1K
 01:29
Drug Distribution: Plasma Protein Binding
5.6K
Drugs predominantly attach to plasma proteins, with only a small percentage remaining unbound. The unbound portion can be calculated as one minus the bound fraction. Acidic drugs form large, inactive complexes by reversibly binding to plasma albumin, which prevents them from diffusing across biological barriers. These drug-protein complexes act as reservoirs for the drugs. As the concentration of unbound drugs decreases, these complexes quickly dissociate to release the free drug, maintaining...
5.6K
 01:14
Time Course of Drug Effect
2.0K
The progression of a drug's impact can be analyzed by examining both the concentration-time course and the effect-time course. The concentration-time course is determined by the drug's half-life and is influenced by factors such as its pharmacokinetics, including absorption, distribution, metabolism, and elimination. The effect of the drug is often related to its concentration in the plasma and is calculated using the maximum drug effect and the plasma concentration that generates 50...
2.0K
 01:22
Nonlinear Pharmacokinetics: Bioavailability and Protein-Drug Binding
176
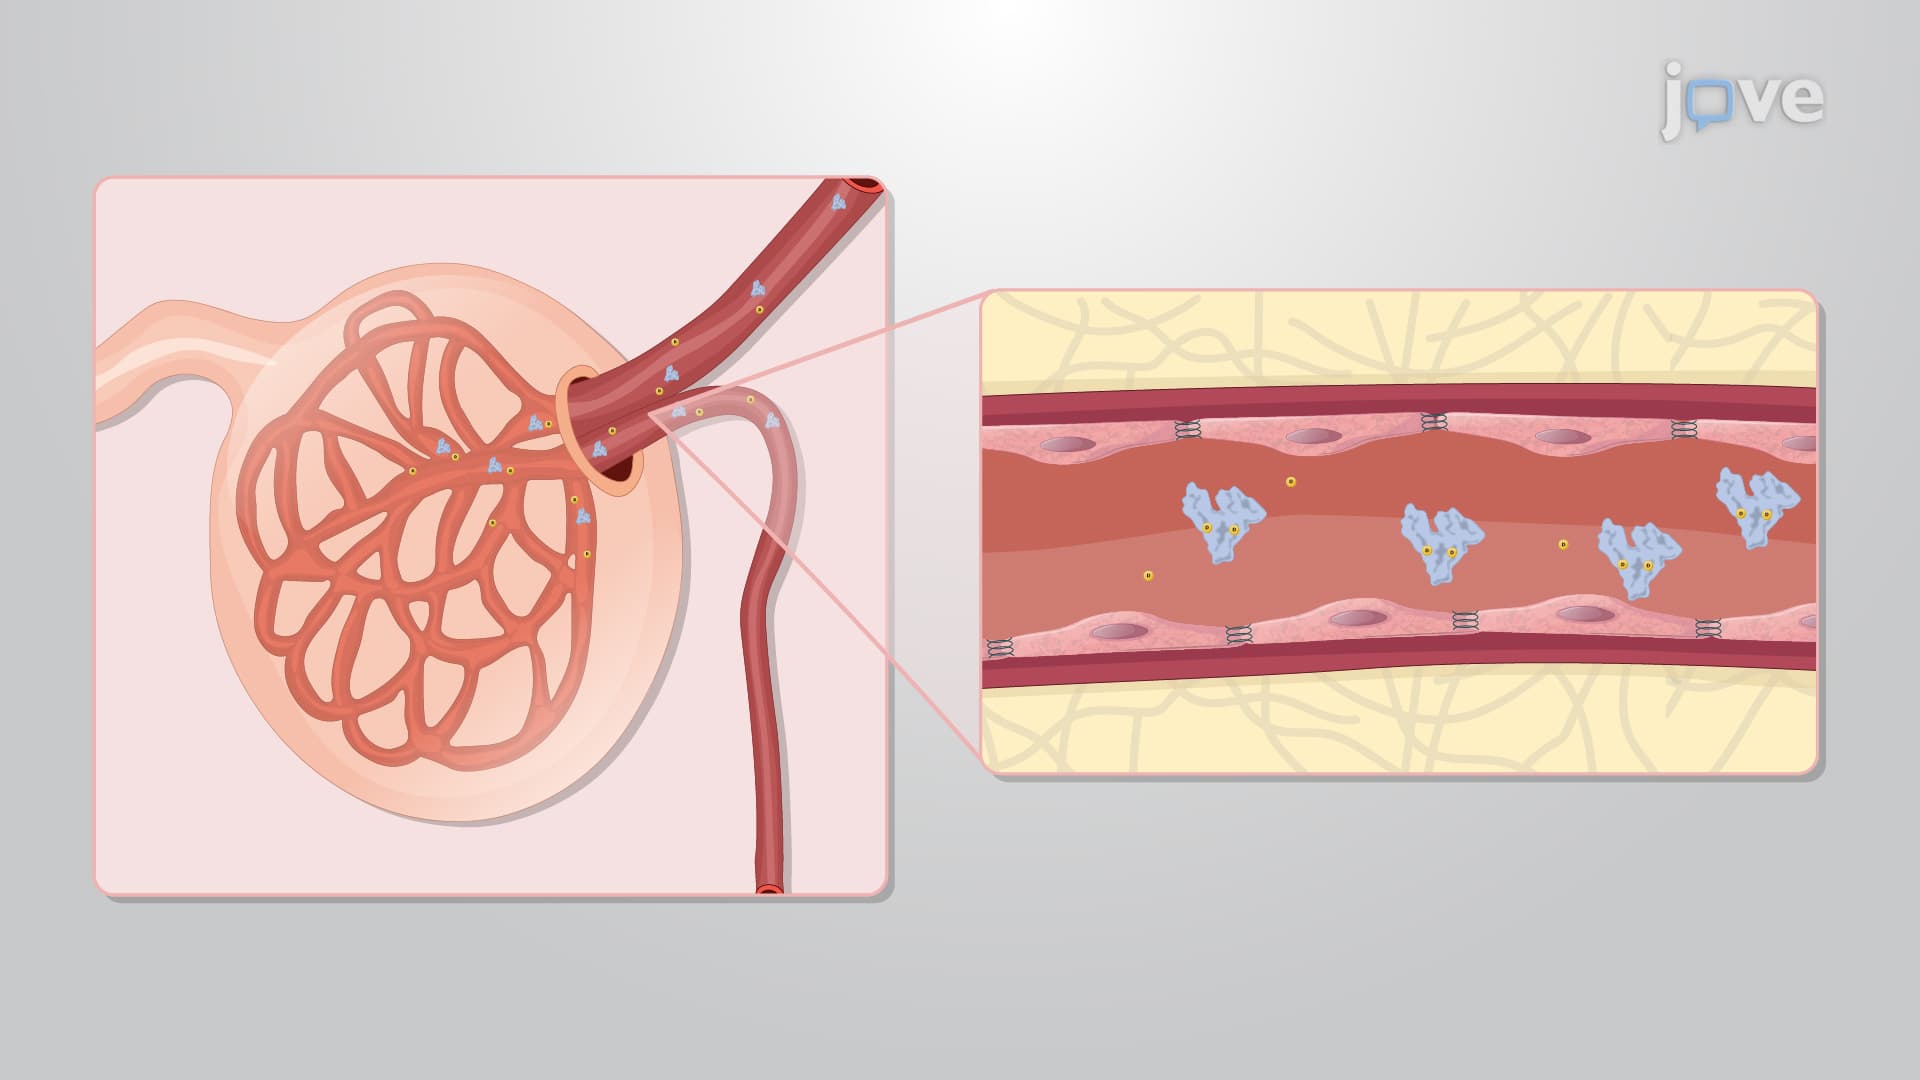
When a drug follows nonlinear pharmacokinetics, its bioavailability, the amount of the drug that reaches the systemic circulation, can change with different doses. This is due to the presence of a saturable pathway. The pathway becomes saturated as the drug concentration increases, decreasing the absorption rate. Consequently, the drug's bioavailability may be lower than expected at higher doses.
To quantify the extent of bioavailability, pharmacologists often use a parameter called .
To quantify the extent of bioavailability, pharmacologists often use a parameter called .
176