
从无形固体分散中同时释放的帕克利塔塞尔和恩塞奎达尔增加了大鼠口服帕克利塔塞尔的生物可用性
Emilie Fynbo Petersen1, Bjarke Strøm Larsen2, Rasmus Blaaholm Nielsen1
1Department of Physics, Chemistry and Pharmacy, University of Southern Denmark, Campusvej 55, DK-5230 Odense M, Denmark.
International journal of pharmaceutics
|March 5, 2024

概括
提高口服帕克利塔塞尔的生物利用性包括通过使用P-glycoprotein (P-gp) 抑制剂创造无形固体分散 (ASD). 从ASD中控制释放显著增加了帕克利塔塞尔的吸收,观察到显著的24倍增加.
科学领域:
- 药理学 药理学是指药理学的学科.
- 材料科学 材料科学 材料科学
- 药物运输 药物运输 药物运输
背景情况:
- 口服帕克利塔塞尔的生物利用性因溶解性差以及P-glycoprotein (P-gp) 的排泄而受到严重限制.
- 无形固体分散 (ASD) 提供了一种改善药物溶解性和吸收的策略.
- 与P-gp抑制剂同时使用可能会增加口服药物暴露.
研究的目的:
- 调查从ASD中同时输送帕克利塔塞尔和P-gp抑制剂恩塞奎达的潜力,以提高口服帕克利塔塞尔的生物可用性.
- 评估不同聚合物矩阵 (PVP-K30,HPMC-5,HPMC-4K) 对药物释放和药物动力学概况的影响.
- 为了确定最大限度地提高口服帕克利塔塞尔吸收的最佳配方.
主要方法:
- 使用PVP-K30,HPMC-5和HPMC-4K的冷干燥来制备帕克利塔塞尔和脑内的ASD.
- 在体外溶解研究以评估不同ASD配方的药物释放率.
- 在Sprague-Dawley大鼠中进行口服药理学研究,以评估各种配方的帕克利塔塞尔生物可用性.
主要成果:
- 药物释放率在聚合物之间差异很大,PVP-K30显示出释放速度最快,HPMC-4K显示出释放速度最慢.
- 从所有ASD中溶解的帕克利塔塞尔维持了比其表面溶解度高的度.
- 与单独的无形帕克利塔塞尔相比,口服帕克利塔塞尔的生物可用性在基于聚合物的ASD中增加了3到4倍.
- 与恩塞奎达尔 (无形粉末或ASD) 的同时使用进一步增加了帕克利塔塞尔的生物可用性2至4倍.
- 在基于HPMC-5的ASD中,帕克利塔塞尔生物可用性显著增加了24倍.
结论:
- 从聚合物基ASD中同时控制释放帕克利塔塞尔和恩塞奎达尔是一种有效的策略,可以提高口服帕克利塔塞尔的生物可用性.
- 配方设计,特别是聚合物的选择和溶解速率的控制,对于优化药物输送至关重要.
- 基于HPMC-5的ASD在增强帕克利塔塞尔口服吸收方面表现优异,突出了临床应用的潜力.
相关概念视频
 01:20
Factors Affecting Dissolution: Drug Permeability, Stability and Stereochemistry
207
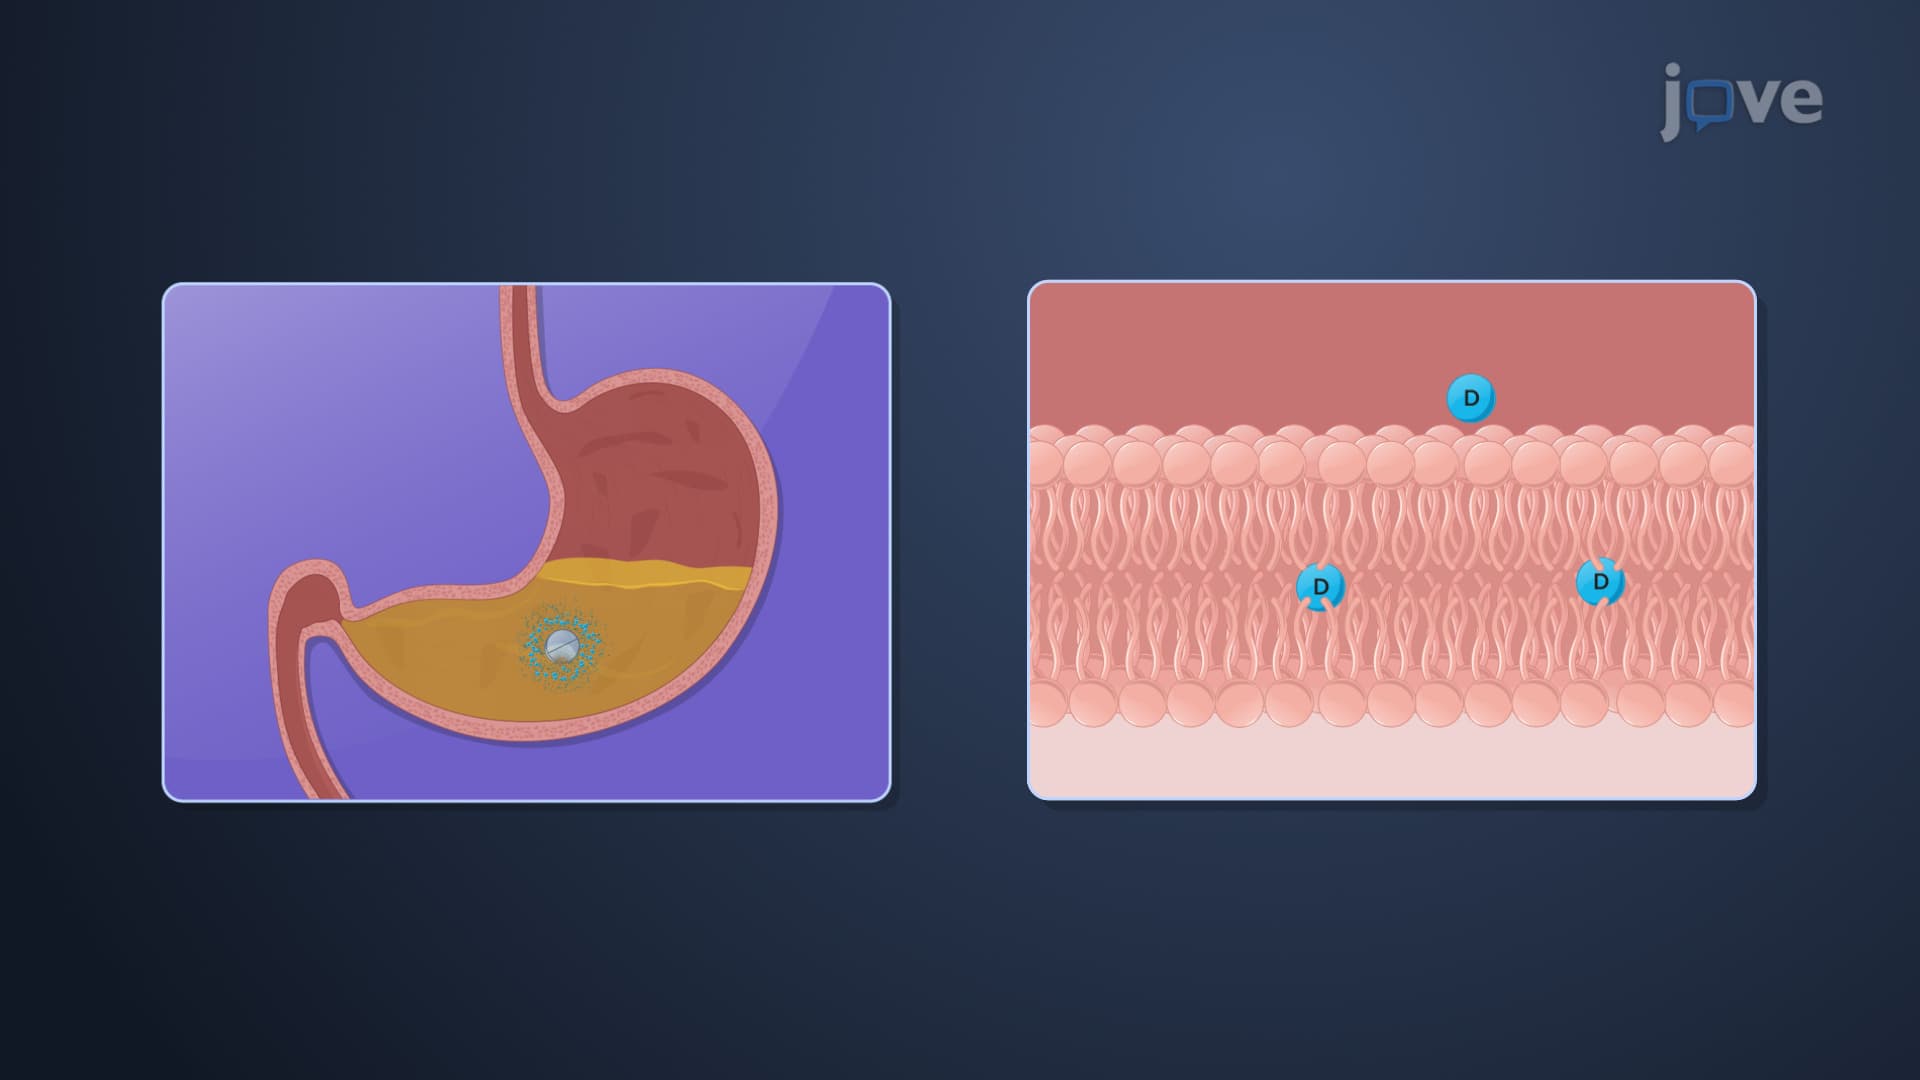
Orally administered drugs primarily enter the systemic circulation via passive diffusion through the intestinal membranes. The drug's absorption is influenced by drug stability in the gastrointestinal GI tract, membrane permeability, the surface area available for absorption, luminal drug concentration, and residence time in the lumen. Drug permeability can be enhanced by adjusting the lipophilicity, polarity, or molecular size of the drug, promoting its passive transport across intestinal...
207
 01:27
Factors Influencing Drug Absorption: Drug Dissolution
493
The pharmacokinetic journey of drugs from solid oral dosage forms into systemic circulation is multifaceted. It begins with disintegration, a prerequisite ensuring a solid dosage form's subdivision into minute particles. Dissolution occurs next as these granulated entities solubilize in gastrointestinal fluids. This solubilization is crucial for the succeeding stage, permeation, which describes the traversal of the drug across the intestinal membrane and its subsequent entry into the blood...
493
 01:23
Factors Affecting Dissolution: Particle Size and Effective Surface Area
840
Dissolution kinetics, an essential aspect of oral drug delivery, is significantly influenced by the drug's particle size. According to the Noyes-Whitney dissolution model, the dissolution rate correlates directly with the drug's surface area. The larger the surface area, the higher the drug's solubility in water, leading to a faster drug dissolution rate. Reducing particle size increases the effective surface area, enhancing the dissolution process. Micronization and nanosizing are...
840
 01:24
Factors Influencing Drug Absorption: Presystemic Elimination
219
The pharmacokinetic journey of oral drugs begins with a crucial first pass through the hepatic portal system, called the first-pass effect. This first pass significantly impacts bioavailability — the proportion of a drug that enters systemic circulation and is available for therapeutic action. The primary route sees the drug absorbed by intestinal membranes and then shunted to the liver via the hepatic portal vein. Here, pre-systemic elimination occurs as drugs face metabolism or biliary...
219
 01:28
Factors Influencing Drug Absorption: Pharmaceutical Parameters
134
Solid dosage forms such as tablets and capsules undergo rigorous manufacturing processes to ensure stability and effectiveness. Their dissolution and absorption properties are influenced significantly by the choice of excipients (inactive ingredients that serve various roles in the formulation), and the methodology applied during production. The manufacturing parameters, such as compression force and granulation techniques, significantly affect dissolution rates. Elevated compression forces...
134
 01:21
Drug Distribution: Tissue Binding
2.6K
Upon entering the systemic circulation, drugs can distribute into the interstitial and intracellular fluid of various tissue cells. This distribution is facilitated by the binding of drugs to different cellular components within tissues, which may lead to drug accumulation in specific areas. Drugs bound to tissue components serve as reservoirs that release free drugs back into the system, prolonging the drug's overall action. However, this accumulation can also result in local toxicity.
For...
For...
2.6K