线粒体功能障碍和溶酶体储存之间的相互作用:遗传代谢肌肉疾病的挑战,重点是婴儿发病的庞培病
Mengjiao Zhang1, Jiechao Niu1, Mengmeng Xu1
1Department of Pediatrics, The First Affiliated Hospital of Zhengzhou University, Zhengzhou, China.
Frontiers in cardiovascular medicine
|March 7, 2024

概括
5个月大的婴儿患有心力衰竭和多变性心肌病变 (hypertrophic cardiomyopathy). 基因分析揭示了GAA基因的突变,突出了改善线粒体功能疗法的需要.
科学领域:
- 生物化学 生物化学
- 遗传学 遗传学 是一个
- 儿科 儿科 儿科
背景情况:
- 庞培病 (PD) 是一种罕见的,渐进的溶酶体储存障碍,影响线粒体功能.
- 它可能导致严重的心脏并发症,包括心力衰竭,其机制尚未完全理解.
研究的目的:
- 在儿童患者中调查庞培病的临床,遗传和线粒体方面.
- 探索PD和线粒体功能障碍之间的关系.
主要方法:
- 一位疑似患有PD的儿科患者的回顾性分析.
- 使用了心声回声图,CT扫描,心电图和遗传/结构分析 (PDB ID 5KZX,VMD软件).
主要成果:
- 一名5个月大的男婴被诊断出患有婴儿发作庞佩病 (IOPD),原因是同卵性GAA基因突变 (c.2662G>T).
- 伴有发烧,心肺功能异常,肝壮病,心力衰竭和缩性心肌病变.
- 尽管初始治疗,心脏异常的改善有限.
结论:
- 虽然酶替代疗法是有益的,但针对线粒体功能为PD提供了一个新的治疗途径.
- 对线粒体增强的进一步研究可以改善PD患者的治疗结果.
相关概念视频
 01:22
Lysosomal Hydrolases
3.8K
Lysosomes are the site for the degradation of macromolecules and biological polymers released during membrane trafficking events such as secretory, endocytic, autophagic, and phagocytic pathways. The membrane-enclosed area of the lysosome, called the lumen, contains hydrolytic enzymes active in an acidic environment. These acid hydrolases are functional at a pH between 4.5 and 5 and are involved in cellular processes such as cell signaling, energy metabolism, restoration of the plasma membrane,...
3.8K
 01:20
Inborn Errors of Metabolism
158
Phenylketonuria (PKU) is a protein metabolism disorder characterized by high blood levels of the amino acid phenylalanine. This results from a mutation in the gene responsible for phenylalanine hydroxylase, an enzyme that converts phenylalanine into tyrosine. When this enzyme is deficient, phenylalanine builds up in the blood, leading to symptoms such as vomiting, rashes, seizures, growth deficiency, and severe mental retardation. An early diagnosis and a diet restricting phenylalanine intake...
158
 01:19
Translocation of Proteins into the Mitochondria
3.1K
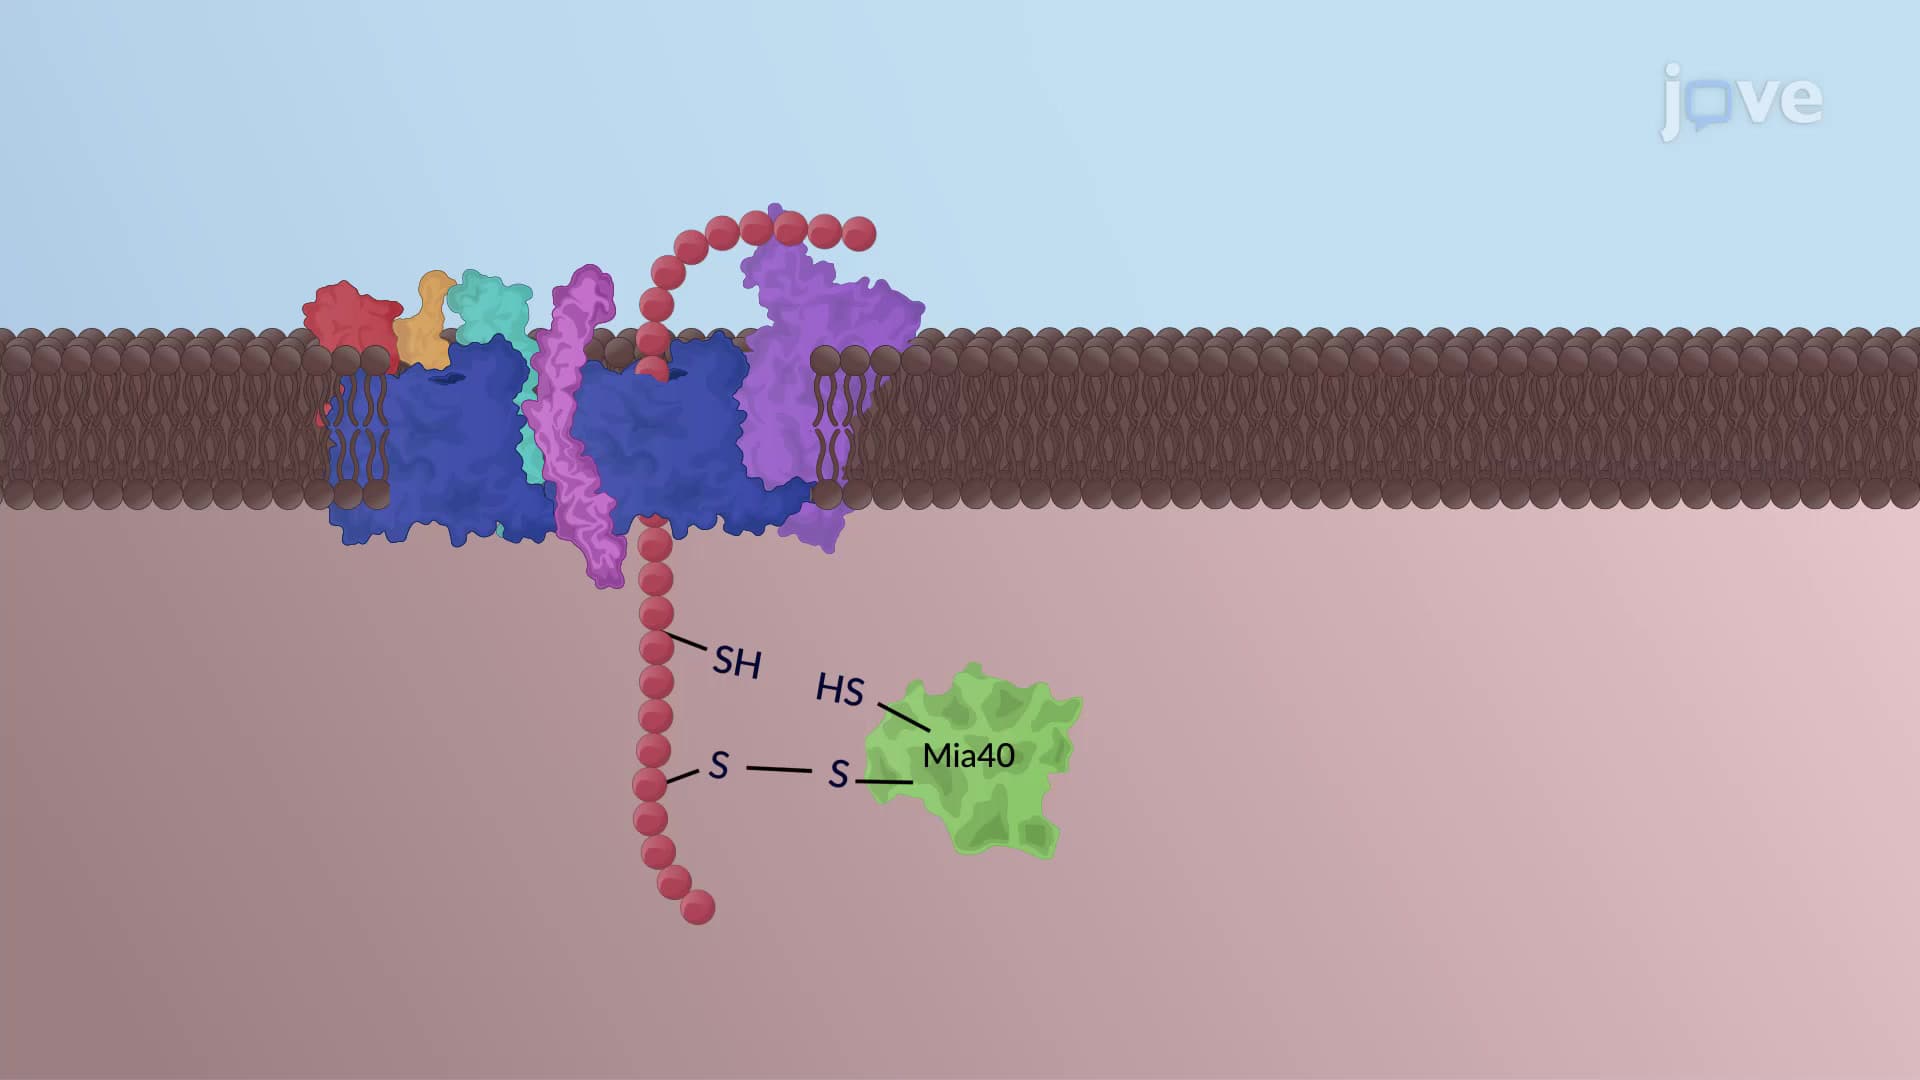
Mitochondrial precursors are translocated to the internal subcompartments via independent mechanisms involving distinct protein machineries called translocases.
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
3.1K
 01:21
Satellite Stem Cells and Muscular Dystrophy
2.0K
Satellite stem cells or myosatellite cells are quiescent stem cells that Alexander Mauro first identified in 1961. These cells are located between the sarcolemma, the plasma membrane of muscle fibers, and the basal lamina, the connective tissue sheath covering it. These mononucleated cells are activated in response to muscle injury, can transform into myoblasts, and may form or repair muscle fibers. Myosatellite cells can provide additional myonuclei for muscle regeneration or return to a...
2.0K
 01:48
ATP Synthase: Mechanism
14.6K
In animals, the mitochondrial F1F0 ATP synthase is the key protein that synthesizes ATP molecules through a complex catalytic mechanism. While the nuclear genome encodes the majority of ATP synthase subunits, the mitochondrial genome encodes some of the enzyme's most critical components. The formation of this multi-subunit enzyme is a complex multi-step process regulated at the level of transcription, translation, and assembly. Defects in one or more of these steps can result in decreased...
14.6K
 01:39
Mitochondrial Protein Sorting
4.3K
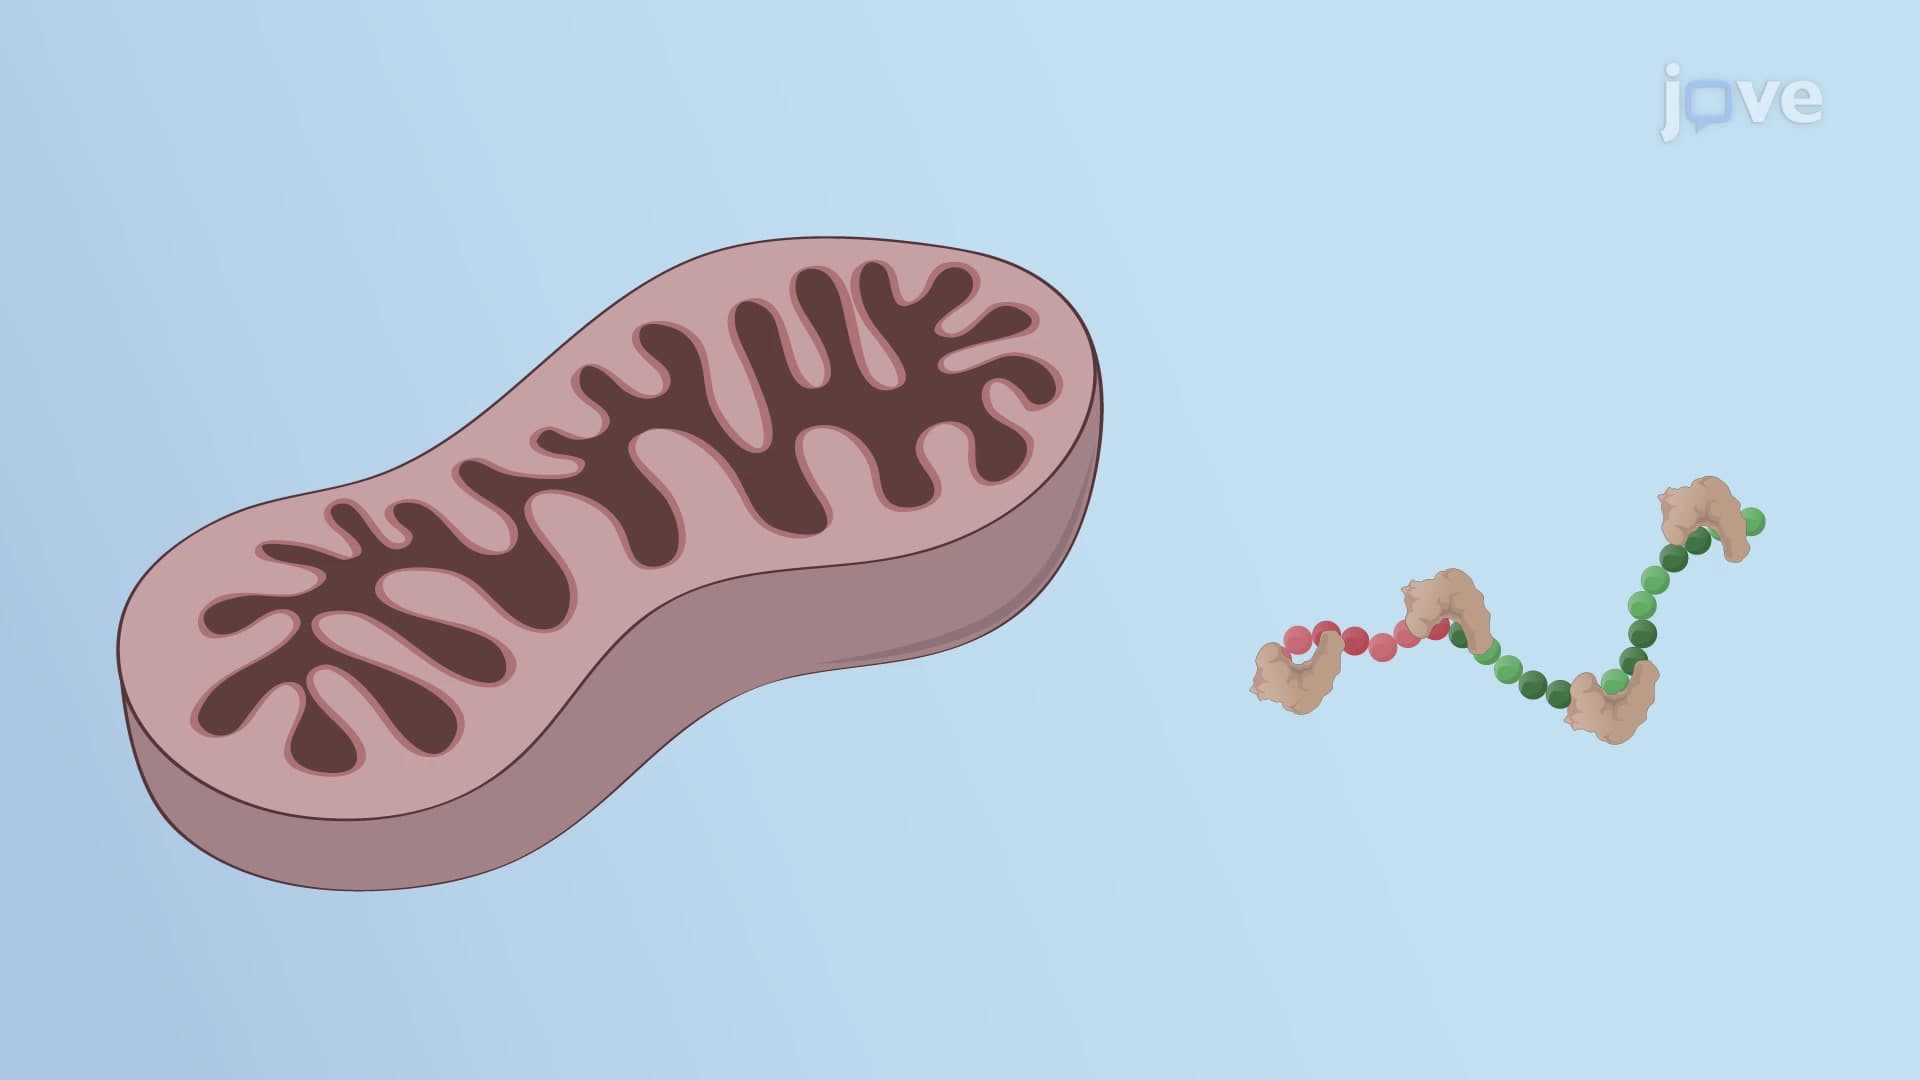
Mitochondria are double-membrane organelles of the eukaryotes involved in cellular metabolism, signaling, ATP synthesis, and programmed cell death. Each of these processes requires specific proteins and enzymes that must be correctly sorted to the right mitochondrial subcompartment for the proper functioning of the organelle.
Most of these mitochondrial proteins are encoded by the nucleus and imported to the mitochondria as unfolded or loosely folded precursors. Mitochondrial precursors...
Most of these mitochondrial proteins are encoded by the nucleus and imported to the mitochondria as unfolded or loosely folded precursors. Mitochondrial precursors...
4.3K