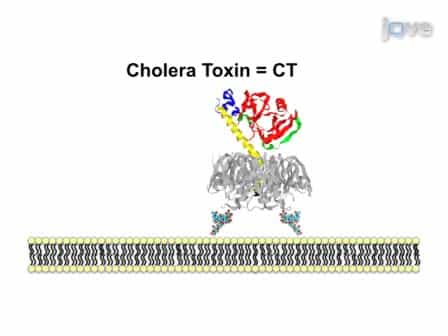
石家庄毒素2的A1亚单元的结构功能分析与类向P-Stalk结合部位并抑制活动的类
Xiao-Ping Li1, Michael J Rudolph2, Yang Chen2
1Department of Plant Biology, Rutgers, The State University of New Jersey, 59 Dudley Road, New Brunswick, New Jersey 08901, United States.
Biochemistry
|March 11, 2024

概括
研究人员确定了Shiga毒素2a (Stx2a) 结合和抑制所必需的最小P-茎蛋白序列 (P8). 这一发现揭示了Stx2a.
科学领域:
- 生物化学 生化学
- 分子生物学分子生物学
- 结构生物学 结构生物学
背景情况:
- 石加毒素2a (Stx2a) 是Sh的一个关键毒性因子.
- Stx2a会引起血溶性尿素性综合征 (HUS),这是儿童功能衰竭的主要原因.
- 这种毒素通过准28S rRNA的素-素循环 (SRL) 来抑制蛋白质合成,但对核糖体蛋白质的确切结合机制尚不清楚.
研究的目的:
- 阐明Stx2a与核糖体P茎蛋白结合的结构和功能基础.
- 为了确定 Stx2a 相互作用和抑制所需的最小 P 茎序列.
- 为了确定Stx2a的X射线晶体结构与已识别的P-stalk碎片复合.
主要方法:
- 核糖体P-茎蛋白的保存的C端域 (CTD) 的删除分析.
- 单独确定Stx2a的X射线晶体结构,并与P-stalk CTD最小序列 (P8) 复合.
- 对结合相互作用的分析和与其他核糖体失活蛋白 (RIP) 的比较.
主要成果:
- P-茎蛋白的最后八个氨基酸 (P8) 构成了最佳 Stx2a 结合和抑制的最小序列.
- 晶体结构显示,P8的C端酸通过关键接触与毒素结合口袋中的氨酸残留物定Stx2a.
- 对于 Stx2a 结合而言,C端的酸更为关键,而不是酸甲子单元 (RTA) 结合,这突出了 RIP-P-茎相互作用的差异.
结论:
- 阻止Stx2a与P茎的相互作用有效地抑制了其催化活性.
- P-茎结合是 Stx2a 招募到 SRL 进行 RNA 清洗的必要初步步骤.
- 对Stx2a-P-stalk相互作用的结构洞察力为开发潜在的解药铺平了道路.
相关概念视频
 00:49
Ligand Binding and Linkage
4.4K
Allosteric proteins have more than one ligand binding site; the binding of a ligand to any of these sites influences the binding of ligands to the other sites. When a protein is allosteric, its binding sites are called coupled or linked. In the case of enzymes, the site that binds to the substrate is known as the active site and the other site is known as the regulatory site. When a ligand binds to the regulatory site, this leads to conformational changes in the protein that can influence...
4.4K
 01:19
Allosteric Proteins-ATCase
4.8K
Binding sites linkages can regulate a protein's function. For example, enzyme activity is often regulated through a feedback mechanism where the end product of the biochemical process serves as an inhibitor.
Aspartate transcarbamoylase (ATCase) is a cytosolic enzyme that catalyzes the condensation of L-aspartate and carbamoyl phosphate to N-carbamoyl-L-aspartate. This reaction is the first step in pyrimidine biosynthesis. UTP and CTP, the end products of the pyrimidine synthesis...
Aspartate transcarbamoylase (ATCase) is a cytosolic enzyme that catalyzes the condensation of L-aspartate and carbamoyl phosphate to N-carbamoyl-L-aspartate. This reaction is the first step in pyrimidine biosynthesis. UTP and CTP, the end products of the pyrimidine synthesis...
4.8K
 01:29
Indirect-Acting Cholinergic Agonists: Chemistry and Structure-Activity Relationship
1.1K
Indirect-acting cholinergic agonists are agents that interact with the acetylcholinesterase enzyme in the synaptic cleft, preventing the breakdown of acetylcholine into choline and acetate. Consequently, the concentration of acetylcholine in the synaptic cleft increases. These agonists can be classified into reversible and irreversible inhibitors based on their duration of action.
Reversible inhibitors display short to medium durations of action. Short-acting agents include simple alcohols with...
Reversible inhibitors display short to medium durations of action. Short-acting agents include simple alcohols with...
1.1K
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
3.9K
Adrenergic agonists' structure-activity relationship (SAR) determines their selectivity and efficacy. These agonists comprise a phenylethylamine moiety with an aromatic ring and an ethylamine side chain.
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of...
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of...
3.9K
 01:17
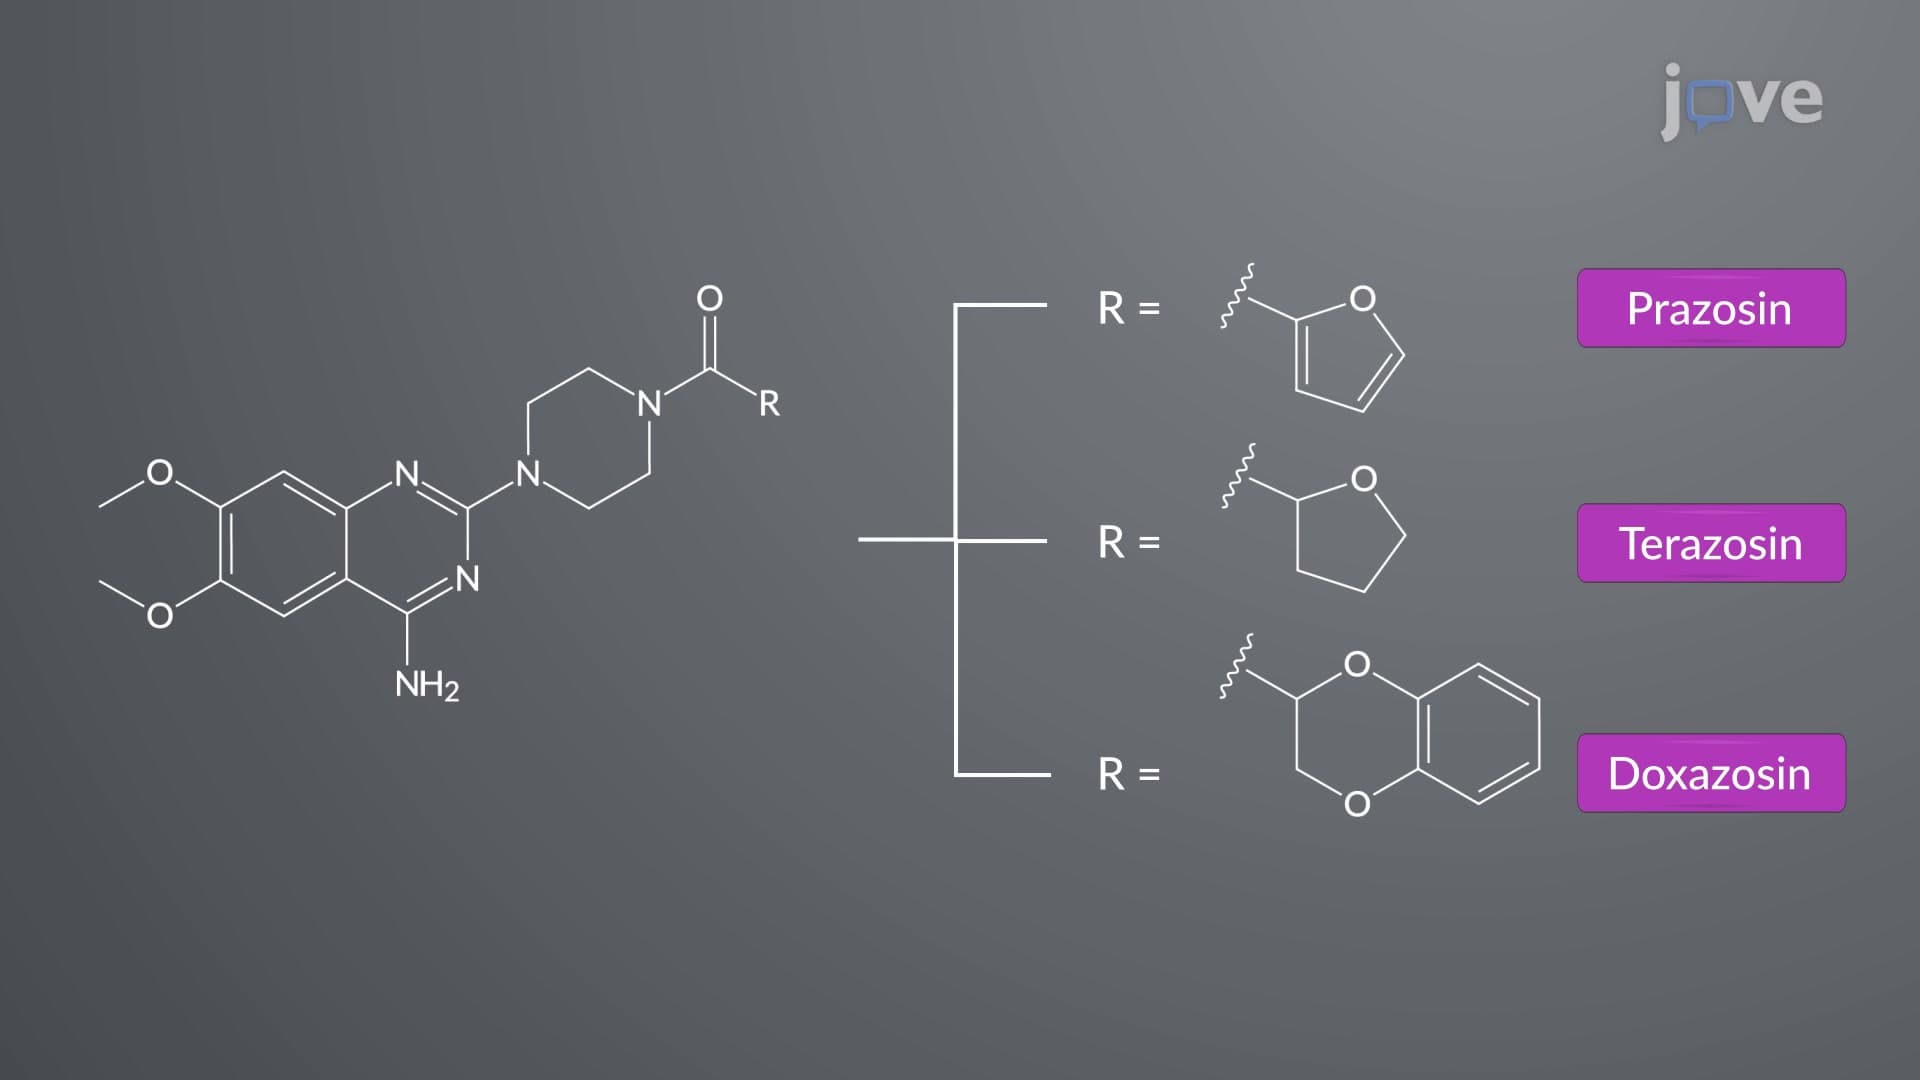
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
1.7K
Adrenergic antagonists, or sympatholytics, inhibit adrenoceptor activation driven by catecholamines or agonists. Based on their adrenoceptor specificity, adrenergic blockers can be categorized into two primary groups: α-adrenergic blockers (α-blockers) and β-adrenergic blockers (β-blockers). α-blockers interact with α1 and α2 subtypes of α-adrenoceptors.
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline...
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline...
1.7K
 01:28
Structure-Activity Relationships and Drug Design
1.9K
Drug design is a dynamic field that involves discovering and developing new medications based on specific biological targets. This process heavily relies on structure-activity relationships (SAR) and quantitative structure-activity relationships (QSAR) to guide the design and optimization of efficient drugs.
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
1.9K