在单核铁分子晶体中低旋转 - 高旋转相互作用的热力学效应的量化(II) 旋转交叉复合体
Juliusz A Wolny1, Konstantin Gröpl1, Jonathan Kiehl2
1Department of Physics, University of Kaiserslautern-Landau, Erwin-Schrödinger-Str. 46, 67663 Kaiserslautern, Germany. wolny@rptu.de.
Dalton transactions (Cambridge, England : 2003)
|April 29, 2024

概括
这项研究模拟了邻近的旋转如何影响旋转交叉复合体,根据周围的旋转状态发现了显著的差异. 邻近的旋转会影响低频振动和协调球体几何,影响旋转过渡.
科学领域:
- 固态化学 固态化学
- 计算材料科学 计算材料科学
- 旋转交叉现象 旋转交叉现象
背景情况:
- 旋转交叉 (SCO) 复合体在低旋转 (LS) 和高旋转 (HS) 状态之间表现出双稳定性.
- 分子间相互作用,特别是分子晶体,显著影响SCO行为.
- 了解邻近旋转的能量和贡献对于SCO材料设计至关重要.
研究的目的:
- 开发和应用一种方法来估计邻近旋转对SCO综合体的能量和效应.
- 使用密度函数理论 (DFT) 建模旋转交叉材料[FeII(Lnpdtz) 2(NCS) 2.
- 研究邻近的旋转状态对旋转过渡的合作性和振动的影响.
主要方法:
- 利用密度函数理论 (DFT) 来建模15个SCO分子的2D排列.
- 对LS和HS邻系矩阵中的分子进行了几何优化和正常模式计算.
- 计算了合作性参数 (Hcoop) 并分析了振动贡献.
主要成果:
- 计算的Hcoop=11kJmol-1对于研究的SCO复合体,可与3D多核SCO材料进行比较.
- 在LS矩阵中,LS向HS过渡的振动比HS矩阵更高 (5JK-1mol-1).
- 邻近的自旋相互作用主要影响低频模式 (<65-70 cm-1) 并导致中央分子协调球的变形.
结论:
- 拟议的DFT方法有效地估计了邻近旋转对SCO过渡的影响.
- 邻近的自旋状态显著影响振动和分子几何学,这对上合组织合作至关重要.
- 这些发现强调了在设计先进的SCO材料时考虑分子间效应的重要性.
相关概念视频
 03:02
Colors and Magnetism
11.6K
Color in Coordination Complexes
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...
11.6K
 02:42
Valence Bond Theory
8.5K
Coordination compounds and complexes exhibit different colors, geometries, and magnetic behavior, depending on the metal atom/ion and ligands from which they are composed. In an attempt to explain the bonding and structure of coordination complexes, Linus Pauling proposed the valence bond theory, or VBT, using the concepts of hybridization and the overlapping of the atomic orbitals. According to VBT, the central metal atom or ion (Lewis acid) hybridizes to provide empty orbitals of suitable...
8.5K
 02:58
Crystal Field Theory - Octahedral Complexes
26.4K
Crystal Field Theory
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
26.4K
 01:22
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
1.1K
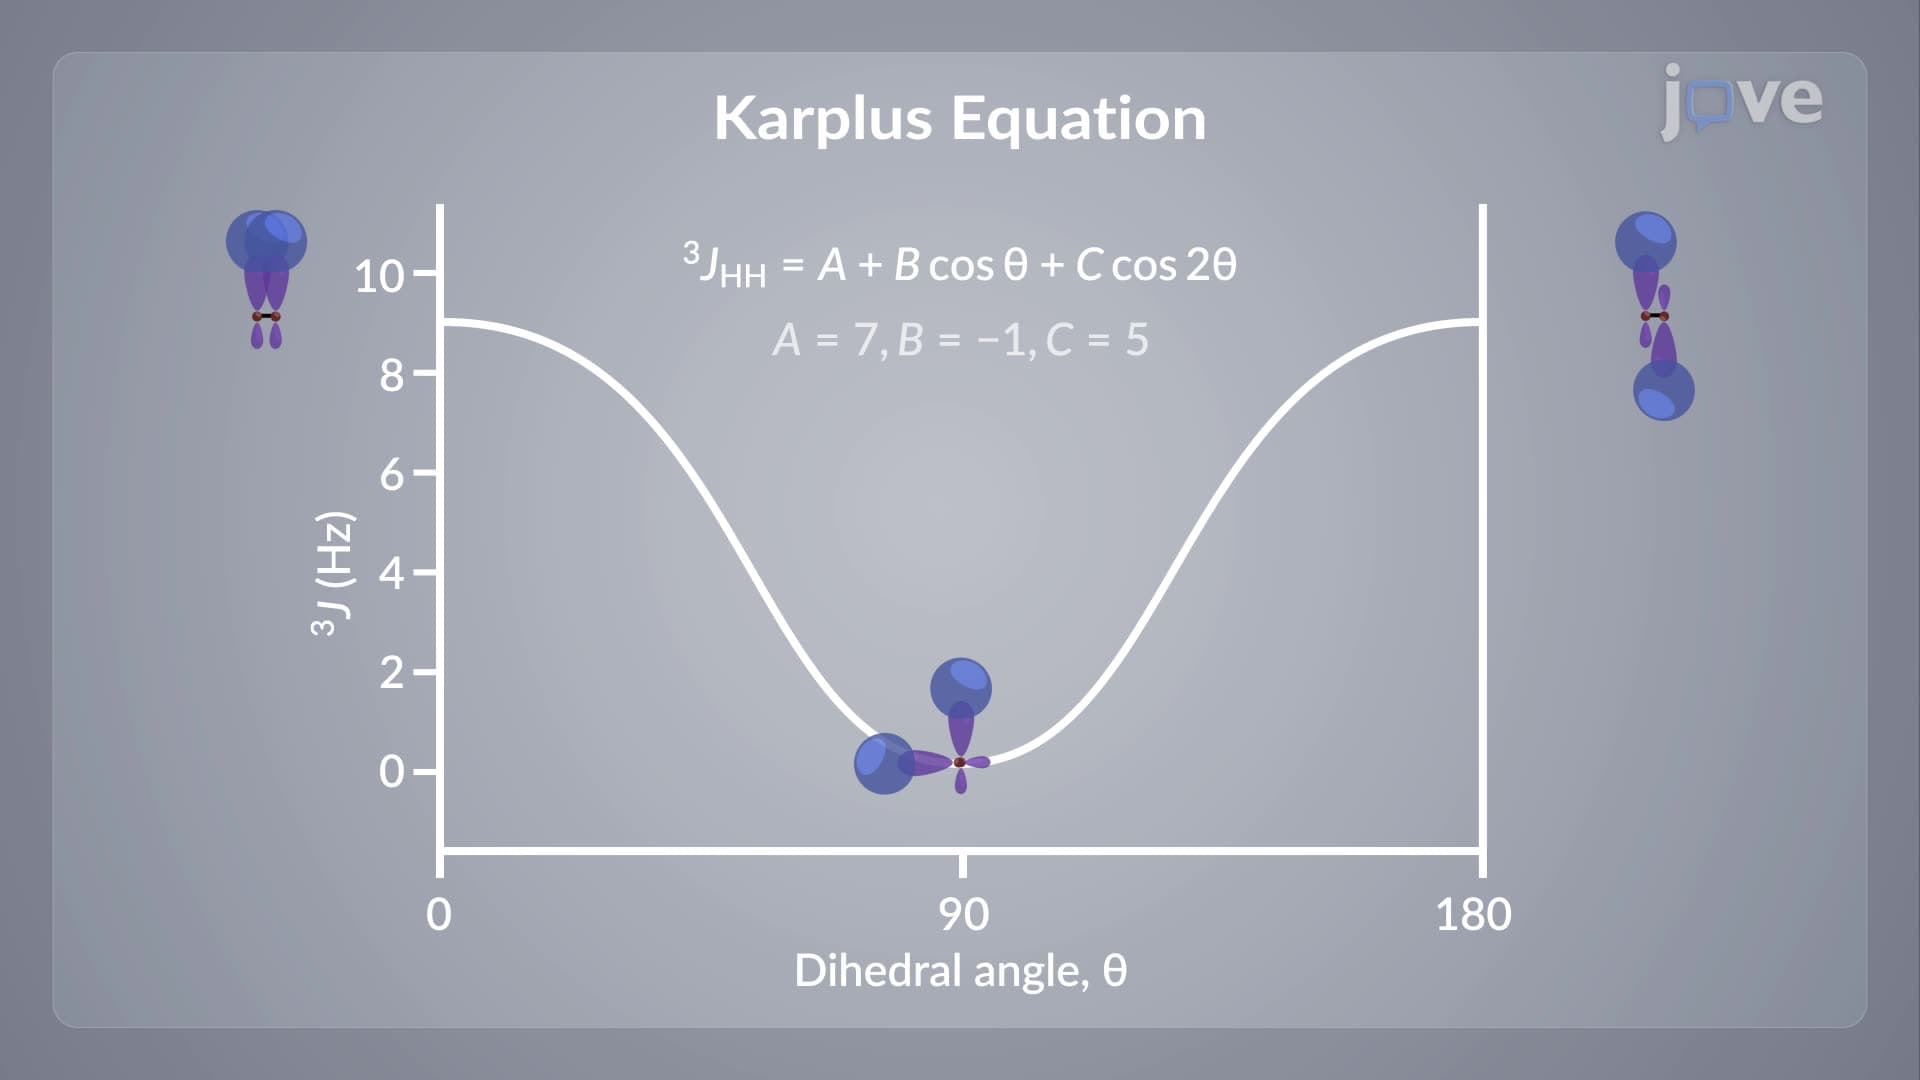
Vicinal or three-bond coupling is commonly observed between protons attached to adjacent carbons. Here, nuclear spin information is primarily transferred via electron spin interactions between adjacent C‑H bond orbitals. This generally favors the antiparallel arrangement of spins, so 3J values are usually positive.
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
1.1K
 01:08
NMR Spectroscopy: Spin–Spin Coupling
1.4K
The spin state of an NMR-active nucleus can have a slight effect on its immediate electronic environment. This effect propagates through the intervening bonds and affects the electronic environments of NMR-active nuclei up to three bonds away; occasionally, even farther. This phenomenon is called spin–spin coupling or J-coupling. Coupling interactions are mutual and result in small changes in the absorption frequencies of both nuclei involved. While nuclei of the same element are involved...
1.4K
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
42.4K
Tetrahedral Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
42.4K