相关实验视频
Updated: Jun 27, 2025
05:46
Rapid Screening of HIV Reverse Transcriptase and Integrase Inhibitors
Published on: April 9, 2014
17.8K
长效卡博特格拉维尔的药理动力学,有或没有口服的HIV PrEP的先导
Kelong Han1, Parul Patel2, Scott McCallister3
1GSK, Collegeville, Pennsylvania, USA.
Antimicrobial agents and chemotherapy
|May 6, 2024

概括
长期起作用的卡博特格拉维尔的药理动力学特征与口服领入剂或不带领的药理动力特征相似. 可选的口服引入可以跳过,因为卡博特格拉维尔暴露在短时间的注射间隔中仍然有效.
科学领域:
- 药理动力学 药理动力学
- 预防和治疗艾滋病病毒
背景情况:
- 长效卡博特格拉维尔已被批准用于HIV暴露前预防 (PrEP) 和治疗.
- 长效卡博特格拉维尔的启动包括一个可选的口服引入 (OLI) 阶段.
- 需要评估OLI对长期起作用的卡博特格拉维尔药理学的影响.
研究的目的:
- 评估OLI对长效卡博特格拉维尔药理动学的影响.
- 为了比较不同研究群体中含有和没有OLI的卡博特格拉维尔血度.
- 在各种OLI注射缺口场景下模拟药物动力学概况.
主要方法:
- 在HIV阳性和HIV阴性参与者中比较卡博特格拉维尔的血度,这些参与者开始注射有或没有OLI.
- 利用了第三阶段治疗研究FLAIR和关键PrEP研究HPTN 083和HPTN 084的数据.
- 在不同的OLI注射缺口场景下,在分配男性出生时和分配女性出生时的种群中模拟的药理动力学概况.
主要成果:
- 观察到的卡博特格拉维尔的最低度与OLI和没有OLI相似 (P > 0.3).
- 模拟的注射前最低度仍然高于药理动力学目标,有3天的OLI-注射间隔.
- 在所有场景中,PK概况在第一次注射后大约1-2周变得几乎相同.
结论:
- 观察到的长效卡博特格拉维尔暴露与OLI或没有OLI相似,支持可选的OLI使用.
- 预计卡博特格拉维尔暴露在OLI注射间隔≤3天的药理动力学目标以上.
- 结果在分配的男性出生和分配的女性出生人群中一致,支持灵活的OLI启动.

相关概念视频
 01:24
Factors Influencing Drug Absorption: Presystemic Elimination
212
The pharmacokinetic journey of oral drugs begins with a crucial first pass through the hepatic portal system, called the first-pass effect. This first pass significantly impacts bioavailability — the proportion of a drug that enters systemic circulation and is available for therapeutic action. The primary route sees the drug absorbed by intestinal membranes and then shunted to the liver via the hepatic portal vein. Here, pre-systemic elimination occurs as drugs face metabolism or biliary...
212
 01:10
Retrovirus Life Cycles
45.9K
Retroviruses have a single-stranded RNA genome that undergoes a special form of replication. Once the retrovirus has entered the host cell, an enzyme called reverse transcriptase synthesizes double-stranded DNA from the retroviral RNA genome. This DNA copy of the genome is then integrated into the host’s genome inside the nucleus via an enzyme called integrase. Consequently, the retroviral genome is transcribed into RNA whenever the host’s genome is transcribed, allowing the...
45.9K
 01:13
Bioavailability: Overview
2.8K
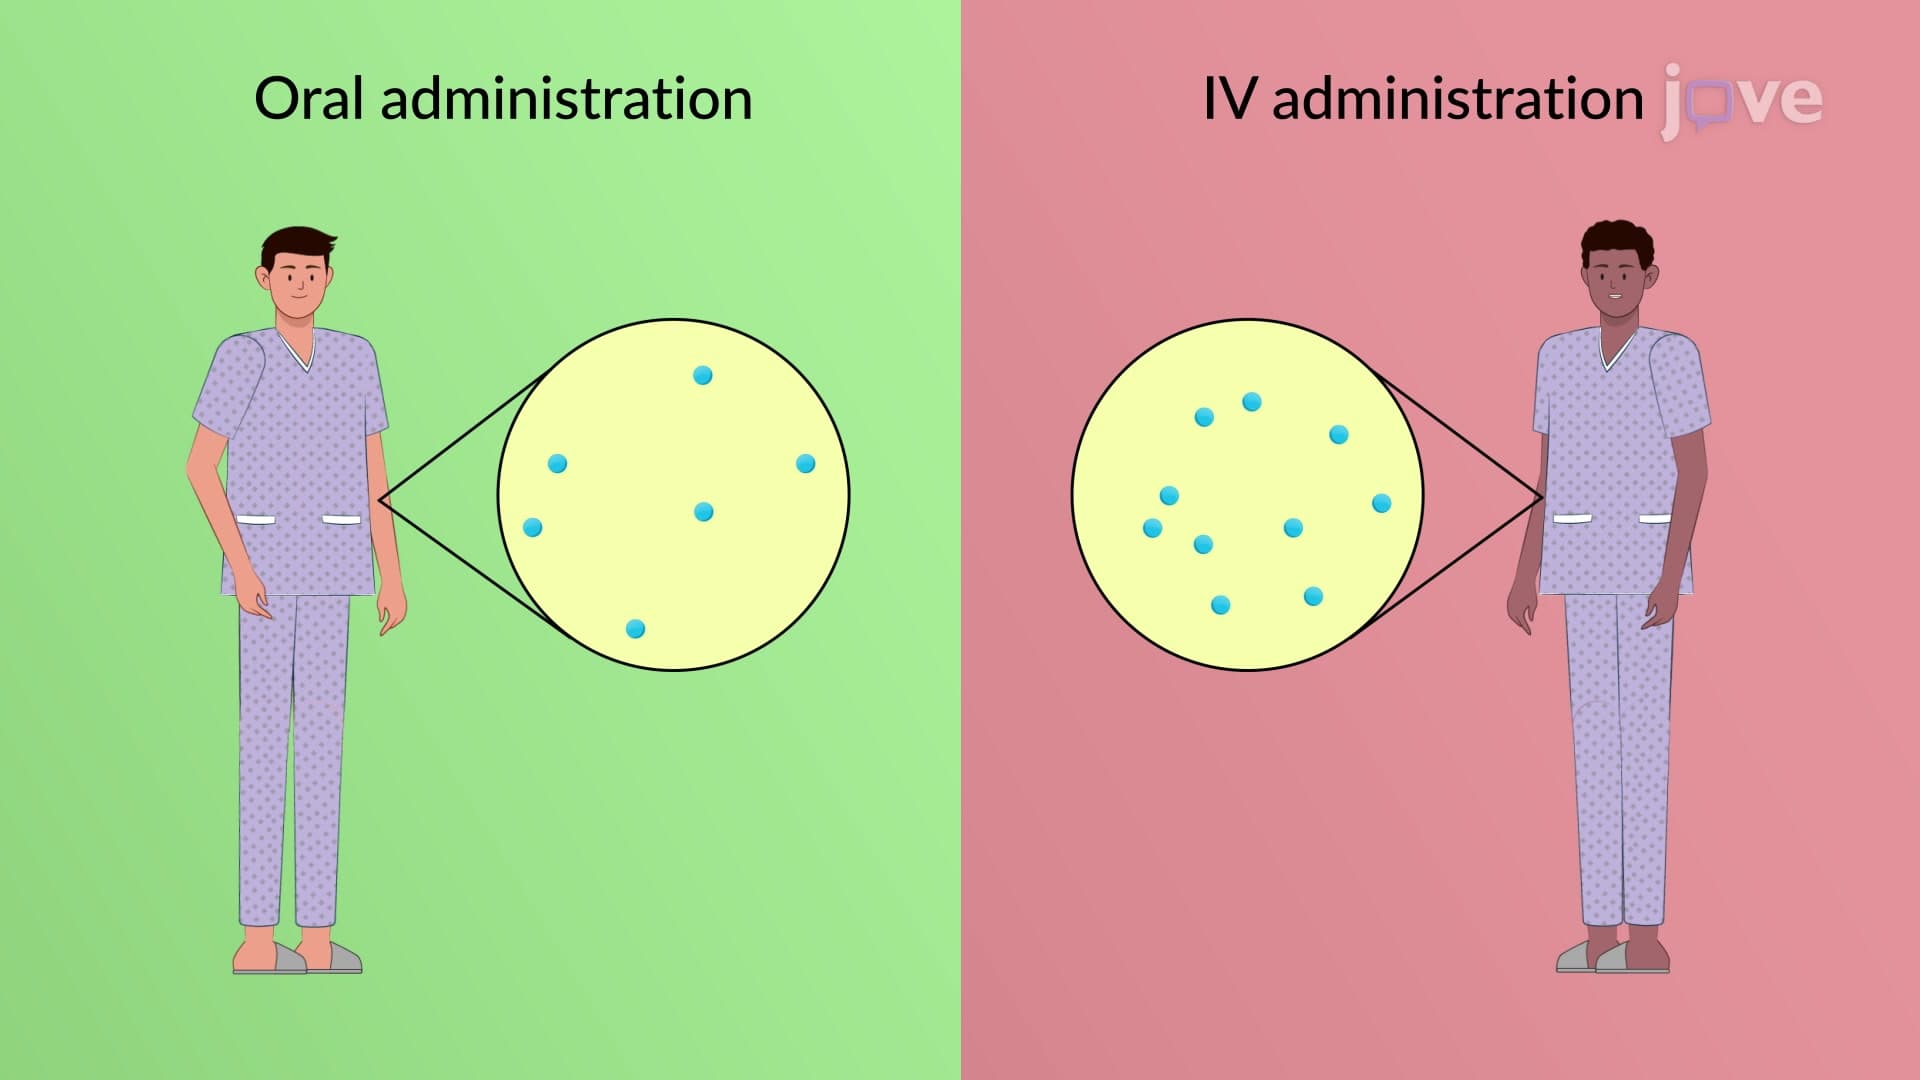
Bioavailability refers to the proportion of an unaltered drug that, after administration, enters the systemic circulation and can be distributed to the desired action site. Factors such as gastrointestinal (GI) absorption and liver biotransformation influence the bioavailability of a drug when it is administered orally. When a drug is administered intravenously, it enters the systemic circulation directly; by definition, its bioavailability is assumed to be 100%. The bioavailability of an...
2.8K
 01:23
Nonlinear Pharmacokinetics: Dependence of Elimination Half-Life and Dose Clearance
126
The elimination half-life and drug clearance of drugs following nonlinear kinetics can vary with dosage. The Michaelis-Menten parameters and drug concentration influence these factors. As the dose increases, the elimination half-life tends to lengthen, resulting in a reduction in clearance and a disproportionately larger area under the curve. The total clearance can be derived from the Michaelis-Menten equation for drugs following a one-compartment model.
A study on guinea pigs examined the...
A study on guinea pigs examined the...
126
 01:14
Time Course of Drug Effect
2.0K
The progression of a drug's impact can be analyzed by examining both the concentration-time course and the effect-time course. The concentration-time course is determined by the drug's half-life and is influenced by factors such as its pharmacokinetics, including absorption, distribution, metabolism, and elimination. The effect of the drug is often related to its concentration in the plasma and is calculated using the maximum drug effect and the plasma concentration that generates 50...
2.0K
 01:22
Nonlinear Pharmacokinetics: Bioavailability and Protein-Drug Binding
157
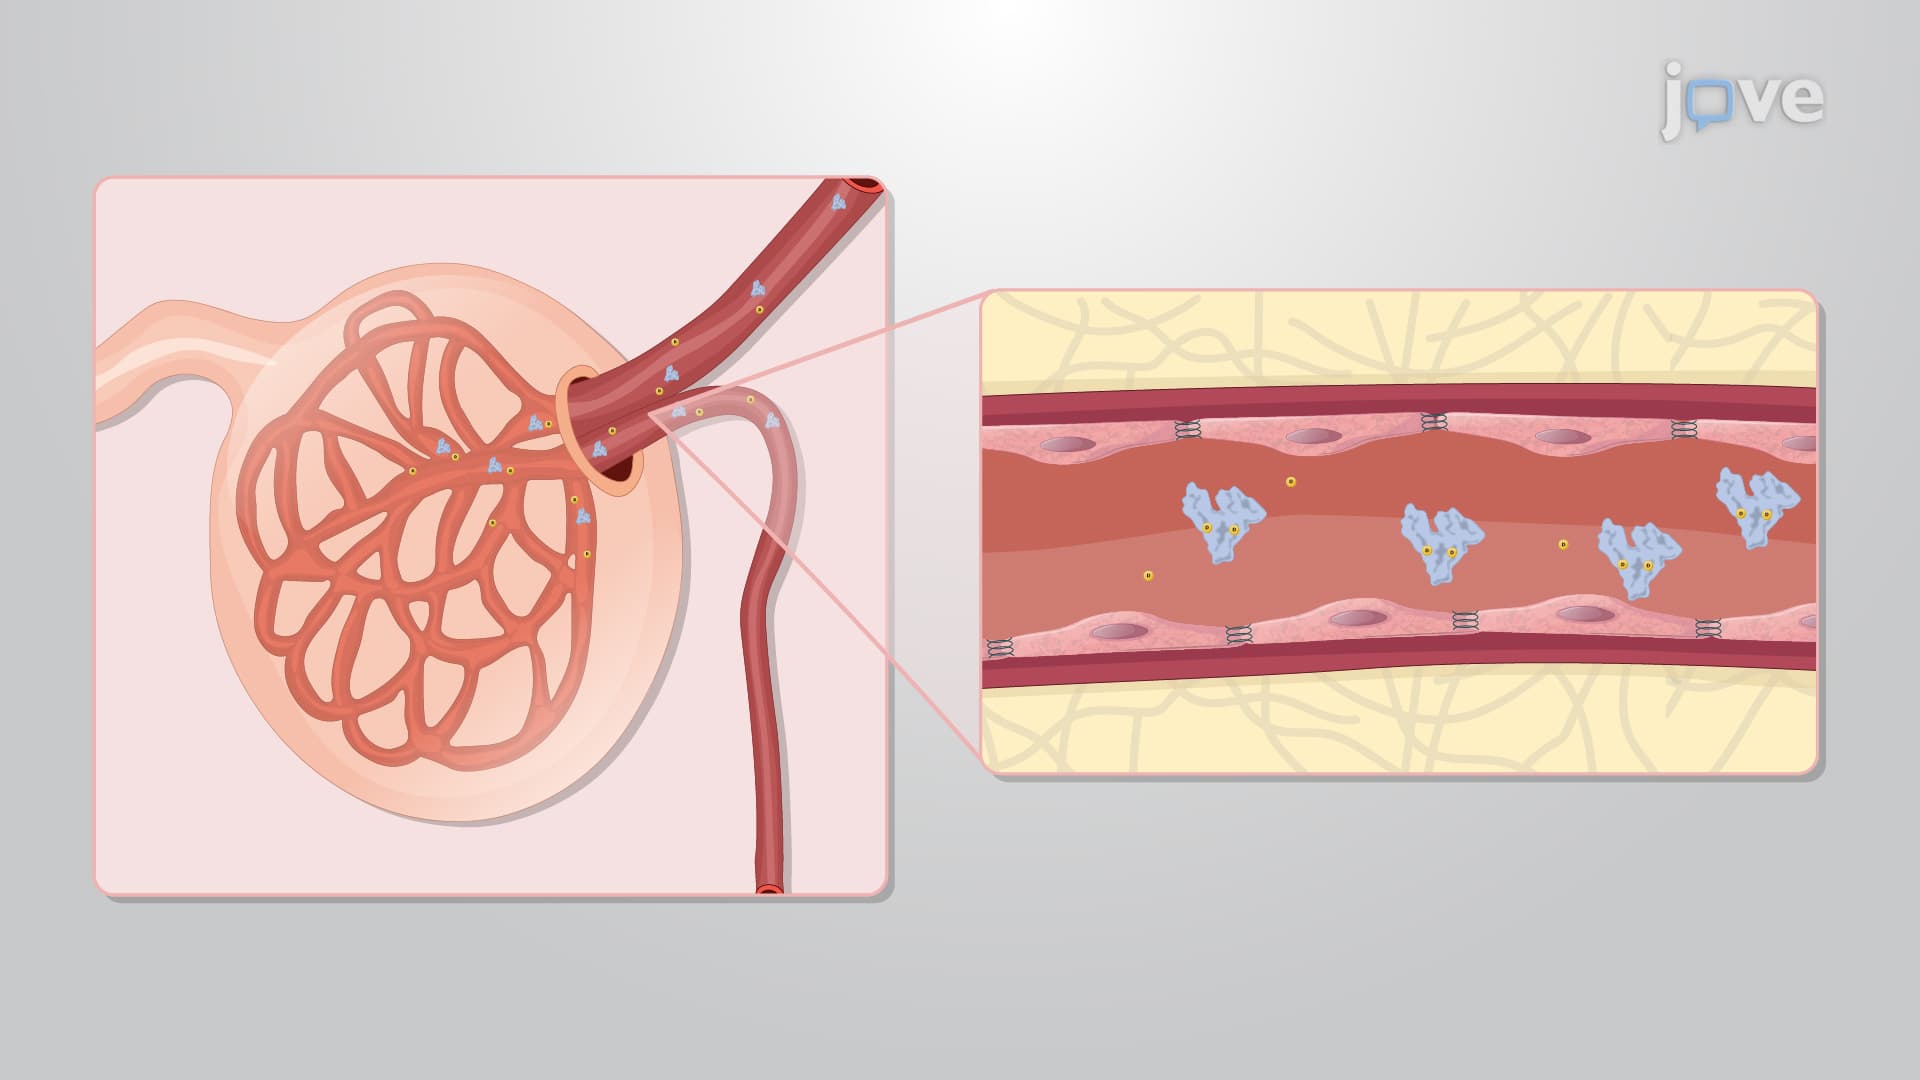
When a drug follows nonlinear pharmacokinetics, its bioavailability, the amount of the drug that reaches the systemic circulation, can change with different doses. This is due to the presence of a saturable pathway. The pathway becomes saturated as the drug concentration increases, decreasing the absorption rate. Consequently, the drug's bioavailability may be lower than expected at higher doses.
To quantify the extent of bioavailability, pharmacologists often use a parameter called .
To quantify the extent of bioavailability, pharmacologists often use a parameter called .
157