

使用多分辨率分析和量子计算的范围分离密度函数理论
Nicolas Poirier1,2, Jakob S Kottmann3, Alán Aspuru-Guzik4,5,6,7
1Department of Mechanical Engineering, McGill University, Montreal, Quebec, Canada.

研究人员通过使用范围分离密度函数理论 (RS-DFT) 和对自然轨道 (PNO) 来分割分子系统来降低量子化学计算的计算成本. 这种方法需要更少的量子比特来准确地计算量子计算机上的基本状态能量.
科学领域:
- 量子计算是一种量子计算.
- 计算化学的计算化学
- 量子化学 是一个量子化学.
背景情况:
- 量子计算机为复杂的量子化学问题提供了与经典计算机相比显著的优势.
- 目前的量子化学计算在计算上昂贵,这限制了它们的应用.
- 分区分子系统是降低计算成本的关键策略.
研究的目的:
- 研究一种用于减少量子化学模拟所需的计算资源的新方法.
- 探索区分密度函数理论 (RS-DFT) 在分离分子系统中的有效性.
- 评估使用对自然轨道 (PNO) 对量子算法的效率的影响.
主要方法:
- 分子系统使用范围分离密度函数理论 (RS-DFT) 进行分区.
- 双自然轨道 (PNO) 是使用基准独立多解析度分析 (MRA) 框架生成的.
- 使用变量量子自溶解器 (VQE) 算法,对各种分子测试该策略.
主要成果:
- 与波函数理论 (WFT) 相比,RS-DFT减少了地面状态能量对基础集和活性空间大小的依赖.
- 使用PNO导致了更紧的量子比特哈密尔顿.
- 提出的方法显著减少了实现目标能量精度所需的量子比特数量.
结论:
- RS-DFT和PNO的组合为量子化学计算提供了一个有效的策略.
- 这种方法降低了精确的能量计算对量子比特的要求,使量子模拟更加可行.
- 这些发现为更容易获得和可扩展的化学量子模拟铺平了道路.
更多相关视频
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
相关概念视频
 02:45
The Quantum-Mechanical Model of an Atom
 01:13
Mass Analyzers: Overview
 01:15
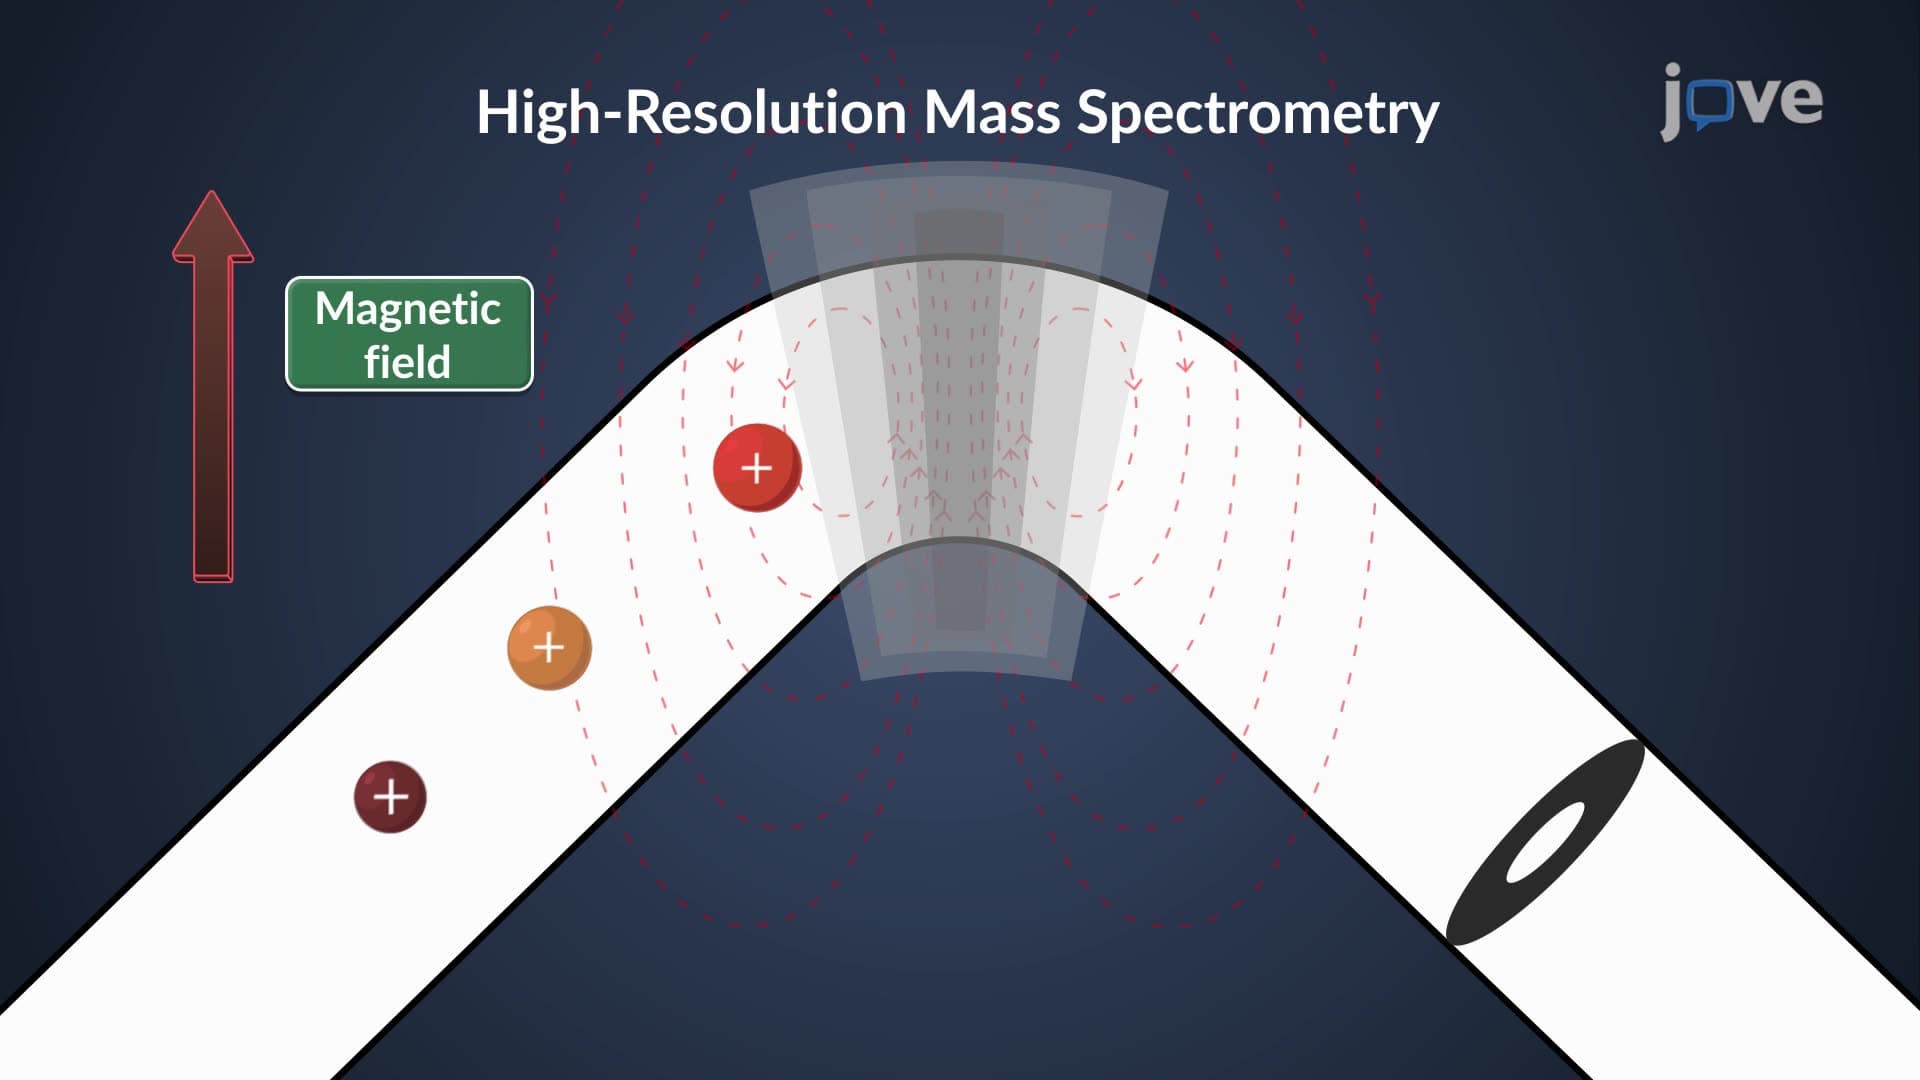
High-Resolution Mass Spectrometry (HRMS)
 02:35
Molecular Orbital Theory I
 01:15
Chromatographic Resolution
The effectiveness of separation can be evaluated by determining the level of separation between two neighboring peaks in a chromatogram, which represents the individual components of a sample.
In chromatography,...
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are...