相关实验视频
Updated: Jun 26, 2025
13:33

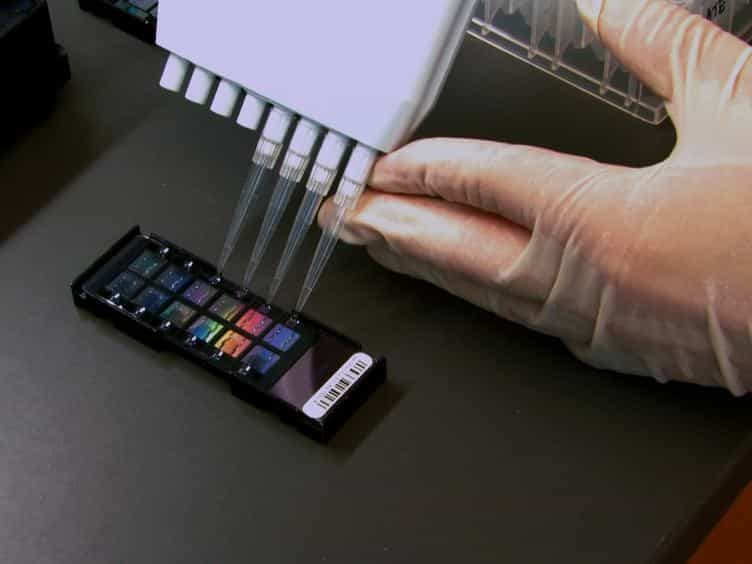
Infinium Assay for Large-scale SNP Genotyping Applications
Published on: November 19, 2013
39.0K
在挪威人口中使用大规模并行测序对自身体STR和身份SNP的概述
Maria Martin Agudo1, Håvard Aanes2, Michel Albert2
1Department of Forensic Sciences, Oslo University Hospital, Oslo, Norway; Department of Forensic Medicine, Institute of Clinical Medicine, University of Oslo, Oslo, Norway.
Forensic science international. Genetics
|May 11, 2024

概括
基于序列的 (SB) 对挪威南部的法医标记的基因频率显示出比基于长度的 (LB) 数据更高的基因多样性. 这些发现支持对挪威和丹麦人口的基因型概率计算中可互换的等位基因频率的使用.
科学领域:
- 法医遗传学 法医遗传学
- 人口遗传学 人口遗传学
- 大规模并行测序 (MPS) 是一种
背景情况:
- 概率基因型软件越来越多地用于分析法医大规模并行测序 (MPS) 数据.
- 概率比率 (LR) 取决于相关人群的精确等位基频率数据.
- 基于序列 (SB) 的数据在法医分析中比基于长度 (LB) 的数据提供了潜在的优势.
研究的目的:
- 在南挪威人群中建立基于序列 (SB) 的基因频率,用于自体短串重复 (aSTR) 和同一单核酸多态 (iSNPs) 的基因频率.
- 为了比较SB和LB数据之间的等位基因多样性和歧视力.
- 评估较小的挪威南部数据集对较大的挪威人口的代表性,并调查与其他人口的遗传差异.
主要方法:
- 在挪威南部的371个人中分析了27个aSTR和94个iSNP,使用ForenSeqTM DNA签名准备套件.
- 在SB和LB数据之间比较等位基频率和多样性指标.
- 评估样本大小对小等位基因频率 (MAF) 值的影响以及人口结构分析 (FST),并与北挪威和丹麦数据集进行比较.
主要成果:
- 与LB等位基相比,SB等位基的罕见等位基频率显著更高 (<0.05) 和等位基多样性更大.
- SB数据显示,随机匹配概率较低,增加了个人歧视.
- 发现了新的等位基因和SNP,包括TH01标记物中一个新的SNP (MAF 0.001).
- 发现南挪威数据集 (n=371) 是较大挪威人口 (n=15,769) 的代表.
- 据FST估计,在挪威南部没有显著的遗传结构.
- 在挪威南部和挪威北部之间观察到最小的遗传差异,而挪威南部和丹麦则显示出一些标志物特定的差异.
结论:
- 基于序列的基因频率为法医应用提供了卓越的区分能力.
- 挪威南部的确定的等位基因频率适用于法医案例工作.
- 来自研究种群 (挪威南部,挪威北部,丹麦) 的等位基因频率可以互换地用于基因型概率计算,在挪威南部和丹麦之间对特定标记的考虑较小.

相关概念视频
 01:05
Single Nucleotide Polymorphisms-SNPs
15.0K
A single nucleotide polymorphism or SNP is a single nucleotide variation at a specific genomic position in a large population. It is the most prevalent type of sequence variation found in the human genome. Point mutations that occur in more than 1% of the population qualify as SNPs. These are present once every 1000 nucleotides on an average in the human genome. Replacement of a purine with another purine (A/G) or a pyrimidine with another pyrimidine (C/T) is known as a transition. In contrast,...
15.0K
 02:26
Comparing Copy Number Variations and SNPs
17.7K
Sequencing of the human genome has opened up several best-kept secrets of the genome. Scientists have identified thousands of genome variations that exist within a population. These variations can be a single nucleotide or a larger chromosomal variation.
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...
17.7K
 03:21
RNA-seq
9.9K
RNA sequencing, or RNA-Seq, is a high-throughput sequencing technology used to study the transcriptome of a cell. Transcriptomics helps to interpret the functional elements of a genome and identify the molecular constituents of an organism. Additionally, it also helps in understanding the development of an organism and the occurrence of diseases.
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
9.9K
 03:00
Next-generation Sequencing
88.7K
The first human genome sequencing project cost $2.7 billion and was declared complete in 2003, after 15 years of international cooperation and collaboration between several research teams and funding agencies. Today, with the advent of next-generation sequencing technologies, the cost and time of sequencing a human genome have dropped over 100 fold.
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
88.7K
 01:11
Genome-wide Association Studies-GWAS
13.3K
Genome-wide association studies or GWAS are used to identify whether common SNPs are associated with certain diseases. Suppose specific SNPs are more frequently observed in individuals with a particular disease than those without the disease. In that case, those SNPs are said to be associated with the disease. Chi-square analysis is performed to check the probability of the allele likely to be associated with the disease.
GWAS does not require the identification of the target gene involved in...
GWAS does not require the identification of the target gene involved in...
13.3K