
N··CO n → π* 相互作用:气相电子和振动光谱与量子化学计算相结合
1Department of Chemistry, Indian Institute of Science Education and Research Pune, Dr. Homi Bhabha Road, Pashan, Pune 411008, India.
The journal of physical chemistry. A
|May 30, 2024

概括
这项研究揭示了N···CO n → π*在乙烯-2-(2-(dimethylamino) phenyl) 乙酸盐中的相互作用,使用气相光谱学. 这一发现对于理解药物设计中的分子相互作用至关重要.
科学领域:
- 物理化学 物理化学
- 频谱学是一种光谱学.
- 计算化学的计算化学
背景情况:
- 非共价相互作用在分子识别和药物设计中至关重要.
- 了解像N···CO n → π*这样的特定相互作用需要先进的光谱方法.
研究的目的:
- 通过气相光谱学研究乙烯基2 - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - - 的乙烯基二- - - - - - - - - - - - - - 的N····CO n → π*相互作用.
- 为了确认单个调节器的存在,并描述其电子和振动特性.
- 为药物设计中的应用提供见解.
主要方法:
- 气相电子光谱学使用共振两光子电离.
- 紫外线-紫外线孔燃烧光谱学.
- 量子化学计算 (DFT). 量子化学计算 (DFT). 量子化学计算 (DFT). 量子化学计算 (DFT). 量子化学计算 (DFT). 量子化学计算 (DFT). 量子化学计算 (DFT). 量子化学计算 (DFT). 量子化学计算 (DFT). 量子化学计算 (DFT). 量子化学计算 (DFT).
主要成果:
- NMe2-Ph-EA碎片的相同电子光谱证实了离子状态解离.
- 紫外线-紫外线孔燃烧光谱检测发现了一个单一的适应器.
- 量子化学计算预测了N···CO n → π*在低能变态体中的相互作用.
- 实验和理论红外光谱表明,在观察到的适应体中存在这种相互作用.
结论:
- 在乙烯-2-(2-(二甲基胺) 乙烯酸中存在N···CO n → π*相互作用.
- 这种相互作用存在于实验观察到的气相适配器中.
- 这些发现对药物设计和分子识别有潜在的影响.
相关概念视频
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
1.4K
In Ultraviolet–Visible (UV–Vis) spectroscopy, the absorption of electromagnetic radiation is used to probe the electronic structure of molecules. This technique provides insights into molecular electronic transitions, particularly the movement of electrons between different molecular orbitals. Radiation is absorbed if the energy of the electromagnetic radiation passing through the molecule is precisely equal to the energy difference between the excited and ground states. During this...
1.4K
 01:27
π Electron Effects on Chemical Shift: Overview
1.1K
An applied magnetic field causes loosely bound π-electrons in organic molecules to circulate, producing a local or induced diamagnetic field over a large spatial volume. As the molecules tumble in solution, the field generated by π-electrons in spherical substituents results in a zero net field. However, the net field generated by π-electrons in non-spherical substituents is not zero. The effect of this induced field depends on the orientation of the molecule with respect to B0,...
1.1K
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
1.3K
A covalently bonded heteronuclear diatomic molecule can be modeled as two vibrating masses connected by a spring. The vibrational frequency of the bond can be expressed using an equation derived from Hooke's law, which describes how the force applied to stretch or compress a spring is proportional to the displacement of the spring. In this case, the atoms behave like masses, and the bond acts like a spring.
According to Hooke's law, the vibrational frequency is directly proportional to...
According to Hooke's law, the vibrational frequency is directly proportional to...
1.3K
 01:14
Molecular Spectroscopy: Absorption and Emission
2.3K
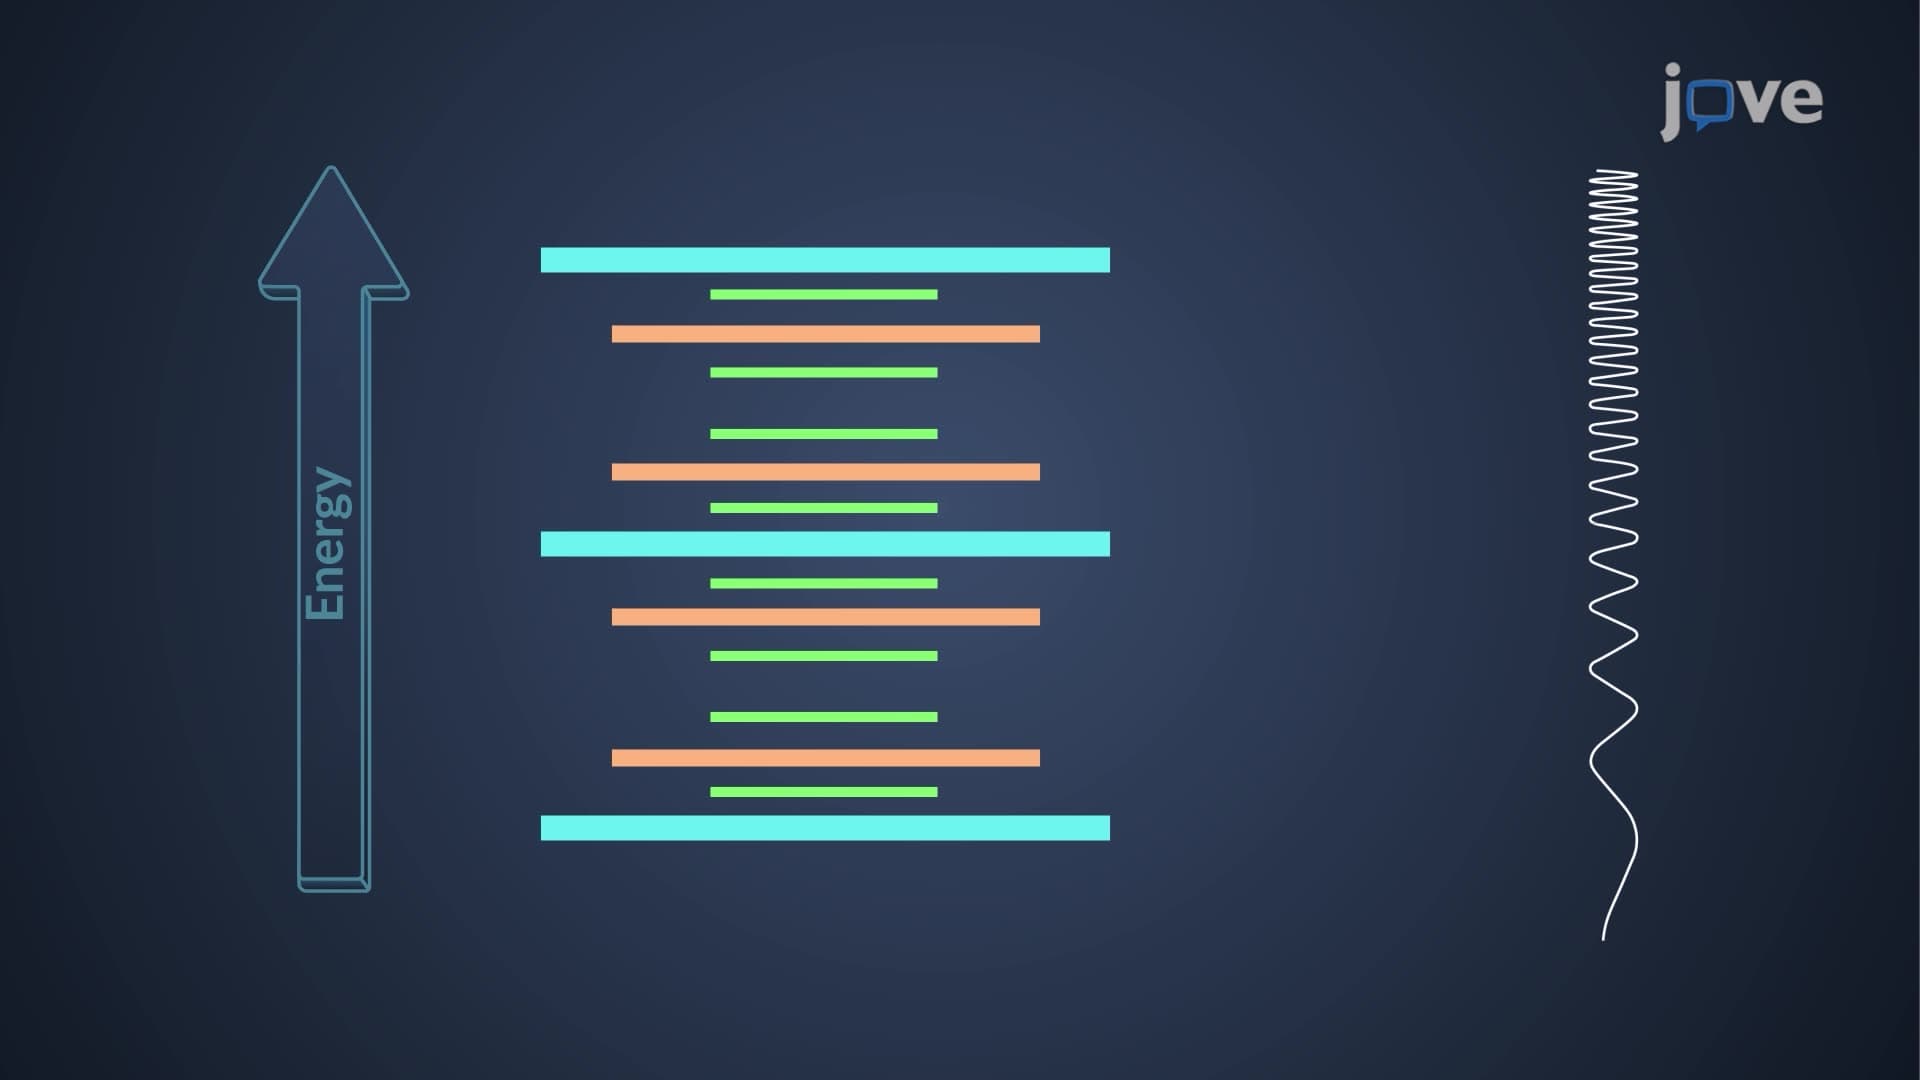
Molecules possess discrete energy levels called quantum states. Unlike atoms, which have simpler energy levels, molecules possess additional rotational and vibrational energy levels. Each energy level is separated by an energy gap, with the gaps between adjacent electronic, vibrational, and rotational levels varying significantly. The three types of energy levels in a diatomic molecule are shown in Figure 1.
2.3K
 02:35
Molecular Orbital Theory I
32.0K
Overview of Molecular Orbital Theory
32.0K
 02:40
MO Theory and Covalent Bonding
10.5K
The molecular orbital theory describes the distribution of electrons in molecules in a manner similar to the distribution of electrons in atomic orbitals. The region of space in which a valence electron in a molecule is likely to be found is called a molecular orbital. Mathematically, the linear combination of atomic orbitals (LCAO) generates molecular orbitals. Combinations of in-phase atomic orbital wave functions result in regions with a high probability of electron density, while...
10.5K