
BaMTeS (M = Fe,Mn,Zn) 和无序结构模拟BaGe0.5TeS的合成和特性
Emil H Frøen1, Domenic Nowak2, Peter Adler3
1Centre for Materials Science and Nanotechnology (SMN), Department of Chemistry, University of Oslo, Sem Sælands vei 26, Oslo N-0371, Norway.

研究人员合成了新的硫化 (BaMTeS) 化合物. BaFeTeS显示了接近200 K的隐藏过渡,可能是动态电荷密度波,尽管缺乏磁性排序.
科学领域:
- 固态化学和材料科学 固态化学和材料科学
- 对新型无机化合物的探索.
- 电子和磁性属性的研究.
背景情况:
- 三级硫化物-化物是一种新兴的材料类.
- 已知相的结构类比 (BaMSO) 提供了一个比较的基础.
- 了解结构属性关系对于材料设计至关重要.
研究的目的:
- 合成和表征新的三级硫化物 tellurides (BaMTeS). 为了合成和表征新的三级硫化物 tellurides (BaMTeS).
- 研究这些化合物的结构,电子和磁性特性.
- 探索BaFeTeS.潜在的相位过渡和潜在的机制.
主要方法:
- 固态合成用于化合物制备.
- 密度函数理论 (DFT) 分析用于财产调查.
- 实验技术:用于BaFeTeS的热容量,磁力测量和莫斯巴乌尔光谱学.
主要成果:
- 合成了四种BaMTeS化合物 (M = Fe,Mn,Zn,Ge),它们具有正方体晶体结构 (Cmcm).
- 对所有类型进行了DFT分析.
- 合成为纯相的BaFeTeS,在2-300K之间没有显示磁性排序或相位过渡.
- 莫斯巴乌尔光谱和电阻数据表明,在BaFeTeS中,有一个接近200K的隐藏过渡.
结论:
- 合成的BaMTeS化合物是BaMSO相的结构模拟物.
- BaFeTeS表现出复杂的行为,表明隐藏的过渡.
- 这种过渡暂时可以通过基于共振价值键 (RVB) 模型的动态电荷密度波机制来解释.
更多相关视频
 09:06
Visualizing Uniaxial-strain Manipulation of Antiferromagnetic Domains in Fe1+YTe Using a Spin-polarized Scanning Tunneling Microscope
Published on: March 24, 2019
 08:00
Chemical Synthesis of Porous Barium Titanate Thin Film and Thermal Stabilization of Ferroelectric Phase by Porosity-Induced Strain
Published on: March 27, 2018
相关概念视频
 02:42
Valence Bond Theory
 02:54
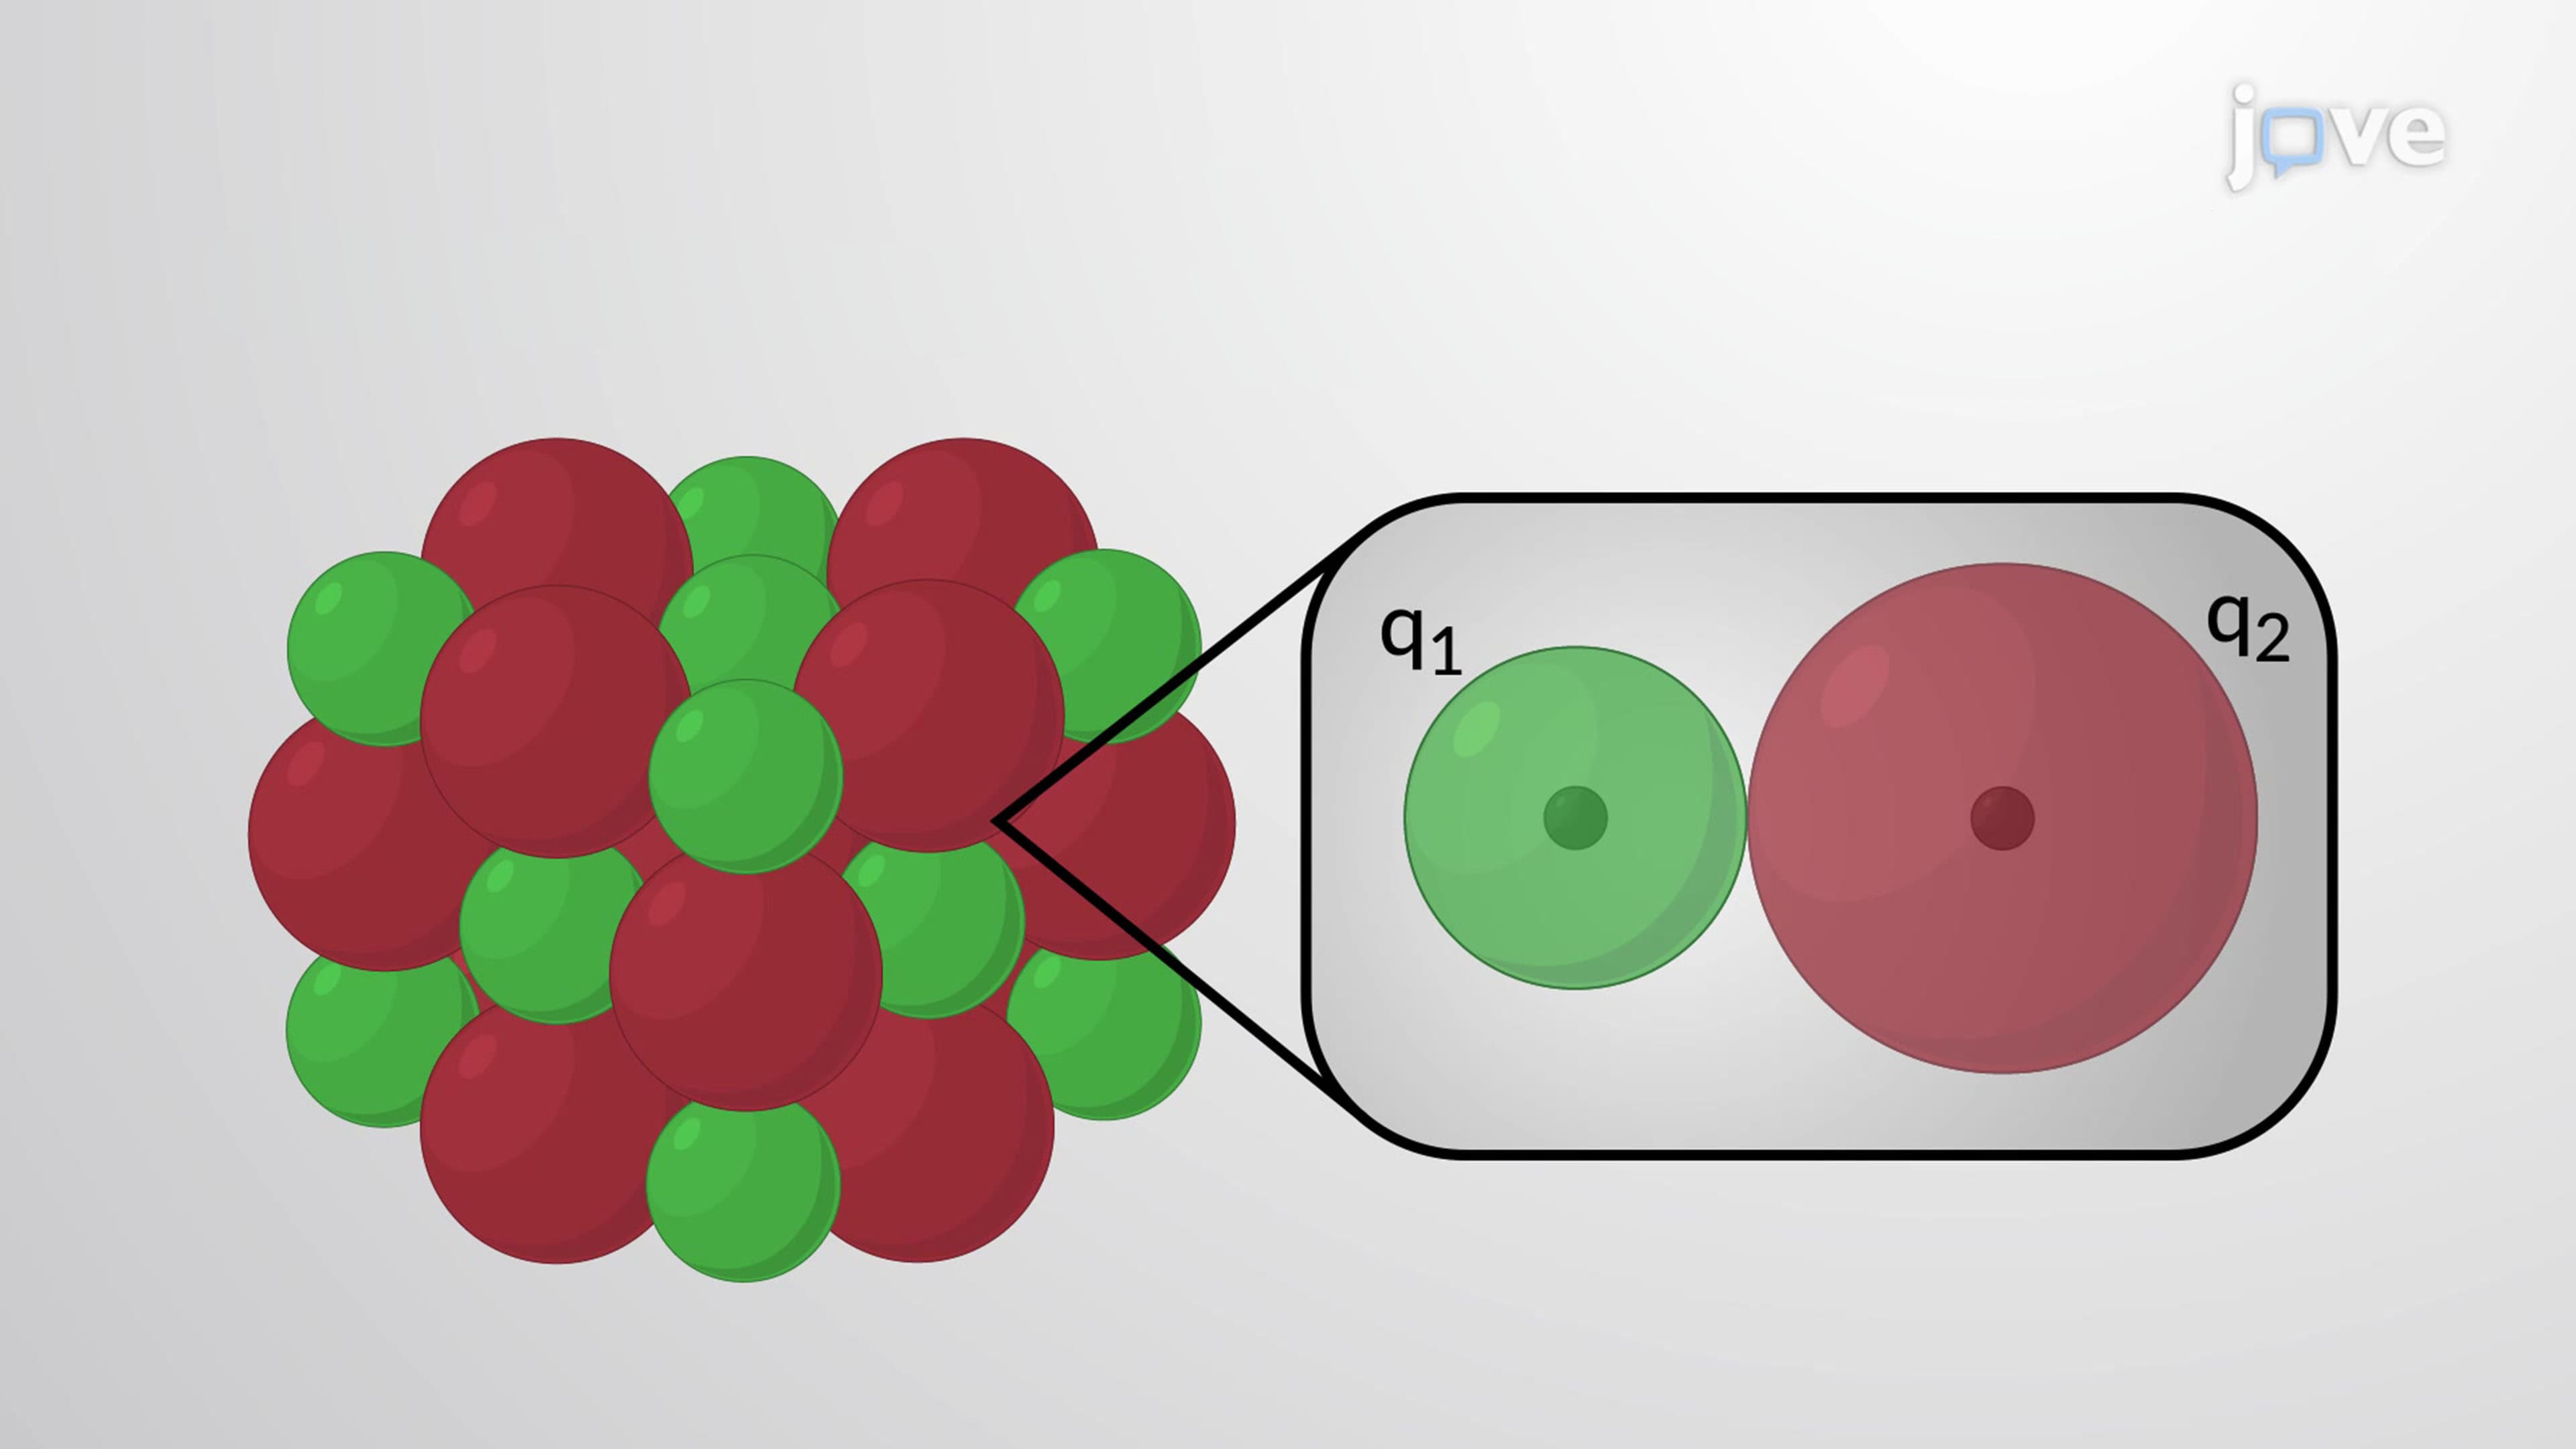
Trends in Lattice Energy: Ion Size and Charge
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...
 02:42
Ionic Crystal Structures
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 02:58
Properties of Transition Metals
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...