
碳硫化物 (OCS) 的高度精确的潜在能量表面:初始计算有多重要?
1Department of Physics and Astronomy, University College London, Gower Street, WC1E 6BT, London, UK. alec.owens.13@ucl.ac.uk.
Physical chemistry chemical physics : PCCP
|June 13, 2024

概括
准确的潜在能量表面 (PES) 对于预测分子振动波数至关重要. 这项研究表明,初始初始计算的质量显著影响了碳硫化物 (OCS) 经验上精制的 PES 的准确性.
科学领域:
- 计算化学计算化学
- 分子光谱学 分子光谱学
- 量子力学就是量子力学.
背景情况:
- 最初的量子化学方法产生了精确的分子潜在能量表面 (PES).
- 需要对PES进行经验性改进,以匹配实验室光谱数据,以便用于高分辨率的应用.
- 最初的ab initio计算对精细的PES准确性的影响需要严格的调查.
研究的目的:
- 调查基础的初始计算对潜在能量表面的经验改进的影响.
- 使用最先进的电子结构计算来构建和完善碳硫化物 (OCS) 的准确 PES.
- 为了证明由于精细的PES而导致计算的旋转振动能量水平的改善.
主要方法:
- 采用最先进的ab initio电子结构计算来生成OCS的高层和低层PES.
- 在生成的 PES 上执行几乎相同的实证精制程序.
- 计算基本的振动波数和反振动能量水平.
主要成果:
- 最初的ab initio计算的质量显著影响最终精细化的PES的准确性.
- 在OCS计算的旋转振动能量水平上实现了数量级的改进.
- 为OCS的电子基底状态生成了最准确的PES,基本值为0.004cm-1的RMSE,884个旋振水平为0.060cm-1.
结论:
- 初始计算的准确性是实现高精度,经验精细的潜在能量表面的关键因素.
- 为OCS开发的PES为光谱学预测提供了前所未有的准确性.
- 这项工作突显了分子光谱学中高级理论计算的重要性.

相关概念视频
 02:52
VSEPR Theory and the Basic Shapes
68.1K
Overview of VSEPR Theory
68.1K
 01:24
π Molecular Orbitals of 1,3-Butadiene
8.9K
Conjugated dienes have lower heats of hydrogenation than cumulated and isolated dienes, making them more stable. The enhanced stabilization of conjugated systems can be understood from their π molecular orbitals.
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
8.9K
 01:29
UV–Vis Spectroscopy: Woodward–Fieser Rules
24.1K
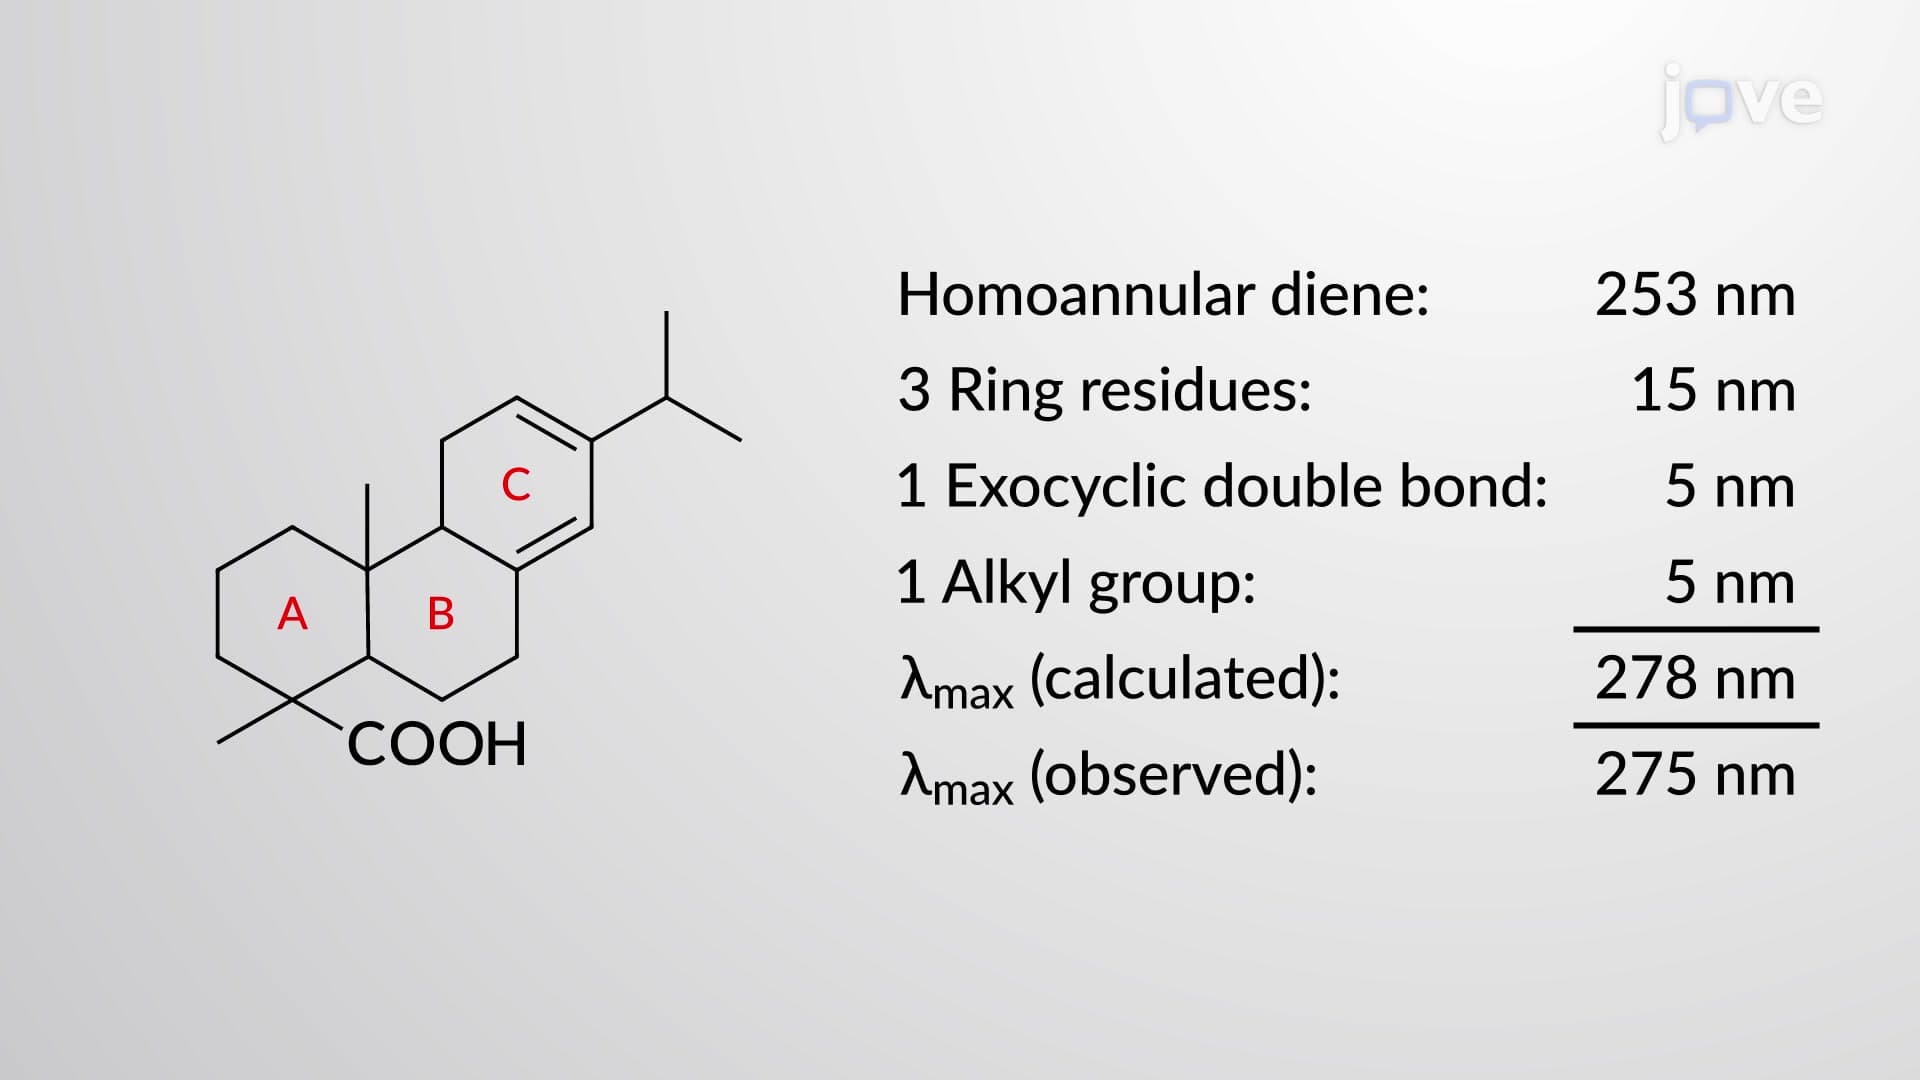
UV–Visible absorption spectra of conjugated dienes arise from the lowest energy π → π* transitions. The light-absorbing part of the molecule is called the chromophore, and the substituents directly attached to the chromophore are called auxochromes. A strong correlation exists between the absorption maxima, λmax, and the structure of a conjugated π system. The Woodward–Fieser rules predict the value of λmax for a given...
24.1K
 02:38
Valence Bond Theory and Hybridized Orbitals
19.3K
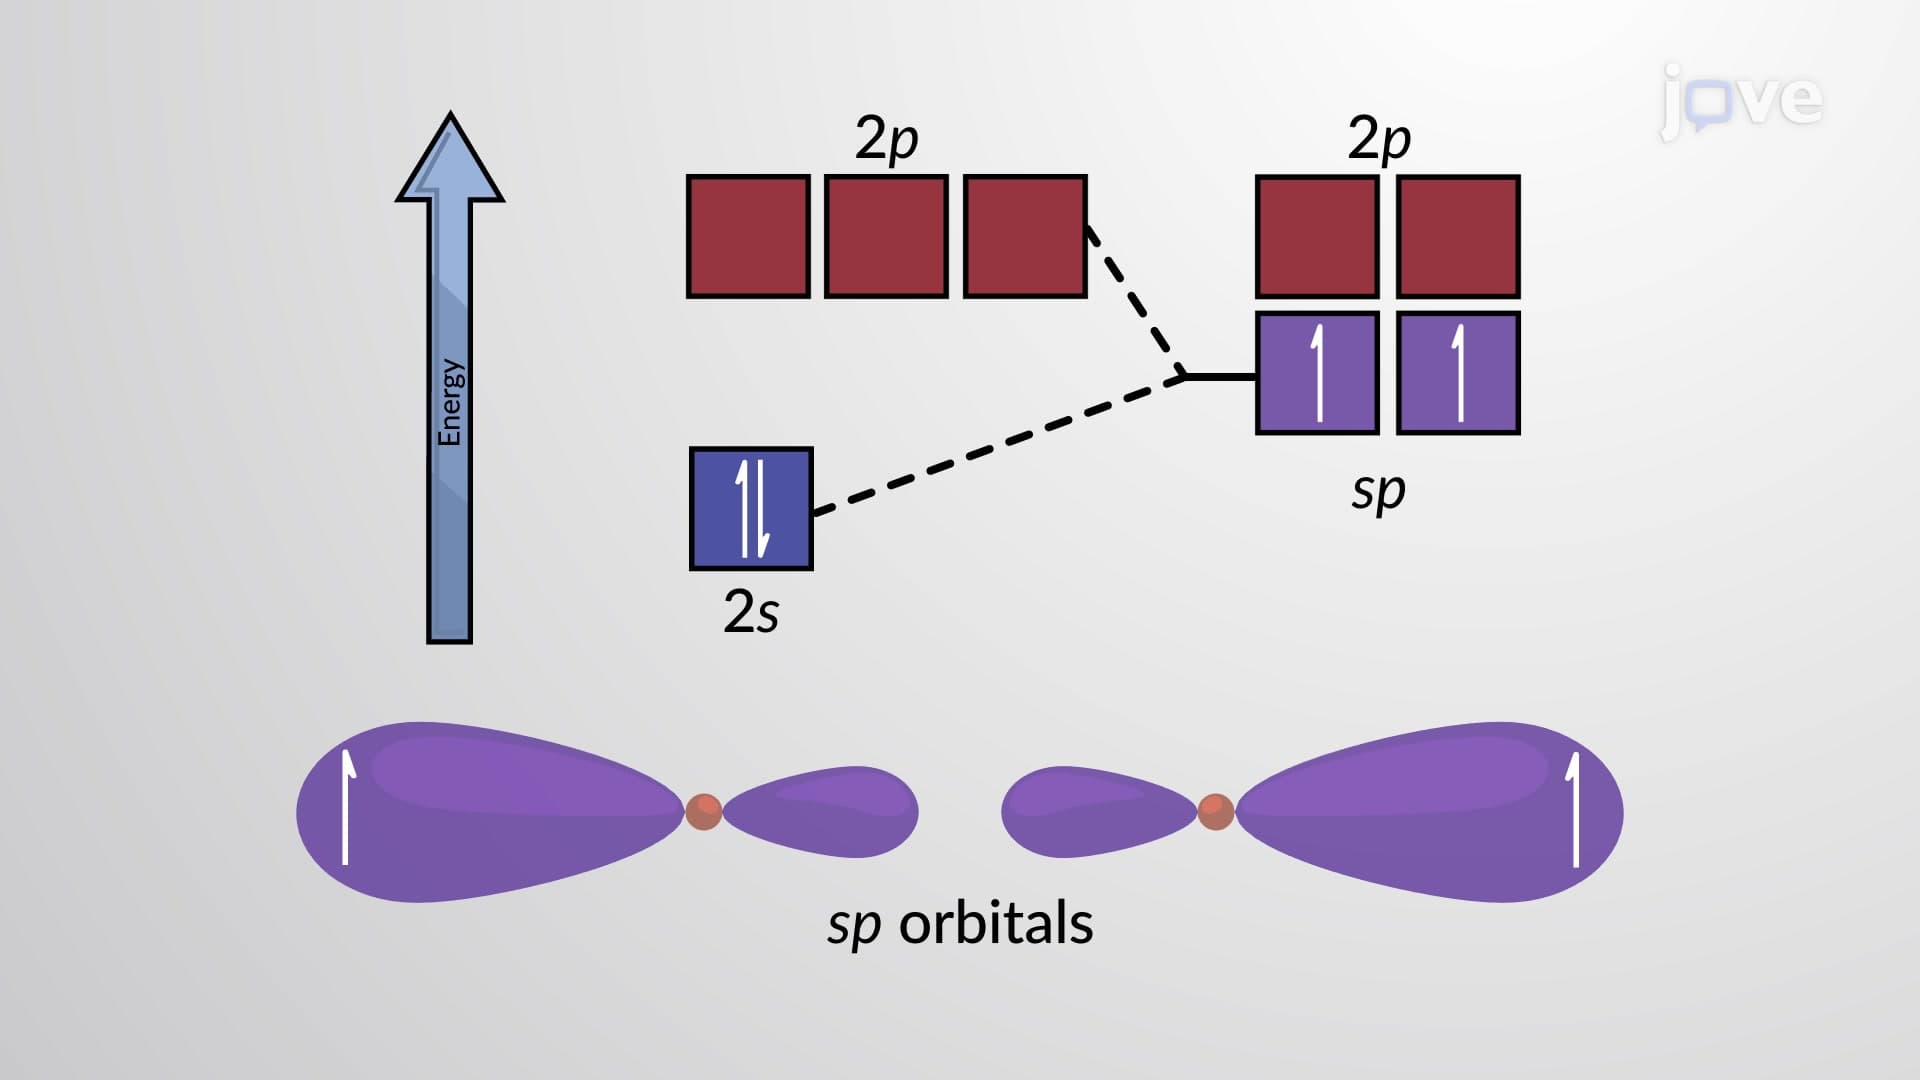
According to valence bond theory, a covalent bond results when: (1) an orbital on one atom overlaps an orbital on a second atom, and (2) the single electrons in each orbital combine to form an electron pair. The strength of a covalent bond depends on the extent of overlap of the orbitals involved. Maximum overlap is possible when the orbitals overlap on a direct line between the two nuclei.
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
19.3K
 02:36
Molecular Geometry and Dipole Moments
12.9K
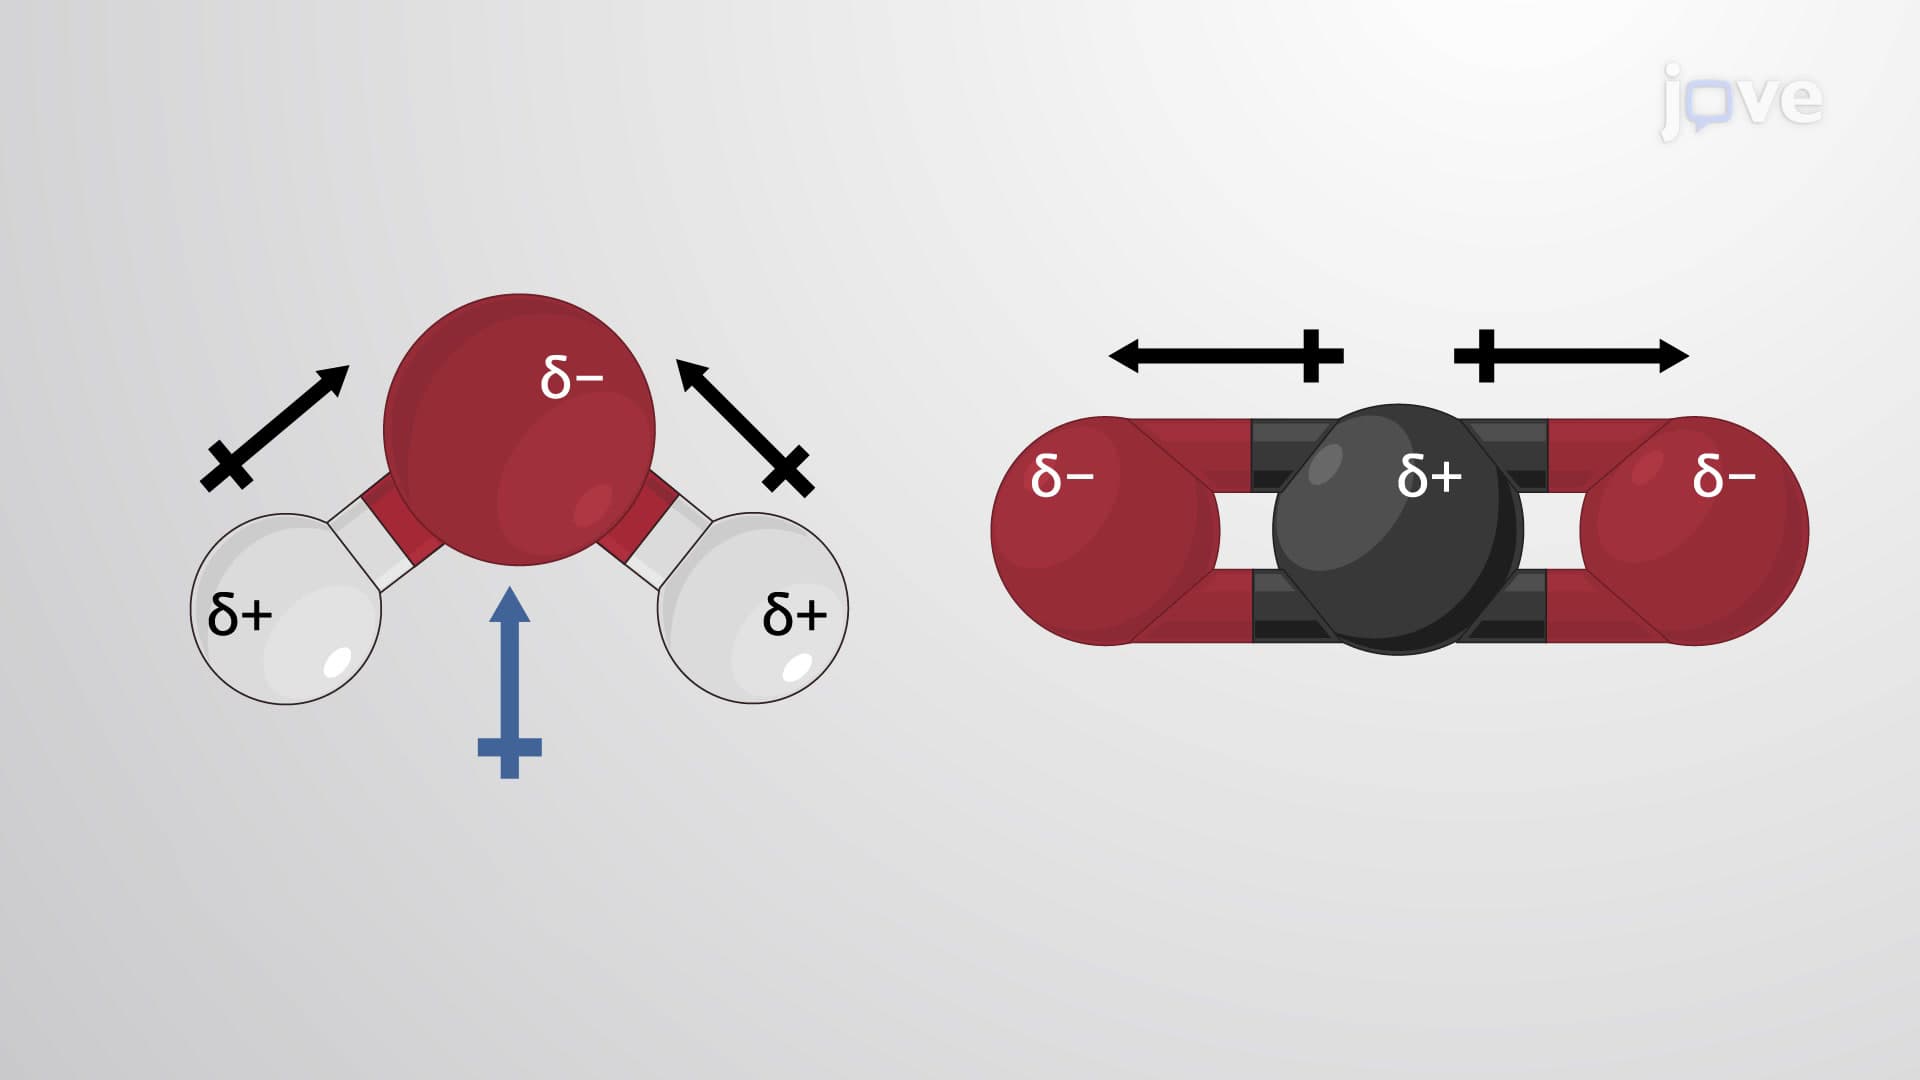
The VSEPR theory can be used to determine the electron pair geometries and molecular structures as follows:
12.9K
 02:34
Arrhenius Plots
39.2K
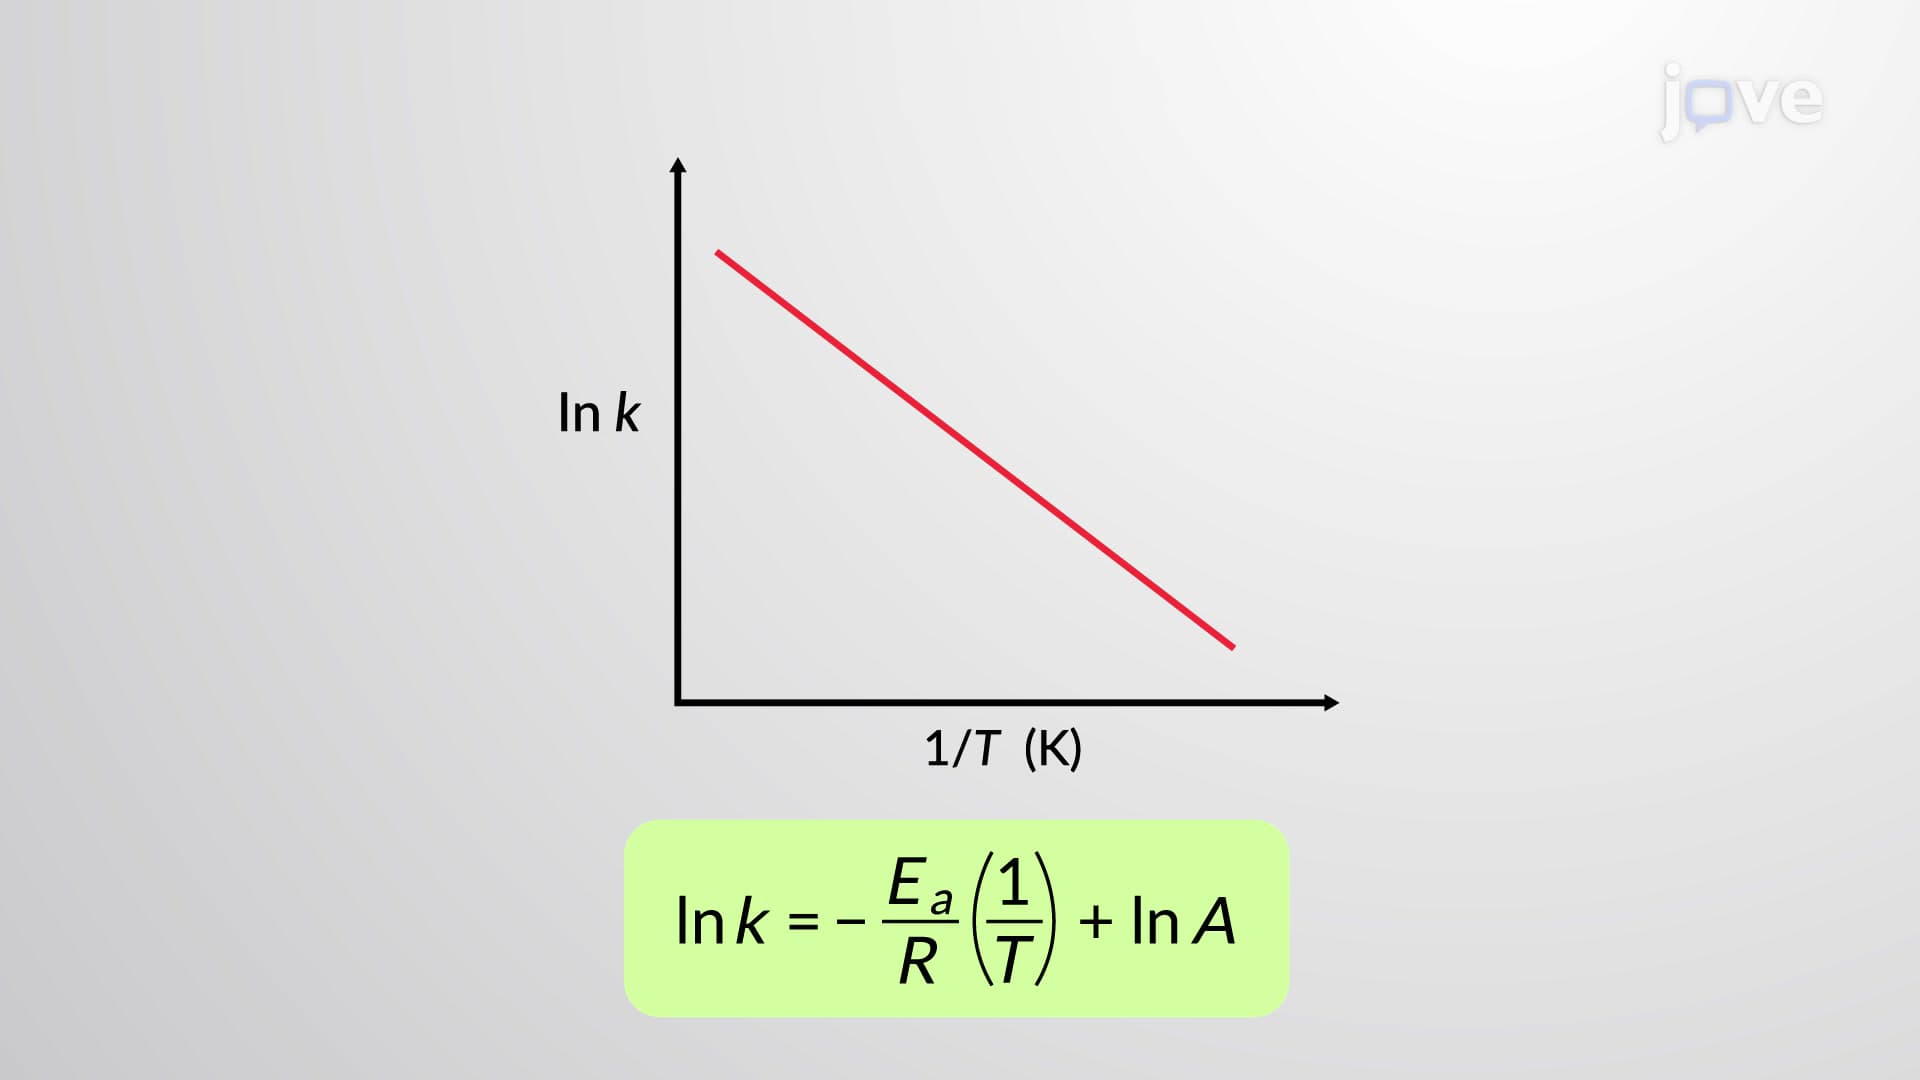
The Arrhenius equation relates the activation energy and the rate constant, k, for chemical reactions. In the Arrhenius equation, k = Ae−Ea/RT, R is the ideal gas constant, which has a value of 8.314 J/mol·K, T is the temperature on the kelvin scale, Ea is the activation energy in J/mole, e is the constant 2.7183, and A is a constant called the frequency factor, which is related to the frequency of collisions and the orientation of the reacting molecules.
The Arrhenius equation can be used...
The Arrhenius equation can be used...
39.2K