
在极化分子动力学力场中,甘氨酸和氨酸的内在形态动力学:与光谱数据的比较
Brian Andrews1, Reinhard Schweitzer-Stenner2, Brigita Urbanc1
1Department of Physics, Drexel University, Philadelphia, Pennsylvania 19104, United States.
The journal of physical chemistry. B
|June 15, 2024

概括
使用偏振力场的分子动力学模拟并没有改善对失序的预测. 与实验数据相比,当前的模型很难准确地捕捉与实验数据相比,内在无序的结构动态.
科学领域:
- 计算化学是一种计算化学.
- 生物物理学的生物物理.
- 分子建模分子建模
背景情况:
- 分子动力学 (MD) 模拟对于在原子水平上理解蛋白质和的动力学至关重要.
- 准确的结构预测严重依赖于MD力场的精度.
- 内在无序的蛋白质和因其灵活的性质而带来了独特的挑战.
研究的目的:
- 为了研究两极分化的分子动力学力场对内在无序的结构动力学的影响.
- 为了比较CHARMM Drude和AMOEBA极化力场与CHARMM36m等添加力场的性能.
- 评估这些力场在复制实验数据中的准确性,用于客氨基酸残留在阴性GxG中.
主要方法:
- 利用了两个可偏化的分子动力学力场:CHARMM Drude和AMOEBA.
- 在水溶液中模拟了客体甘氨酸和氨酸残留物在酸性GxG中的构造动力学.
- 分析并将得到的Ramachandran分布与CHARMM36m力场的实验数据和结果进行比较.
主要成果:
- 与CHARMM36m相比,CHARMM Drude和AMOEBA极化力场都没有显著改善Ramachandran对糖氨酸和氨酸残留物的分布预测.
- 对于研究的系统,AMOEBA力场与实验数据的一致性比CHARMM Drude更为接近.
- 尽管有进步,但极化力场并没有完全解决这些无序模型的形状分布的差异.
结论:
- 力量场中的极化性虽然在概念上很重要,但本质上并没有提高本研究中对内在无序的构造预测的准确性.
- 进一步开发分子动力学力场是必要的,以准确地建模无序系统的行为.
- 当前的极化力场可能不足以克服预测内在无序的构造组合的局限性.
相关概念视频
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
829
At room temperature, the chair conformer of cyclohexane undergoes rapid ring flipping between two equivalent chair conformers at a rate of approximately 105 times per second. These two chair conformers are in equilibrium. The rapid ring flipping results in the interconversion of the axial proton to an equatorial proton and an equatorial to the axial proton. Such interconversions are too rapid and cannot be detected on the NMR timescale. Hence, the NMR spectrometer cannot distinguish between the...
829
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
1.1K
The axial and equatorial protons in cyclohexane can be distinguished by performing a variable-temperature NMR experiment. In this process, except for one proton, the remaining eleven protons are replaced by deuterium. The deuterium substitution avoids the possible peak splitting caused by the spin-spin coupling between the adjacent protons. The remaining proton flips between the axial and equatorial positions.
1.1K
 01:19
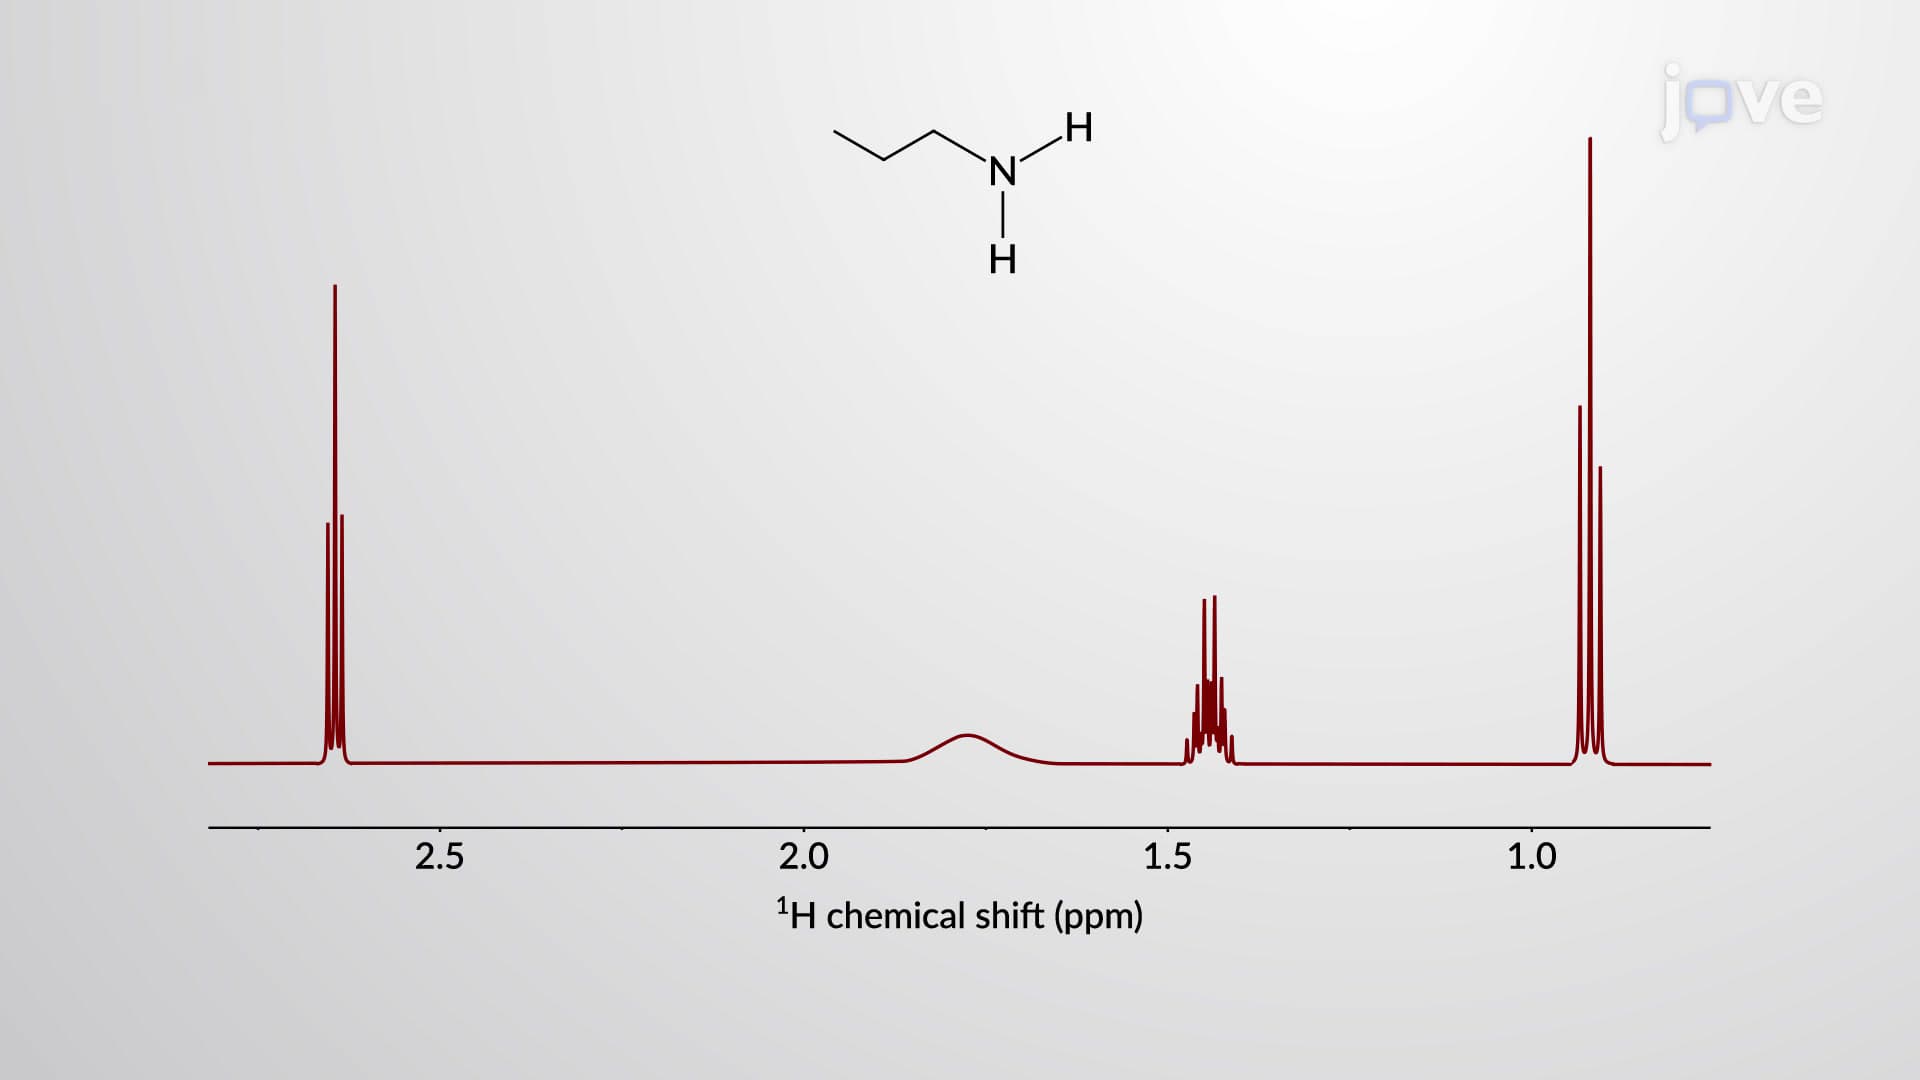
NMR Spectroscopy Of Amines
8.7K
In proton NMR spectroscopy, primary amines and secondary amines showcase their N–H protons as a broad signal in the chemical shift range between δ 0.5 and 5 ppm. The exact position in this range depends on several factors, including sample concentration, hydrogen bonding, and the type of solvent used. Since amine protons undergo fast proton exchange in solution, the protons are labile and therefore do not participate in any splitting with adjacent protons. Thus, the observed peak is...
8.7K
 02:35
Noncovalent Attractions in Biomolecules
50.2K
Noncovalent attractions are associations within and between molecules that influence the shape and structural stability of complexes. These interactions differ from covalent bonding in that they do not involve sharing of electrons.
Four types of noncovalent interactions are hydrogen bonds, van der Waals forces, ionic bonds, and hydrophobic interactions.
Hydrogen bonding results from the electrostatic attraction of a hydrogen atom covalently bonded to a strong-electronegative atom like oxygen,...
Four types of noncovalent interactions are hydrogen bonds, van der Waals forces, ionic bonds, and hydrophobic interactions.
Hydrogen bonding results from the electrostatic attraction of a hydrogen atom covalently bonded to a strong-electronegative atom like oxygen,...
50.2K