相关实验视频
Updated: Jun 23, 2025
10:36

Rare Event Detection Using Error-corrected DNA and RNA Sequencing
Published on: August 3, 2018
12.1K
通过大规模并行测序,改善了不相关或相关贡献者的DNA混合物中的个体识别
Zhiyong Liu1, Enlin Wu1, Ran Li2
1Faculty of Forensic Medicine, Zhongshan School of Medicine, Sun Yat-sen University, Guangzhou 510080, China; Guangdong Province Translational Forensic Medicine Engineering Technology Research Center, Sun Yat-sen University, Guangzhou 510080, China.
Forensic science international. Genetics
|June 18, 2024

概括
研究与相关贡献者的DNA混合物揭示了亲属关系不会阻碍主要贡献者识别,但基于序列的基因型定型可以提高准确性. 这项研究验证了大规模并行测序 (MPS) 面板用于法医DNA混合分析.
科学领域:
- 法医遗传学 法医遗传学
- 分子生物学分子生物学
- 生物统计学 生物统计学
背景情况:
- 法医DNA混合分析通常在计算概率比率 (LR) 时假定不相关的贡献者.
- 在DNA混合物中的相关贡献者在确定贡献者数量 (NOC) 和构建概率基因型化模型方面存在挑战.
- 了解亲属关系对个人识别的影响对于复杂的法医案件至关重要.
研究的目的:
- 评估潜在亲属关系对DNA混合物中感兴趣的人 (POI) 个体识别的影响.
- 评估大规模并行测序 (MPS) 使用MGIEasy签名识别库准备包在分析相关和非相关DNA混合物的性能.
- 为了比较基于长度和基于序列的短并列重复 (STR) 基因型鉴定在混合物解释中的有效性.
主要方法:
- 模拟两个和三个人的DNA混合物,不同比例的无关和相关的贡献者 (父母-孩子,兄弟姐妹,叔叔-子).
- 在MGI平台上使用MGIEasy签名识别库准备套件进行了大规模并行测序 (MPS).
- 在模拟中进行并分析了多个遗传标记,亲属假设和STR基因定型方法对LR值和NOC推断的影响.
主要成果:
- 多个遗传标记在DNA混合物中改善了亲属关系和NOC推断.
- 对感兴趣的人 (POI) 的概率比率 (LR) 值高度依赖混合比率.
- 亲属关系考虑没有显著影响主要POI识别,尽管正确的亲属关系产生了保守的LR,而不正确的亲属关系并不能保证识别失败;小贡献者识别显示不确定性.
- 基于序列的STR基因型定型增强了POI识别能力,与基于长度的基因型定型相比,改进了混合比率推断准确度.
结论:
- MGIEasy签名识别库准备套件是用于法医DNA混合物解释的强大的MPS面板.
- 该套件证明了与无关和相关贡献者的混合物分析的可行性.
- 基于序列的STR基因型定型为法医DNA混合分析提供了准确性和识别能力的优势.
相关概念视频
 03:00
Next-generation Sequencing
88.6K
The first human genome sequencing project cost $2.7 billion and was declared complete in 2003, after 15 years of international cooperation and collaboration between several research teams and funding agencies. Today, with the advent of next-generation sequencing technologies, the cost and time of sequencing a human genome have dropped over 100 fold.
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
88.6K
 01:57
Sanger Sequencing
754.0K
DNA sequencing is a fundamental technique that is routinely used in the biological sciences. This method can be applied to a range of questions at different scales - from the sequencing of a cloned DNA fragment or the study of a mutation in a gene up to whole-genome sequencing. However, despite the widespread use of sequencing today, it was not until 1977 that Fredrick Sanger and his collaborators developed the chain-termination method to decode DNA sequences. It relies on the separation of a...
754.0K
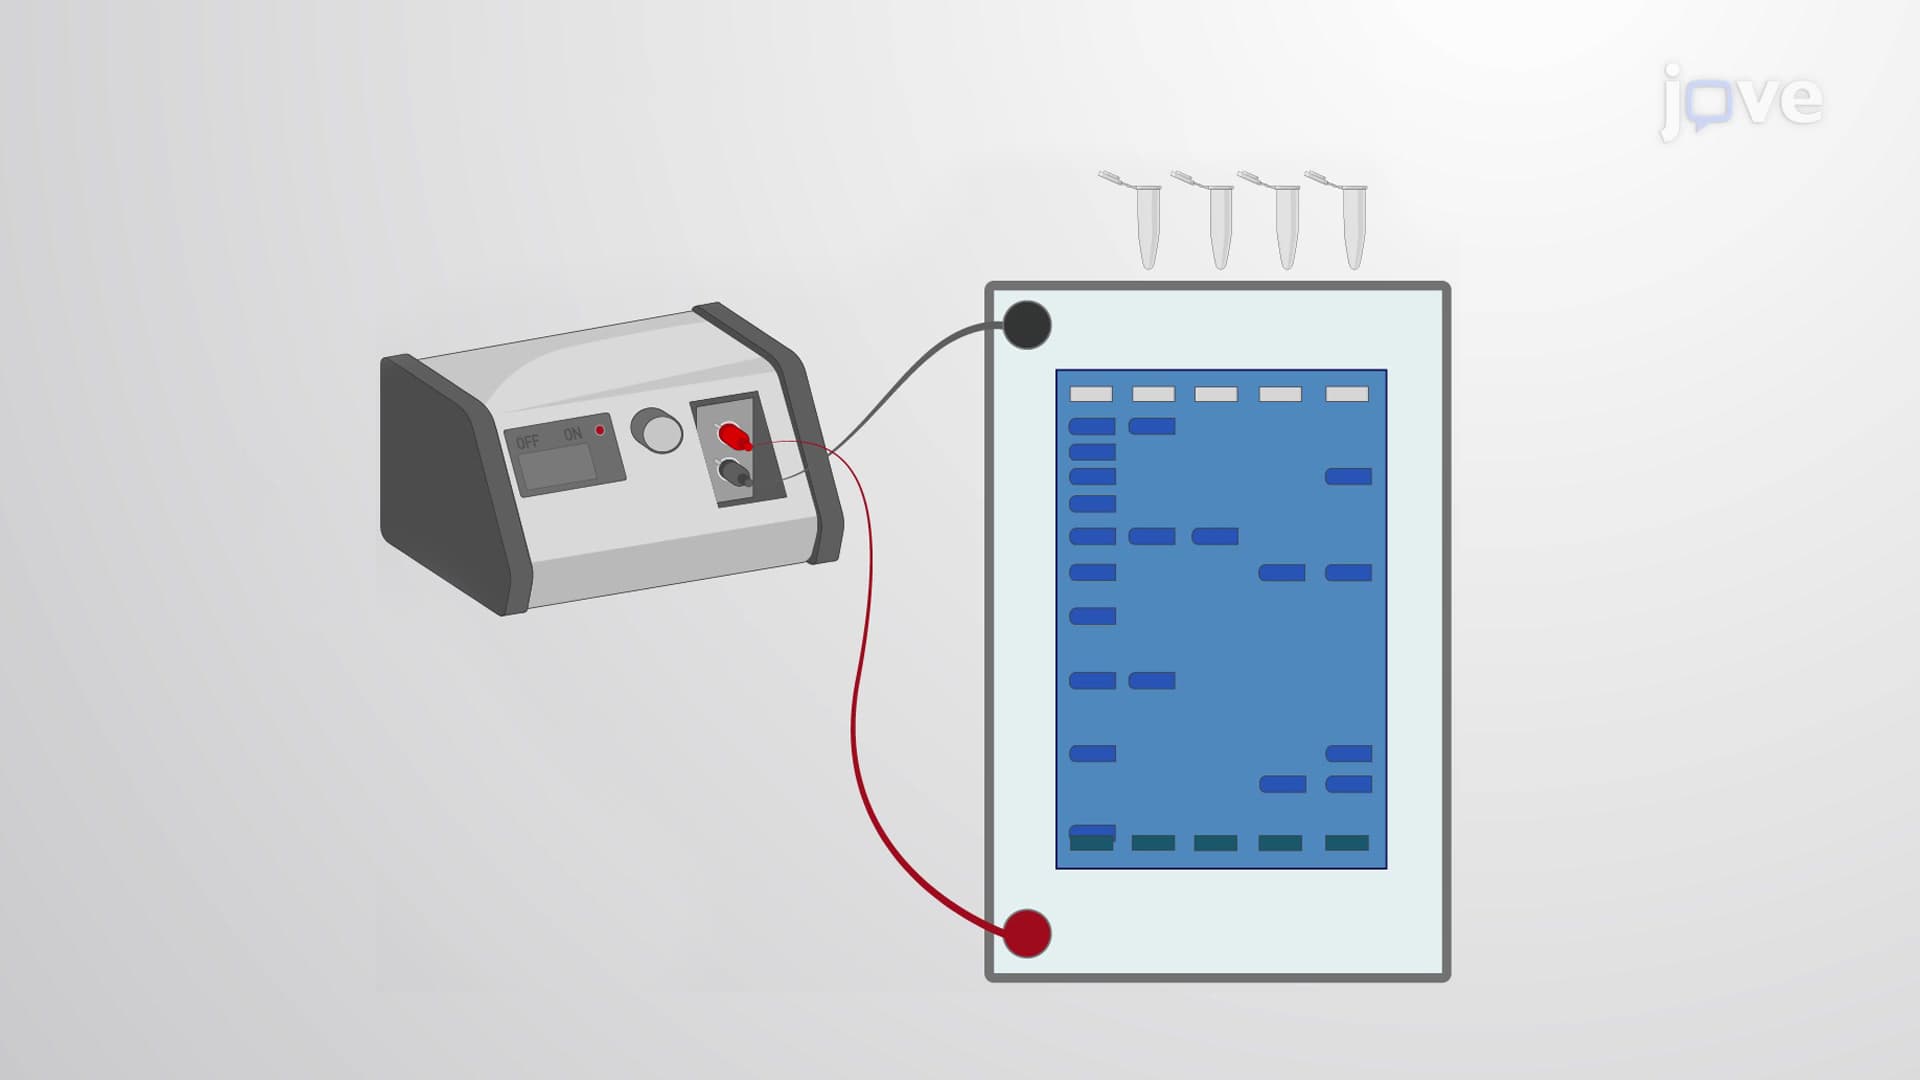 01:05
Maxam-Gilbert Sequencing
11.2K
In the same year as the discovery of the Sanger sequencing method, another group of scientists, Allan Maxam and Walter Gilbert, demonstrated their chemical-cleavage method for DNA sequencing. The Maxam-Gilbert method relies on using different chemicals that can cleave the DNA sequence at specific sites, the separation of resulting DNA fragments of variable size using electrophoresis, and deciphering the DNA sequence from the resulting gel bands.
Challenges of the Maxam-Gilbert Method
The...
Challenges of the Maxam-Gilbert Method
The...
11.2K
 03:21
RNA-seq
9.9K
RNA sequencing, or RNA-Seq, is a high-throughput sequencing technology used to study the transcriptome of a cell. Transcriptomics helps to interpret the functional elements of a genome and identify the molecular constituents of an organism. Additionally, it also helps in understanding the development of an organism and the occurrence of diseases.
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
9.9K