
SATIN:全基因组和编码区域的微型和迷你卫星挖掘工具,分析编码区域的完美重复多态
Carlos Willian Dias Dantas1, Sebastião Rodrigues da Costa Neto2, Sandy Ingrid Aguiar Alves2
1Institute of Biological Sciences, Federal University of Minas Gerais, Belo Horizonte, Brazil. cwillian@ufmg.br.
BMC bioinformatics
|June 18, 2024

概括
SATIN是微观卫星采矿的新软件,可以有效地识别DNA中的并列重复,包括编码区域中的重复. 该工具有助于发现各种应用的潜在遗传标记.
科学领域:
- 基因组学就是基因组学.
- 生物信息学是一种生物信息学.
- 分子生物学分子生物学
背景情况:
- 串联重复 (TRs),包括微卫星,是所有生物体中发现的关键DNA序列.
- 小卫星和微卫星特别有趣,因为它们的不稳定性和识别多态的潜力.
- 当前的SSR挖矿工具往往忽略了编码区域,限制了它们的范围.
研究的目的:
- 开发一种用于高效微卫星 (SSR) 挖矿的新型软件工具.
- 解决现有工具的局限性,特别是在编码区域内识别SSR方面.
- 为分析SSR多态性和编码区域的差异丰度提供一个工具.
主要方法:
- 开发SATIN (微型和迷你卫星识别工具),一个使用新的滑动窗口算法的软件.
- 在C和Python中实现,以获得高效的性能.
- 使用大肠杆菌K12和O157数据集进行验证,将结果与已知的编码区域SSR进行比较.
主要成果:
- SATIN展示了高速,在完美和复合SSR挖矿方面排名第二.
- 该工具成功地在编码区域和图案大小大于6的区域中识别了SSR.
- 在237个大肠杆菌基因的验证组中,SATIN确定了所有5个已知的编码区域SSR.
结论:
- SATIN是一种新且高效的微卫星搜索软件,采用创新的滑窗技术.
- 它支持FASTA和GenBank格式,在编码区域中识别SSR.
- 预计SATIN将促进发现潜在的SSRs作为遗传标记物的使用.
相关概念视频
 01:57
Sanger Sequencing
754.0K
DNA sequencing is a fundamental technique that is routinely used in the biological sciences. This method can be applied to a range of questions at different scales - from the sequencing of a cloned DNA fragment or the study of a mutation in a gene up to whole-genome sequencing. However, despite the widespread use of sequencing today, it was not until 1977 that Fredrick Sanger and his collaborators developed the chain-termination method to decode DNA sequences. It relies on the separation of a...
754.0K
 01:05
Single Nucleotide Polymorphisms-SNPs
15.0K
A single nucleotide polymorphism or SNP is a single nucleotide variation at a specific genomic position in a large population. It is the most prevalent type of sequence variation found in the human genome. Point mutations that occur in more than 1% of the population qualify as SNPs. These are present once every 1000 nucleotides on an average in the human genome. Replacement of a purine with another purine (A/G) or a pyrimidine with another pyrimidine (C/T) is known as a transition. In contrast,...
15.0K
 02:34
DNA Microarrays
17.3K
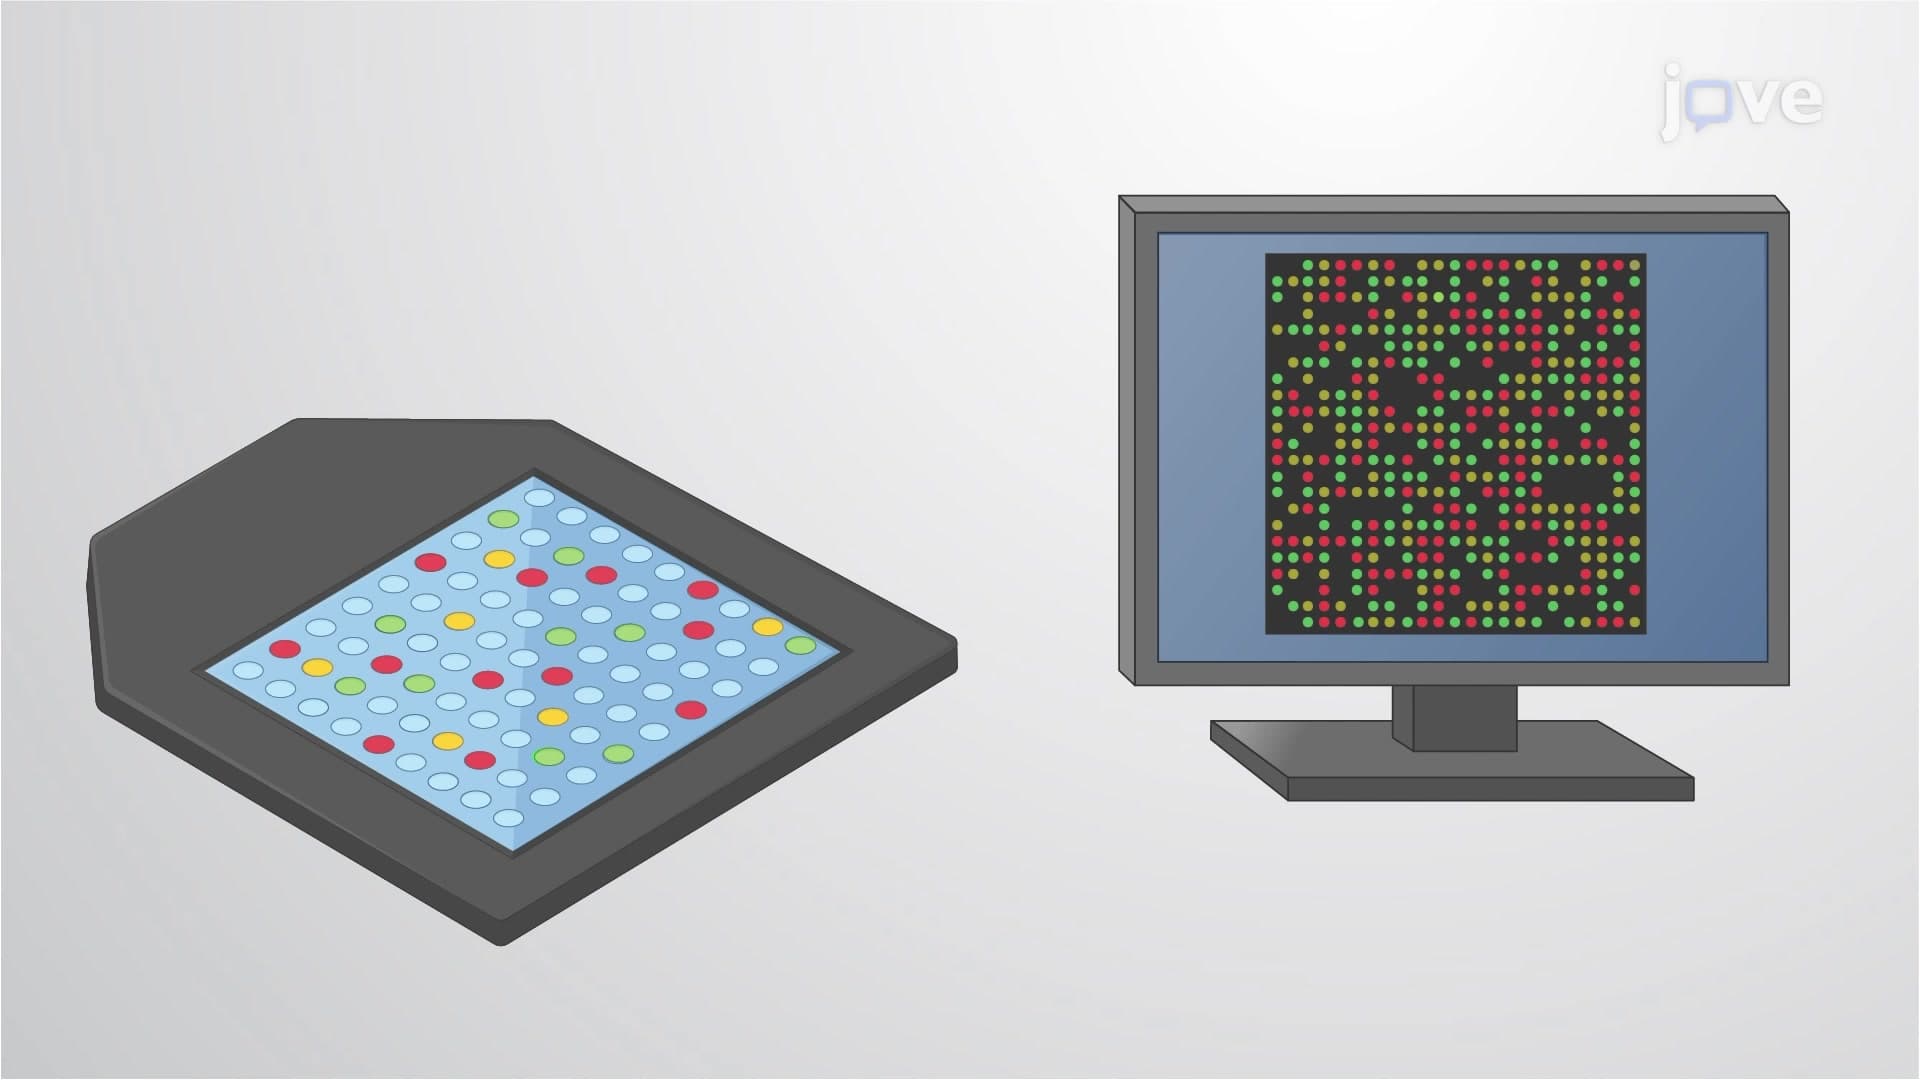
Microarrays are high-throughput and relatively inexpensive assays that can be automated to analyze large quantities of data at a time. They are used in genome-wide studies to compare gene or protein expression under two varied conditions, such as healthy and diseased states. Microarrays consist of glass or silica slides on which probe molecules are covalently attached through surface functionalization. Most commonly, the slides are prepared through the chemisorption of silanes to silica...
17.3K
 03:21
RNA-seq
9.9K
RNA sequencing, or RNA-Seq, is a high-throughput sequencing technology used to study the transcriptome of a cell. Transcriptomics helps to interpret the functional elements of a genome and identify the molecular constituents of an organism. Additionally, it also helps in understanding the development of an organism and the occurrence of diseases.
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
9.9K