
使用DFT方法预测铁氨酸的旋转状态,包括晶体包装效应和热力学校正
1Faculty of Chemistry, Jagiellonian University, Kraków, Poland. mradon@chemia.uj.edu.pl.
Physical chemistry chemical physics : PCCP
|June 20, 2024

概括
使用密度函数理论 (DFT) 准确预测铁酸中的自旋状态是具有挑战性的. 双混合功能表现出最高的准确性,在这些关键的生物无机模型中表现优于混合功能.
科学领域:
- 量子生物无机化学 量子生物无机化学
- 计算化学是一种计算化学.
- 材料科学是一种材料科学.
背景情况:
- 精确计算处理过渡金属复合体中的自旋状态,如铁,对于量子生物无机化学至关重要.
- 大致密度函数理论 (DFT) 方法在精确计算这些自旋状态时面临挑战.
研究的目的:
- 评估DFT方法的准确性,以预测铁二烯的旋转状态分裂.
- 量化氨酸替代剂和晶体包装效应 (CPE) 对旋转状态确定的影响.
- 建立一个基准数据集,用于评估该领域的DFT方法.
主要方法:
- 采用分子和周期 DFT 计算,对六种经过实验特征的 Fe (III) 或 Fe (II) 氨酸复合物进行了测量.
- 将晶体包装效应 (CPE) 分为直接和结构部件.
- 使用实验性地面状态数据对电子能量差异推导的定量约束.
主要成果:
- 双混合函数 (例如,B2PLYP-D3,DSD-PBEB95-D3) 在旋转状态分裂中显示出最高的精度.
- 混合函数,特别是那些精确交换减少的函数,倾向于过度稳定中间旋转状态,导致Fe (III) 甲的基态预测不正确.
- 该研究建立了晶体铁二烯 (SSCIP6) 基准数据集的Spin状态.
结论:
- 开发的方法提供了一种可靠的方法,用于对过渡金属复合物的DFT函数进行基准测试.
- 这些发现为解释金属氨酸的实验数据提供了洞察力,这是理解血红蛋白的关键模型.
- 在类似的系统中,建议使用双混合函数来准确计算旋转状态.
更多相关视频
 05:51
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
6.2K
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
5.7K
相关概念视频
 02:58
Crystal Field Theory - Octahedral Complexes
26.3K
Crystal Field Theory
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
26.3K
 02:42
Valence Bond Theory
8.5K
Coordination compounds and complexes exhibit different colors, geometries, and magnetic behavior, depending on the metal atom/ion and ligands from which they are composed. In an attempt to explain the bonding and structure of coordination complexes, Linus Pauling proposed the valence bond theory, or VBT, using the concepts of hybridization and the overlapping of the atomic orbitals. According to VBT, the central metal atom or ion (Lewis acid) hybridizes to provide empty orbitals of suitable...
8.5K
 03:02
Colors and Magnetism
11.6K
Color in Coordination Complexes
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...
11.6K
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
42.0K
Tetrahedral Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
42.0K
 02:27
Predicting Molecular Geometry
34.2K
VSEPR Theory for Determination of Electron Pair Geometries
34.2K
 01:22
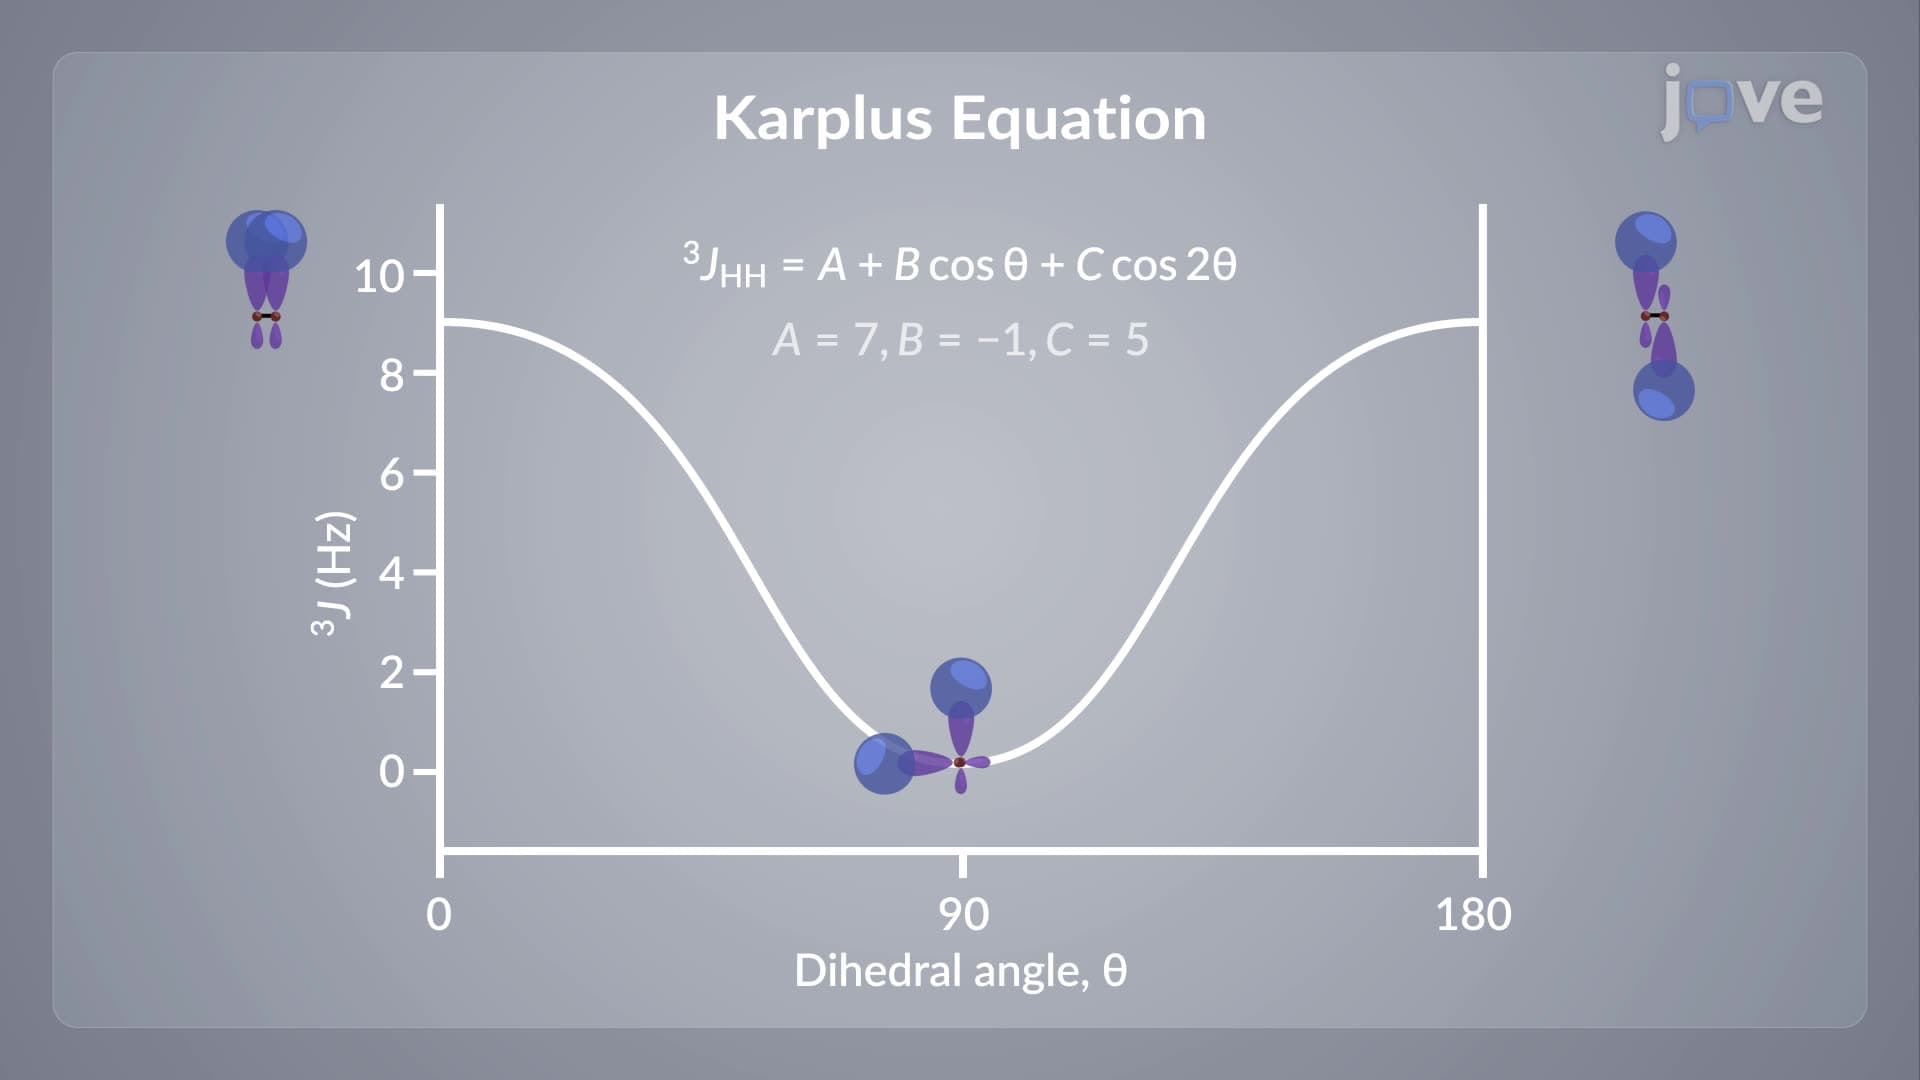
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
1.1K
Vicinal or three-bond coupling is commonly observed between protons attached to adjacent carbons. Here, nuclear spin information is primarily transferred via electron spin interactions between adjacent C‑H bond orbitals. This generally favors the antiparallel arrangement of spins, so 3J values are usually positive.
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
1.1K