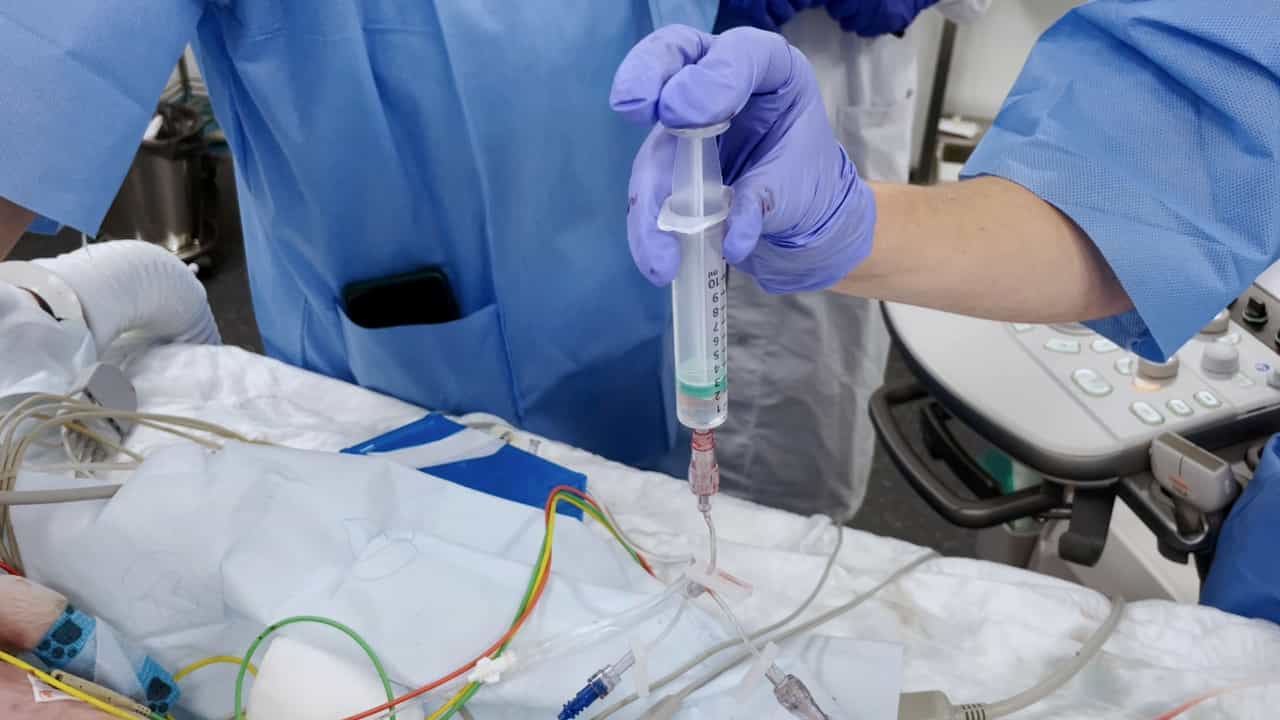
在受体外膜氧化支持的儿科患者中,人体血衍生抗血的药理动力学
Dawoon Jung1, David Procaccini2, Jennifer Roem3
1Center for Translational Medicine, University of Maryland School of Pharmacy, Baltimore, MD, USA.
Journal of clinical pharmacology
|July 2, 2024

概括
抗血素 (AT) 替代对于由于肝素耐药性导致体外膜氧化 (ECMO) 的儿科患者至关重要. 这项研究特征AT的药理动力学,告知剂量维持治疗水平.
科学领域:
- 儿科重症监护医学 儿科重症监护医学
- 药理动力学 药理动力学
- 血液学 血液学 血液学
背景情况:
- 在重症儿童体外膜氧化 (ECMO) 具有高血栓栓塞事件的风险.
- 未分离的氨酸是标准的抗凝药,但获得的抗血素 (AT) 缺乏症可能会导致氨酸耐药性.
- 对AT活性监测和非标签的AT替代是常见的,但缺乏最佳剂量指南.
研究的目的:
- 在ECMO治疗的儿科患者中描述人体血衍生AT的药理动力学 (PK).
- 为接受ECMO的不同儿科年龄组开发基于证据的AT替代疗法.
- 在这个人群中建立最佳的治疗药物监测策略.
主要方法:
- 在单个学术中心对儿科ECMO患者 (0至<18岁) 的回顾性队列研究.
- 使用两部分的营业额模型来描述AT的药理动力学.
- 分析了PK参数,包括清除率,分布量和基底AT输入.
主要成果:
- 一个双隔间模型准确地描述了儿科ECMO患者的AT PK.
- 在ECMO的支持下,生物可利用的AT减少了50%,清除和分销量翻了一番.
- 在非ECMO条件下建立了PK参数估计.
结论:
- 建议针对新生儿和年龄较大的儿童使用ECMO的特定年龄的AT替代疗法,以保持目标AT活性水平.
- 强调治疗药物监测对于优化AT替代疗法的重要性.
- 提供了改善儿科ECMO抗凝管理的基础.
相关概念视频
 01:15
Drug Concentration Versus Time Correlation
721
The plasma drug concentration-time curve is a crucial tool in pharmacokinetics, representing the drug's concentration in plasma at different time intervals post-administration. This curve illustrates the drug's journey from absorption into the systemic circulation, distribution to body tissues, and eventual elimination through excretion or biotransformation.
Two pivotal parameters are the minimum effective concentration (MEC) and the minimum toxic concentration (MTC). The MEC is the...
Two pivotal parameters are the minimum effective concentration (MEC) and the minimum toxic concentration (MTC). The MEC is the...
721
 01:30
Anticoagulant Drugs: Low-Molecular-Weight Heparins
675
Hemostasis is a crucial process that prevents excessive blood loss from damaged blood vessels. It involves various mechanisms such as vasoconstriction, platelet adhesion and activation, and fibrin formation. The importance of each mechanism depends on the type of vessel injury. In contrast, thrombosis is the abnormal formation of a blood clot within the blood vessels, leading to potential complications if the clot obstructs blood flow. Thrombosis can be caused by increased coagulability of the...
675
 01:29
Drug Distribution: Plasma Protein Binding
5.5K
Drugs predominantly attach to plasma proteins, with only a small percentage remaining unbound. The unbound portion can be calculated as one minus the bound fraction. Acidic drugs form large, inactive complexes by reversibly binding to plasma albumin, which prevents them from diffusing across biological barriers. These drug-protein complexes act as reservoirs for the drugs. As the concentration of unbound drugs decreases, these complexes quickly dissociate to release the free drug, maintaining...
5.5K
 01:12
Two-Compartment Open Model: Extravascular Administration
176
The two-compartment model for extravascular administration represents a drug's absorption and distribution process. It features a central compartment, where the drug is first absorbed, and a peripheral compartment, which illustrates the drug's distribution throughout the body. The rate of change in drug concentration in the central compartment is calculated by three exponents: absorption, distribution, and elimination.
The absorption exponent (ka) indicates the speed at which the drug...
The absorption exponent (ka) indicates the speed at which the drug...
176
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
38
Drug transporters are critical in drug absorption, distribution, and excretion processes. They should be included in physiological-based pharmacokinetic (PBPK) models, which help predict human drug disposition. However, predicting this is challenging during drug development, especially when liver transport is involved. However, with a realistic representation of body transport processes, an accurate model may be possible.
A recent model describes pravastatin's hepatobiliary excretion,...
A recent model describes pravastatin's hepatobiliary excretion,...
38
 01:15
Factors Affecting Drug Distribution: Organ Perfusion Rate
123
Drug distribution within the body is a complex process influenced by several factors, including perfusion rate, the rate at which the bloodstream transports drugs to tissue. This limitation becomes particularly significant when dealing with highly lipophilic drugs. In such cases, the rate at which the drug can move across membranes is crucial, and if the membrane is highly permeable to the drug, distribution becomes rate-limited by perfusion.
Perfusion rate-limited distribution relies on the...
Perfusion rate-limited distribution relies on the...
123